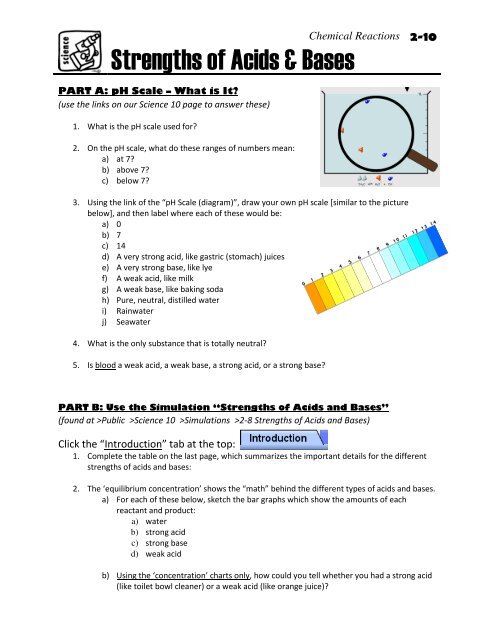
Working With Acid-Base Strengths in Practice
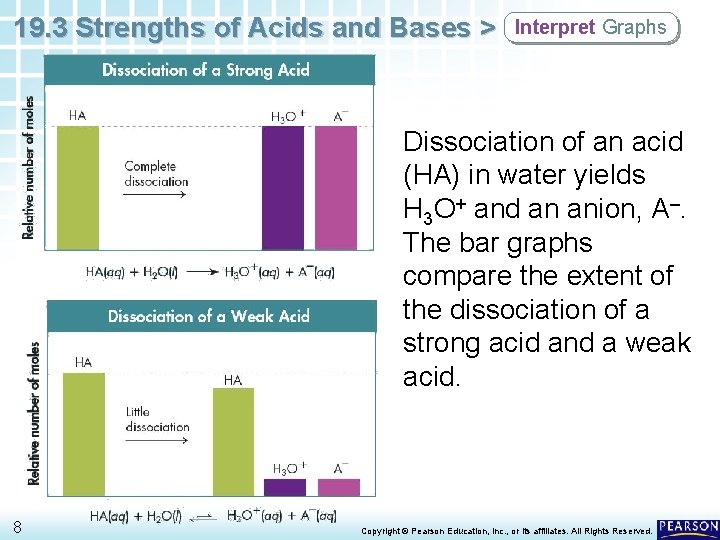
The thing most people miss when they first tackle 18 2 Review And Reinforcement Determining The Strengths Of Acids And Bases is that pKa values aren't just numbers you memorize — they're predictions. A strong acid donates its proton because the resulting conjugate base is stable. That stability comes from electronegativity, size, resonance, and inductive effects, usually acting together. When you can read those factors in sequence, you don't need to look up every single value. I spent a lot of time going through practice problems where students would correctly identify that oxygen is more electronegative than sulfur, then incorrectly conclude that an O–H bond is always more acidic than an S–H bond. It isn't. Size dominates when you move down a column. Hydroiodic acid is one of the strongest simple acids you'll encounter in an introductory course precisely because iodide is huge and spreads the negative charge over a large volume. The periodic trend down a group overrides the trend across a row every time those two compete. Here's how I approach any comparison now without second-guessing myself. First, locate the atom holding the acidic proton. Second, compare atoms across the same row using electronegativity and resonance. Third, if the atoms are in different rows, switch to size and polarizability. Fourth, check for any nearby electron-withdrawing groups that could exert an inductive effect. This takes about thirty seconds per problem once it's automatic.
I ran into a specific case last semester that still comes up in office hours. A student was comparing the acidity of 2,2,2-trichloroethanol versus plain ethanol and kept getting tripped up because the textbook answer involved a Hammett-style reasoning that felt disconnected from what we'd practiced. The workaround was straightforward — I had them draw the conjugate bases and count the sigma bonds between the chlorine atoms and the oxygen bearing the negative charge. Three chlorines, three bonds away. The inductive pull is real but decays rapidly with distance. Once they visualized the sigma framework, the ~10^6 difference in Ka stopped feeling like a magic number and started feeling like geometry.
Conjugate Base Stability Is The Actual Shortcut
Everything in acid-base strength circles back to the conjugate base. If the anion is stable, the acid is strong. That sounds tautological, but the practical payoff is that you can rank acids by drawing their deprotonated forms instead of trying to remember which bond is weaker. A carboxylate with two equivalent resonance structures beats an alkoxide with none. A phenoxide with delocalization into the ring beats an alkoxide. An enolate beats both when you count the carbonyl participation. One counter-intuitive detail that trips people up regularly involves nitromethane. Its pKa is around 10, which puts it in the same ballpark as phenol despite the nitrogen being less electronegative than oxygen. The stability here comes from resonance between two equivalent oxygen atoms on the nitro group after deprotonation. Nitrogen handles the formal charge distribution elegantly, and beginners who focus only on atom electronegativity will misrank it every time.
Get the Full Details

What The Numbers Actually Tell You
A pKa difference of one unit means roughly a tenfold difference in acidity. A difference of three units means a thousandfold. When you see something listed as pKa 4.76 for acetic acid and pKa 1.4 for trichloroacetic acid, that's not a small gap. The chlorines withdraw electron density through the sigma bonds, stabilizing the conjugate base enough to shift equilibrium dramatically. In the lab, this is why trichloroacetic acid can protonate species that acetic acid leaves largely untouched. The common pitfall is assuming pKa values are fixed constants independent of environment. They shift with solvent. Water stabilizes ions through solvation, but in DMSO the same compounds can have pKa values that differ by several units. If you're working with organolithium bases or non-aqueous titrations, your aqueous pKa table becomes a rough guide at best. Gas-phase acidities follow different trends entirely, where polarizability dominates even more heavily.
Applying This To Titration And Equilibrium Problems
When you move from ranking acids to calculating pH, the framework stays the same but the arithmetic changes. For a weak acid, you set up the ICE table, write the Ka expression, and decide whether the x is small approximation holds. The rule of thumb is that if the initial concentration divided by Ka is greater than about four hundred, the approximation introduces less than five percent error. Below that, you solve the quadratic. I found that students who skip the reasoning behind the approximation end up applying it everywhere and getting wrong answers on dilute solutions. A 0.01 M solution of a acid with Ka = 1e-4 fails the test. The quadratic gives you a pH that's noticeably different. Working through the derivation once makes the threshold feel less arbitrary. For polyprotic acids, the key is recognizing when the steps separate enough to treat them independently. Phosphoric acid has pKa values of roughly 2.1, 7.2, and 12.3. The gaps are large enough that at the first equivalence point you can use the standard amphiprotic salt formula, pH (pKa1 + pKa2)/2, without worrying much about the third dissociation. Sulfuric acid is different because the first proton is strong and the second has pKa2 around 1.9. The overlap means you can't isolate the steps cleanly, and you need a simultaneous equilibrium treatment or a numerical approach.
Strong Bases, Weak Bases, And What Confuses Everyone
Strong bases like NaOH and KOH dissociate completely in water. Their conjugate acids are water and the corresponding metal hydroxides, which don't really exist as discrete molecular species in solution. That's why we say the conjugate acid of hydroxide is water — and why hydroxide can't act as an acid in aqueous solution. It's already fully deprotonated. Weak bases follow the same logic in reverse. Ammonia accepts a proton to become ammonium, and Kb tells you how far that goes. The relationship Ka × Kb = Kw holds for any conjugate pair, which means knowing one value gives you the other instantly. This is useful when a problem gives you the pKa of an acid and asks for the pH of its conjugate base solution, or vice versa. Multiplying by 1e-14 and taking the negative log is faster than looking up a separate table entry. Amines are where this gets messy in practice. Aliphatic amines like methylamine have pKb values around 3.4, making them moderately strong weak bases. Aryl amines like aniline drop to pKb values near 9.4 because the lone pair participates in resonance with the aromatic ring. Students sometimes rank aniline as more basic than methylamine because nitrogen is the same atom, but the resonance delocalization removes electron density from the nitrogen lone pair and makes it significantly less available for protonation.
When The Model Breaks Down
The electrostatic and resonance framework works remarkably well for most compounds you'll encounter in a general chemistry or organic chemistry sequence. It breaks down when you hit hypervalent species, transition metal complexes, or systems where solvation effects dominate over intrinsic electronic structure. Aluminum fluoride in water doesn't behave like a simple Brønsted acid at all. You also lose predictive power with extremely dilute solutions where water autoionization contributes a non-negligible fraction of the hydrogen ion concentration, or with extremely concentrated acids where activity coefficients diverge significantly from one. If your problem involves a compound outside the typical organic and main-group pool, the pKa table approach becomes unreliable. In those cases, computational chemistry or direct experimental measurement is the only honest path. I've seen graduate students waste weeks trying to force a resonance argument onto a coordination compound where crystal field stabilization and ligand field effects control the chemistry instead. It's easier to admit the model doesn't apply than to keep stretching it.
Practical Takeaways For The Next Problem Set
Start every ranking question by drawing the conjugate base. Count resonance structures. Check for inductive donors and withdrawers. Verify whether you're comparing atoms in the same row or the same column. Apply the size-over-electronegativity rule when rows and columns conflict. Use the pKa difference to gauge how much a substituent actually shifts acidity rather than treating every electron-withdrawing group as equally powerful. And remember that solvent choice can flip your expectations entirely if you're working outside aqueous conditions. The 18 2 Review And Reinforcement Determining The Strengths Of Acids And Bases material isn't about accumulating facts. It's about building a decision tree you can run through quickly under test conditions. Once the tree is automatic, you stop wasting time searching for the right formula and start seeing the structure of the problem directly.