How CRISPR-Cas9 Actually Works in the Lab
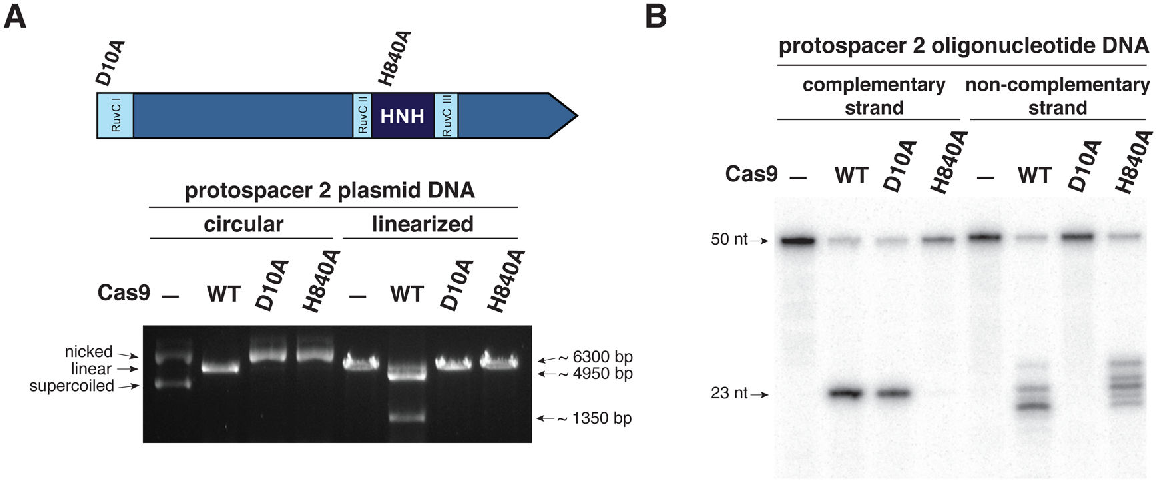
The system is straightforward on paper. A bacterium integrates a short stretch of viral DNA into its own CRISPR array. When the same virus attacks again, that array gets transcribed into a precursor crRNA, which is processed into individual guide RNAs. Each guide pairs with a tracrRNA and binds to the Cas9 protein, forming a complex that scans for matching DNA sequences. When it finds the right target, Cas9 cuts both strands of the DNA. That's it. Simple mechanism. Brutally effective editing once you get it working outside a cell. I spent about six months before I got clean edits on my first cell line. The literature makes it sound like you design a guide, order the oligos, clone into a vector, transfect, and within a week you have your knockout. That is not how it goes. The gap between theory and practice is mostly noise — off-target cuts, low HDR efficiency, and guides that look perfect on paper but do nothing in a real nucleus. Start with guide design. There are dozens of algorithms out there, and they will all give you different scores for the same target. I stopped trying to pick the "best" one and just designed five guides for each locus. The most commonly recommended feature is a PAM sequence — NGG for SpCas9 — located 3 prime to the target. The 20-nucleotide spacer adjacent to the PAM is what directs specificity. Pick your five, verify they don't share obvious secondary structures, and order them as IDT crRNA and tracrRNA pairs if you're doing in vitro transcription, or clone them into a U6-driven gRNA vector if you're doing plasmid-based expression.
Here's something most protocols gloss over: the PAM-proximal region of your target matters more than the distal end for cutting efficiency, but the distal end is where most off-targets sneak in. I ran a side-by-side comparison once where two guides had identical on-target activity but wildly different off-target profiles because their seed regions differed by a single nucleotide from an unrelated gene. That unintended cut showed up in the sequencing data as a messy indel pattern at the wrong locus. It took me three months to trace it back. Check your guides against the whole genome with a tool like Cas-OFFinder or CHOPCHOP before you order anything. Don't skip this step. For the actual delivery, I use electroporation with pre-complexed ribonucleoprotein (RNP) rather than plasmid DNA. The difference is night and day in terms of timing. Plasmid-based Cas9 expression means the protein is being made inside the cell over hours, which increases the window for off-target activity. With RNP delivery, you're adding the protein and guide RNA directly, and the cutting happens within minutes. The transfection efficiency drops slightly compared to plasmid, but the specificity gain is worth it. Mix your Cas9 protein with your crRNA and tracrRNA at a 1:2 molar ratio, incubate at room temperature for ten minutes, and then electroporate into your cells using a program tuned for your specific cell line. Jumpstart Biotechnologies makes a good RNP kit if you want something that just works out of the box. After transfection, you need to assess whether editing actually happened. The quickest method is T7E1 mismatch cleavage. Extract your genomic DNA three days post-transfection, amplify the target region by PCR, denature and reanneal the fragments, treat with T7 endonuclease I, and run on an agarose gel. If you see bands below the full-length product, you have indels. This tells you the cut happened but not what the edit actually is. For that, you need deep sequencing or Sanger sequencing with a tool like TIDE or Synthego's ICE. TIDE is free and fast but it only estimates the edit spectrum — it doesn't give you individual clone sequences. If you need precise characterization, clone your PCR products into a TOPO vector and sequence at least twenty clones per sample.
Homology-directed repair is where things get difficult. If you're trying to knock in a point mutation or a small tag rather than just break the gene, you're relying on HDR, which in most mammalian cell lines operates at less than 10% efficiency. I tried raising HDR efficiency by synchronizing cells in S phase — the logic being that HDR machinery is most active then — and it worked marginally. More importantly, I found that blocking the non-homologous end joining pathway with a small molecule like SCR7 or KU-0060648 pushed the balance toward HDR significantly. You get cleaner knock-ins and fewer random indels. The tradeoff is that cells grow slower and some lines die outright under NHEJ inhibition, so you need to titrate the dose carefully. One edge case that cost me real time: when your target site sits in a highly repetitive region, your PCR primers will amplify multiple loci simultaneously. The T7E1 assay still looks positive, which makes you think you edited your intended target, but the sequencing reveals a mixture of products from five or six different genomic locations. I caught this by running a high-fidelity PCR with primers that span a much wider amplicon than usual, then checking the gel for unexpected bands before proceeding to sequencing. If you see smearing or extra bands, redesign your primers or switch to a different target site. There is no workaround for repetitive-region editing with standard Sanger sequencing — you'd need long-read sequencing or a targeted capture approach, and neither of those is practical for a quick knockout screen. Another practical detail: the of Cas9 protein matters more than people admit. At high concentrations, you get efficient on-target cutting but also a dramatic increase in off-target activity. I've seen guides with theoretically zero off-target matches produce detectable edits at unrelated loci when the RNP concentration was too high. Start with 50 to 100 nM final concentration in your electroporation mix and work up only if your on-target efficiency is unacceptably low. Most of the time, lower is better.
Get the Full Details

If you're working with primary cells or hard-to-transfect lines, consider using a lentiviral vector instead of RNP. The advantage is stable Cas9 expression and the ability to select for transduced cells. The disadvantage is that you're stuck with plasmid-driven expression, which brings back the off-target window problem I mentioned earlier. I use lentivirus only when electroporation simply won't work, and even then I combine it with a self-excising system like Flp-In or a Cre-lox arrangement so I can remove Cas9 after editing is complete. The economics are worth noting. A single round of RNP-based editing with five guides, electroporation, and deep sequencing runs roughly $400 to $800 depending on your sequencing depth. Plasmid-based approaches are cheaper upfront but you'll burn through more money on failed attempts because you're guessing at guide efficiency. Budget accordingly and plan for at least two rounds of guide testing before you commit to a final construct. For anyone just starting out, I'd recommend ordering a commercial CRISPR knockout kit from a supplier like IDT, SnapGene, or Synthego. They come with pre-designed guides that have been empirically validated in your cell line of interest. It costs more than designing guides yourself, but it eliminates the first two months of failure that most people go through. Once you've nailed the workflow and know your cell line's quirks, you can move to custom design and save money on subsequent projects.
CRISPR isn't a finished tool. The basic mechanism is solid and well-understood, but the practical application is still full of variables that depend entirely on your specific targets, cell types, and delivery methods. The guidelines above are based on what I've learned through trial and error, and they'll need adjustment for your particular setup. That's normal. Nobody gets this right on the first try.