Getting a peptide out of a flask isn't magic. It's tedious chemistry done right.
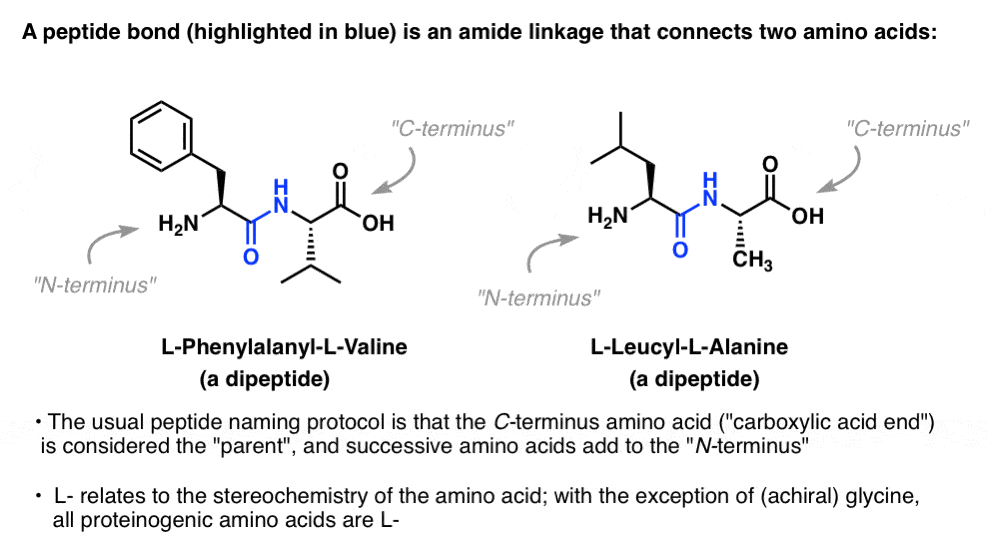
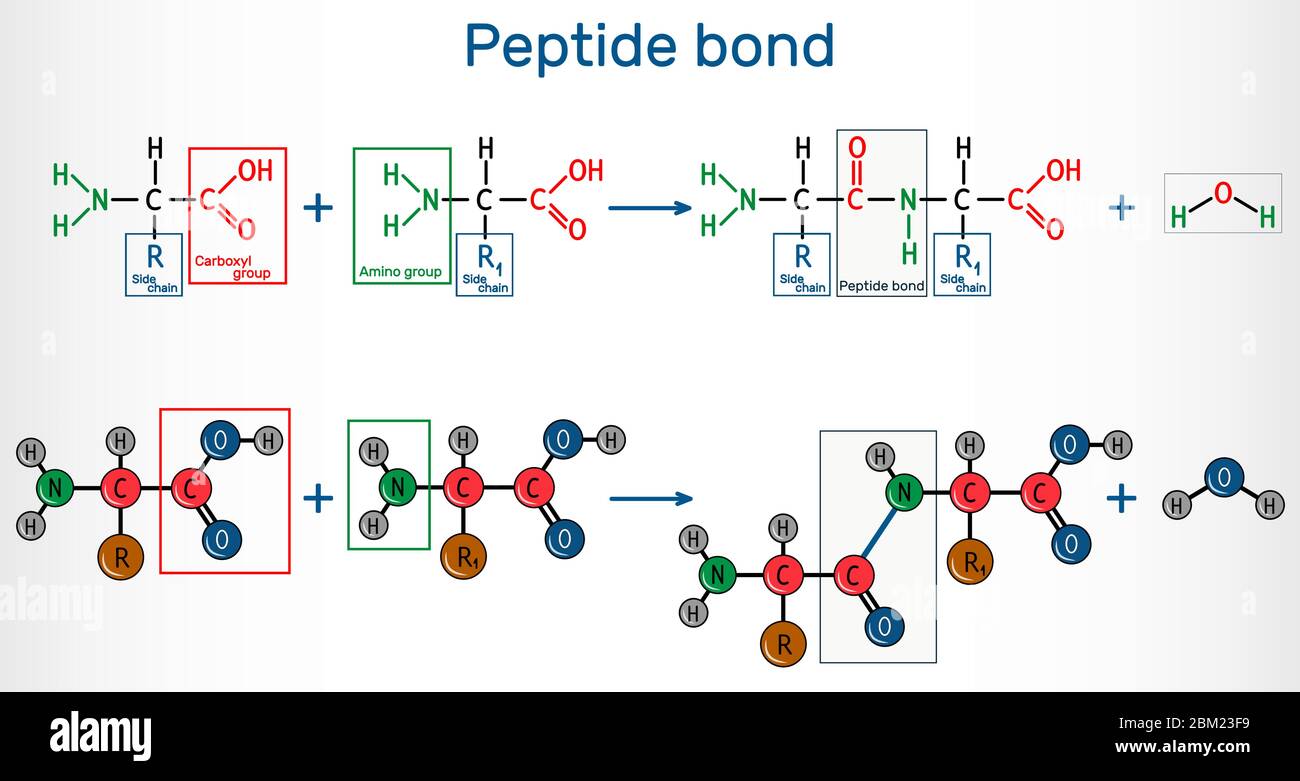
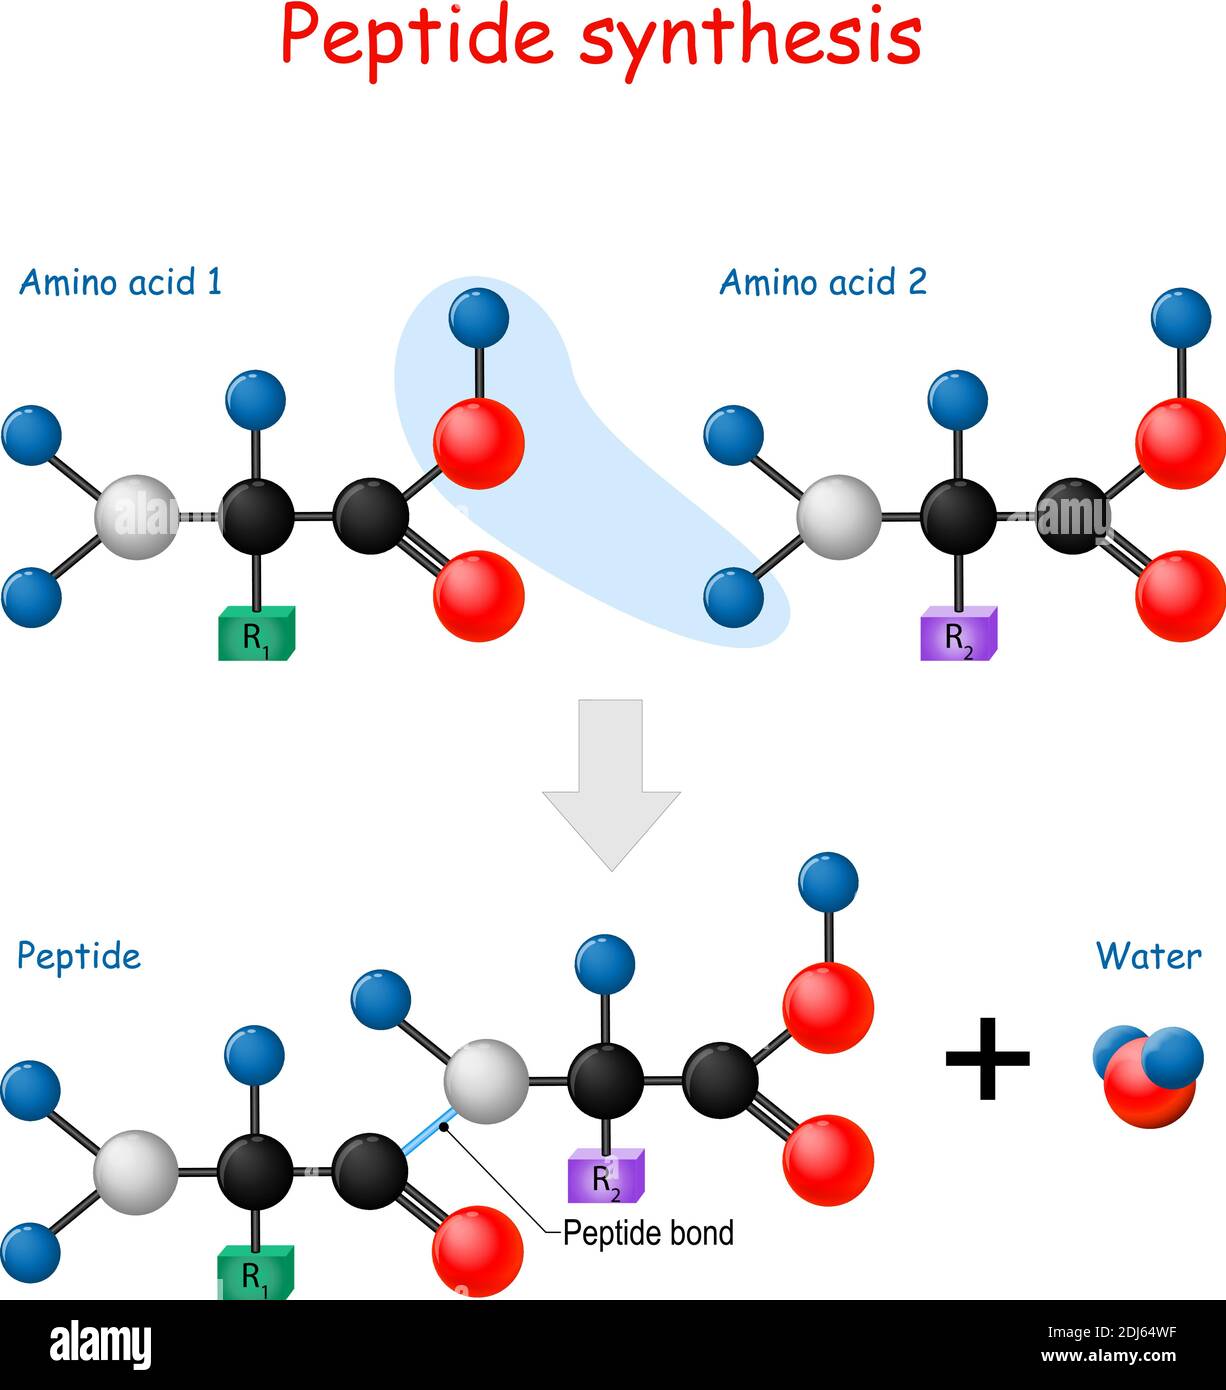
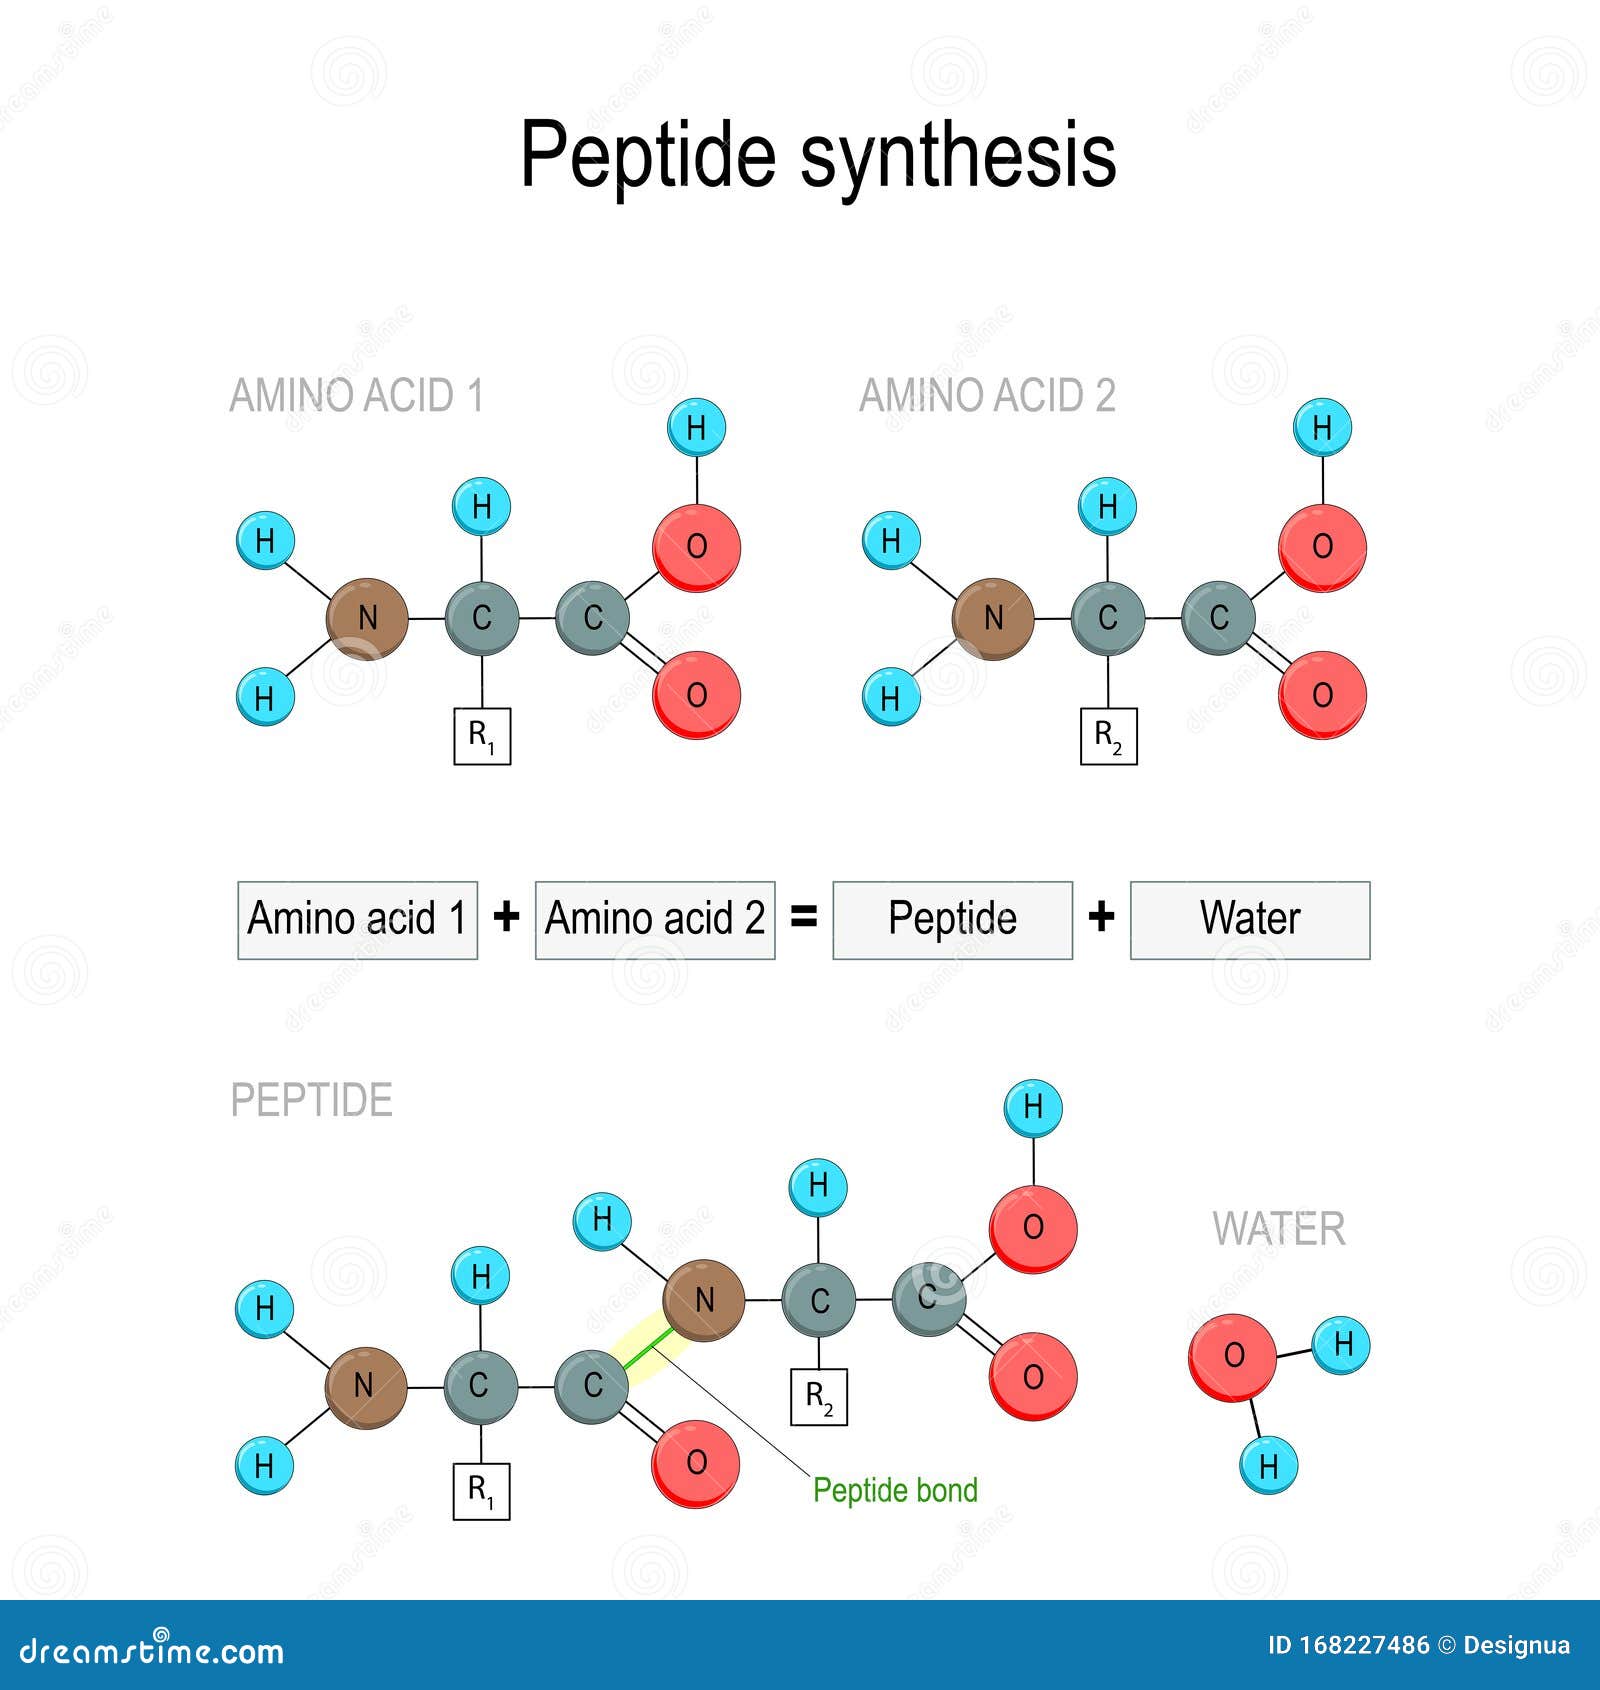
Solid-phase peptide synthesis is the standard method for chaining amino acids together in the lab. You start with the C-terminal amino acid anchored to a resin bead, then cycle through deprotection, coupling, and washing steps while building the sequence from the carboxyl end toward the amine end. Most people use Fmoc chemistry these days rather than the older Boc method because it avoids hydrofluoric acid, which is genuinely dangerous and requires specialized equipment to handle safely. The side chains stay protected with groups like tBu, Trt, or Boc depending on the residue, and the alpha-amine gets temporary protection from the Fmoc group, which comes off with a secondary amine base. You load your first amino acid onto the resin first. If you are using Wang resin, you typically activate the carboxyl group with DIC and a catalyst like DMAP, then swell the resin in DCM and let it sit for a couple hours with gentle rotation. The loading capacity varies by resin batch, so checking the actual loading by a test cleave and HPLC quantification saves you from building on a resin that turned out to be half-active. I learned that the hard way with a batch of 2-chlorotrityl chloride resin that had been sitting on a shelf too long and was nearly dead. Just run a small test coupling and check it. Once your first residue is anchored, each cycle goes like this. Deprotect the Fmoc group using 20 percent piperidine in DMF, usually two separate treatments, the first for ten minutes and the second for twenty. You can monitor deprotection by the UV absorbance at 301 nanometers of the cleared solution, which is useful if you are doing manual synthesis and want to catch incomplete deprotection before moving on. Then wash the resin thoroughly. DMF, DCM, methanol, back to DMF. Rushing the washes is one of the most common mistakes I see, and it shows up later as deletion sequences on your HPLC trace.
Coupling comes next. You activate the incoming Fmoc-protected amino acid with a coupling reagent. HBTU or HATU with DIPEA in DMF is the standard combination for most residues. You typically use three to five equivalents of the amino acid and two to four equivalents of the coupling reagents relative to the resin loading. Shake or rotate the resin for thirty to sixty minutes. For sterically hindered couplings likeVal-Val or Ile-Ile sequences, you might need double couplings, longer reaction times, or switch to PyBOP as the activator. Some people also add a small amount of HOAt to the mix, which can make a real difference with difficult sequences because it reduces racemization and speeds up the coupling. After coupling, wash again and check whether the previous step actually worked. Kaiser test or chloranil test depending on the N-terminus. These colorimetric tests tell you pretty quickly if there is leftover free amine, which means your coupling failed and you need to either re-couple or troubleshoot what went wrong. After the full sequence is assembled, you cleave the peptide from the resin using a cocktail that removes both the resin linker and the side chain protecting groups. For Wang-anchored peptides, standard TFA with scavengers like water, triisopropylsilane, and ethanedithiol is typical. The exact ratios depend on how many sensitive residues you have in your sequence. Cysteine and methionine need extra protection from alkylation by carbocations generated during cleavage, so you will see more thiol scavengers in recipes for sequences containing those amino acids.
Precipitate the crude peptide in cold ether or ethyl acetate, centrifuge, dry under vacuum, then dissolve in water or water-acetonitrile with a little TFA and run it through reverse-phase HPLC. Analytical HPLC first to check purity, then prepareative if you need milligram quantities. Lyophilize and store at minus twenty or lower. That is the basic workflow from start to finish. I ran into a specific problem a few years ago with a twelve-residue peptide that kept coming out with a persistent deletion at position seven, which turned out to be a proline. The standard double coupling wasn't touching it. The issue was that the previous residue was also proline, and the Fmoc-Pro-Pro segment is notoriously difficult. I ended up switching to a longer coupling time with Oxyma Pure instead of HOBt, using DIC as the activator rather than the uranium salts, and pre-activating the amino acid in solution for five minutes before adding it to the resin. That combination worked. The key insight is that proline is a secondary amine and doesn't deprotect and couple the same way the other amino acids do. You have to treat it differently or it drags the whole cycle down. Another thing people miss is that cumulative yield loss is brutal in SPPS. If each coupling step goes at ninety-eight percent efficiency, a twenty-residue peptide ends up with a theoretical yield around sixty-six percent even before you account for deprotection inefficiencies, side reactions, or losses during workup. At ninety-five percent per step you are down to about thirty-six percent. This is why going longer than roughly fifty residues with standard Fmoc SPPS is basically an exercise in frustration. You get a messy mixture and very little product.
Get the Full Details
There are strategies to deal with longer sequences. Pseudoproline dipeptides inserted at certain positions can help with aggregation. Using different resin types or microwave-assisted synthesis changes the kinetics enough to sometimes break through problem regions. But at some point you are better off using fragment condensation or expressed protein ligation rather than pushing a single solid-phase synthesis further than it should go. One counter-intuitive point about reagents: HATU is often marketed as universally superior, but for certain sequences it actually causes more racemization than HBTU or PyBOP, especially with cysteine and histidine residues. If you care about stereochemistry, you should verify it by chiral HPLC after coupling, particularly if your peptide has multiple chiral centers introduced sequentially. Racemization doesn't always show up as a major peak, sometimes it is a small shoulder, but it matters if you are doing anything where biological activity depends on the correct configuration. The biggest bottleneck nobody warns you about is peptide aggregation on resin. When the growing chain starts forming beta-sheet-like structures with the resin-bound neighbors, coupling reagents physically cannot reach the N-terminus. This manifests as truncations, but they show up late in the sequence where the problematic motif occurs. The workaround is usually changing the solvent system, sometimes using NMP instead of DMF, adding chaotropic agents like lithium chloride to the coupling mixture, or running the synthesis at elevated temperature if your resin and protecting groups allow it. Some resins degrade above sixty degrees Celsius, so check the manufacturer specs before heating things up.
If you need longer peptides or are working with difficult sequences that resist standard protocols, automated synthesizers are worth considering. They give better reproducibility and reduce the hands-on time significantly, but they are expensive and the method development is still mostly manual for non-standard sequences. The instruments are fine once you have optimized the conditions, but getting there without doing it by hand at least once teaches you less about what is actually happening in the flask. For most labs doing routine peptides up to twenty or twenty-five residues, a modest setup with a proper fume hood, a rotator, basic glassware, an HPLC system, and a lyophilizer is sufficient. The reagents are not prohibitively expensive if you buy them from major suppliers. The real cost is time and the willingness to troubleshoot when something goes wrong.