Building the MO Diagram for Ammonia
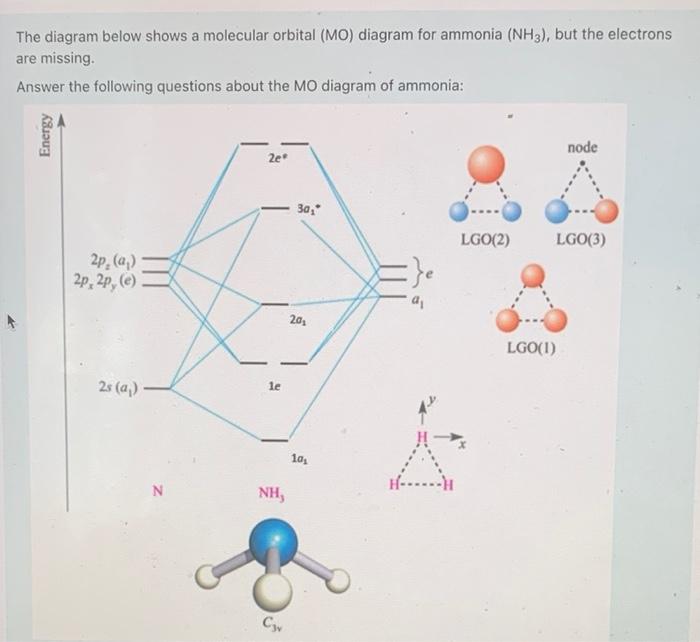
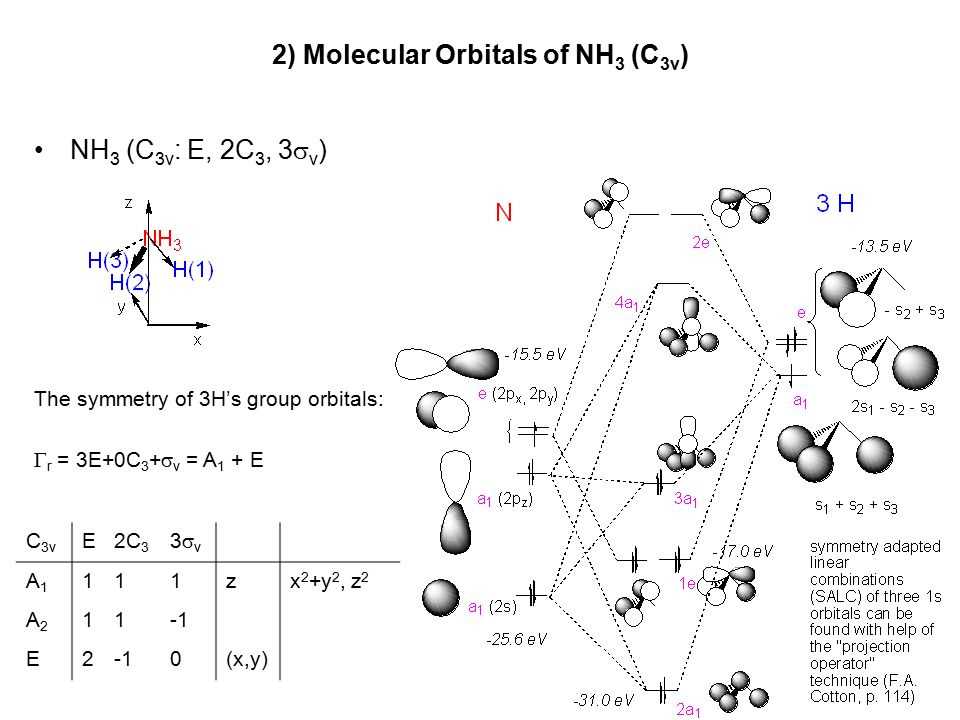
The ammonia MO diagram comes together once you stop treating it like water or methane and just let the C3v symmetry do the work. I spend most of my time running Hartree-Fock jobs on simple molecules, and every now and then a student sends me a diagram where the 2a1 and 1b2 are switched, or worse, they label the lone pair as a bonding orbital. It happens. The group theory itself is straightforward but the cartoon diagrams you find everywhere gloss over the actual energy ordering, and that gap is where people trip up. What you actually need to draw is a correlation between the atomic orbitals on nitrogen and the symmetry-adapted linear combinations of the three hydrogen 1s orbitals. Nitrogen brings a 2s and three 2p orbitals. The hydrogens, taken as a set, generate orbitals that transform as a1 plus a doubly degenerate e set under C3v. That gives you four occupied molecular orbitals and a few empty ones above, and the lowest three are bonding while the highest occupied one is the nonbonding lone pair.
Reading an Ammonia Molecular Orbital Diagram Correctly
I ran into a real problem last month when a researcher wanted to use the NH3 MO diagram as a reference for a ligand field calculation on a cobalt complex. They had pulled a textbook figure where the 2a1 was drawn below the 1b2, which is the standard textbook convention, but their computational output from a DFT run showed the 1b2 coming out lower. The mismatch wasn't a mistake on either side. It's the difference between the qualitative LCAO picture and what actually comes out of an eigenvalue solver when you let the basis set do its thing. I told them to trust the numbers for the complex but keep the textbook ordering as a shorthand for talking about the frontier orbitals. Both are useful if you know which pile you're standing in. The nitrogen 2s orbital is a1 symmetry. It overlaps with the a1 combination of the three H 1s orbitals, which is the in-phase sum of all three hydrogens. That produces a bonding 1a1 and an antibonding 2a1. The nitrogen 2pz orbital, if you put the z-axis along the C3 symmetry axis, is also a1, and it mixes with that same hydrogen a1 combination. That mixing pushes the 1a1 down and the 2a1 up, and it is exactly why the 2a1 ends up being the lone pair orbital in the usual picture. The 2px and 2py orbitals on nitrogen are e symmetry. They overlap with the e pair of hydrogen combinations, giving you a bonding 1b2 set and an antibonding 2b2 set. The bonding 1b2 orbitals are the two N-H bonds that sit in the plane roughly perpendicular to the symmetry axis. Orbital energies in the diagram run from about minus 40 eV for the 1a1 core-like nitrogen 2s-derived orbital up to the 2a1 lone pair near minus 10 eV, with the 1b2 bonding orbitals somewhere in between. The exact numbers shift depending on your method, but the relative ordering is stable: 1a1, then 1b2, then 2a1, then the empty 2b2 and higher antibonding orbitals. That ordering is what determines the photoelectron spectrum, and if you look at experimental HEISE data for ammonia, the first ionization comes from the 2a1, which confirms the lone pair sits highest among the occupied orbitals.
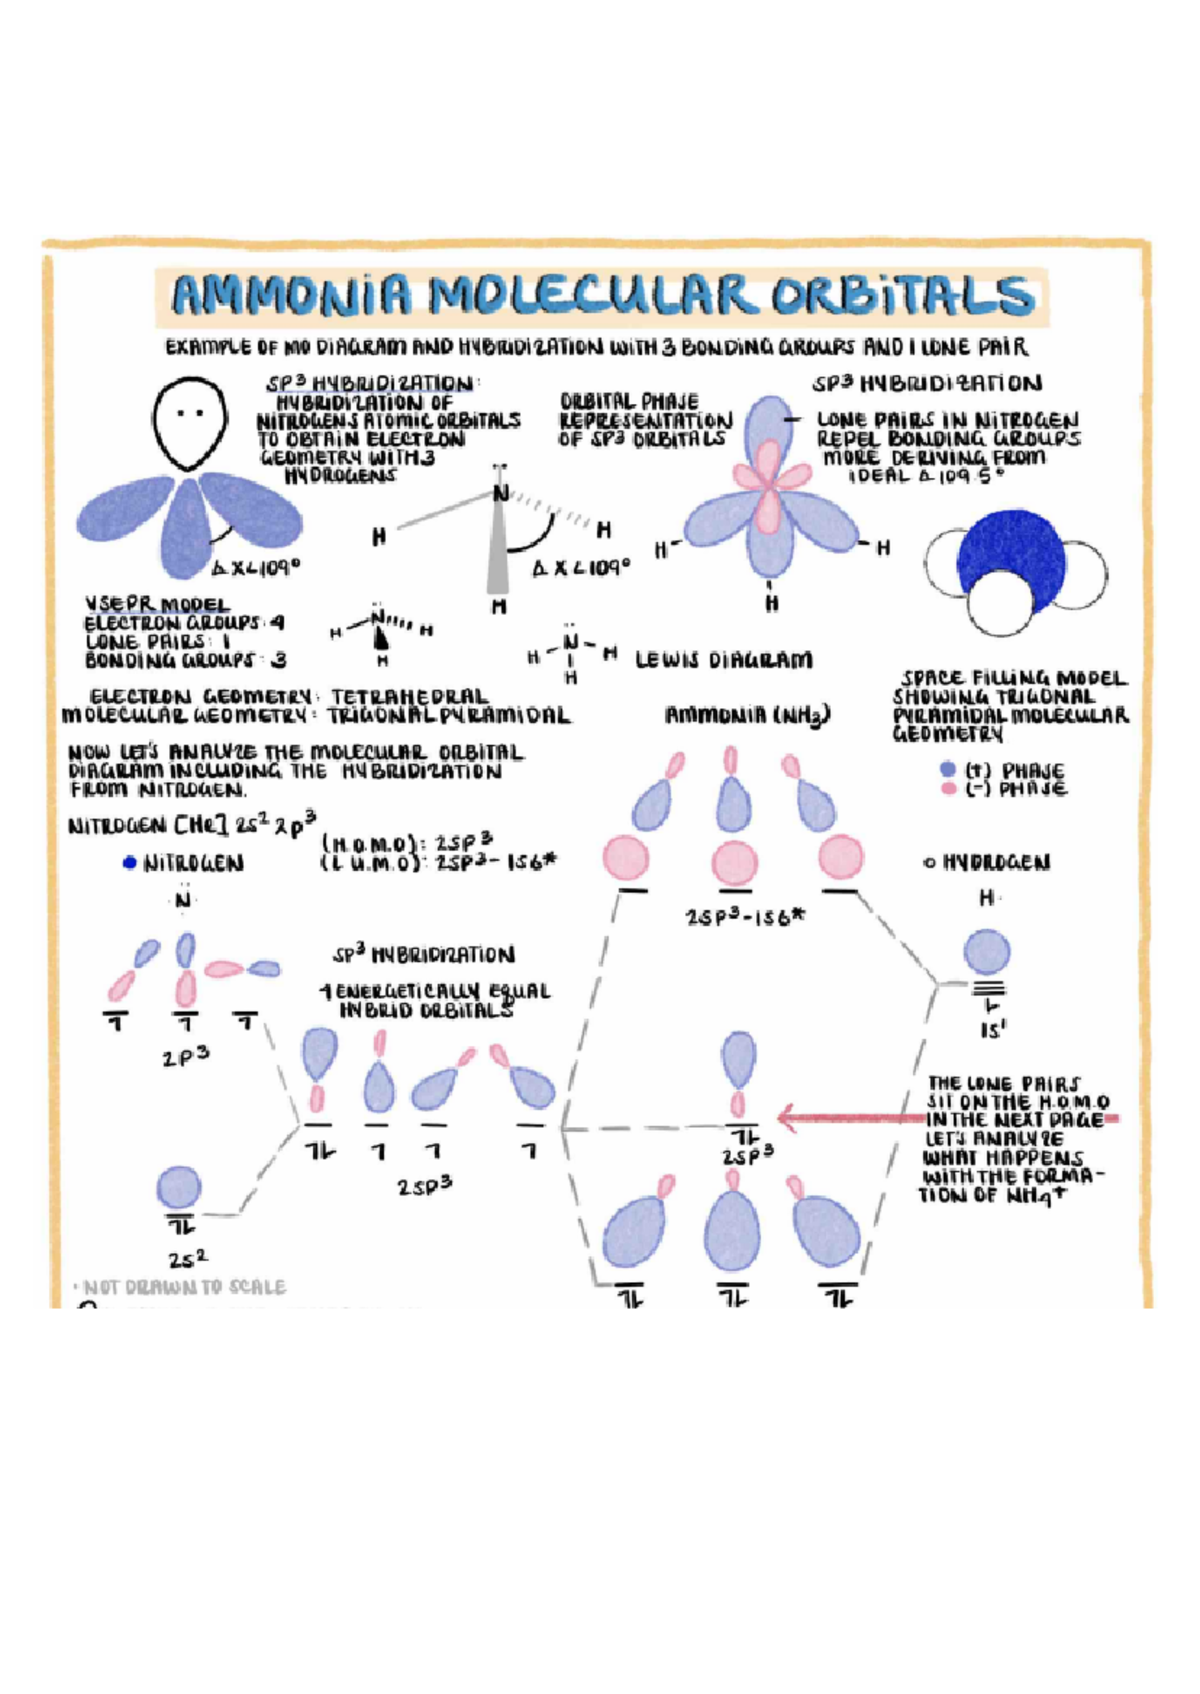
One thing people routinely miss is that the lone pair is not a pure sp3 hybrid. The 2a1 orbital has substantial s-character because the nitrogen 2s contributes directly to it, and the geometry of ammonia, with its 107-degree bond angle, reflects that. If you force a pure sp3 model onto the diagram, you get the angle wrong and you also misinterpret what the lone pair actually is. It is an a1 orbital, yes, but it is better described as a nitrogen-centered orbital with mixed s and pz character rather than a tetrahedral hybrid pointing straight up. The distinction matters when you are doing anything beyond drawing a pretty picture, like calculating dipole moments or modeling how ammonia binds to a metal center. The antibonding 2b2 orbitals are the next step up in energy, and they are e symmetry. They are unoccupied in the ground state but show up clearly in UV absorption and in core-level excitation spectra. If you are fitting a simple Hamiltonian or building a tight-binding model for ammonia adsorbed on a surface, those 2b2 orbitals are the ones that accept electron density from the metal, and their e symmetry means they interact differently with surface orbitals depending on the adsorption site. I learned that the hard way when someone tried to use a C2v-symmetric model for ammonia on a stepped copper surface and got the binding geometry wrong because the symmetry was incompatible with the actual C3v frontiers. The diagram itself has limits. It assumes a static, isolated molecule in its equilibrium geometry. Real ammonia in solution, or under pressure, or vibrationally excited, does not sit still long enough for the diagram to stay perfectly accurate. The 2a1 orbital shape changes as the molecule inverts, and that inversion doublet is not captured by a single equilibrium MO picture. If you need dynamics, you run a molecular dynamics simulation and project the orbitals onto the trajectory, or you use a vibronic coupling model. The MO diagram is a snapshot, not a movie.
Get the Full Details
Another practical limitation is that the simple LCAO-MO picture here does not include electron correlation explicitly. Post-Hartree-Fock methods like MP2 or CCSD(T) will shift the orbital energies by a few electron volts compared to a basic HF calculation, and the ordering can change for certain excited-state configurations. For ground-state chemistry and basic spectroscopy, the qualitative diagram is fine. For quantitative work, you should rely on the computed Kohn-Sham or Hartree-Fock orbitals and treat the textbook diagram as a mnemonic rather than a prediction engine. If you want to generate your own diagram quickly, a standard Gaussian or ORCA job with a modest basis set like 6-31G* will output the orbital energies and symmetries directly. You can then plot the isosurfaces to verify that the 1a1 is mostly nitrogen 2s, the 1b2 orbitals are the bonding pi-like combinations, and the 2a1 is the lone pair with the expected pz mixture. I usually export the .fchk file and use Molden or Multiwfn to visualize the orbitals alongside the energy level table. That takes about ten minutes for a molecule this small, and it saves you from arguing with a textbook figure that may or may not match the convention you are using. For reference data, the NIST Webbook and the HITRAN database have spectroscopic constants that anchor the experimental energy levels, and the classic papers by Herzberg on molecular spectra contain the original band assignments that validate the orbital ordering. If you are teaching this material, I recommend showing both the textbook diagram and a computed one side by side, and explicitly pointing out where they agree and where they diverge. That tends to prevent the kind of confusion that shows up in exam answers and in the first round of computational projects.