Understanding Trigonal Pyramidal Geometry in Practice
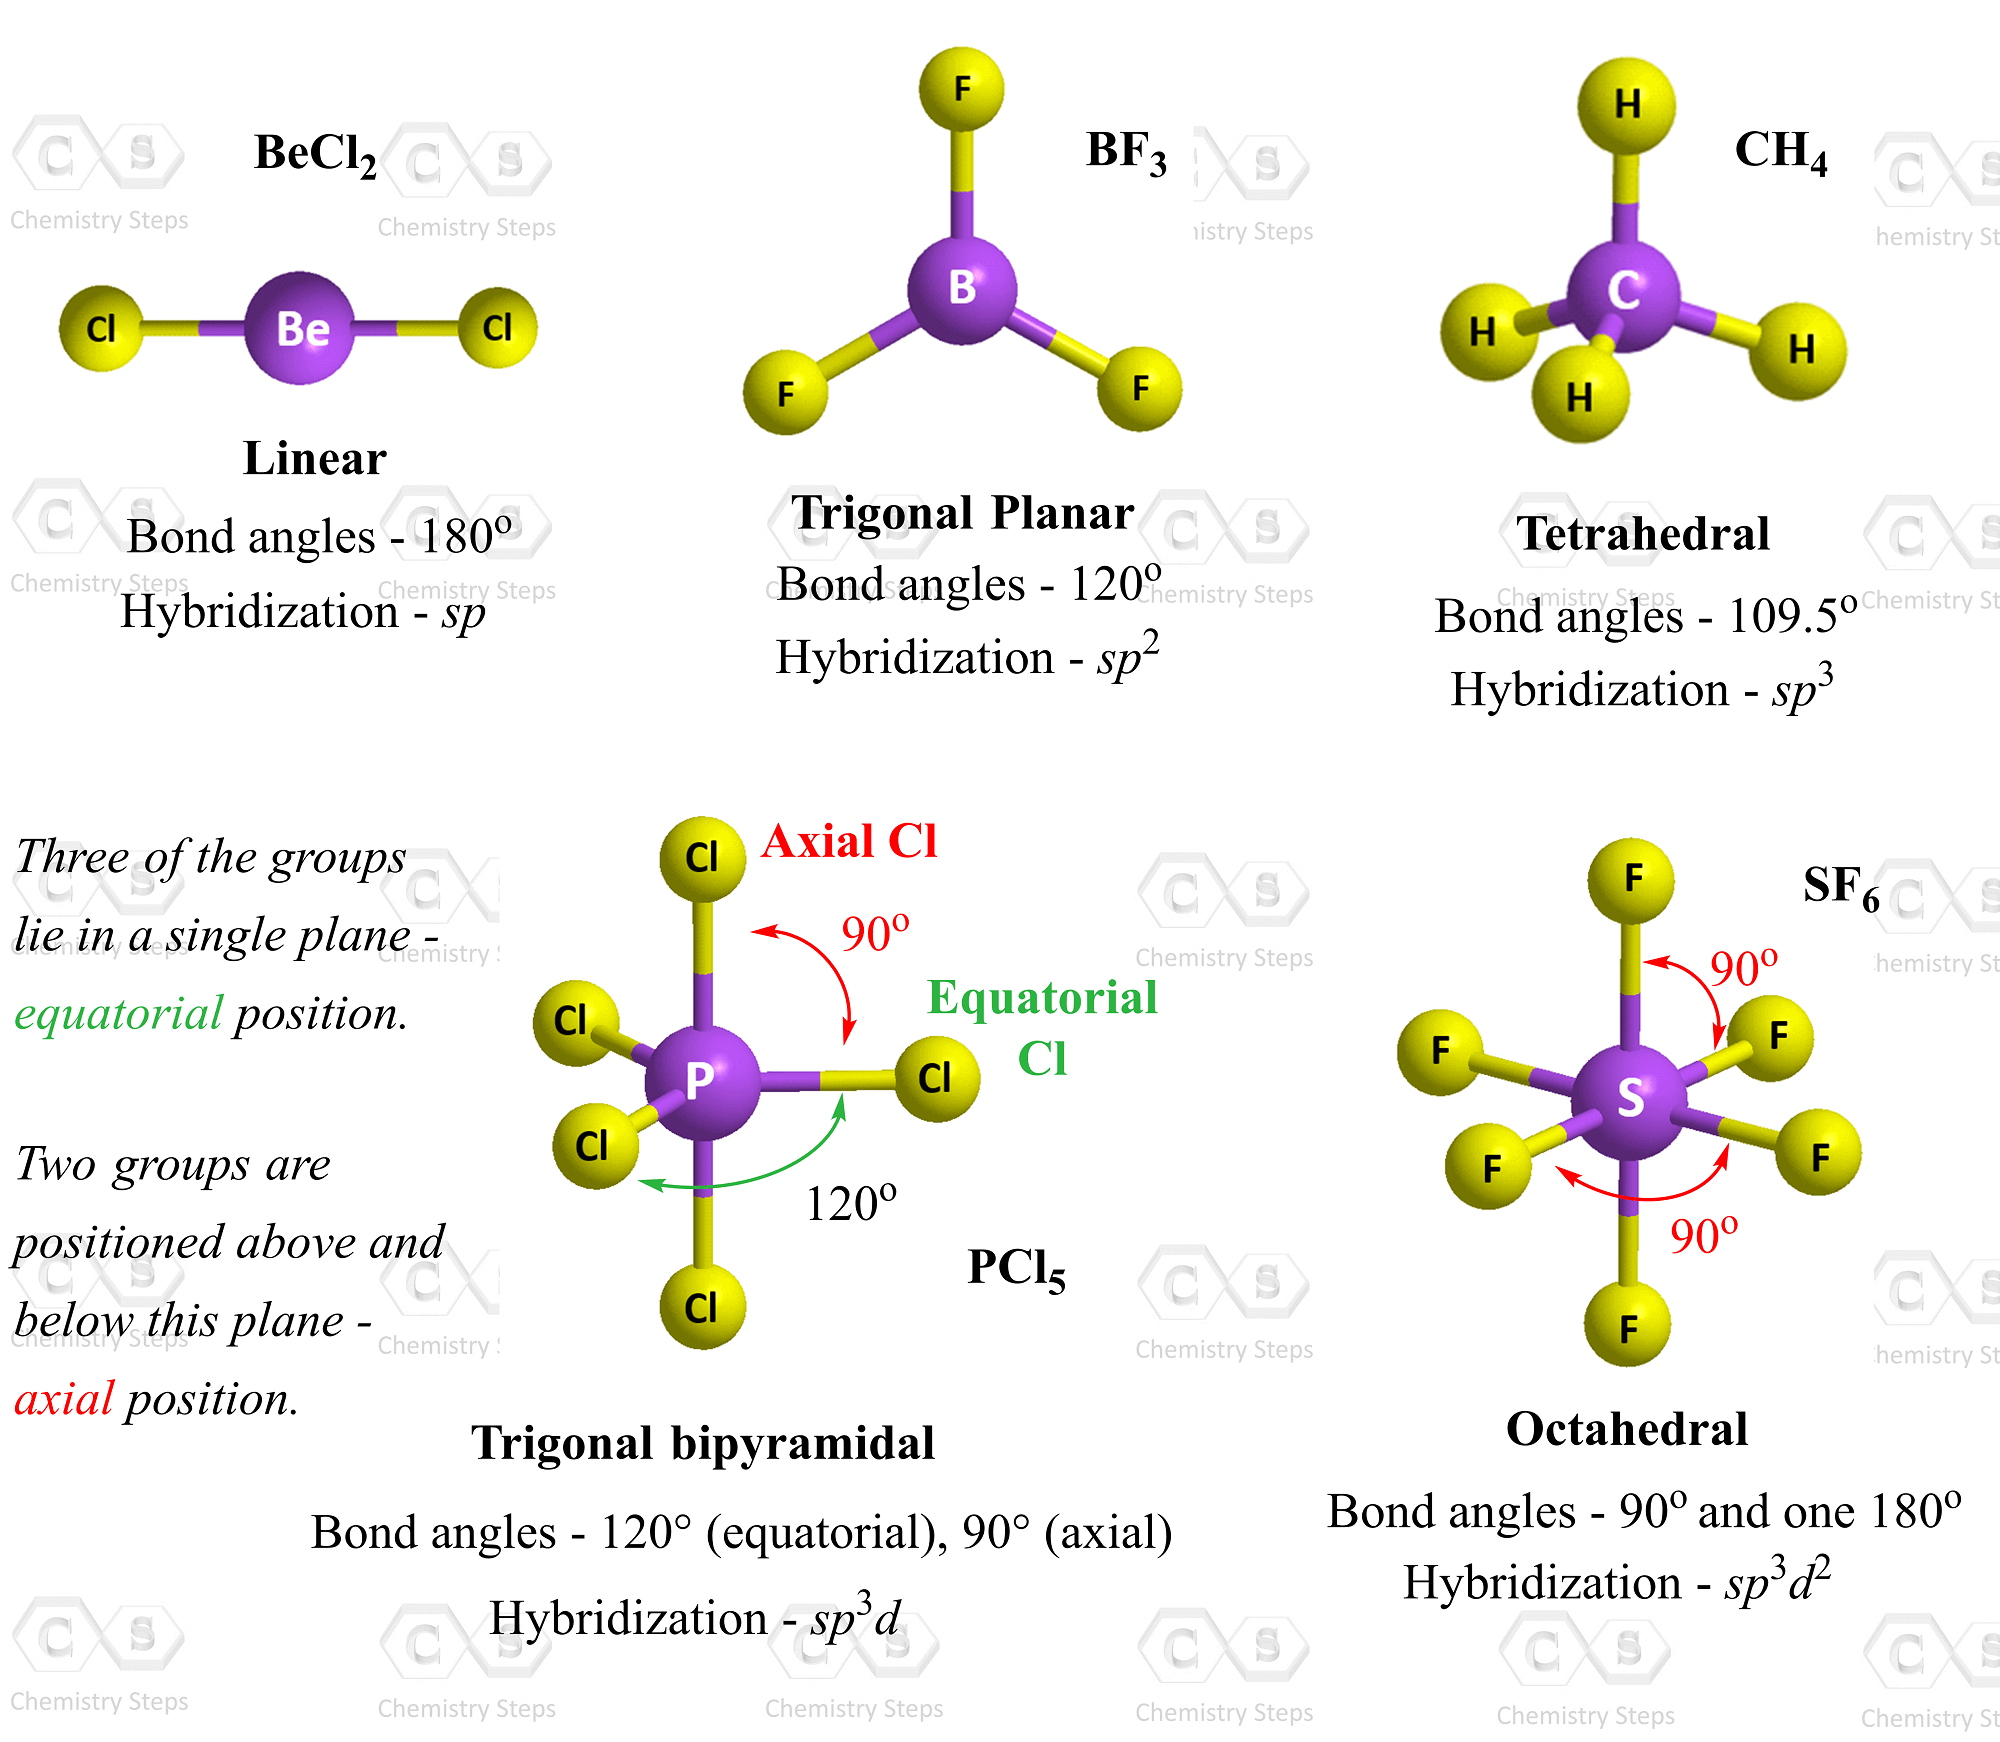
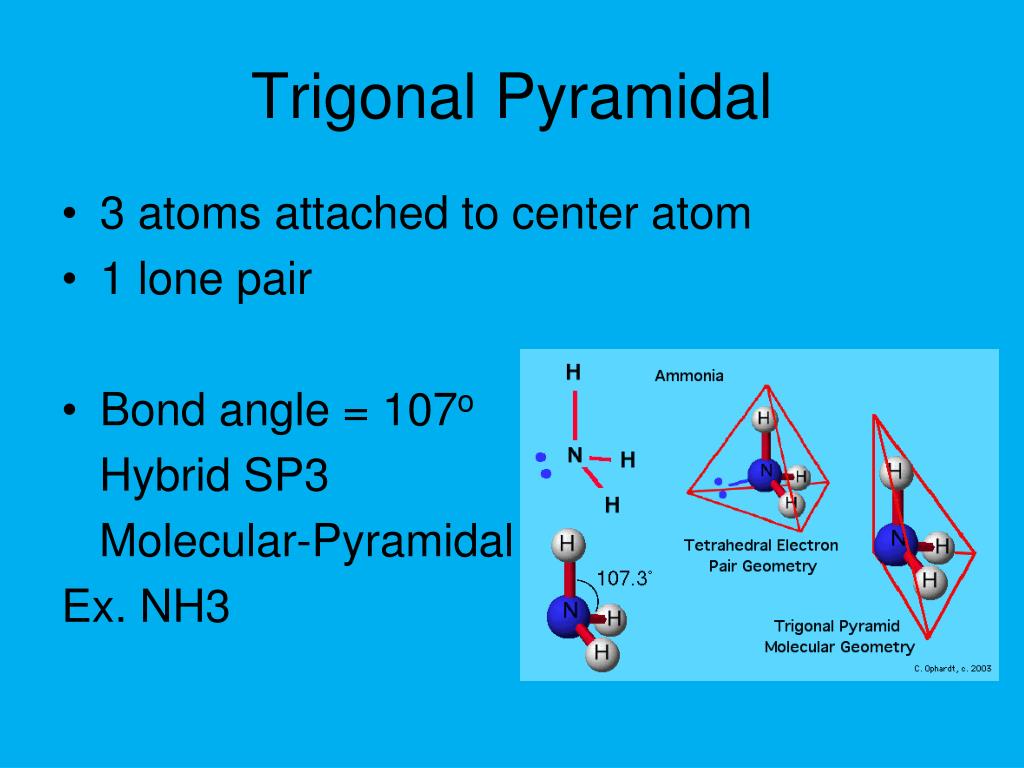
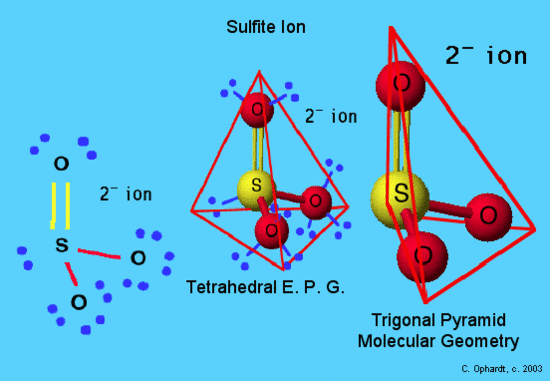
Most people learn about trigonal pyramidal shapes in their first chemistry course and move on. The real picture is messier than the textbook version. When you actually work with molecules like ammonia, phosphine, or certain coordination compounds, the bond angles don't behave exactly as simple VSEPR theory predicts. I've spent years dealing with compounds where the geometry looked right on paper but the angles came out somewhere unexpected in the lab data. At its core, trigonal pyramidal geometry arises when a central atom has three bonding pairs and one lone pair of electrons. The electron geometry is tetrahedral, but the molecular shape is pyramidal because the lone pair occupies one vertex. The ideal tetrahedral angle is 109.5 degrees. With one lone pair present, the bond angle compresses below that value. Ammonia (NH3) is the standard reference point at about 107 degrees. That two-and-a-half degree compression comes from the lone pair exerting greater repulsion than bonding pairs. VSEPR theory treats lone pairs as taking up more space, which pushes the bonding pairs closer together. This works well for light main-group elements but falls apart when you start mixing in heavier atoms or transition metals.
Here's where it gets interesting. Phosphine (PH3) has a bond angle of only about 93.5 degrees. That's nearly 14 degrees smaller than ammonia, even though both have the same basic electron configuration. The difference comes down to the central atom's size and orbital hybridization. As you go down the group, the s-character of the bonding orbitals increases and the p-character decreases. The bonds essentially become nearly pure p-orbital interactions, which naturally sit at 90 degrees to each other. The VSEPR model doesn't predict this well at all. It would suggest the angle should stay roughly similar or decrease only slightly. I once spent three weeks troubleshooting X-ray crystallography data on a phosphonium salt where the reported P-C bond angles were clustering around 96 degrees instead of the expected 100-plus range. The compound was a standard reagent, nothing exotic. The issue turned out to be crystal packing forces distorting the geometry, something that never shows up in gas-phase calculations. I had to run computational models at the B3LYP/6-311+G(d,p) level with solvent corrections to get the angles to match the experimental data within acceptable error margins. Simple gas-phase optimization gave me angles that were 4 to 6 degrees too wide.
The Practical Side of Bond Angle Predictions
If you need to estimate bond angles for trigonal pyramidal molecules without running computations, there are rough heuristics you can use. For first-row elements, subtract roughly 2 to 3 degrees from the tetrahedral angle for each lone pair. That puts you in the 106 to 108 degree range for species like NH3 or the ammonium cation minus one hydrogen. For second-row and heavier elements, this rule breaks down fairly quickly. PH3, AsH3, and SbH3 all show significantly compressed angles that have nothing to do with lone pair repulsion and everything to do with poor orbital overlap and increased p-character in bonding. Ligand electronegativity also plays a role that students often overlook. When the substituents attached to the central atom are highly electronegative, they pull electron density away from the bonding region. This reduces the repulsion between bonding pairs and allows the lone pair to compress the angle further. NF3 has a bond angle of about 102 degrees, noticeably smaller than NH3's 107 degrees. The fluorine atoms are dragging electron density outward, which weakens the bonding pair-bonding pair repulsion and lets the lone pair squeeze the structure more. Here's a counter-intuitive point that catches people off guard. Adding electron-withdrawing groups doesn't always decrease the bond angle in every context. In some coordination complexes and hypervalent systems, the trend reverses depending on whether the central atom can access d-orbitals or engage in back-bonding. I've seen cases where replacing hydrogen with trifluoromethyl groups actually opened the angle slightly because the steric bulk of the CF3 groups overrode the electronic effects. The angle went from about 107 to roughly 109 degrees despite the increased electronegativity of the substituents.
Get the Full Details
When Standard Models Fail Completely
Don't trust VSEPR for anything involving transition metals with trigonal pyramidal geometry. The model assumes pure electrostatic repulsion between electron pairs, which is a reasonable approximation for main-group molecules but meaningless when d-orbitals, ligand field effects, and Jahn-Teller distortions are in play. A square planar complex with a missing ligand site might look pyramidal on a casual glance, but the electronic structure driving the geometry is entirely different from NH3. Crowded molecules with bulky substituents are another failure mode. Steric pressure from large groups can force bond angles wider than electronic effects alone would predict. I've worked with amines bearing triisopropylsilyl groups where the N-C-C angles were pushed out to 112 degrees despite the presence of a lone pair that should be compressing them. The steric repulsion between the bulky groups dominated completely over the electronic repulsion effects that VSEPR describes. Computational chemistry tools like Gaussian, ORCA, or even lighter programs like Avogadro with semi-empirical methods can give you reasonable angle estimates quickly. For most organic molecules, HF/3-21G or PM6 optimization will get you within a few degrees of the experimental value in under a minute on a modern laptop. If you need higher accuracy, DFT with a triple-zeta basis set will get you there but might take 15 to 30 minutes depending on system size. For routine work, the semi-empirical route is usually sufficient unless you're publishing structural data that needs to survive peer review.
The most common mistake I see people make is treating the bond angle as a fixed constant rather than a value that shifts with environment. Gas-phase ammonia is 107.8 degrees. In liquid water, hydrogen bonding can shift that by a degree or two. In a crystal lattice with strong intermolecular contacts, deviations of 3 to 5 degrees from the gas-phase value are routine and not necessarily a sign of anything wrong with the structure. If you're comparing computed angles to experimental ones, always make sure you're comparing the same phase and conditions, or at least acknowledge the difference when it exists.