
How to Calculate Bond Order Without Getting Confused
Bond order tells you how many chemical bonds exist between two atoms. It is not just a number you look up. You calculate it from the electronic structure of the molecule. Most people try to skip that step and go straight to a Bond Order Calculator Chemistry tool because it is faster. That works for simple diatomics. It falls apart quickly when you start dealing with ions, resonance, or transition metals. Let me explain what the calculation actually looks like under the hood before we get to the tools, because knowing that will save you time when the online calculators give you an answer you do not trust.
What a Bond Order Calculator Chemistry Tool Actually Does
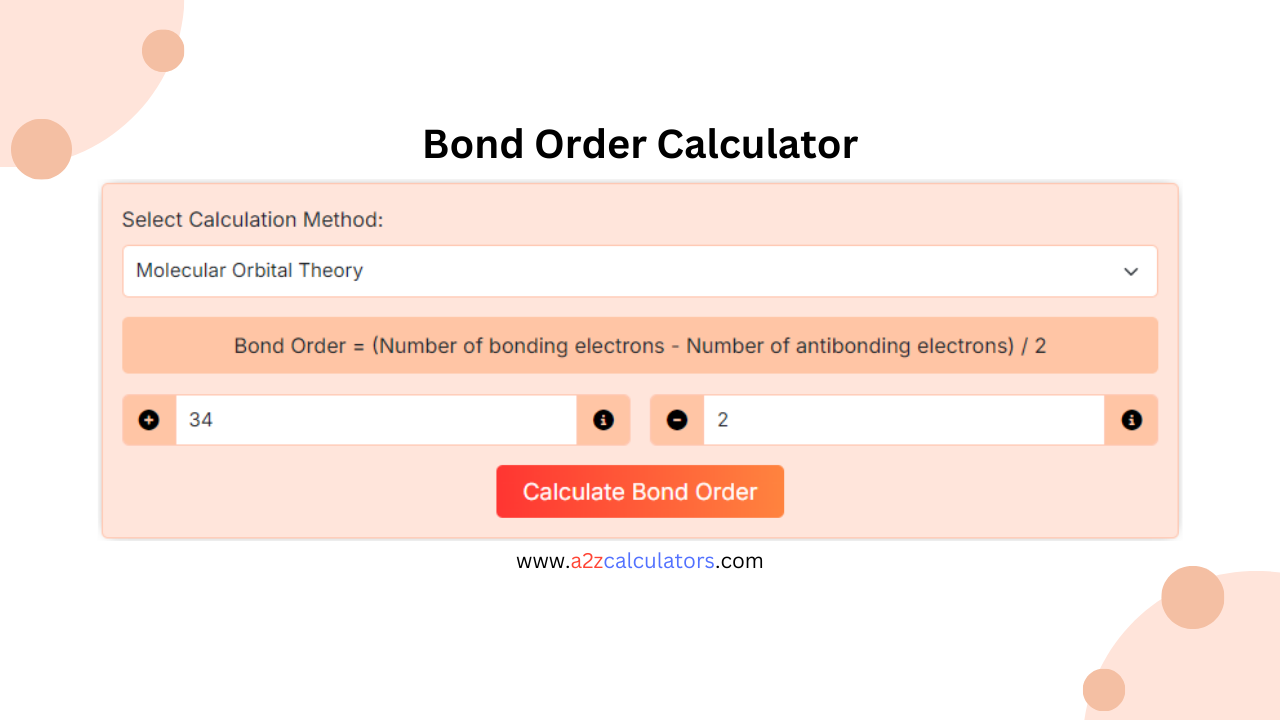
Most bond order calculators operate on one of two approaches: molecular orbital theory or Lewis structure averaging. The MO method counts bonding electrons and antibonding electrons, then applies a simple formula. The Lewis method identifies all valid resonance structures and averages the bond multiplicity across them. Some calculators do both and show you the result from each approach. This can reveal discrepancies that tell you something important about the molecule. The standard MO formula is: Bond Order = (Number of bonding electrons Number of antibonding electrons) / 2
This is correct for homonuclear diatomic molecules. It works for heteronuclear diatomics too, provided the MO diagram is built properly. It does not work well for anything with three or more atoms unless the software handles the full MO construction automatically. Here is the thing most introductory textbooks leave out. The formula is not the hard part. Getting the electron configuration into the correct MOs is where everything goes wrong. I have seen students input the total valence electron count but fill the orbitals in the wrong order, particularly for second-row diatomics where the 2p and 2p energy levels cross over between B2/C2 and N2/O2/F2. Put those levels in the wrong order and your bond order will be off by one. A Bond Order Calculator Chemistry interface might flag this with a warning, or it might just give you a confidently wrong answer.
Get the Full Details

Resonance and the Averaging Problem
For polyatomic molecules with resonance, bond order is calculated by averaging bond multiplicities across all significant resonance contributors. Consider ozone, O3. You draw two valid resonance structures. Each has one single bond and one double bond. The actual bond order for each OO bond is (1 + 2) / 2 = 1.5. Any decent calculator should return 1.5 for both bonds. If it returns 1.0 for one and 2.0 for the other, the program is not handling resonance correctly, or it is treating one Lewis structure as the only valid one. I ran into this exact issue last year when a student submitted results from an online calculator for the carbonate ion, CO3². The tool returned bond orders of 1.0, 1.0, and 2.0 instead of the correct 1.33 for all three CO bonds. I traced it back and the calculator was simply taking the first valid Lewis structure it generated and not averaging across all resonance forms. That is a structural limitation in a lot of free online tools. They are fast but shallow.
Carbon Monoxide Is a Trap
CO is one of the most common gotchas. The Lewis structure shows a triple bond with a formal charge separation: CO with a lone pair on carbon and a lone pair on oxygen. The bond order from Lewis theory is 3. The MO calculation also gives a bond order of 3. But here is the nuance that most calculators and textbooks gloss over. The highest occupied molecular orbital in CO is slightly antibonding in character, concentrated mostly on the carbon atom. This means the true bond order is somewhere between 2 and 3, not exactly 3, and the bonding is highly polarized. Experimental data supports a bond length consistent with a bond order near 2.6 to 2.8, not a clean triple bond. If your calculator says exactly 3 for CO, it is running the simplified MO model, not a computed one. That is fine for an intro course. Do not rely on it for research-level work. Counting total electrons instead of valence electrons. This is the single most frequent error. The bond order formula requires valence electrons only. Neon has 10 electrons total but 8 valence electrons. Using the wrong count shifts every orbital assignment by one or two electrons. Ignoring charge on ions. NO has 10 valence electrons. NO has 11. NO has 12. Each step changes the bond order by 0.5. NO has a bond order of 3. NO has 2.5. NO has 2. A calculator that does not ask for the ion charge will return the neutral molecule result every time.
Assuming bond order equals bond multiplicity from a single Lewis structure. Benzene has alternating single and double bonds in any one Kekulé structure, but the actual bond order for every CC bond is approximately 1.67. If you only draw one resonance structure, your answer will be wrong. The same applies to nitrate, nitrite, sulfite, and most conjugated systems. Treating fractional bond orders as impossible. Bond orders of 1.5, 2.5, and 3.5 are completely normal in stable molecules. O has a bond order of 1.5. O has 2.5. These are real, measurable species with real bond lengths between the single and double (or double and triple) bond references.
When Bond Order Predictions Break Down
Bond order correlates well with bond length and bond strength for simple molecules. Higher bond order means shorter, stronger bonds. This correlation holds for diatomics and small polyatomics. It breaks down in several specific scenarios: Transition metal complexes. The d-orbital splitting patterns and variable oxidation states make bond order calculations ambiguous. Different ligand field models give different answers. A calculator based on simple MO theory will not handle this correctly. You need crystal field theory or ligand field theory computations instead. DFT software is the standard approach for transition metal bond orders, not any simple calculator. Delocalized systems beyond simple resonance. Aromatic compounds, conjugated polymers, and metallic bonding cannot be captured by counting electrons in localized bonds. The concept of a single bond order between two atoms loses meaning in these systems. The electrons are shared across the entire framework.
Weak interactions. Hydrogen bonds, van der Waals forces, and coordination bonds do not fit the bond order framework at all. You cannot use this method to calculate a bond order for something like the interaction between water and a sodium ion in solution.
Practical Steps for Reliable Results
Draw the Lewis structure first. Identify all resonance forms. Count the valence electrons carefully, including any ionic charge. For diatomics, construct or look up the correct MO diagram for your specific molecule, paying attention to whether the 2p and 2p ordering applies. Feed the numbers into a Bond Order Calculator Chemistry tool that shows its work. If it only shows a final number without the electron count or orbital diagram, verify the result independently. For benzene, the bond order is 1.67, not 1.5. The six electrons are delocalized across six carbon atoms, giving each CC bond a contribution of one extra half-bond on top of the sigma framework. This is different from ozone, where the system is confined to three atoms and gives a bond order of 1.5. Do not conflate the two cases. For NO, the nitrogen dioxide radical, you have 17 valence electrons. The unpaired electron goes into an antibonding orbital. The bond order comes out to approximately 1.5 for each NO bond, but the molecule is bent with a bond angle near 134°, which is wider than you would expect from simple VSEPR because the unpaired electron in the antibonding orbital reduces the electron repulsion in the bonding region. A calculator will give you the number. It will not explain the geometry.

Limitations You Should Know About
The biggest limitation of any bond order calculation is that it is a model, not a measurement. It gives you a useful approximation. It does not give you the true electron distribution. Experimental bond lengths and vibrational frequencies are the real data. Bond order is derived from theory to help you predict and interpret that data. When the prediction disagrees with experiment, trust the experiment. Free online calculators are adequate for homework problems involving diatomics and simple ions. They are not adequate for publication-quality work. If you need accurate bond orders for anything beyond the simplest molecules, you need to run a quantum chemistry calculation. Gaussian, ORCA, or even the free package Psi4 will give you population analyses that report Mayer bond orders or Wiberg bond indices, which are more rigorous than the simple electron-counting method. These programs take longer to set up. A typical calculation for a medium-sized organic molecule takes maybe 10 to 30 minutes on a standard laptop. That is slower than clicking a web calculator, but the results are credible. I stopped trying to use web calculators for anything past sophomore-level chemistry about two years ago. They fail silently on edge cases, and the failures are the ones that matter most. A manual calculation with a properly drawn MO diagram takes five minutes for O and gives you an answer you actually understand.


