Understanding The Proton Transfer That Runs Half Of Chemistry
When I first started running acid-base titrations in a teaching lab, I kept getting weird inflection points that didn't match the textbook curves. It turned out I had a weak polyprotic system where two pKa values were too close together, and the usual assumption of isolated proton transfers broke down completely. That was the moment I stopped treating Bronsted acid-base chemistry like a set of rules and started treating it like a map of actual proton movements. The Bronsted Acid And Bronsted Base framework is one of those models that sounds simple on paper and then reveals hidden complexity the first time you try to use it outside of ideal conditions. It is not wrong. It is just incomplete by design, and that incompleteness is what makes it useful for most practical work while creating real headaches at the edges.
How The Bronsted Acid And Bronsted Base Model Actually Works
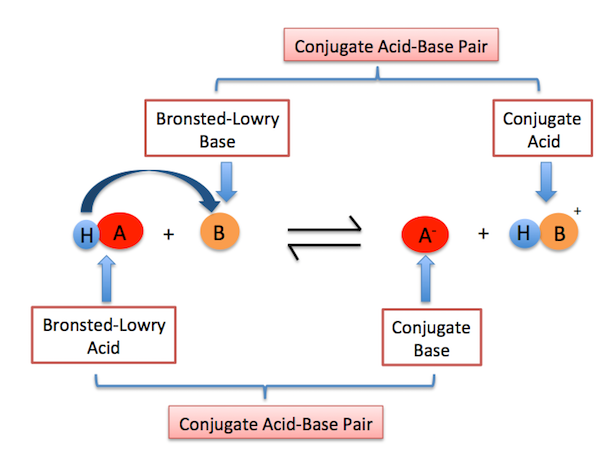
A Brønsted acid is any species that can donate a proton. A Brønsted base is any species that can accept a proton. That is the entire definition. Everything else follows from tracking where the proton goes and how much energy it takes to move it. The model treats every acid-base reaction as a proton transfer between two partners. The acid loses a proton and becomes its conjugate base. The base gains a proton and becomes its conjugate acid. You write the equation, identify both conjugate pairs, and then compare their pKa values to predict which direction the equilibrium favors. If the acid on the left has a lower pKa than the conjugate acid on the right, the reaction proceeds forward. If it has a higher pKa, it sits on the left side and barely reacts. The math is straightforward. The chemistry underneath it is not always obvious. What most students miss is that the Brønsted model does not care about metals, orbitals, or electron pairing. It cares about protons and their affinity for different molecular environments. Lewis acid-base theory handles the stuff Brønsted leaves out, but you rarely need the Lewis framework for routine organic and aqueous chemistry. The Brønsted model covers the territory that matters 90 percent of the time.
Reading pKa Tables Like A Practical Chemist
A pKa table is not a reference library. It is a prediction engine. When you know the pKa of your acid and the pKa of the conjugate acid formed by your base, you can calculate the equilibrium constant in your head. The difference in pKa values multiplied by roughly 2.3 gives you the log of K. A difference of 3 pKa units means the equilibrium favors products by about a thousand to one. A difference of 1 unit means you are sitting at roughly three-to-one. A difference of zero means you have a 50-50 mixture and no useful reaction driving force. I learned this the hard way when I was trying to deprotonate a phenol with sodium bicarbonate in a synthesis step. The pKa of carbonic acid is about 6.4. The pKa of phenol is about 10. The bicarbonate was not strong enough to pull the proton off the phenol in any meaningful concentration. I wasted two days watching the reaction sit there, convinced my substrate was unreactive, before realizing the base was simply the wrong choice. Switching to sodium hydroxide or potassium carbonate fixed it immediately. The model predicted this perfectly. I just did not trust the numbers before I checked them. Another thing people do not absorb quickly enough is that pKa values are solvent-dependent. The numbers you see in tables are almost always measured in water. Once you move to dimethyl sulfoxide, acetonitrile, or methanol, the ordering of acidity can shift dramatically. Dimethyl sulfoxide in particular makes even weak C-H acids look much more acidic because it stabilizes the resulting anion far better than water does. If you are running reactions in non-aqueous solvents, your pKa table is a starting point, not the final word. You need to account for solvation effects or find experimental data measured in the solvent you are actually using.
Get the Full Details
Conjugate Pairs And The Myth Of Isolation
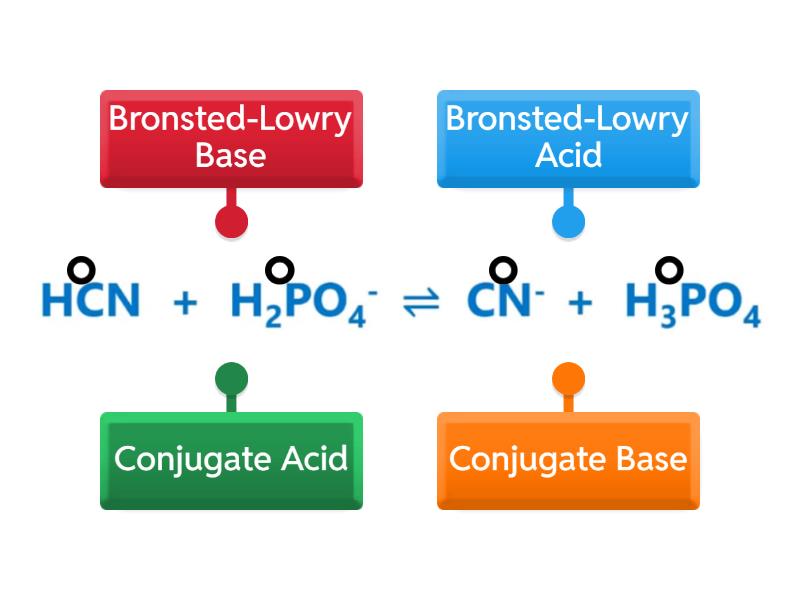
Every Brønsted acid comes with a conjugate base, and every Brønsted base comes with a conjugate acid. The strength of one dictates the weakness of the other in a reciprocal way. A strong acid has a weak conjugate base. A weak acid has a relatively stronger conjugate base. This relationship is not a suggestion. It is a mathematical consequence of the equilibrium expression. The mistake beginners make is treating conjugate pairs as if they exist in isolation. They do not. In solution, all four species are present simultaneously, and their concentrations shift according to pH. The Henderson-Hasselbalch equation describes this relationship, but even that equation breaks down when concentrations get extreme or when activity coefficients matter. In dilute aqueous solutions, Henderson-Hasselbalch works fine. In concentrated electrolyte solutions or non-aqueous media, you are better off using a speciation calculator or measuring the actual pH and adjusting from there. Here is a practical rule I rely on: if you need to maintain a stable pH around a specific value, choose a weak acid whose pKa is within about one unit of your target pH. That gives you a buffer capacity that is actually useful. Outside that range, the buffer becomes mostly ineffective, and adding more of it does not help. I once tried to buffer a reaction at pH 9 using an acetate system with a pKa of 4.76. The buffer capacity was essentially zero at that pH, and the pH drifted all over the place during the reaction. Switching to a borate or carbonate system at the appropriate pKa stabilized everything within minutes.
Edge Cases Where The Bronsted Model Stumbles
The Brønsted model assumes that proton transfer is the only relevant process. In many systems, this assumption holds. In others, it does not. Here are the cases where the model gives you the wrong answer if you apply it blindly. Autoprotolysis of the solvent matters when you are working in very dilute acids or bases. Water self-ionizes to about 10 to the minus 7 M in both H3O+ and OH-. Below that concentration, the solvent's own ions contribute significantly to the measured pH. This is why extremely dilute strong acid solutions do not have the pH you would calculate from simple stoichiometry. You have to solve the full equilibrium including water's contribution. The calculation adds about three lines to your algebra but prevents a significant error in the result. Polyprotic acids create overlapping equilibria that the simple Brønsted picture does not capture cleanly. Phosphoric acid has three pKa values spaced well enough apart that you can treat each deprotonation step separately in most practical situations. Carbonic acid, on the other hand, has pKa values that are close enough that the intermediate species dominates over a wide pH range, and assuming sequential independent deprotonations introduces noticeable error. When I worked on a water treatment project involving carbonate systems, I had to switch from a hand-calculation approach to a full speciation model because the approximations accumulated into errors that mattered at the scale we were operating at.
Amphoteric species are another category where the model works but requires careful bookkeeping. Water, bicarbonate, and amino acids can act as either acids or bases depending on what they are paired with. Bicarbonate is the classic example. It accepts a proton to form carbonic acid and donates a proton to form carbonate. Which behavior dominates depends entirely on the pH of the solution relative to the relevant pKa values. If the pH is below 6.4, bicarbonate acts primarily as a base. If the pH is above 10.3, it acts primarily as an acid. Between those values, it sits in the middle, and both equilibria are active.

Why This Framework Still Matters
The Brønsted model persists because it maps directly onto observable behavior. You can measure pH. You can predict direction of reaction. You can choose the right base for a deprotonation. You can design a buffer. You can explain why certain reagents work and others do not. It is not the most fundamental description of acid-base chemistry, but it is the most useful one for everyday laboratory work. Lewis theory is deeper. It explains reactions that involve no proton transfer at all, like the addition of ammonia to boron trifluoride. It unifies metal coordination chemistry with acid-base chemistry. But for organic synthesis, biochemistry, environmental chemistry, and most analytical work, the Brønsted framework is faster and sufficient. You reach for Lewis when Brønsted cannot describe the phenomenon you are observing. That happens, but not as often as textbooks imply. The practical takeaway is to learn the Brønsted model thoroughly first. Master pKa comparisons, conjugate pair relationships, buffer design, and the limitations of the Henderson-Hasselbalch equation. Then learn where it breaks. The gap between knowing the model and knowing its failure modes is where actual competence lives.