Why Everyone Messes Up Nucleophilic Acyl Substitution
Nucleophilic acyl substitution is the single most important reaction type in organic chemistry, and it trips people up constantly. The mechanism isn't actually that complicated, but most textbooks present it in a way that makes you memorize four separate reactions instead of understanding one unified process. I spent three semesters watching students fail the same problem over and over because they couldn't see the pattern. Carboxylic acids have that familiar carboxyl group: a carbonyl carbon bonded to a hydroxyl group. What makes this functional group behave the way it does is that the hydroxyl oxygen can stabilize the carbonyl through resonance, but not enough to make it inert. The carbon is electrophilic, yes, but it is also significantly less reactive than a ketone or aldehyde carbonyl. That difference matters enormously when you are trying to predict whether a nucleophile will attack the carbonyl or do something else entirely.
Understanding Carboxylic Acids And Acid Derivatives Through Reactivity
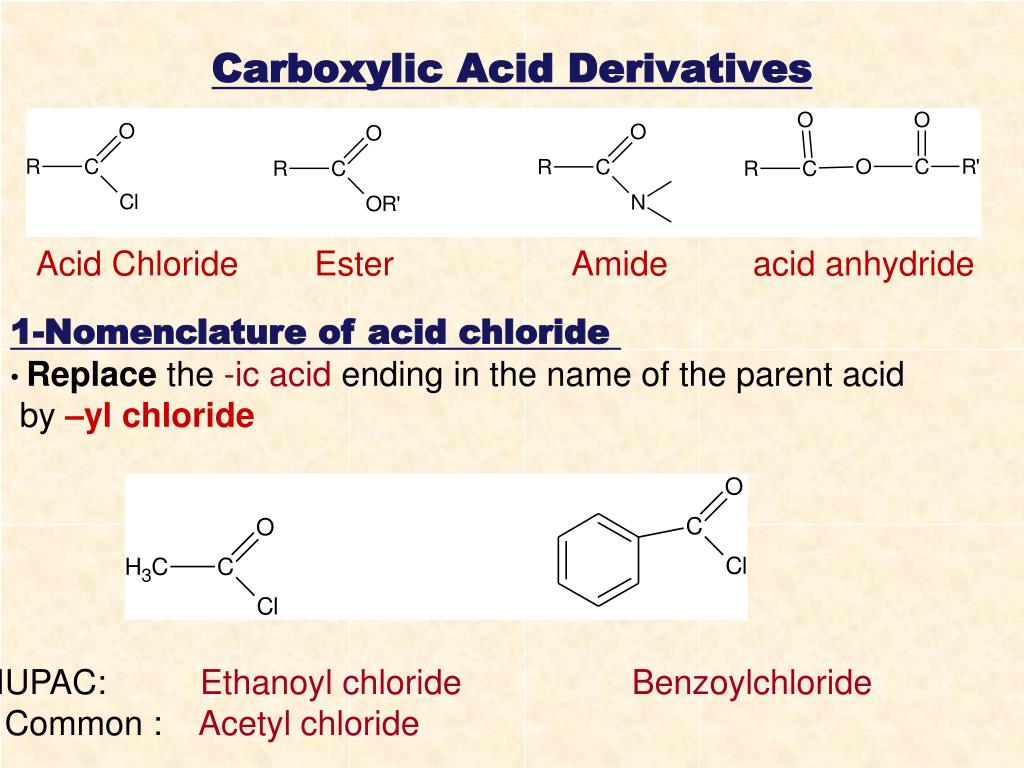
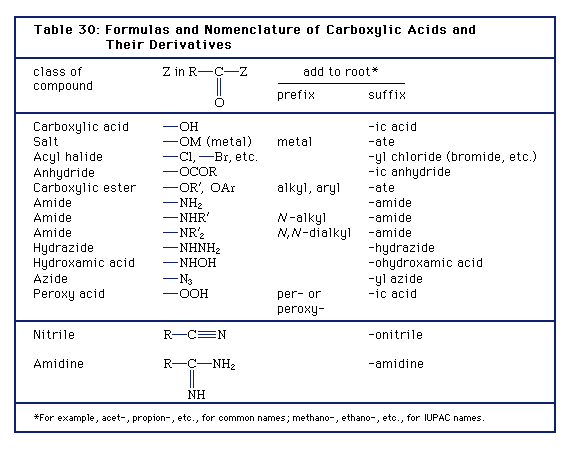
The acid derivatives exist on a reactivity ladder, and knowing where each one sits is more useful than any individual reaction mechanism. At the top of the ladder, meaning the most reactive, you have acyl chlorides. Below those are acid anhydrides. Esters sit in the middle, and amides are at the bottom as the least reactive derivatives. This ordering is not arbitrary. It comes down to two factors: the quality of the leaving group and the degree of resonance stabilization around the carbonyl. Chloride is a weak base and an excellent leaving group. The chlorine atom also does not donate electron density into the carbonyl through resonance to any meaningful degree, so the carbonyl carbon stays highly electrophilic. Amides are the opposite. The nitrogen lone pair participates strongly in resonance with the carbonyl, which dramatically reduces the electrophilicity of the carbon and makes the amino group a terrible leaving group under normal conditions. You can convert a more reactive derivative into a less reactive one without much difficulty. Going from an acyl chloride to an ester happens readily with an alcohol and a weak base like pyridine. Going from an ester to an amide requires heating with ammonia or an amine, sometimes under pressure. But you cannot simply reverse the ladder. Trying to turn an amide back into an acyl chloride with thionyl chloride will not work the way you might expect. The amide nitrogen gets protonated or alkylated first, and the carbonyl survives mostly unchanged.
The Mechanism That Actually Works in Practice
Every nucleophilic acyl substitution follows the same three-step pathway, regardless of which derivative you start with. First, the nucleophile attacks the carbonyl carbon and forms a tetrahedral intermediate. Second, the carbonyl reforms as the intermediate collapses. Third, the leaving group departs. That is the entire sequence. It sounds simple because it is simple, but students frequently forget that the tetrahedral intermediate is a real, identifiable species and not just a conceptual placeholder. When I was working in a synthesis lab, a colleague once tried to run a Schotten-Baumann esterification on a substrate bearing a free amine group. The reaction proceeded, but the product was a mess of N-acylated and O-acylated species because the amine competed as a nucleophile faster than the alcohol did. The workaround was straightforward: protect the amine as a Boc carbamate before running the esterification, then remove the protecting group afterward with trifluoroacetic acid. This added two extra steps to the sequence but cut the purification time from overnight column chromatography to a single recrystallization. Time saved: roughly two hours of hands-on work versus forty-five minutes of actual reaction time. The tetrahedral intermediate is also the reason why certain reactions proceed with retention of configuration at a chiral center adjacent to the carbonyl, while others lead to racemization. If the leaving group is poor and the intermediate lives long enough, bond rotation can occur before collapse, scrambling stereochemistry. This is a well-known problem when you are trying to convert a chiral carboxylic acid derivative without losing optical purity. Using DMAP as a catalyst in esterification reactions can sometimes accelerate the collapse of the intermediate faster than rotation occurs, which helps preserve stereochemistry, but it is not a guarantee. The substrate structure matters more than the catalyst.
Get the Full Details

Common Pitfalls That Do Not Make It Into Textbooks
One thing that beginners consistently miss is the behavior of carboxylic acids themselves in substitution reactions. You cannot directly convert a carboxylic acid into an ester or an amide by simply adding an alcohol or amine and waiting. The acid will protonate the amine nucleophile, killing its reactivity, and the hydroxyl group is a poor leaving group. You need either a strong acid catalyst for Fischer esterification, which operates through a different mechanistic pathway involving protonated intermediates, or you need to activate the acid first by converting it to an acyl chloride or an anhydride. Another overlooked detail is how steric hindrance affects reactivity across the derivative series. An sterically hindered acid chloride, like pivaloyl chloride, will react sluggishly with a bulky nucleophile even though chloride is theoretically an excellent leaving group. The initial nucleophilic attack becomes the rate-determining step, and the geometry around the carbonyl carbon simply blocks approach. I encountered this specifically when attempting to couple 2,2-dimethylpropanoic acid derivative with a substituted aniline. The reaction stalled completely at room temperature and required heating to reflux in toluene with a Dean-Stark trap to drive off the HCl byproduct. Even then, the yield was only around sixty percent after twelve hours. Switching to the corresponding anhydride instead of the acid chloride improved the yield to nearly eighty-five percent because the anhydride is less sterically encumbered at the carbonyl carbon being attacked. Esters also have a reputation for being stable, but that stability is conditional. A methyl ester will survive basic aqueous conditions reasonably well if the temperature is kept low, but once you heat it in aqueous hydroxide, saponification proceeds rapidly and irreversibly. The carboxylate product cannot be re-esterified under those basic conditions because the equilibrium lies entirely toward the deprotonated acid. This is why ester protection strategies in multi-step synthesis require acidic conditions for removal, not basic ones, if you need to regenerate the carboxylic acid later in the sequence.
What This Approach Does Not Handle Well
The reactivity ladder model works exceptionally well for simple aliphatic and aromatic substrates. It breaks down when you introduce electron-rich heterocycles adjacent to the carbonyl, conjugated systems with extended pi networks, or substrates containing multiple competing functional groups that interact through intramolecular hydrogen bonding. In those cases, the textbook reactivity order becomes unreliable, and you end up relying more on empirical testing than on prediction. There is also the issue of side reactions that the model does not account for. Acyl chlorides can undergo alpha-halogenation under certain conditions if trace amounts of enolizable protons are present. Acid anhydrides can promote Friedel-Crafts acylation on aromatic substrates present in the reaction mixture, which is usually undesirable unless explicitly intended. Esters can undergo Claisen condensation if a strong base and alpha protons are both available. These are not minor concerns when you are working at scale or when purification is difficult. If your substrate contains a free hydroxyl group alongside a carboxylic acid, you will likely get lactonization rather than the intermolecular product you intended, especially if the resulting ring would be five or six members. This happens automatically under acidic conditions and cannot be prevented simply by controlling the stoichiometry. You need to protect the hydroxyl group first or work under conditions that suppress intramolecular cyclization, which usually means high dilution. High dilution is effective but impractical for reactions larger than a few grams.
Practical Guidance for Working With These Compounds
When you need to prepare an ester from a carboxylic acid and the substrate is sensitive to acid, use DCC coupling with a catalytic amount of DMAP rather than Fischer esterification. DCC activates the carboxylic acid through an O-acylisourea intermediate, and DMAP accelerates the nucleophilic attack by the alcohol. The byproduct, dicyclohexylurea, precipitates out of most organic solvents and can be removed by filtration. This method typically runs at room temperature in two to four hours and gives yields in the eighties to low nineties for unhindered substrates. For hindered substrates, expect the reaction to take longer and the yield to drop into the sixties. If you need an amide and the carboxylic acid is expensive or, convert it to the acyl chloride using oxalyl chloride with a catalytic amount of DMF, then add the amine directly without isolating the acid chloride. The DMF catalyst forms the Vilsmeier-Haucard reagent in situ, which activates the carboxylic acid far more efficiently than thionyl chloride alone. The oxalyl chloride byproducts are gases and leave the reaction mixture cleanly. This avoids the isolation step, which is often where material is lost, and typically improves overall yield by fifteen to twenty percent compared to the thionyl chloride route followed by separate amide formation. For large-scale work where acyl chlorides are problematic due to moisture sensitivity and HCl evolution, acid anhydrides remain the practical choice. Acetic anhydride is inexpensive and easy to handle. The acetylated byproduct is water-soluble and removes easily during aqueous workup. The main drawback is that anhydrides can acetylate other nucleophilic sites on your molecule, so chemoselectivity can be an issue if your substrate contains free hydroxyl or amine groups elsewhere. In those situations, you need to rely on differential reactivity or protection strategies, which adds complexity back into the synthesis.

Amide bonds are remarkably stable once formed, which is why they are useful as protecting groups and permanent structural features in drug molecules. The stability is also why reducing an amide to an amine requires a strong reducing agent like lithium aluminum hydride. Borane reduces amides selectively in the presence of esters, which is a useful distinction if you need to differentiate between the two. Sodium borohydride will not reduce either group under normal conditions, which makes it safe to use in molecules containing both functional groups when you are targeting a different reducible site elsewhere. The reactivity relationships among Carboxylic Acids And Acid Derivatives are consistent enough that you can navigate most synthetic problems with the ladder model as your starting point. It will not solve every edge case, and you will encounter substrates where the predictions fail. That is normal. The useful skill is recognizing when the model is likely to break down and having a practical fallback ready, whether that means switching to a different activating agent, changing the solvent system, or going through a protection-deprotection sequence. Most of the time, the difference between a good reaction and a failed one comes down to choosing the right derivative for the job rather than forcing a reaction that the substrate is not suited for.