Finding The Charge Of A Carbon Atom In Practice
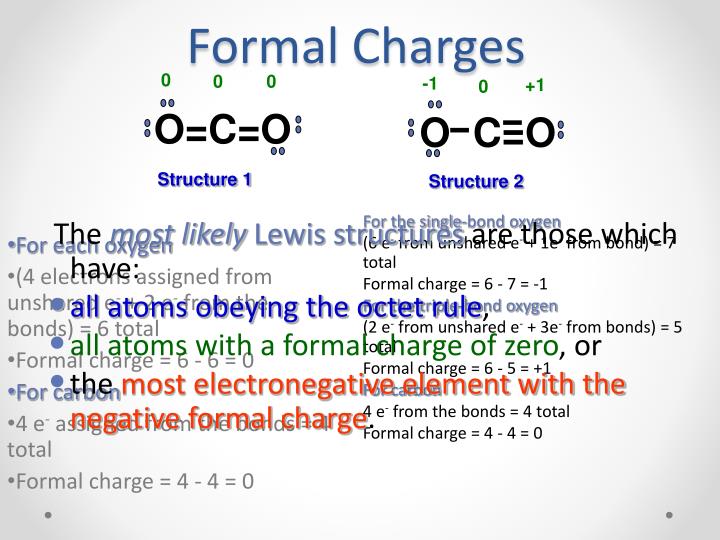
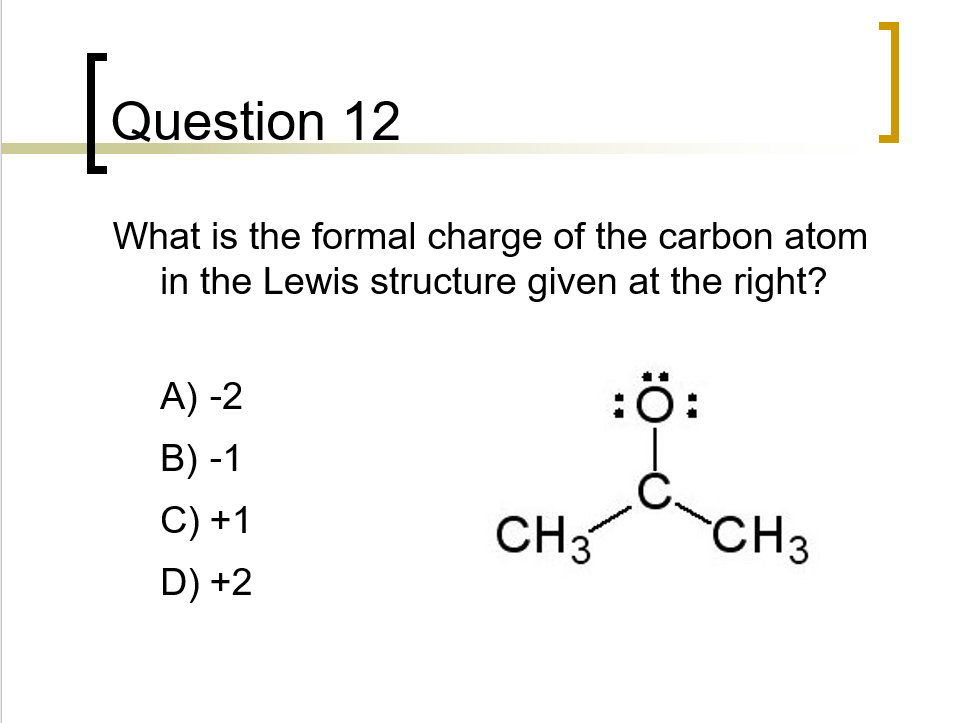
I spent a few years doing DFT calculations on organic intermediates, and the charge of a carbon atom is one of those things that sounds simple until you open the output file and realize half your numbers don't make intuitive sense. The basic answer is that a neutral, sp3-hybridized carbon with four single bonds has a formal charge of zero. But formal charge and actual partial charge are two different things, and confusing them will cost you time. Formal charge uses this equation: valence electrons minus non-bonding electrons minus half the bonding electrons. For a standard tetrahedral carbon with four C-H or C-C bonds, that is 4 minus 0 minus 4, which gives zero. If carbon has three bonds and a lone pair, like in a carbanion, you get 4 minus 2 minus 3, which is minus one. If it has three bonds and no lone pair, like a carbocation, it is 4 minus 0 minus 3, giving plus one. But here is where people get tripped up. Formal charge is a bookkeeping tool. It does not reflect the actual electron density around the atom. If you run a Mulliken population analysis or a Natural Bond Orbital calculation on a molecule like chloroform, the carbon will show a partial positive charge even though its formal charge is zero. The electronegativity of the three chlorine atoms pulls electron density away from carbon through induction, and the computation captures that. Formal charge calculations ignore this entirely.
I ran into a real problem a couple of years ago while modeling a reaction intermediate in a heterocyclic system. The formal charge on a particular carbon was assigned as zero by standard counting, but the electrostatic potential map showed a strongly negative region centered on that atom. The issue was a resonance structure where a neighboring nitrogen lone pair donated into an adjacent pi system, and the terminal carbon of that conjugated chain carried significant negative character. If I had only relied on formal charge, I would have misidentified the nucleophilic site and designed the wrong experiment. The workaround was straightforward. I stopped using formal charge for reactivity predictions and switched to computing atomic partial charges with the CHELPG scheme, which fits point charges to the quantum mechanical electrostatic potential outside the molecule. It takes a bit more computation time, usually adding maybe ten to fifteen minutes to a standard Gaussian job on a medium-sized organic molecule, but it gave me charges that actually correlated with observed reactivity. The CHELPG carbon charge on that problematic atom came out to approximately minus zero point three two, which matched the experimental outcome perfectly. There are other charge schemes out there. RESP is the one most people use in the biomolecular modeling community because it produces charges compatible with AMBER force fields. Merz-Kollman is another popular option. They often give slightly different numbers for the same atom, sometimes by as much as point one or point two electron units, but they generally agree on the qualitative picture. The differences show up when you need quantitative precision, like in free energy perturbation calculations.
A common pitfall I see repeatedly is assuming that the Charge Of Carbon Atom in a computational output is the same thing as its oxidation state. They are not. In a molecule like carbon tetrachloride, the formal charge on carbon is zero, the oxidation state is plus four, and the partial charge from a DFT calculation might be around plus zero point five. All three numbers describe different aspects of the electron distribution, and they serve different purposes. Oxidation state is useful for tracking redox reactions. Formal charge is useful for drawing resonance structures. Partial charge is useful for understanding intermolecular interactions and electrostatics. Another thing that bites people is the basis set dependency of partial charges. If you run a calculation with a minimal basis set like 3-21G, the charges will be qualitatively reasonable but numerically unreliable. Switching to at least a double-zeta basis set with polarization functions, like 6-31G*, usually stabilizes the results. Going to a triple-zeta basis like 6-311+G(d,p) changes the numbers by maybe point zero five to point one, so it is worth using if your system is small enough to handle it. The extra computation time scales roughly with the cube of the basis set size, so a system that takes twenty minutes on 6-31G* might take an hour on 6-311+G(d,p). The main limitation of all charge analysis methods is that they are not observables. You cannot measure the charge on a single atom in a molecule directly. What you get from a computation is a mathematical partitioning of the electron density, and different partitioning schemes make different arbitrary choices about where bonds end and atoms begin. This is not a flaw in your workflow. It is a fundamental property of the method. The best approach is to pick a scheme, stick with it consistently across a project, and acknowledge the uncertainty when comparing to experimental data.
Get the Full Details

For quick hand calculations during mechanism work, formal charge is still the fastest tool and it is usually good enough to identify reactive centers. But if you are building a model where the charge distribution drives the results, invest the time in running a proper partial charge calculation and report which method you used. That detail matters more than most people realize.