Understanding How Collisions Drive Chemical Reactions
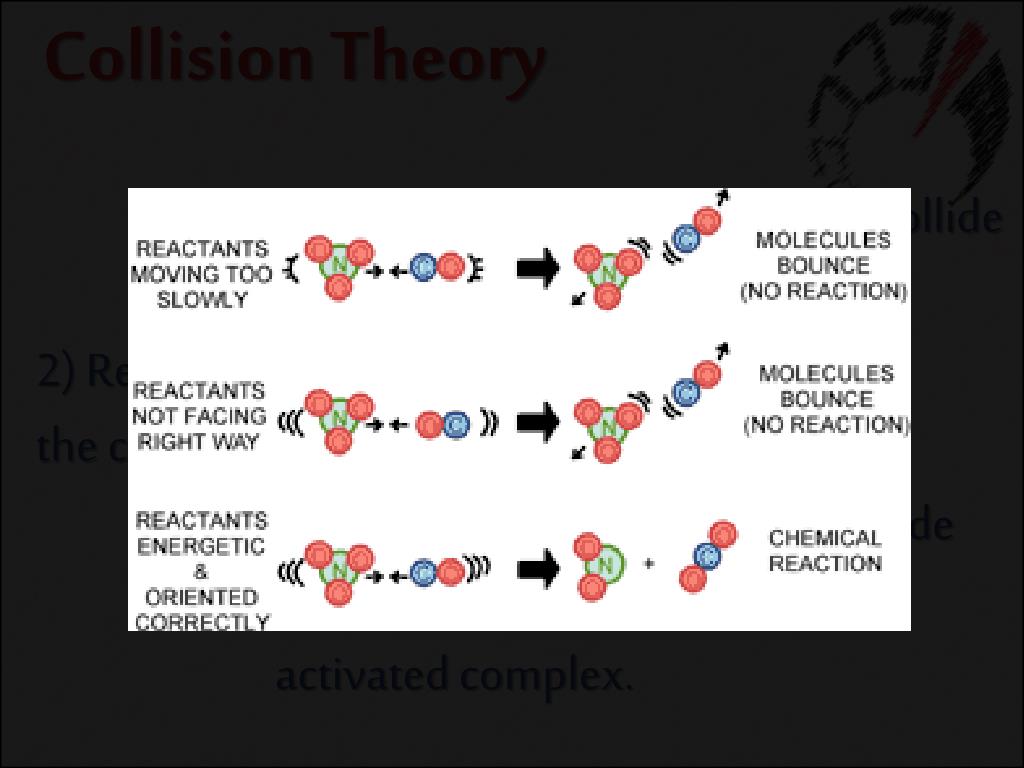
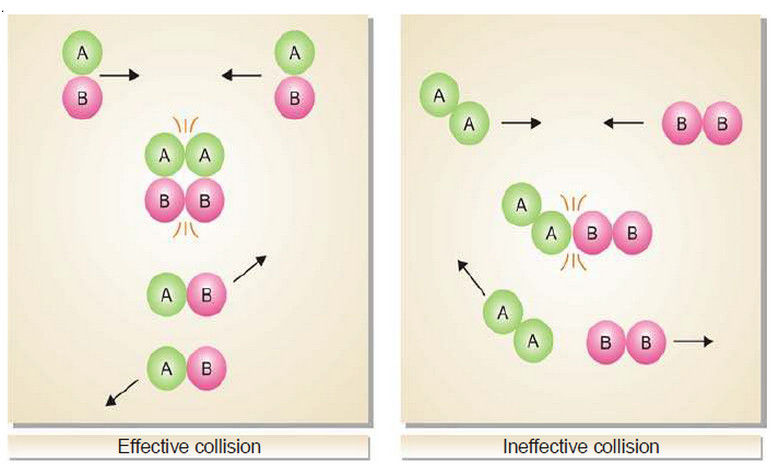
Most people learn collision theory as a set of neat bullet points: particles must collide, they need enough energy, and the orientation matters. The reality is messier than that. When you actually work with kinetic data in a lab or try to model reaction rates computationally, you quickly realize that the textbook version leaves out a lot of the friction. The collision theory impact for a chemical reaction is real and measurable, but applying it correctly requires understanding where the model works and where it quietly fails. At its simplest, collision theory says a reaction happens when reactant particles hit each other with enough energy to overcome the activation barrier and in the right geometric alignment. The rate of reaction is proportional to the collision frequency multiplied by two correction factors: the fraction of collisions that have energy above Ea, and the steric factor p, which accounts for orientation requirements. The full expression looks like this: rate = Z × e^(-Ea/RT) × p, where Z is the collision frequency from kinetic molecular theory. The exponential term comes from the Maxwell-Boltzmann distribution. Only a certain fraction of molecules at any given temperature have kinetic energy exceeding the activation energy. Raise the temperature, and that fraction grows exponentially, not linearly. That is why a 10-degree increase often doubles reaction rates for reactions with activation energies in the 50 to 80 kJ/mol range. It is not a small effect. It is a steep climb.
I spent several months trying to reconcile experimental rate data for an ester hydrolysis reaction with the predictions from basic collision theory, and the numbers did not line up. The measured rate was roughly a third of what the collision frequency and Boltzmann factor predicted. The problem was the steric factor. Ester hydrolysis requires the water molecule to approach the carbonyl carbon from a specific angle, and the surrounding alkyl groups blocked a lot of the geometric space. Once I introduced an experimentally derived p value of about 0.33, the calculated and observed rates matched within experimental error. That is a detail you will not find emphasized in most introductory textbooks.
Practical Factors That Modify Reaction Rates
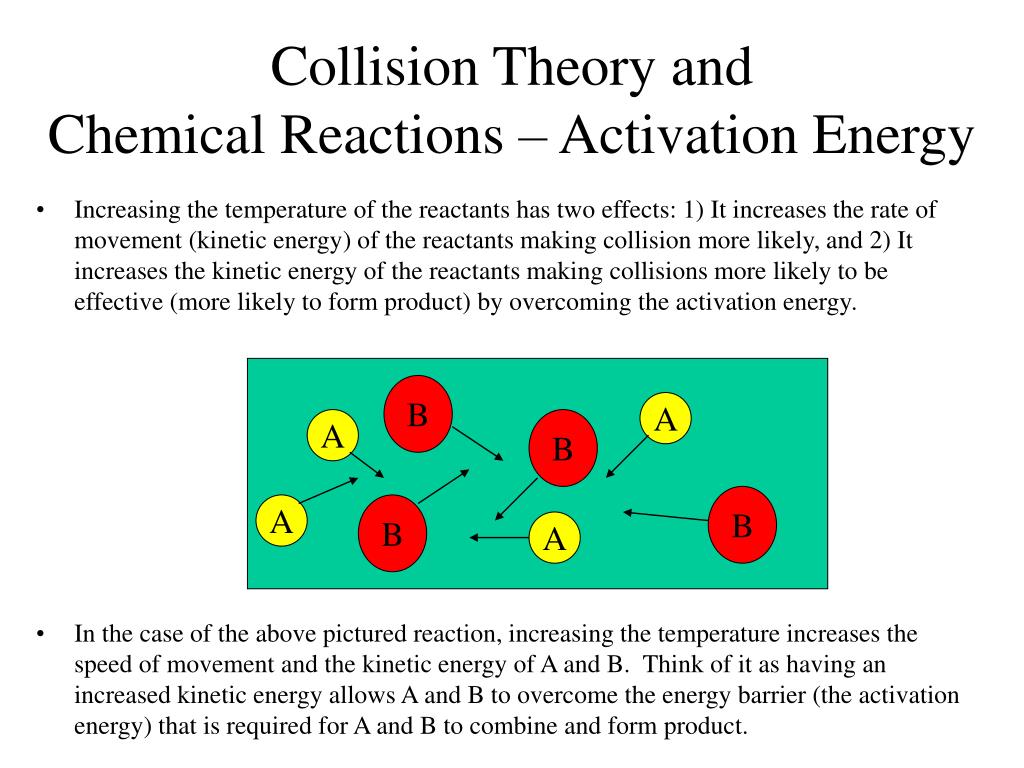
Temperature is the most straightforward variable. Increasing temperature does two things simultaneously: it increases the total number of collisions per second and, more importantly, it increases the proportion of those collisions that exceed the activation energy. The second effect dominates. You can calculate the exact change using the Arrhenius equation, but the takeaway is simple. Small temperature changes produce large rate changes for reactions with high activation energies. Concentration and pressure work by increasing collision frequency. In a gas phase reaction, doubling the pressure roughly doubles the collision frequency and typically doubles the rate for a second-order reaction. In solution, doubling concentration has a similar effect, but you have to account for solvent effects. Diffusion limits how fast molecules can reach each other in a liquid, and at high concentrations, intermolecular forces start interfering with the ideal behavior that collision theory assumes. I ran into this when working with a concentrated sulfuric acid system where activity coefficients deviated significantly from unity. Using molar concentration instead of activity gave rates that were off by a factor of four. Surface area matters for heterogeneous reactions. A solid reacting with a gas or liquid only reacts at its surface, so grinding a solid into a fine powder increases the available collision sites dramatically. This is why industrial processes often use catalysts in pellet or powder form rather than solid blocks. The collision frequency per unit volume increases, and the reaction proceeds faster. This is well understood and widely applied.
Get the Full Details
Catalysts lower the activation energy by providing an alternative reaction pathway. They do not change the collision frequency. They change the energy requirement. This means more of the existing collisions become effective. A catalyst can increase a reaction rate by orders of magnitude without changing temperature or concentration. The trade-off is that catalysts can be expensive, sensitive to poisoning, and sometimes difficult to separate from the product. I worked on a hydrogenation process where the palladium catalyst was temporarily deactivated by trace sulfur in the feedstock. The reaction rate dropped by about 60 percent over six hours. Switching to a sulfur-tolerant catalyst and adding a guard bed upstream resolved the issue, but the downtime cost far outweighed the price of the better catalyst.
Where Collision Theory Falls Apart
The simplest form of collision theory assumes hard sphere particles colliding in a vacuum. Real molecules are not hard spheres. They have electronic clouds, dipole moments, and complex shapes. The theory also assumes that every collision with sufficient energy results in reaction, which is clearly wrong. The steric factor can range from nearly 1 for simple atom recombination to less than 0.001 for complex organic reactions with strict geometric requirements. For reactions in solution, solvent effects are significant. Solvent molecules form shells around reactants, and breaking those interactions requires energy. The theory does not account for this. Transition state theory, which incorporates the concept of an activated complex and uses equilibrium statistical mechanics, handles solution phase reactions much better. If you are working with liquid-phase kinetics, switching from collision theory to transition state theory will give you more accurate predictions. Another limitation is that collision theory treats all molecules of a given species as identical. In reality, rotational and vibrational energy states matter. A molecule might have enough translational energy but be in a vibrational state that prevents bond formation. Modern computational chemistry methods, such as molecular dynamics simulations with ab initio potentials, capture these details but require significant computing resources. For quick estimates, collision theory is still useful. For accurate predictions, you need something more sophisticated.
The theory also breaks down for chain reactions and reactions involving radicals. The simple collision model does not account for the branching and termination steps that dominate radical mechanisms. Polymerization kinetics, combustion, and atmospheric chemistry are examples where collision theory gives at best a rough qualitative picture. Rate equations derived from steady-state approximations are the standard approach there.

Using the Theory in Practice
If you need to estimate how a change in conditions will affect a reaction rate, collision theory gives you a useful framework. Calculate the collision frequency using the kinetic theory expression Z = n²(8kT/), where n is the number density, is the collision cross-section, k is Boltzmann's constant, T is temperature, and is the reduced mass. Multiply by the Boltzmann factor and an estimated steric factor. This approach works reasonably well for gas-phase bimolecular reactions between small molecules. For anything more complex, measure the rate constant at a few temperatures and use the Arrhenius plot to extract Ea and the pre-exponential factor A empirically. This is faster and more reliable than trying to calculate everything from first principles. Most published kinetic data sets include Arrhenius parameters precisely because the theoretical calculation is unreliable for real systems. The main practical insight is this: collision theory explains why the factors it identifies affect rates, but it should not be treated as a predictive tool for complex reactions. It is a conceptual framework, not a calculation method. Use it to understand trends, not to predict exact rates. When accuracy matters, rely on experimental data or transition state theory calculations.