Working With Chemical Bonds In Practice
I spent about three years troubleshooting protein crystallization problems before I really understood how covalent And Noncovalent Interactions actually behave under experimental conditions. The textbook definitions are fine, but they don't prepare you for the moments when your hydrogen bonding network collapses during a pH shift or your disulfide bridges rearrange in ways that weren't supposed to happen. Here's what actually works. Start with the covalent framework. These are the strong bonds where atoms share electron pairs. Carbon-carbon bonds, carbon-nitrogen bonds, the peptide backbone connections. They're relatively stable once formed. Breaking them requires either significant energy input or specific enzymatic catalysis. In my work with peptide synthesis, I learned that protecting group strategy matters more than most people realize. A Boc group on a lysine side chain behaves completely differently from an Fmoc group when you're trying to form a selective amide bond later in the sequence.
Understanding the Full Scope of Covalent And Noncovalent Interactions
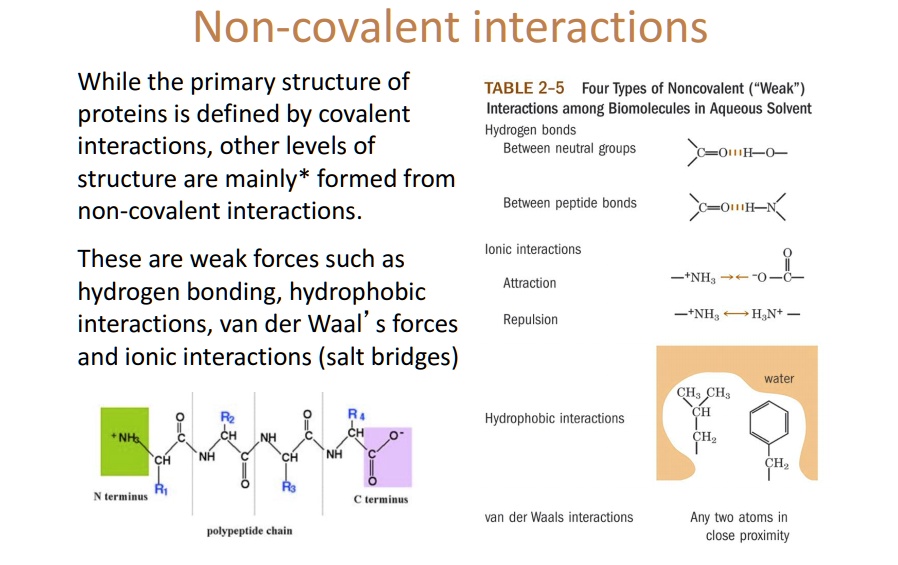
The noncovalent stuff is where things get complicated. Hydrogen bonds, ionic interactions, van der Waals forces, pi-stacking, hydrophobic effects. Individually weak, collectively decisive. A single hydrogen bond might contribute only 1 to 5 kcal/mol of stability. But you're rarely dealing with just one. In a protein active site, you might have six or eight hydrogen bonds oriented in specific geometries, plus some ionic interactions from charged residues, plus van der Waals contacts from the packing core. I remember working on an enzyme engineering project where we needed to improve thermal stability. We identified a surface salt bridge that looked promising in the crystal structure. When we mutated the residues to strengthen that ionic interaction, the protein actually became less stable above 45 degrees Celsius. The issue was that the rigid electrostatic attraction reduced conformational flexibility needed for proper folding kinetics at higher temperatures. We ended up weakening that interaction instead and gained about 8 degrees of melting temperature by allowing more adaptive packing during the folding process.
Measuring And Quantifying These Forces
Differential scanning calorimetry gives you thermodynamic parameters, but it's expensive and requires relatively pure samples. Isothermal titration calorimetry works well for binding studies where you can observe the heat change as one molecule interacts with another. Surface plasmon resonance lets you measure association and dissociation rates, which tells you something about the strength and lifetime of the interaction without needing to know the exact thermodynamics. For quick assessments, circular dichroism spectroscopy will show you secondary structure changes as you vary temperature or denaturant concentration. The trick is knowing what signals to look for. Alpha helices show characteristic double minima at 208 and 222 nanometers. Beta sheets show a single minimum around 215 nanometers. When you see those peaks shift or change intensity, you're watching the balance between covalent and noncovalent interactions being disrupted. I once spent two weeks trying to figure out why my protein sample kept aggregating during purification. The SEC traces showed a broad shoulder at the void volume, indicating large species formation. The concentration was fine, the buffer was appropriate, nothing obvious. Eventually I ran a small-angle X-ray scattering experiment and realized the protein had a low-affinity dimer interface that only became significant above 10 micromolar concentration. Dropping the concentration during later purification steps solved the problem completely. Sometimes the noncovalent interactions are there, just below the threshold where they normally cause trouble.
Get the Full Details
Common Approaches For Manipulating Bond Strengths
If you need to strengthen a specific interaction, consider the geometry. Hydrogen bonds are highly direction-dependent. A donor-acceptor distance around 2.8 angstroms with a bond angle near 180 degrees is optimal. Deviate from that geometry and the interaction energy drops significantly. When redesigning binding interfaces, maintaining proper hydrogen bond geometry often matters more than simply adding more potential interaction partners. For covalent modifications, crosslinking reagents like DSS or BS3 introduce stable links between proximal amino groups. These can lock protein complexes together or stabilize specific conformations. The caveat is that crosslinking changes the system. Added mass, potential steric interference, possible perturbation of the very interactions you're trying to study. Use it sparingly and validate that your modified system still behaves like the native one. Disulfide bonds present an interesting case. They're covalent but can be reversible under reducing conditions. I've used engineered disulfides to trap protein intermediates during mechanistic studies. The reduction step had to be carefully controlled because over-reduction led to complete unfolding in some cases. Adding 5 millimolar dithiothreitol gradually over thirty minutes worked better than a single large addition. The kinetics matter even when breaking bonds.
What Fails And When To Expect Problems
Computational prediction of interaction strengths remains unreliable outside narrow parameter spaces. Molecular dynamics simulations can give reasonable estimates for hydrogen bonding networks in well-folded proteins, but they struggle with conformational sampling and force field accuracy. AutoDock and similar tools might score a binding mode correctly but assign interaction energies that are off by 30 percent or more. Treat computational predictions as hypotheses requiring experimental validation, not as measurements. Surface plasmon resonance data interpretation has its own pitfalls. Mass transport limitation can make a fast-binding interaction appear slower than it is. If your analyte concentration is high relative to the flow rate through the chip flow cell, the binding event becomes limited by diffusion rather than by the actual on-rate. Running experiments at multiple flow rates and checking for consistency helps catch this issue. Another failure mode involves ignoring solvent effects. Water molecules participate in hydrogen bonding networks constantly. A buried water molecule in a protein active site might be integral to the catalytic mechanism, bridging substrate and residue through coordinated hydrogen bonds. Removing that water in a computational model can make the predicted binding affinity look completely different from reality. Structural water is structural, not solvent, when it's playing a functional role.
The hydrophobic effect doesn't follow simple rules about nonpolar surface area. It depends on temperature, pressure, and the specific atomic composition of the interface. The transfer free energy of a methylene group from water to a nonpolar environment isn't constant across different proteins or different locations within the same protein. Parameterize carefully if you're building quantitative models.
Practical Tips From Working With These Systems Daily
Keep your samples away from unnecessary surface area. Protein adsorption to plastic or glass is a noncovalent interaction problem that affects concentration measurements, activity assays, and crystallization trials. Silanized glassware or treatment with blocking agents like BSA or casein can reduce losses. The amount lost to surface binding scales with concentration in unpredictable ways, making accurate quantification difficult when working with dilute samples. When screening buffer conditions, vary pH, ionic strength, and additive concentration systematically rather than randomly. pH affects protonation states, which changes both covalent geometry through altered bond angles and noncovalent interactions through charge state changes. Ionic strength screens electrostatic interactions according to Debye-Huckel theory, but the exact screening length depends on ion valency and concentration. Monovalent salts at 150 millimolar typically screen charges within about 8 angstroms in aqueous solution at room temperature. Temperature control matters more than people sometimes acknowledge. Both enthalpy and entropy contributions to binding free energy change with temperature. A favorable enthalpy-driven interaction might become unfavorable at higher temperatures if the entropic term dominates. Van't Hoff analysis of binding constants measured at multiple temperatures can reveal whether an interaction is enthalpy or entropy driven, which guides rational optimization strategies. The crossover point where enthalpy and entropy contributions balance is often where small perturbations have the largest effects on binding affinity.