How Exothermic Reactions Actually Work in Practice
Most people learn the textbook definition in high school chemistry and never really think about it again, but if you work with these reactions outside a controlled lab environment, you quickly learn that the simple "heat is released" description doesn't capture half the problems that can go wrong. When I first Define Exothermic Chemical Reaction for a batch process I was troubleshooting, I was working with a nitrated intermediate in a solvent system that was supposed to be stable at reflux temperature. It wasn't. The reaction ran hot enough to exceed the designed operating window by about forty degrees Celsius within twelve minutes, and we lost a full reactor charge to thermal runaway before I could get the cooling system responding fast enough. At its core, an exothermic chemical reaction is one where the total enthalpy of the products is lower than the total enthalpy of the reactants. The difference has to go somewhere, and in most practical cases it leaves as heat energy that raises the temperature of the reaction mixture and its surroundings. That basic fact sounds harmless until you realize that most exothermic reactions speed up as temperature increases, and faster reactions produce heat faster, which speeds them up even more. This is what the literature calls positive feedback, and it is the single most important concept to understand when dealing with anything beyond textbook examples.
Define Exothermic Chemical Reaction in Technical Terms
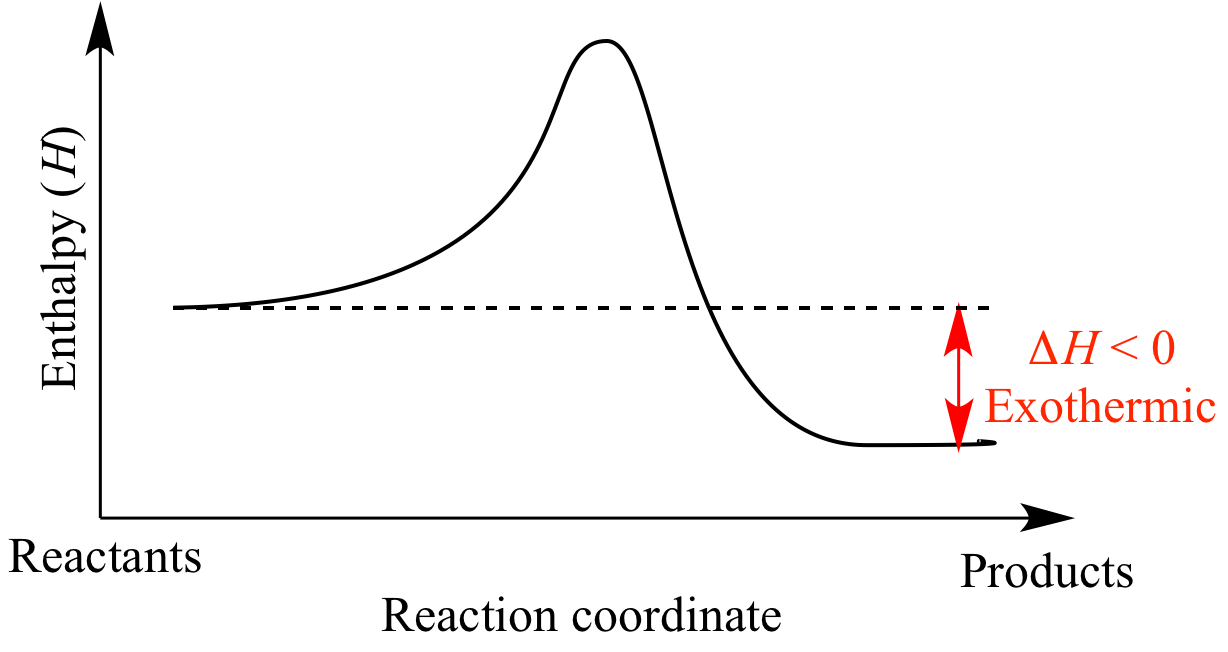
The formal definition involves the change in enthalpy, Delta H, being negative. The bond energies of the products exceed the bond energies of the reactants, and that excess energy manifests primarily as kinetic energy of the molecules in the system. In practical terms, this means you need to manage that heat deliberately or things get out of hand very quickly. I spent months working on a continuous flow process where an exothermic esterification had to be kept under tight thermal control. The problem was that the reaction heat was so significant relative to the volume of the mixture that even small fluctuations in feed rate caused temperature spikes that altered the product distribution. We ended up designing a system with a static mixer and a segmented cooling zone that kept the reaction within two degrees of the setpoint across the entire residence time. Without that level of control, the selectivity dropped from about ninety-three percent to something barely acceptable for commercial use.
What the Textbooks Don't Tell You
One thing beginners consistently miss is that not all the energy comes out as usable heat. A significant portion can go into phase changes, especially if you are running a reaction at or near the boiling point of your solvent. I once spent three days trying to figure out why our calorimetry data didn't match the theoretical Delta H value for a hydrogenation reaction, only to realize that roughly thirty percent of the released energy was being consumed as latent heat of vaporization of the solvent. The actual temperature rise was much smaller than the stoichiometry suggested it should be, which initially looked like the reaction was sluggish when it was just boiling away some of its own energy. Another counter-intuitive point is that some exothermic reactions become less favorable as temperature rises, even though they are technically still exothermic. Le Chatelier's principle applies here in a way that people don't always expect. For a highly exothermic equilibrium process, raising the temperature shifts the equilibrium back toward the reactants. So there is a genuine tradeoff: higher temperatures increase the reaction rate but decrease the equilibrium conversion. In practice, this often means running these reactions at the lowest temperature that still gives you an acceptable rate, then letting the exotherm itself do some of the heating work once you get going. The workaround I used for the esterification issue mentioned earlier was basically running the reaction semi-batch, feeding one reagent slowly enough that the heat generation stayed within the removal capacity of the cooling system. It cut throughput by about forty percent compared to the original batch design, but it eliminated the thermal runaway risk entirely and gave consistent product quality. Sometimes you accept lower productivity because losing a reactor charge and shutting down for a week costs a lot more than running slower.
Get the Full Details

Common Pitfalls and Where These Reactions Fail
The biggest practical problem is heat removal capacity. Every exothermic reaction generates heat at a certain rate, and your cooling system has a maximum rate at which it can remove that heat. If the generation rate exceeds the removal rate, temperature climbs, rate increases, generation climbs further, and you end up with a runaway scenario that can lead to vessel rupture or a release of hazardous materials. This is not theoretical. I have seen multiple incidents in pilot plants where operators underestimated the adiabatic temperature rise and learned the hard way that a jacket cooling water supply can only remove so many kilojoules per hour regardless of how hot the reaction mixture gets. A related issue is that the heat transfer coefficient changes during a reaction. As viscosity increases with conversion, as solids precipitate out, or as the mixture becomes heterogeneous, your effective heat transfer drops. A reaction that was comfortably under control at the start can become impossible to cool as it progresses. We saw this with a polymerization reaction where the viscosity increased by a factor of five hundred over the course of the run. The cooling jacket was adequate at the beginning and completely inadequate by the end, so we switched to internal coils and reduced the initial charge size by half to buy ourselves more thermal margin during the high-viscosity phase. If you are working with highly exothermic chemistry at scale, the absolute best thing you can do is run a differential scanning calorimetry test before committing to any new process. It takes a few hours and costs maybe two hundred dollars in sample preparation time, but it will tell you the onset temperature, the adiabatic temperature rise, and the heat release rate. Without that data, you are essentially flying blind and hoping nothing goes wrong. I used to skip it to save time on small-scale exploratory work and got burned twice in the same month. After that, I never skip it again regardless of how straightforward the chemistry looks on paper.
Some reactions are exothermic but practically safe because the heat release is mild and easy to manage. Others are borderline dangerous even at small scale. The difference often comes down to the magnitude of Delta H relative to the heat capacity of the reaction mass. A reaction with a Delta H of negative fifty kilojoules per mole in a dilute solution behaves very differently from one with the same Delta H concentrated in a small volume with minimal solvent. When I calculated the maximum temperature rise for a proposed chlorination step, the number came out to over two hundred degrees Celsius under adiabatic conditions. We redesigned the process to use a much larger solvent volume and a lower concentration of the chlorinating agent, which brought the theoretical maximum temperature down to something manageable, though still requiring careful monitoring.