Chemical Bonds, Explained Without the Textbook Fluff
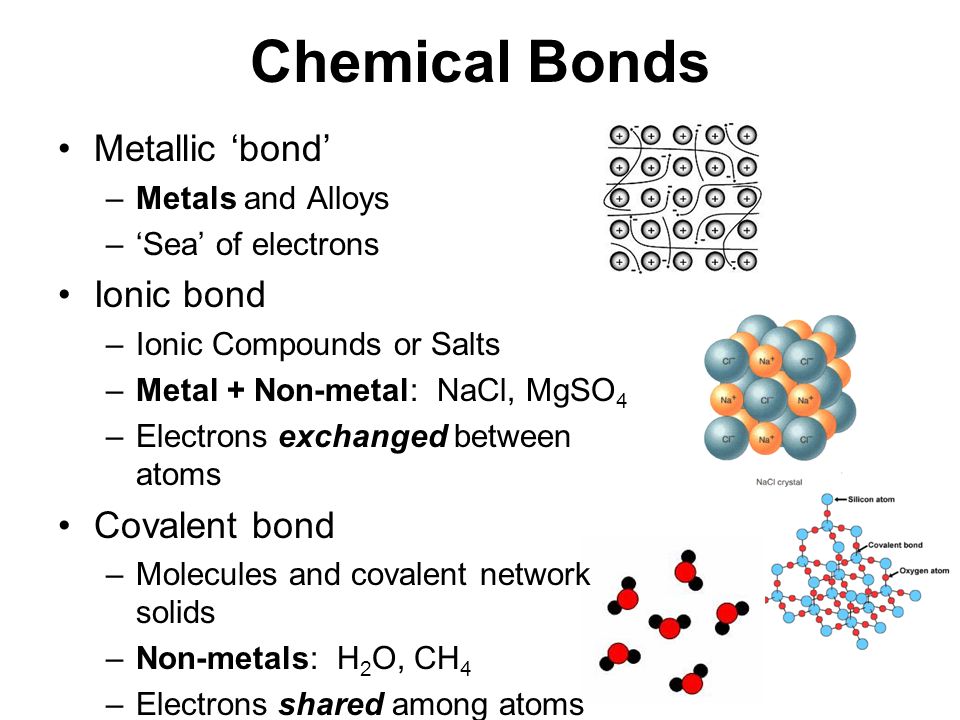
Most people learn about chemical bonds as if they are three separate topics that have nothing to do with each other. Ionic. Covalent. Metallic. They are not separate topics. They are points on a single spectrum of electron behavior, and treating them as isolated categories is why students get confused when a molecule like aluminum chloride starts misbehaving. Let me start with the mechanism because definitions become useless without understanding what actually moves. Electrons in the outermost shell of an atom are either shared, transferred, or delocalized. That is it. Everything else is just a more detailed description of which of those three things is happening under what conditions. The driving force is always the same: atoms adjust their electron configurations to reach a lower energy state. Lower energy means more stable. Nature does not care about your Lewis diagrams. It cares about potential energy. I ran into this the hard way when I was modeling reactions involving beryllium compounds. Beryllium has two valence electrons and a very small atomic radius. The textbook definition says it should form purely ionic bonds with highly electronegative elements like fluorine. In practice, BeF2 has significant covalent character because the beryllium ion is so small and highly charged that it polarizes the electron cloud of the fluoride ion. Fajan's rules describe this precisely, but the simplified introductory definition does not prepare you for it. I spent two days debugging a simulation that kept predicting incorrect bond lengths until I stopped treating the bond type as a binary label and started calculating the polarization contribution directly. The workaround was to apply a correction factor based on ionic potential rather than relying on electronegativity difference alone.
The Actual Definition Of Chemical Bonds and Why It Matters
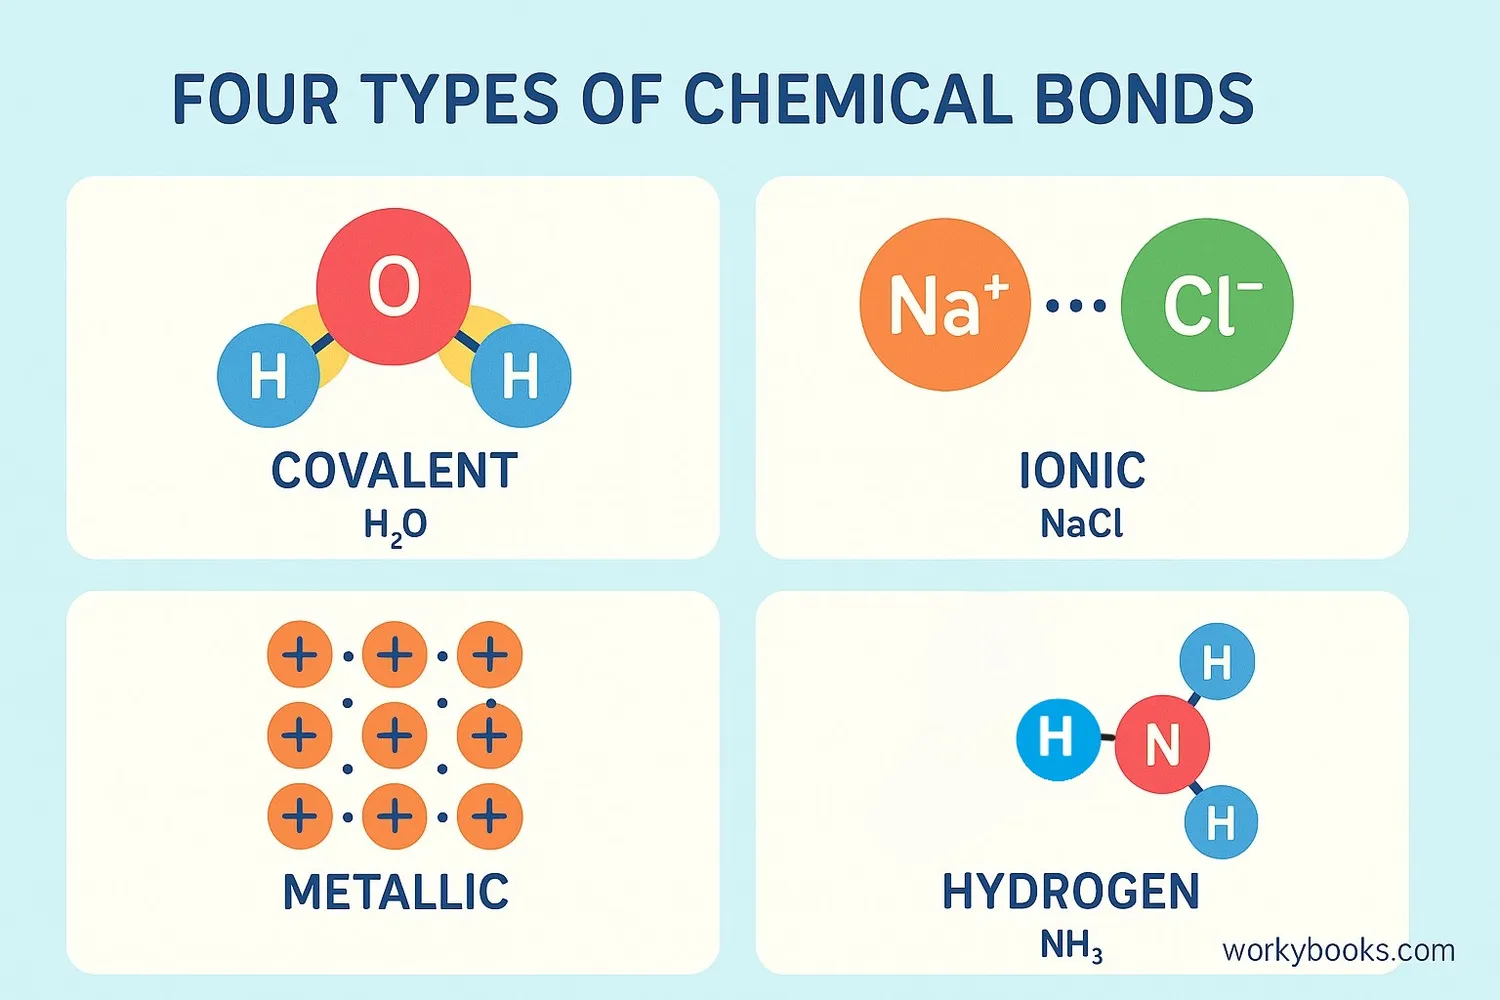
A chemical bond is a persistent attractive interaction between two or more atoms or ions that holds them together in a structured arrangement. The attraction arises from the electrostatic forces between electrons and nuclei, modified by quantum mechanical effects that determine how electron orbitals overlap and distribute charge. Electrostatic forces alone would predict that every negative electron would simply collapse into the positive nucleus. Quantum mechanics prevents that collapse and introduces the concept of bonding and antibonding orbitals, which is where the real work happens. Here is what most sources leave out. Bond type is not an intrinsic property of a pair of atoms. It depends on the environment. Sodium chloride is ionic in a crystal lattice. It is molecular and covalent in the gas phase at high temperature. The same atoms, different conditions, different bond classification. If you are working with computational chemistry or interpreting spectroscopic data, assuming a fixed bond type will give you wrong answers every time. The Coulombic model works fine for ionic interactions at long range, where the charge separation is clear and the distance is large enough that orbital overlap is negligible. For covalent bonds, you need molecular orbital theory or valence bond theory because the electrons are shared and the orbital overlap determines bond strength, bond length, and bond angle. Metallic bonding is best described by the free electron model or band theory because the valence electrons are delocalized across the entire lattice, not attached to any specific atom.
Electronegativity difference is the most commonly used shortcut for predicting bond character, but it is blunt and often misleading. A difference of 1.7 or above on the Pauling scale is the traditional cutoff for ionic character, but that number was derived from a handful of compounds and does not hold universally. Compounds with differences above 2.0 can still show covalent behavior if the cation is small and highly charged. Compounds with differences below 0.5 can behave ionically in certain crystal structures. The cutoff exists because it is useful, not because it is fundamentally correct. I once had to explain to a colleague why his DFT calculations on a supposed ionic solid were producing band structures with finite density of states at the Fermi level. The compound was supposedly insulating based on textbook ionic bonding rules. The issue was that the anion was large and polarizable, and the cation had enough charge density to induce covalent mixing in the valence band. The solid was not ionic in the way he assumed. Switching from a standard generalized gradient approximation to a hybrid functional with a portion of exact Hartree-Fock exchange corrected the band gap and matched the experimental optical absorption edge within 0.1 electron volts. That was a three-week delay I did not recover from. Bond energy is another area where the simple definition falls apart quickly. The bond dissociation energy of a particular bond depends on what else is attached to the atoms. Breaking the O-H bond in water costs 492 kilojoules per mole for the first bond and 424 for the second. They are the same bond type in the same molecule, but the energies are different because the remaining fragment after the first break is no longer the same chemical species. Average bond energies listed in reference tables are exactly that, averages across many different molecules, and using them for precise enthalpy calculations can introduce errors of 10 to 15 percent depending on the reaction.
Get the Full Details

Bond length correlates inversely with bond order, but the relationship is not linear. A double bond is shorter than a single bond, but it is not half the length. In carbon-carbon bonds, a single bond is about 154 picometers, a double bond is about 134, and a triple bond is about 120. The increments decrease as bond order increases because each additional bond involves a different set of orbitals with different overlap characteristics. Sigma bonds and pi bonds contribute differently to bond strength and length, and pi bonds are more sensitive to steric and electronic perturbations than sigma bonds. The coordination number of a metal center in a complex is often treated as a simple counting exercise, but it is really a balance between steric constraints of the ligands and electronic requirements of the metal. Six-coordinate octahedral geometry is common for transition metals, but four-coordinate complexes can be tetrahedral or square planar depending on the d-electron count and the ligand field strength. Crystal field theory and ligand field theory handle this, but the simplified definition of a coordinate covalent bond as just a shared electron pair does not capture why one geometry is preferred over another in a given case. I should note where this framework breaks down. Hydrogen bonding is not a true chemical bond in the same sense as ionic or covalent interactions. It is an electrostatic attraction with some partial covalent character in strong cases. Calling it a bond is convenient but imprecise. Van der Waals interactions are even weaker and entirely non-specific. These forces dominate in large biomolecules and supramolecular assemblies where the actual covalent and ionic bonds are few relative to the total number of interacting atoms. If you try to model protein folding using only bond definitions, you will get nothing useful.
Another limitation is that the concept of a discrete bond does not apply well to electron-deficient compounds. Diborane B2H6 has bridging hydrogen atoms that participate in three-center two-electron bonds. There is no way to draw a conventional two-center bond structure for diborane without violating the octet rule or creating impossible formal charges. The molecular orbital description works cleanly here, but the basic bond definition fails completely. For practical purposes, you need to know when the simplified model is sufficient and when you need to move to a more rigorous treatment. The simplified bond classification works for predicting stoichiometry, general reactivity patterns, and qualitative properties like solubility and melting point trends. It does not work for quantitative thermochemistry, spectroscopic prediction, or anything involving transition metals with partially filled d-orbitals. When you cross into that territory, you need ligand field theory, molecular orbital diagrams, or computational methods, and the idea of a bond as a simple connection between two atoms becomes inadequate. Orbital hybridization is another useful concept that people treat as more fundamental than it actually is. sp3, sp2, sp notation is a mathematical construction that approximates the shape of bonding orbitals. It works well for main group elements in simple molecules. It breaks down for molecules where the bonding cannot be described by localized two-center orbitals, and for transition metal complexes where d-orbital participation changes everything. The hybridization concept is a tool, not a law of nature.
If you want a single operational definition that covers most cases, a chemical bond exists when the total energy of the bonded system is lower than the total energy of the separated atoms or ions by an amount that exceeds the thermal energy at the conditions of interest. The energy threshold is arbitrary but practical. At room temperature, thermal energy is about 2.5 kilojoules per mole. Interactions weaker than that do not produce persistent bonding. That is why noble gases do not form bonds under standard conditions, and why van der Waals complexes are transient and weak. The takeaway is straightforward. Chemical bonds are electrostatic attractions modified by quantum mechanics, ranging from complete electron transfer to complete electron sharing to complete electron delocalization, with most real bonds falling somewhere in between. The classification system is a human invention for organizing observations, not a fundamental feature of nature. Bonds exist on a continuum, and the labels you assign depend on which properties you are interested in and how much accuracy you need.