The Core Postulates You Actually Need to Remember
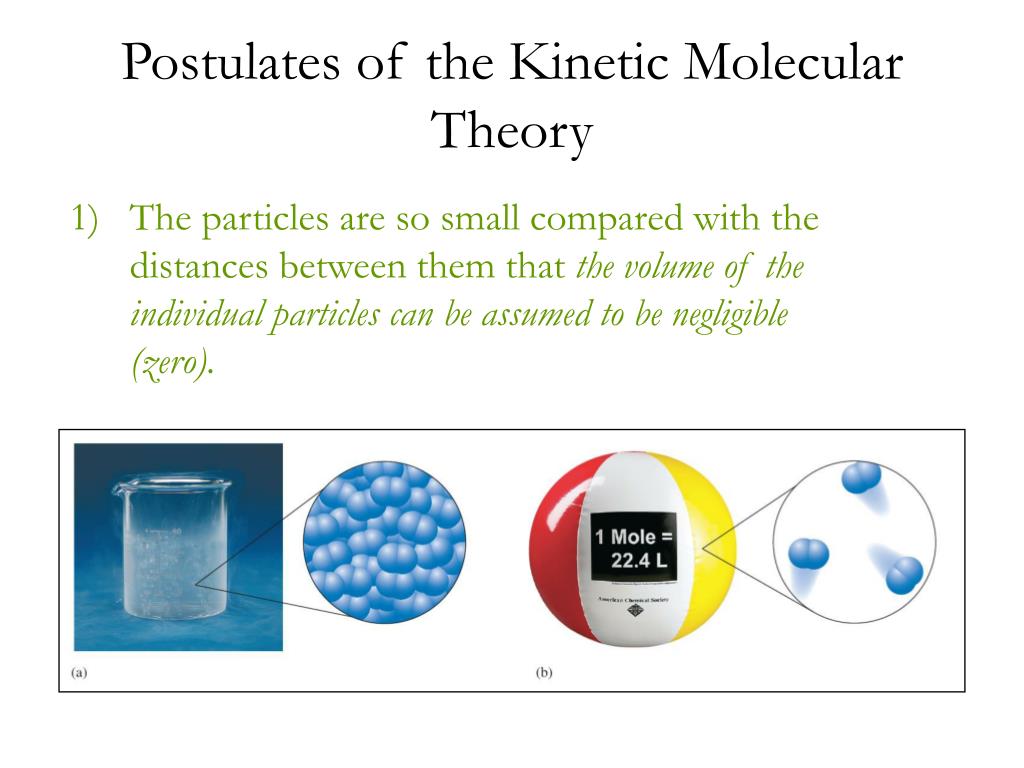
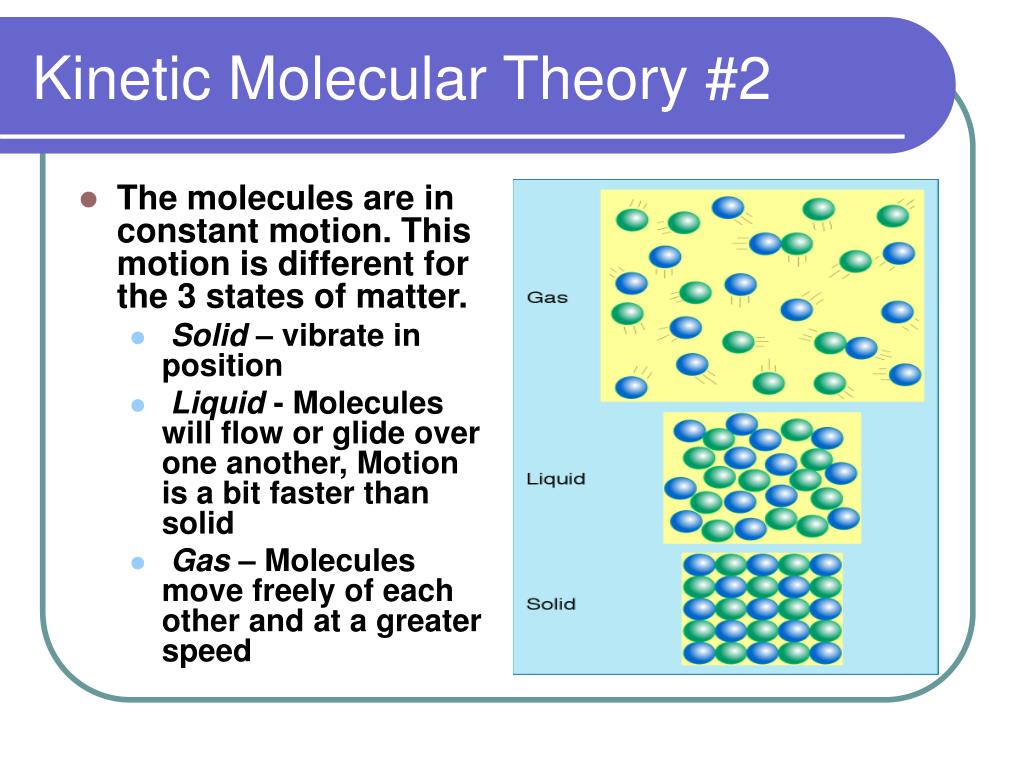
Most textbooks present the Definition Of Kinetic Molecular Theory as a neat list of five bullet points. That is accurate but incomplete. The theory makes several simplifying assumptions about how gas particles behave, and understanding where those assumptions break is what separates someone who can plug numbers into a formula from someone who actually knows what is happening. The core idea is straightforward: all matter is made of tiny particles in constant random motion, and the temperature of a gas is directly proportional to the average kinetic energy of those particles. Assumption one: gas consists of a large number of tiny particles separated by large distances. This means most of the volume of a gas is empty space. Assumption two: collisions between particles and between particles and container walls are perfectly elastic, meaning no kinetic energy is lost to friction or deformation. Assumption three: the volume of the individual particles themselves is negligible compared to the volume of the container. Assumption four: there are no intermolecular forces between particles. Assumption five: the average kinetic energy depends only on temperature, not on the identity of the gas. These assumptions let you derive the ideal gas law and connect macroscopic measurements to microscopic behavior. You get equations like KE_avg = 3/2 kT, where k is Boltzmann's constant and T is absolute temperature. You also get the root mean square speed equation, v_rms = sqrt(3RT/M), which tells you how fast gas molecules are actually moving at a given temperature. For nitrogen at room temperature, that comes out to roughly 517 meters per second. They are moving incredibly fast, but they collide with other molecules so frequently that their net displacement is small.
Definition Of Kinetic Molecular Theory and Where It Falls Apart
I learned the theoretical framework fine. What I did not learn until I was actually running gas-phase experiments was how quickly the model desynchronizes from reality when conditions shift. The theory assumes point particles with no attraction. That works reasonably well for argon at standard temperature and pressure. It does not work for water vapor near condensation. I spent a week debugging pressure readings that consistently ran about eight percent lower than the ideal gas law predicted for a nitrogen sample at around 150 atmospheres. The particles were close enough together that intermolecular forces started mattering, and the volume of the molecules themselves was no longer negligible. Switching to the van der Waals equation with the appropriate constants for nitrogen brought my predictions within one percent of the observed values. This is the practical edge case nobody emphasizes enough. The kinetic molecular theory gives you a baseline, not a universal answer. When pressure climbs above roughly 10 atmospheres or temperature drops below roughly twice the boiling point of the gas, the ideal assumptions start producing systematic errors. You need to know when to stop using PV = nRT and switch to a real gas equation of state. For rough calculations at moderate conditions, the error is usually acceptable. For anything requiring precision, it will bite you. Another thing that trips people up is the relationship between temperature and molecular speed. Temperature is proportional to average kinetic energy, not average speed. Since kinetic energy is 1/2 mv^2, lighter molecules must move faster than heavier ones at the same temperature. A molecule of hydrogen at 300 Kelvin has roughly four times the rms speed of a molecule of oxygen. This is why hydrogen escapes Earth's atmosphere more readily. The distribution of speeds matters too. Not every molecule moves at v_rms. The Maxwell-Boltzmann distribution describes the spread, and it has a long tail toward higher speeds. That tail is functionally important for reaction rates and for understanding why some molecules escape gravitational binding even when the average speed is well below escape velocity.
The theory also explains diffusion and effusion in a way that simple intuition does not. Graham's law states that the rate of effusion is inversely proportional to the square root of the molar mass. This is a direct consequence of the kinetic molecular framework and has been verified experimentally to high precision. You can separate isotopes of uranium using this principle, which is how enrichment plants historically operated. The theory connects the microscopic picture to measurable macroscopic behavior without requiring any additional ad hoc assumptions. One counter-intuitive point worth noting: the pressure of a gas does not depend on the size or mass of individual particles in the ideal model, only on their number, their average kinetic energy, and the container volume. Two different gases at the same temperature and volume exert the same pressure if they have the same number of moles, regardless of whether one is helium and the other is xenon. Heavier molecules hit the walls less frequently but each collision transfers more momentum. Lighter molecules hit more frequently but each collision transfers less. The effects cancel exactly in the ideal approximation. The kinetic molecular theory is not wrong. It is an idealization with a well-defined domain of applicability. Inside that domain, it is remarkably predictive and internally consistent. Outside it, you need corrections. Knowing the correction terms and when to apply them is the actual skill. The definitions are easy to memorize. Understanding the breakdown is harder and more useful.
Get the Full Details