Getting Your Head Around Acid-Base Definitions Without Losing It
I've been teaching general chemistry for twelve years and watching students struggle with the same three definitions keeps getting old fast. The problem isn't that the concepts are hard. It's that every textbook presents them as separate topics instead of showing how they connect and where each one breaks down. Let me explain this the way I wish someone had explained it to me back when I was learning.Definitions For Acids And Bases From Different Angles
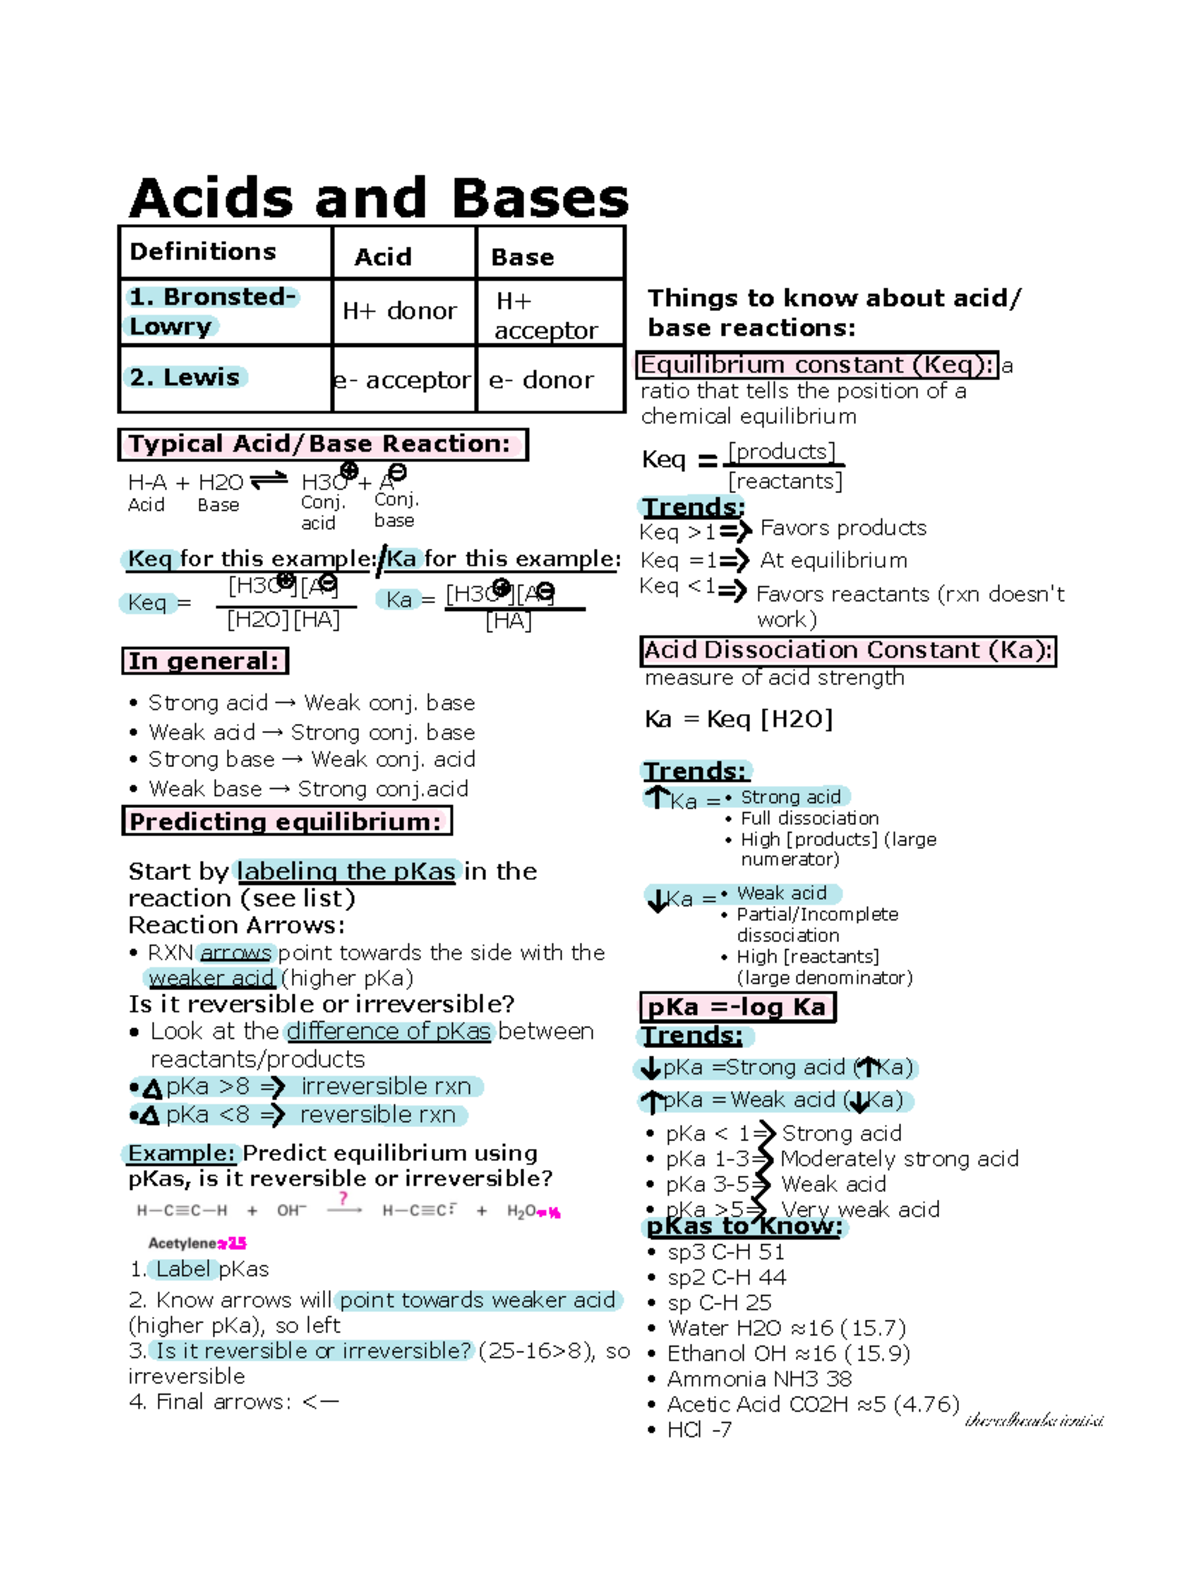
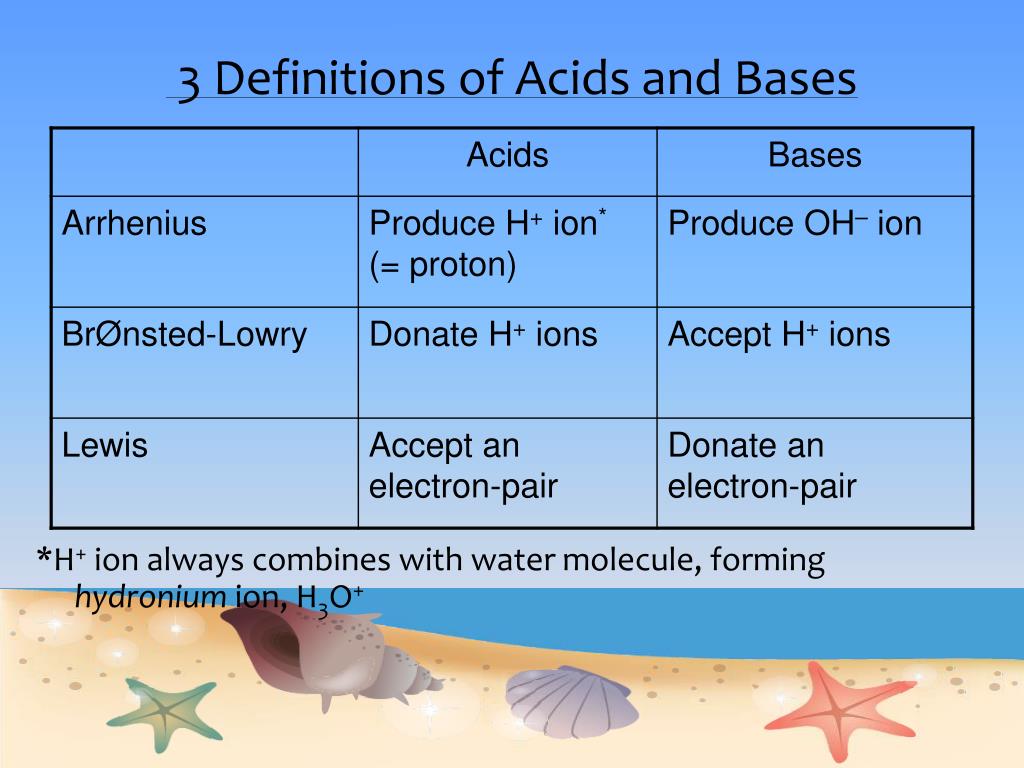
The Arrhenius definition is the simplest and the most limited. Acids produce hydrogen ions in water. Bases produce hydroxide ions in water. That's it. Strong acids like HCl and strong bases like NaOH fit perfectly. Weak acids like acetic acid work too if you account for partial dissociation. But the moment you step outside aqueous solution, this definition hits a wall. Anhydrous ammonia is a base. It doesn't contain OH groups at all. The Arrhenius framework can't handle it without stretching the definition until it snaps. The Brønsted-Lowry definition fixes that gap. Acids donate protons. Bases accept protons. This works in water, in organic solvents, even in the gas phase. You can explain why ammonia acts as a base without referencing hydroxide. You can explain why ammonium chloride is acidic without needing water as the medium. The proton transfer concept unifies everything. It also introduces conjugate acid-base pairs, which is where students usually get confused but shouldn't. The Lewis definition goes even further. Acids accept electron pairs. Bases donate electron pairs. This covers reactions that don't involve protons at all. Boron trifluoride acting as an acid toward ammonia is a classic example. No hydrogen transfer. Just electron pair sharing. The Lewis framework includes both Brønsted acids and bases as special cases. It's the most general definition we have.
How These Definitions Actually Work In Practice
When I first ran a lab experiment with Friedel-Crafts acylation, I saw aluminum chloride acting as a Lewis acid accepting electron pairs from the acyl chloride. That reaction doesn't move a single proton. The Arrhenius and Brønsted definitions would call it something else entirely or fail to classify it. The Lewis definition handles it without a second thought. That's when the hierarchy clicked for me. Here's the practical way to think about this. Start with Arrhenius when you're doing basic aqueous solutions in high school chemistry. It's sufficient for strong acids and bases at reasonable concentrations. Move to Brønsted-Lowry when you encounter weak acids, buffer systems, or non-aqueous solvents. Switch to Lewis when you're dealing with metal complexes, organometallic reactions, or any situation where proton transfer isn't the mechanism. I keep a decision tree on my office board. If the reaction happens in water with simple ionic compounds, Arrhenius works. If protons move between species, Brønsted-Lowry applies. If electrons move without proton transfer, Lewis is your framework. Most reactions fall into one category. Some require switching between definitions depending on how you describe the mechanism.
Common Pitfalls That Waste Student Time
The biggest mistake I see is treating these definitions as competing theories instead of complementary tools. Students think they need to pick one and stick with it. That's wrong. Each definition has a scope where it works best. Arrhenius is convenient for quick calculations with strong electrolytes. Brønsted-Lowry handles equilibrium calculations across different solvents. Lewis explains reactivity patterns that proton-based definitions can't touch. Another trap is assuming that Lewis acids are always stronger than Brønsted acids. That's not necessarily true. Aluminum chloride is a powerful Lewis acid. Hydrochloric acid is a powerful Brønsted acid. They operate in different regimes. Comparing their strength directly depends on what you're trying to accomplish in the reaction. Conjugate pairs cause confusion too. When HCl donates a proton, chloride is its conjugate base. When ammonia accepts a proton, ammonium is its conjugate acid. Students often mix up which species is which because the labels depend entirely on which direction you're considering the reaction. I tell them to write the equilibrium both ways and label each species relative to the reaction direction they're analyzing.
Get the Full Details

Where The Definitions Fail Completely
No acid-base definition handles superacid media well. In magic acid (HSO3F-SbF5), the concept of a solvent cage breaks down. Proton activity becomes so high that traditional pH measurements lose meaning. The Hammett acidity function replaces pH in these conditions, but that's a different topic entirely. Gas-phase acidities don't map cleanly onto solution-phase measurements. Trifluoromethanesulfonic acid is one of the strongest Brønsted acids in water. In the gas phase, the ordering changes because solvation effects disappear. Hydrogen bonding patterns shift. Students who memorize pKa tables without understanding what those numbers represent will make wrong predictions when they leave aqueous solution behind. Amphoteric substances trip people up. Water, aluminum hydroxide, amino acids, hydrogen carbonate. These can act as acids or bases depending on what they encounter. The definitions handle this fine. The intuition takes practice. I recommend writing out every reaction as a proton transfer and identifying which species donates and which accepts in each direction.
A Specific Problem I Ran Into
Last semester I was preparing students for an exam question about why pyridine is a weaker base than pyrrole despite both being nitrogen heterocycles. Students immediately reached for electronegativity arguments. That's the wrong approach. The real issue is orbital hybridization and lone pair availability. Pyridine's nitrogen is sp2 hybridized with the lone pair in an sp2 orbital perpendicular to the pi system. Pyrrole's nitrogen is also sp2, but the lone pair participates in aromaticity. Removing it to accept a proton would break the aromatic system. I spent twenty minutes drawing orbital diagrams on the board showing electron density distribution. That helped. The definition-based explanation alone wouldn't have captured the energetic penalty of disrupting aromaticity. Lewis theory gives you the framework. Molecular orbital theory gives you the reason.
What To Focus On Instead of Memorizing
Understanding proton transfer mechanisms matters more than memorizing which definition to apply. When you can trace electron movement through a reaction arrow by arrow, the classification becomes obvious. Strong acids push protons off easily. Strong bases grab protons eagerly. Weak acids and bases sit in equilibrium somewhere in the middle. For lab work, I always ask students to identify the acid and base in every reaction before calculating anything. That habit catches errors early. A student who misidentifies the acid-base pair will get the wrong conjugate, the wrong equilibrium expression, and the wrong pH calculation. Starting with correct identification saves time downstream. The trend tables for acid strength exist for a reason. Oxidation state matters. Electronegativity matters. Atomic size matters. Resonance stabilization matters. Inductive effects matter. Each factor shifts equilibrium positions in predictable directions. Once you internalize those trends, you don't need to look up every pKa value. You can estimate whether an acid is strong enough to protonate a particular base.
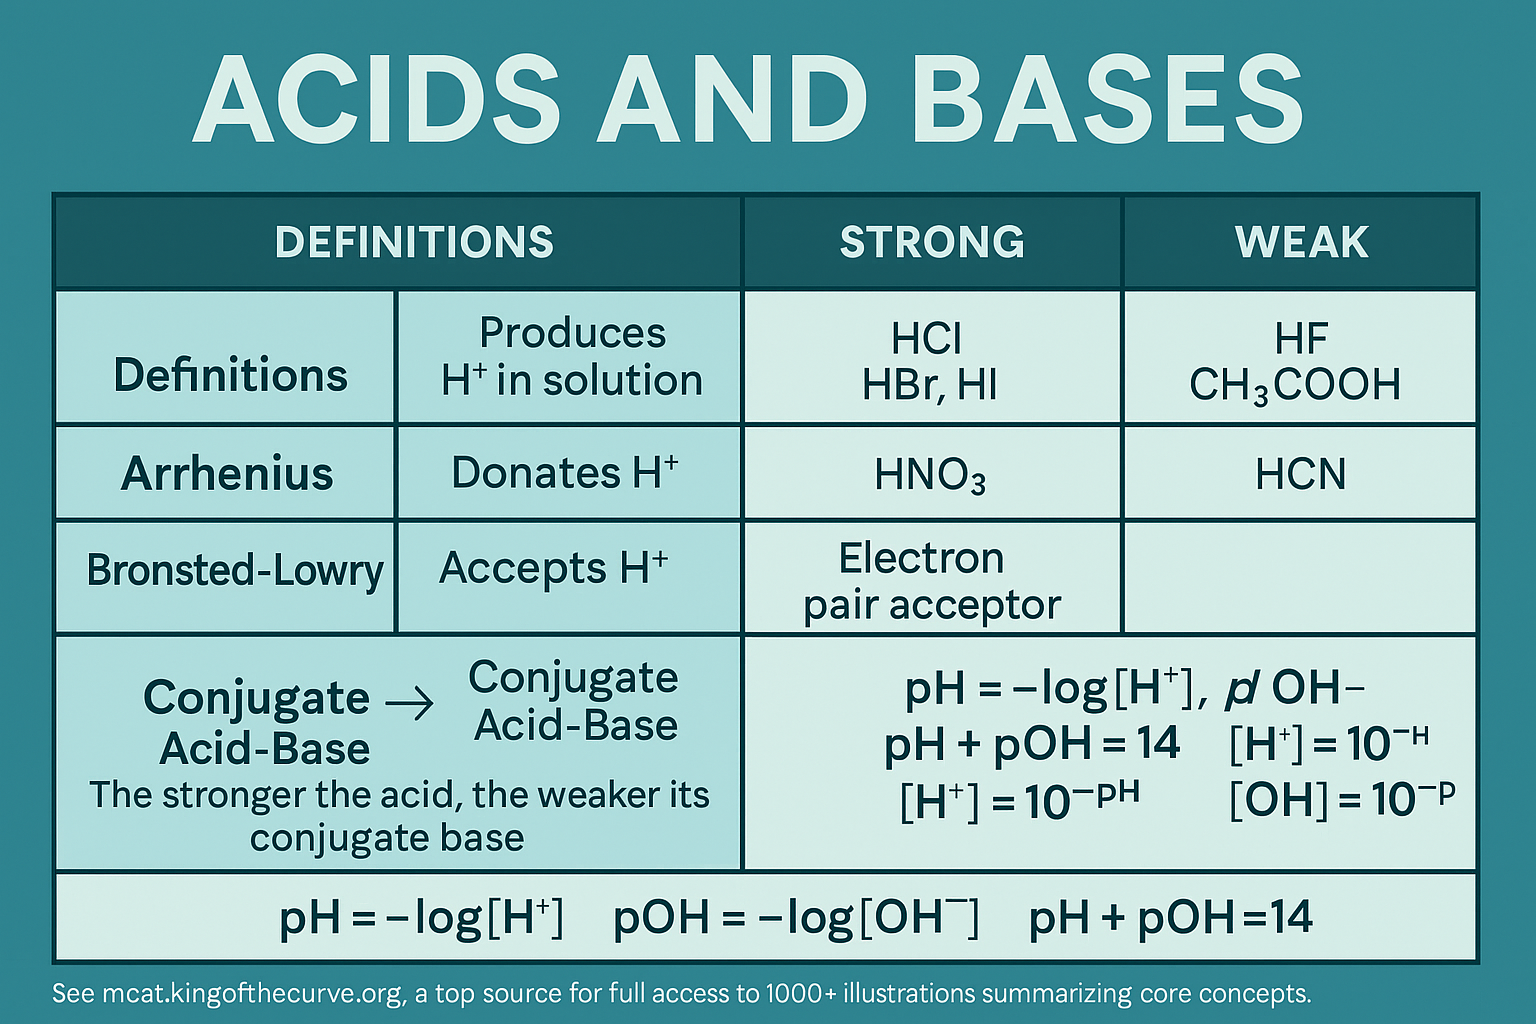
I've found that students who draw out the full mechanism with all lone pairs and formal charges make far fewer mistakes on exam questions about acid-base chemistry. The visual representation forces you to confront where electrons are going. Definitions alone leave too much room for hand-waving.