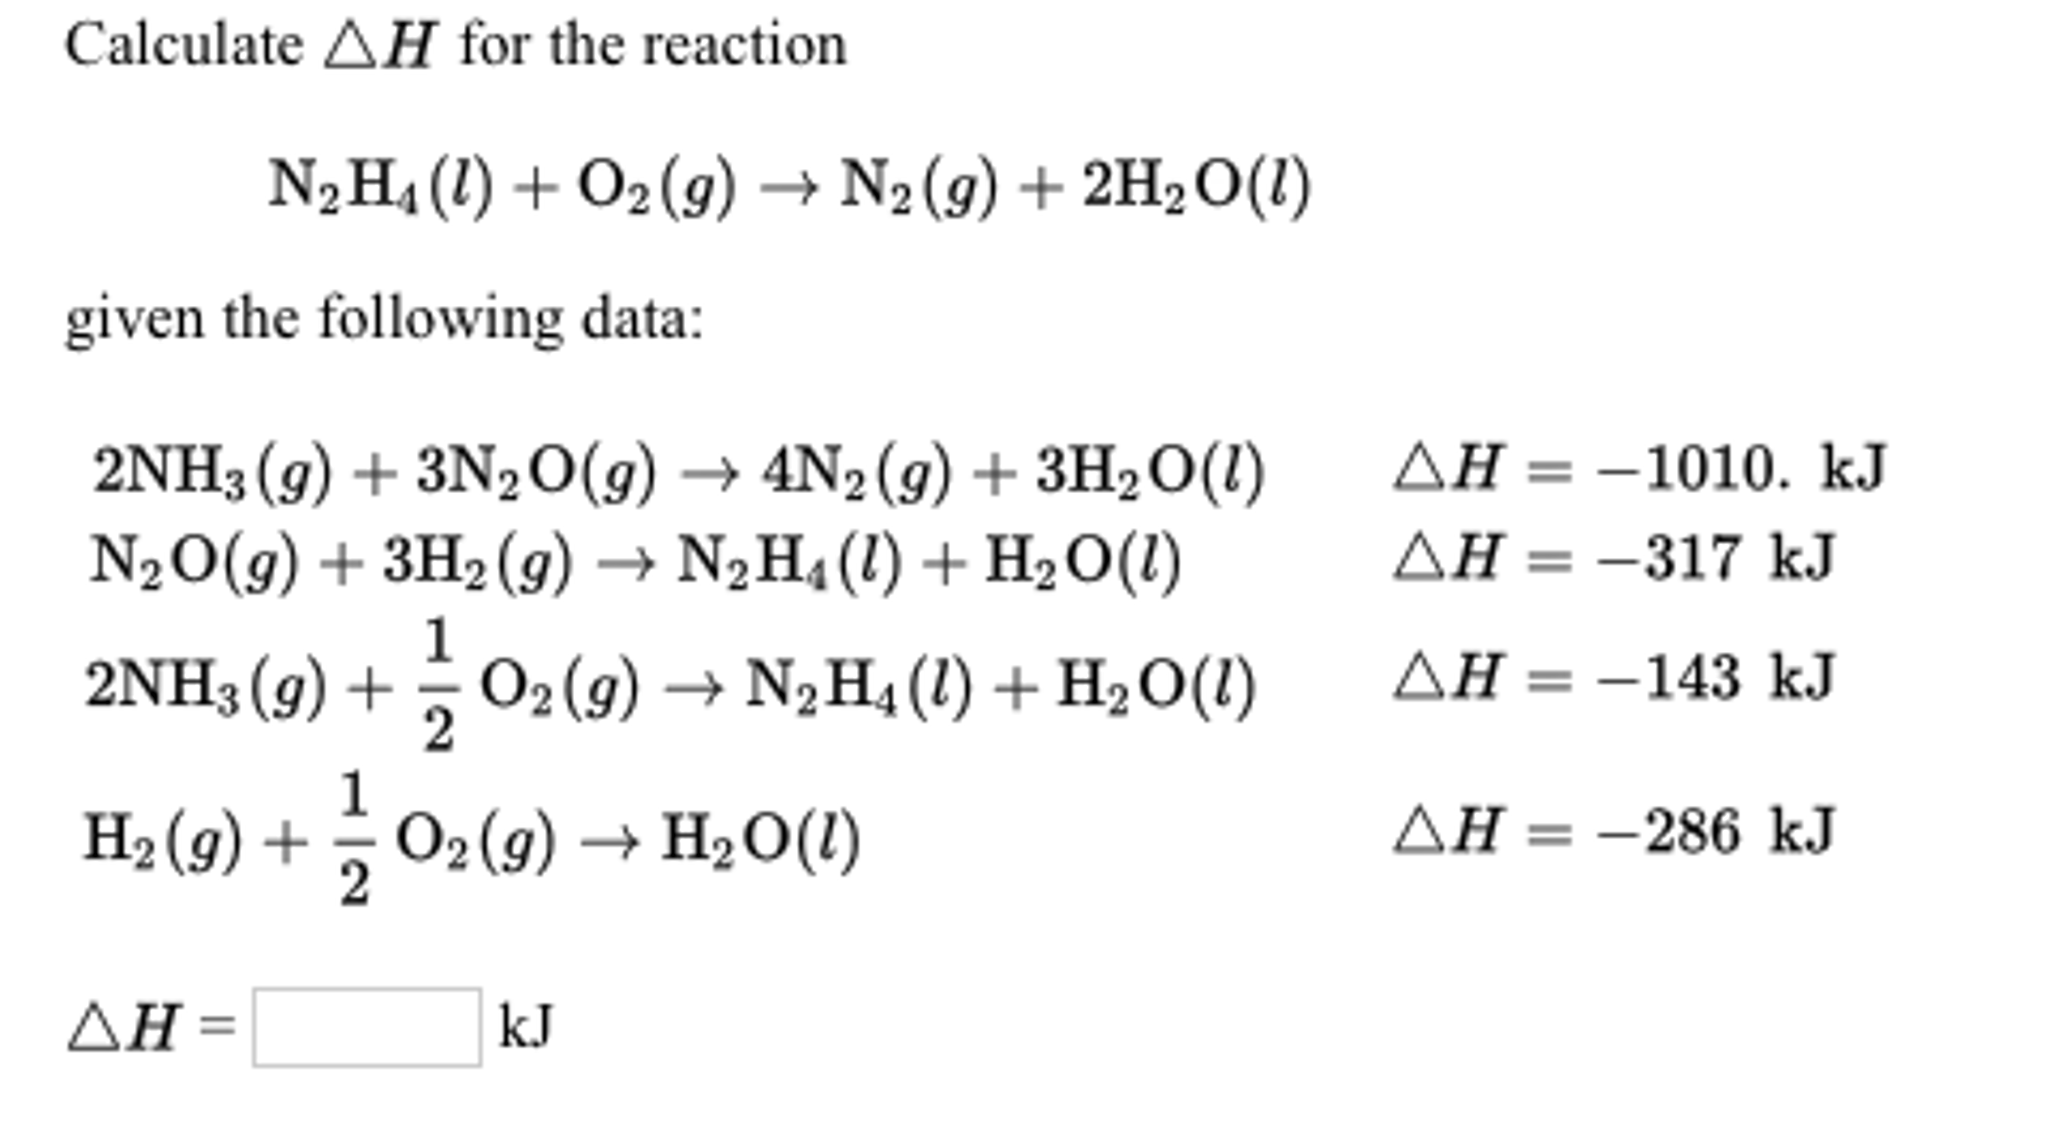
Why Your Delta H Of Formation Numbers Keep Looking Wrong
You look up a table. You subtract products from reactants. You get an answer. It's off by twelve kilojoules per mole and you can't find the mistake. This happens all the time. Here is what actually goes wrong and how to fix it. The standard formula is straightforward: delta H reaction equals the sum of formation enthalpies for products minus the sum for reactants, each multiplied by their stoichiometric coefficients. The trouble starts immediately after you write that formula because the numbers in your table are not all created equal. First, verify the phase. A table entry for water vapor carries a significantly different value than liquid water. Students regularly pull the gas value when the reaction produces liquid, or vice versa, and blame arithmetic. Check the phase label next to every single number before you do anything else. Second, verify the state is standard state. Some values are given at one bar, older tables at one atmosphere. The difference is small for condensed phases but not zero, and if you are working across multiple sources it compounds.
Third, remember that elements in their standard states are defined as having zero formation enthalpy. That means oxygen gas, nitrogen gas, solid graphite, and liquid bromine are all zero. Things like O2 gas at standard conditions, sulfur as rhombic solid, and phosphorus as white P4 are zero. But oxygen as O3, carbon as diamond, and phosphorus as red P are not zero. Look them up individually. I once lost forty minutes rechecking my work on a problem where the answer turned out to be wrong because I had assigned zero to red phosphorus instead of looking up its actual formation enthalpy. For the calculation itself, I recommend writing out each term explicitly on paper rather than compressing it into a single line. You need to show the coefficient, the value, and the phase for every species. When you are working through five or six compounds, a compressed line makes it nearly impossible to spot which number you pulled from the wrong row.
A Real Problem I Hit Last Semester
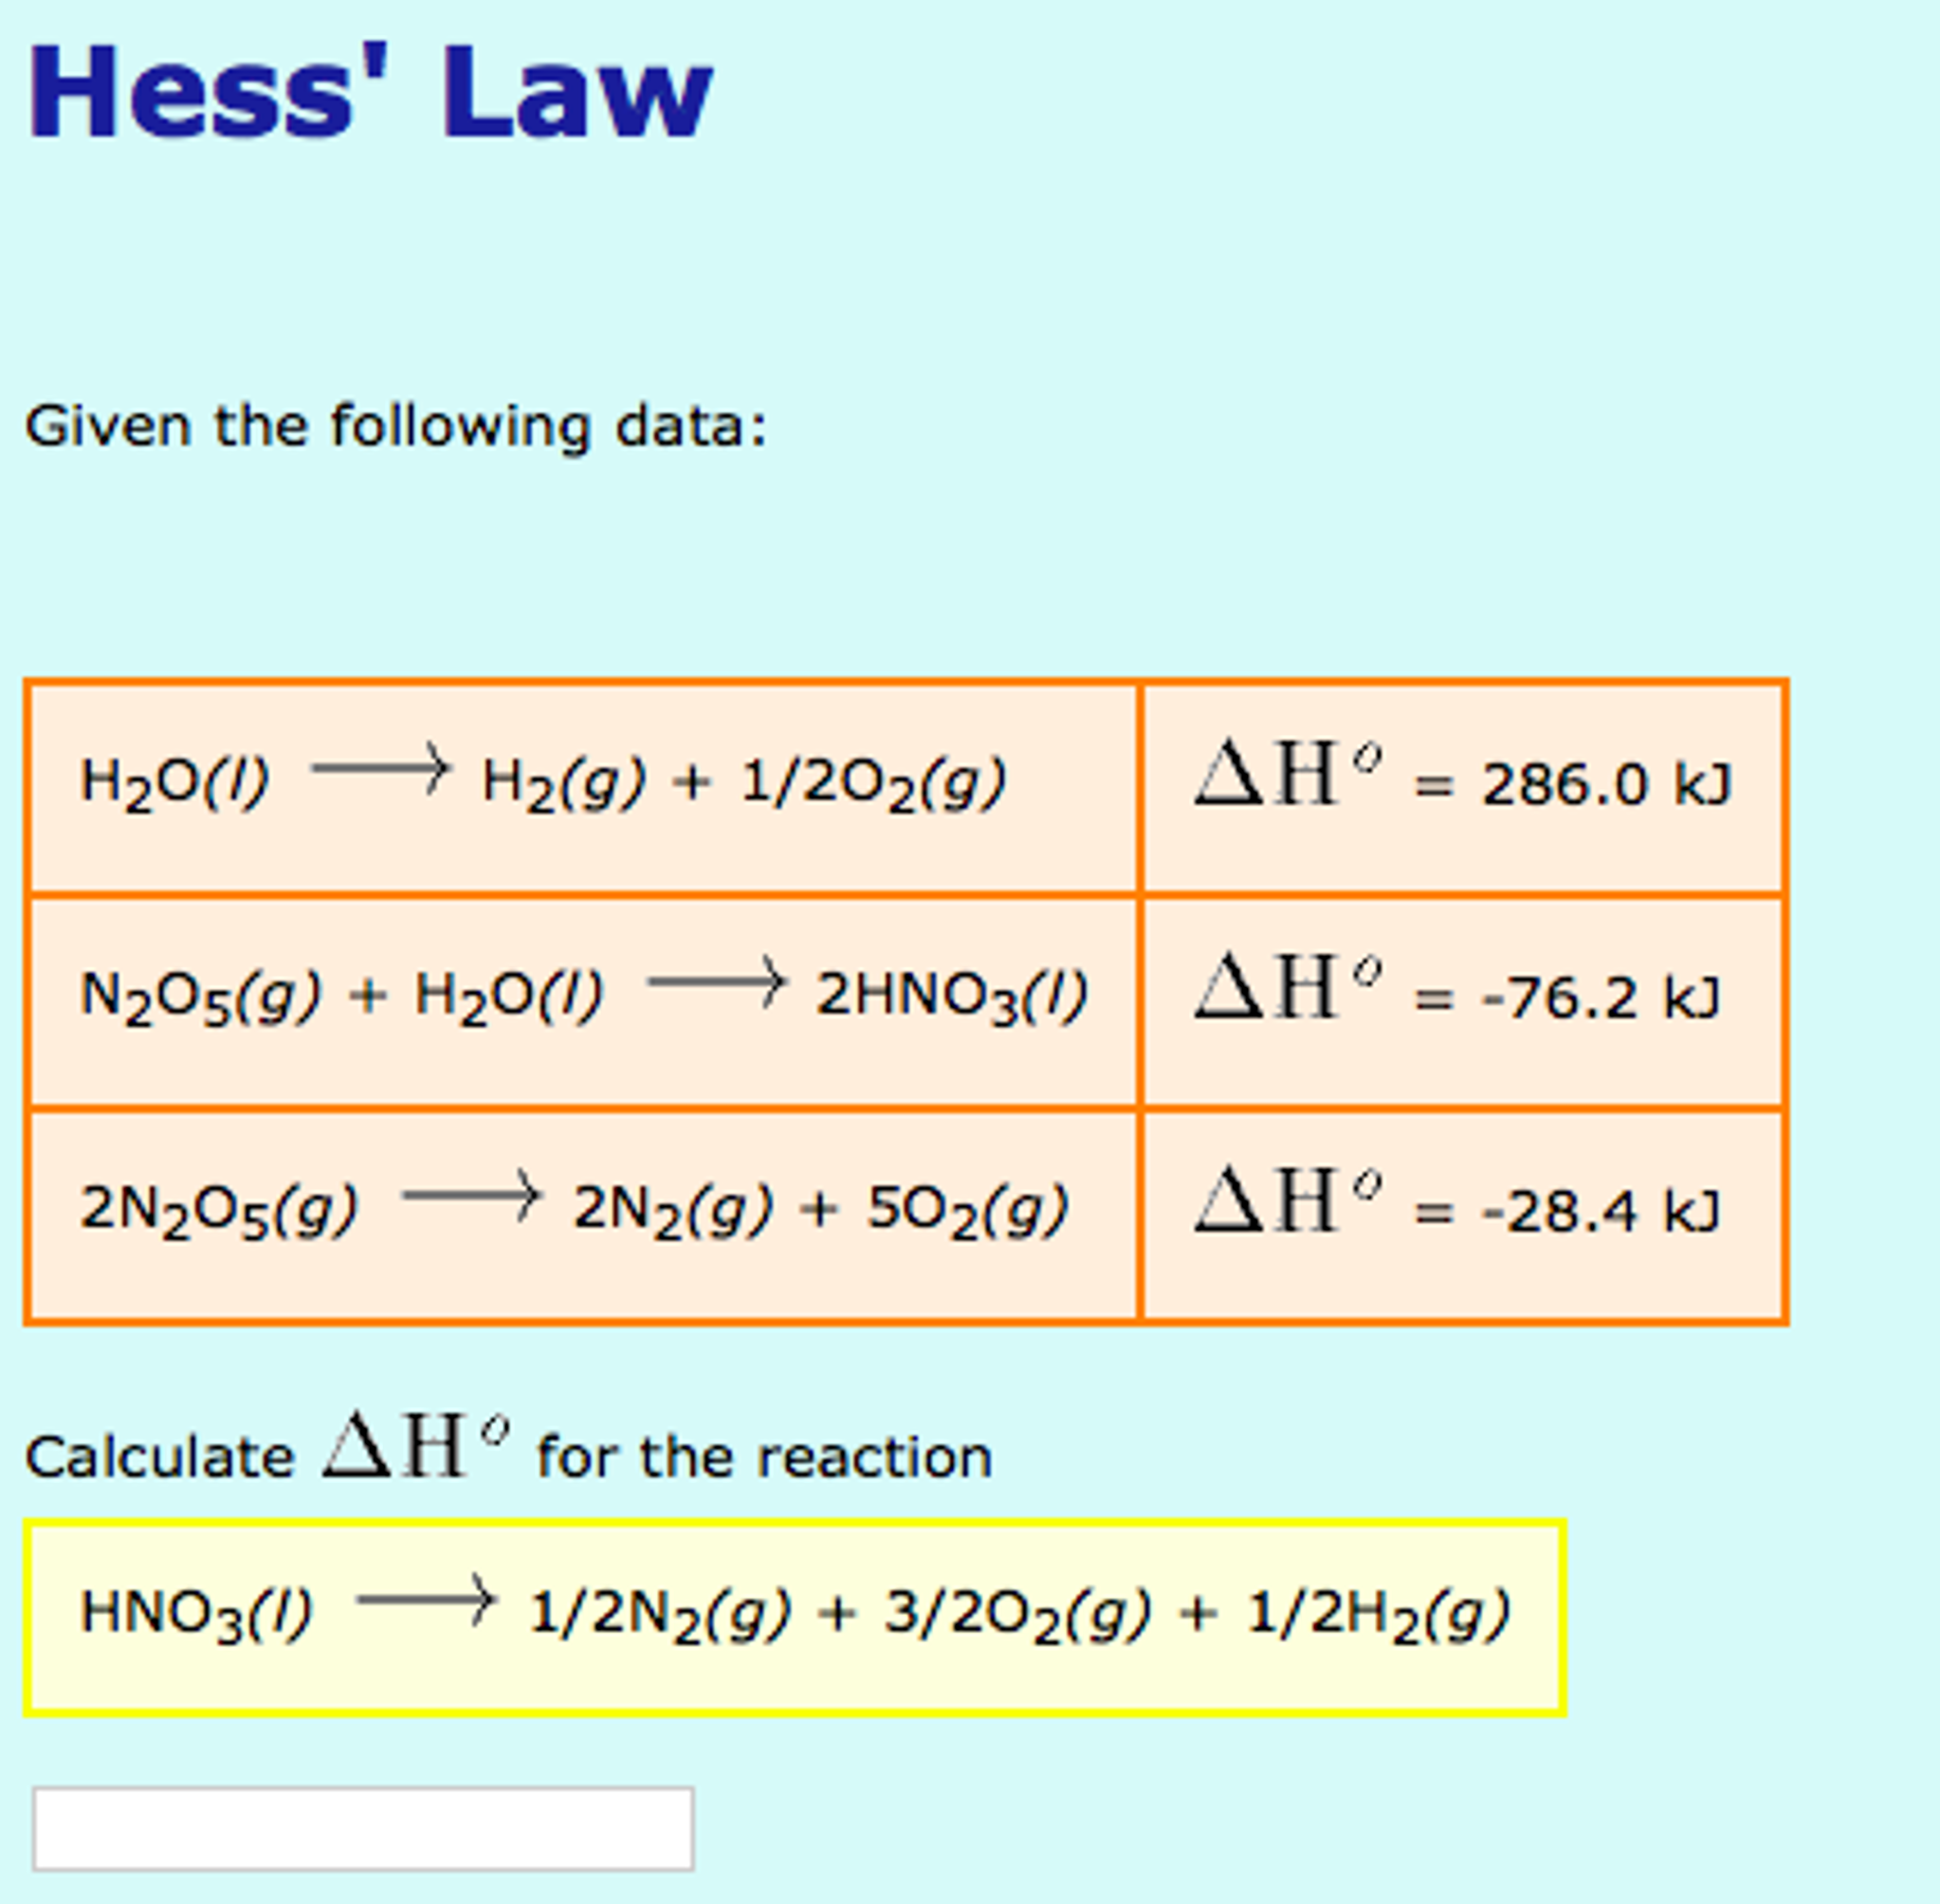
I was working with a compound that did not appear in any of the standard tables we had available. It was not a common organic molecule, just some substituted derivative we synthesized for a lab project. There was no tabulated standard enthalpy of formation. The straightforward subtraction method was impossible because one term was missing. The workaround is to use Hess's law indirectly. If the compound participates in a combustion reaction or another reaction where all other enthalpies are known, you can measure or calculate the overall reaction enthalpy and solve for the unknown formation enthalpy algebraically. In practice this usually means running a combustion calorimetry experiment or finding a published reaction enthalpy and rearranging. I used a known combustion enthalpy from the literature and back-calculated the formation value. It added about an hour of work compared to a simple table lookup but it gave a result that was consistent within experimental uncertainty. If you cannot find a literature reaction to anchor the calculation, this method stops working. There is no way to derive a formation enthalpy from nothing. In those cases you either need to measure it yourself or accept that the value is unknown and leave that species out of your tabular approach entirely.
Get the Full Details

Counter-Intuitive Details Beginners Miss
Formation enthalpies are not temperature independent. The values you see in tables are almost always at 298.15 kelvin. If your reaction occurs at a substantially different temperature, you need to apply heat capacity corrections. The adjustment uses the difference in heat capacities between products and reactants, integrated over the temperature range. For reactions spanning more than fifty kelvin from standard conditions, skipping this correction can introduce errors larger than typical experimental uncertainty. The correction is usually small at first but it grows linearly with temperature difference for reactions with significant heat capacity mismatches. Another thing that trips people up is the sign convention for endothermic formations. A positive formation enthalpy means the compound is thermodynamically less stable than its constituent elements. That does not mean the compound will decompose spontaneously. Kinetics matter. Many compounds with positive formation enthalpies are perfectly stable at room temperature because the activation barrier for decomposition is high. The enthalpy value alone does not tell you whether a substance will fall apart on a shelf.
Limits of the Tabular Method
The standard formation enthalpy approach has real bottlenecks. It requires complete tabular data for every species in the reaction. Gaps in the table make the method unusable unless you can reconstruct missing values through indirect calorimetry or quantum chemical calculations. It assumes ideal standard states, which breaks down at high pressures or in non-ideal solutions where activity coefficients become important. For electrolyte solutions in particular, the standard state conventions vary between sources and mixing tables from different authors without checking the reference state is a reliable way to introduce systematic error. When the tabular method fails, the practical alternatives are solution calorimetry for compounds in solution, bomb calorimetry for combustion reactions, or computational chemistry estimates. The computational route is fast but the accuracy depends heavily on the method and basis set you use. DFT values can be within ten kilojoules per mole for well-behaved organic molecules but may drift further for transition metal complexes. It is better than nothing when experimental data does not exist, but treat those numbers as estimates until someone verifies them. The biggest single cause of wrong answers is still pulling the wrong table entry. Spend the first five minutes of any problem verifying phases, standard states, and element definitions before you start crunching numbers. The rest of the work usually follows cleanly from there.