The Diels Alder Reaction Mechanism
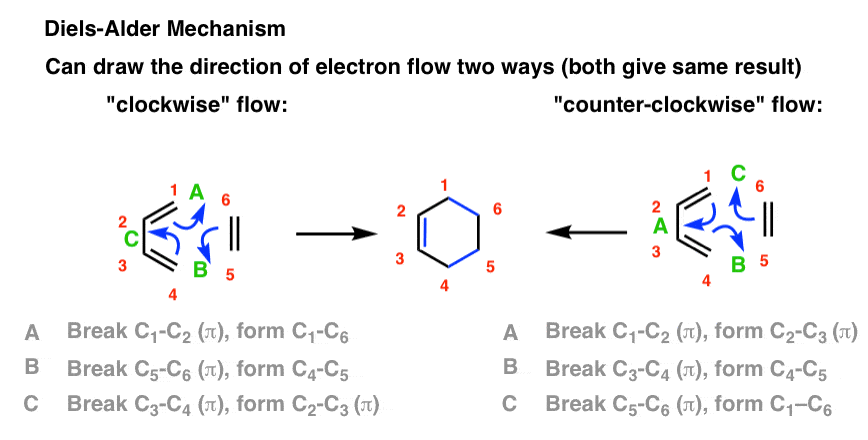
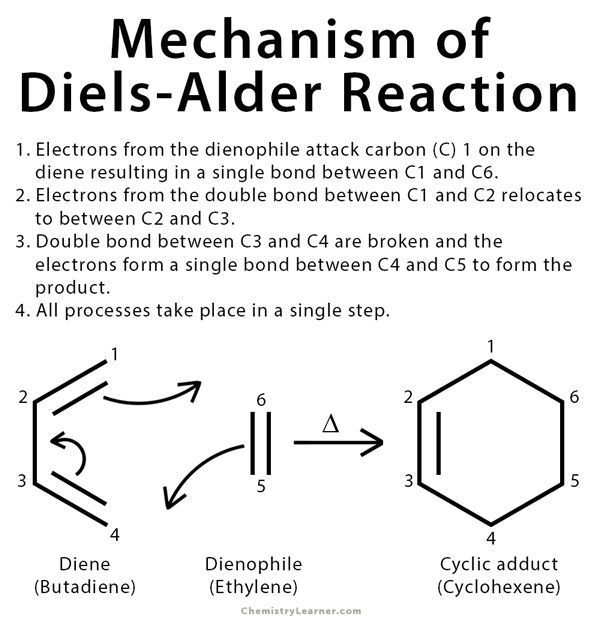
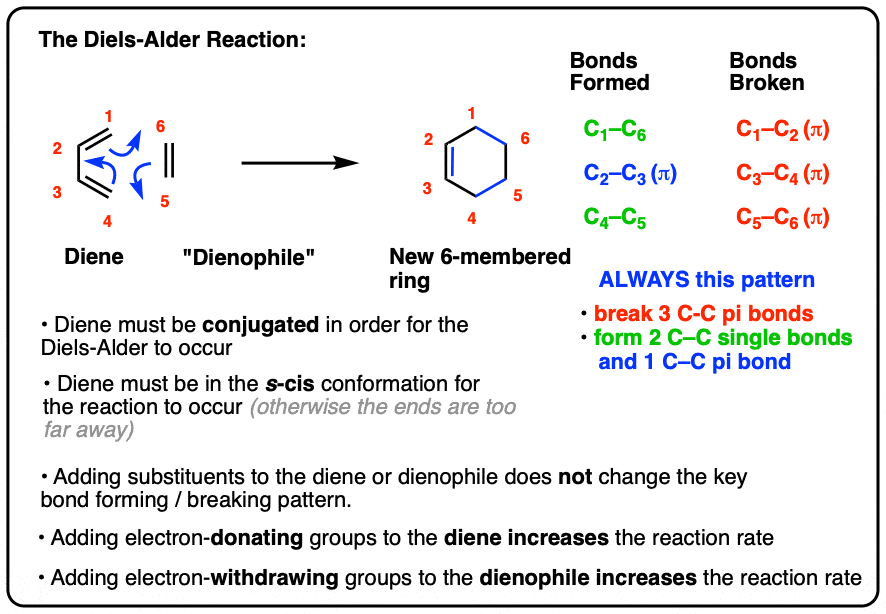
The Diels Alder is a [4+2] cycloaddition between a conjugated diene and a substituted alkene, commonly called the dienophile. It forms a six-membered ring in a single step. The new bonds form at the ends of the diene system and at the double bond of the dienophile. Two sigma bonds replace three pi bonds. No intermediates. No ions. Just a concerted rearrangement of electrons through a cyclic transition state. Most textbooks draw this with curly arrows showing six electrons moving in a circle. That diagram is correct but incomplete for anyone who has actually tried to run the reaction and gotten something else entirely. The mechanism is pericyclic and suprafacial on both components under normal thermal conditions. That means the orbital symmetry match is everything. The HOMO of one reactant has to overlap properly with the LUMO of the other, and if the energy gap is too large, nothing happens at useful temperatures.
What the Diels Alder Reaction Mechanism Actually Looks Like in Practice
Here is the straightforward part. You pick a diene that can adopt the s-cis conformation and a dienophile with an electron-withdrawing group. Acrolein and cyclopentadiene react readily at room temperature. Butadiene and maleic anhydride need mild heating, maybe 60 to 80 degrees Celsius in a sealed tube. Electron-rich dienes paired with electron-poor dienophiles is the standard combination because it minimizes the HOMO-LUMO gap and speeds things up considerably. When both partners are unactivated, you are looking at 150 to 200 degrees or a Lewis acid catalyst, sometimes both. I spent a week last year trying to force a reaction between 1,3-cyclohexadiene and methyl acrylate without any catalyst. The literature said it should work at 100 degrees in toluene. It did not work at all after six hours. I ran thin-layer chromatography, NMR, nothing showed product. Only the starting materials. Then I switched to using titanium tetrachloride at -10 degrees Celsius in dichloromethane and got quantitative conversion in 45 minutes. The Lewis acid coordinates to the carbonyl oxygen of the ester, drops the LUMO energy of the dienophile, and the reaction becomes fast even with a moderately reactive diene. That was a useful reminder that textbook conditions and actual lab conditions are not always the same thing.
Regioselectivity and Stereochemistry
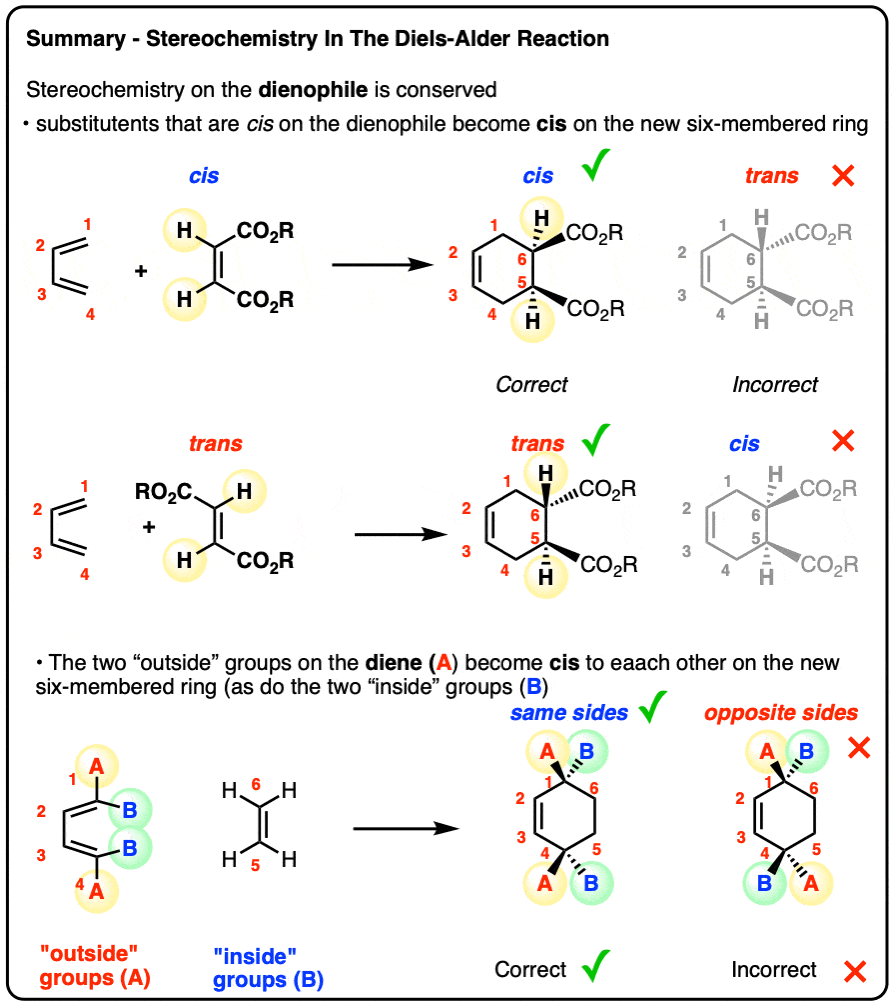
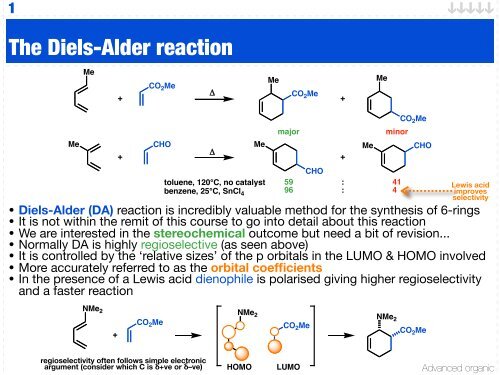
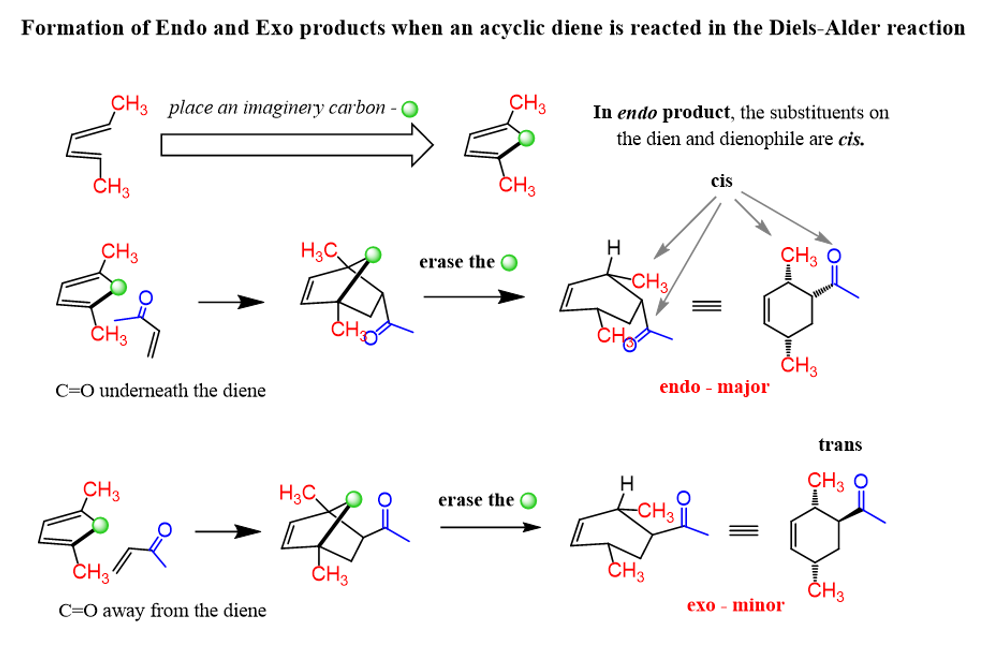
The Diels Alder Reaction Mechanism is stereospecific. The geometry of the dienophile is preserved in the product. A trans-disubstituted dienophile gives a trans-disubstituted ring. A cis one gives cis. This is not approximate, it is exact because the reaction is concerted. The diene also maintains its stereochemistry, and any substituents that were E or Z stay that way relative to the new ring. Regioselectivity follows from the electronic character of the substituents. With an electron-donating group on the diene and an electron-withdrawing group on the dienophile, the product distribution is usually predictable using the ortho and para rules. An ortho-like product dominates when the substituents are in the 1,2-relationship of the resulting cyclohexene, and a para-like product dominates for the 1,4-relationship. These are not absolute rules but they work reliably for simple systems. Complicated dienes with multiple substituents sometimes give mixtures that require chromatography to separate, and that can cost you 20 to 30 percent of your material. The endo rule matters here too. The endo product is usually the kinetic product because secondary orbital interactions stabilize the transition state. The exo product is more stable thermodynamically but forms more slowly. At lower temperatures, endo selectivity is high. At higher temperatures, you may get a mixture or even predominant exo product as the reaction approaches equilibrium. I had a case where running a Diels Alder at 120 degrees instead of 80 degrees switched the major product from endo to exo, and I wasted about three hours analyzing the wrong isomer before realizing what happened.
Get the Full Details
When the Diels Alder Fails Completely
There are several scenarios where this reaction simply will not work. If the diene is locked in the s-trans conformation by a rigid ring system, you cannot get the required geometry for the reaction. (E,E)-1,4-diphenyl-1,3-butadiene is unreactive under normal conditions because the phenyl groups create too much steric strain in the s-cis conformation. Some bicyclic dienes suffer from this problem routinely. You can sometimes overcome it with extreme pressure or by using a highly activated dienophile, but those are special cases. Polymerization is another real issue. Conjugated dienes can undergo radical or ionic polymerization, especially under heat. If your reaction mixture turns into a thick, insoluble gum instead of giving you a clean cycloadduct, the diene is polymerizing faster than it is reacting with the dienophile. Switching to a Lewis acid catalyst at lower temperature usually solves this because it accelerates the Diels Alder selectively while suppressing radical pathways. Running the reaction in a concentrated solution also helps, since intermolecular Diels Alder is second-order overall while polymerization often has higher kinetic order. Stretched dienophiles like tetrafluoroethylene are highly reactive but extremely difficult to handle safely. I once saw a student attempt a Diels Alder with hexafluorobut-2-yne in a standard Schlenk line setup. The pressure gauge spiked unexpectedly and the reaction vessel needed immediate cooling. High-pressure equipment is the standard for gaseous or volatile dienophiles, not improvisation.
Practical Setup Notes
If you are setting this up for the first time, start with a known system. Cyclopentadiene and maleic anhydride in xylene at reflux is about as reliable as it gets. It gives you a clean solid product that precipitates as it forms, which drives the equilibrium forward and makes isolation trivial. From there, you can explore variations. For substrate scopes that matter, consider whether your diene needs activation. Danishefsky's diene, which has a silyloxy group and a methoxy group, is highly reactive and tolerates a range of conditions including aqueous workup. It is the go-to choice when you need a hetero-Diels Alder variant that still delivers a useful cyclohexenone after workup. The mechanism involves an oxygen heteroatom in the diene, and the product contains a ketone that can be further functionalized. Inverse electron demand Diels Alder reactions flip the usual electronics. Here the diene is electron-poor and the dienophile is electron-rich. Tetrazines reacting with strained alkenes like trans-cyclooctene are the canonical example. These reactions are extraordinarily fast, sometimes with rate constants above 1000 per molar per second, and they are widely used in bioconjugation because they proceed cleanly in aqueous media at room temperature. They are not suitable for every synthesis, but they solve problems that traditional Diels Alder chemistry cannot touch.
The Diels Alder Reaction Mechanism remains one of the most dependable C-C bond-forming methods available, provided you respect the constraints. The geometry requirement, the electronic matching, the temperature dependence of selectivity, and the real possibility of competing side reactions are all factors you need to plan for before you mix the reagents. Good reactions are predictable. Bad ones usually are not, and the difference is almost always one of those factors being wrong.