So you need to actually distinguish these two nucleic acids in practice
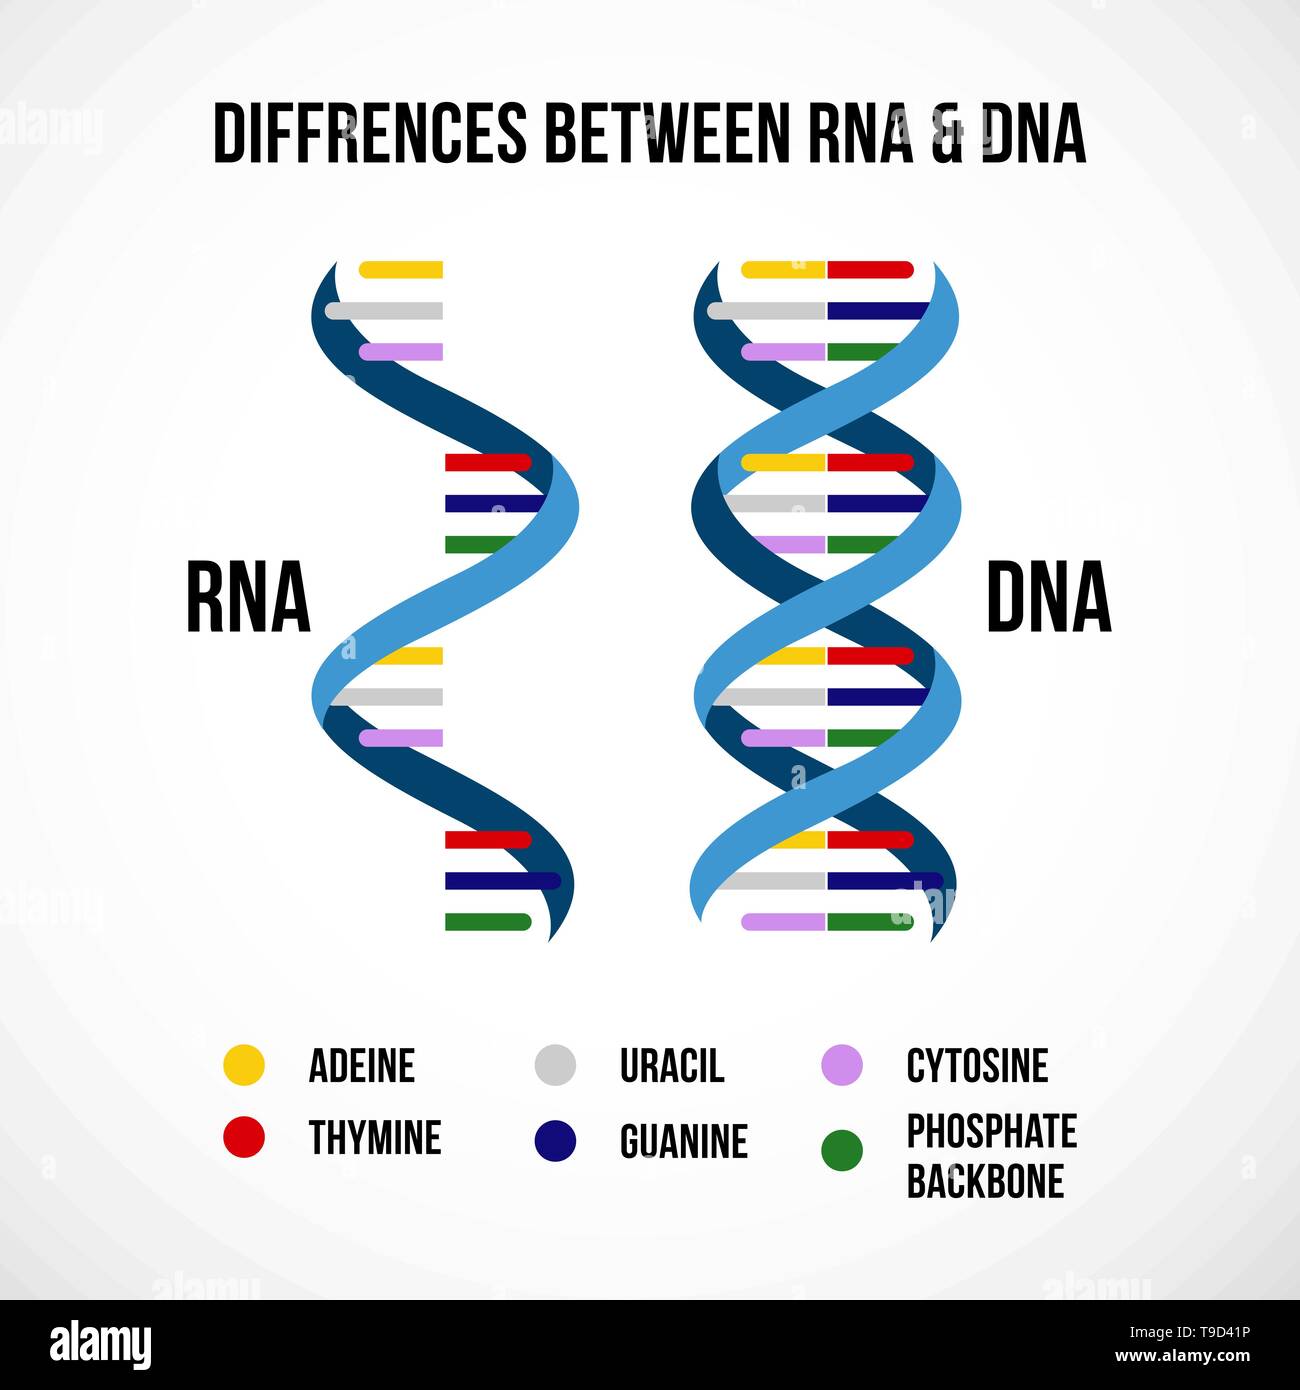
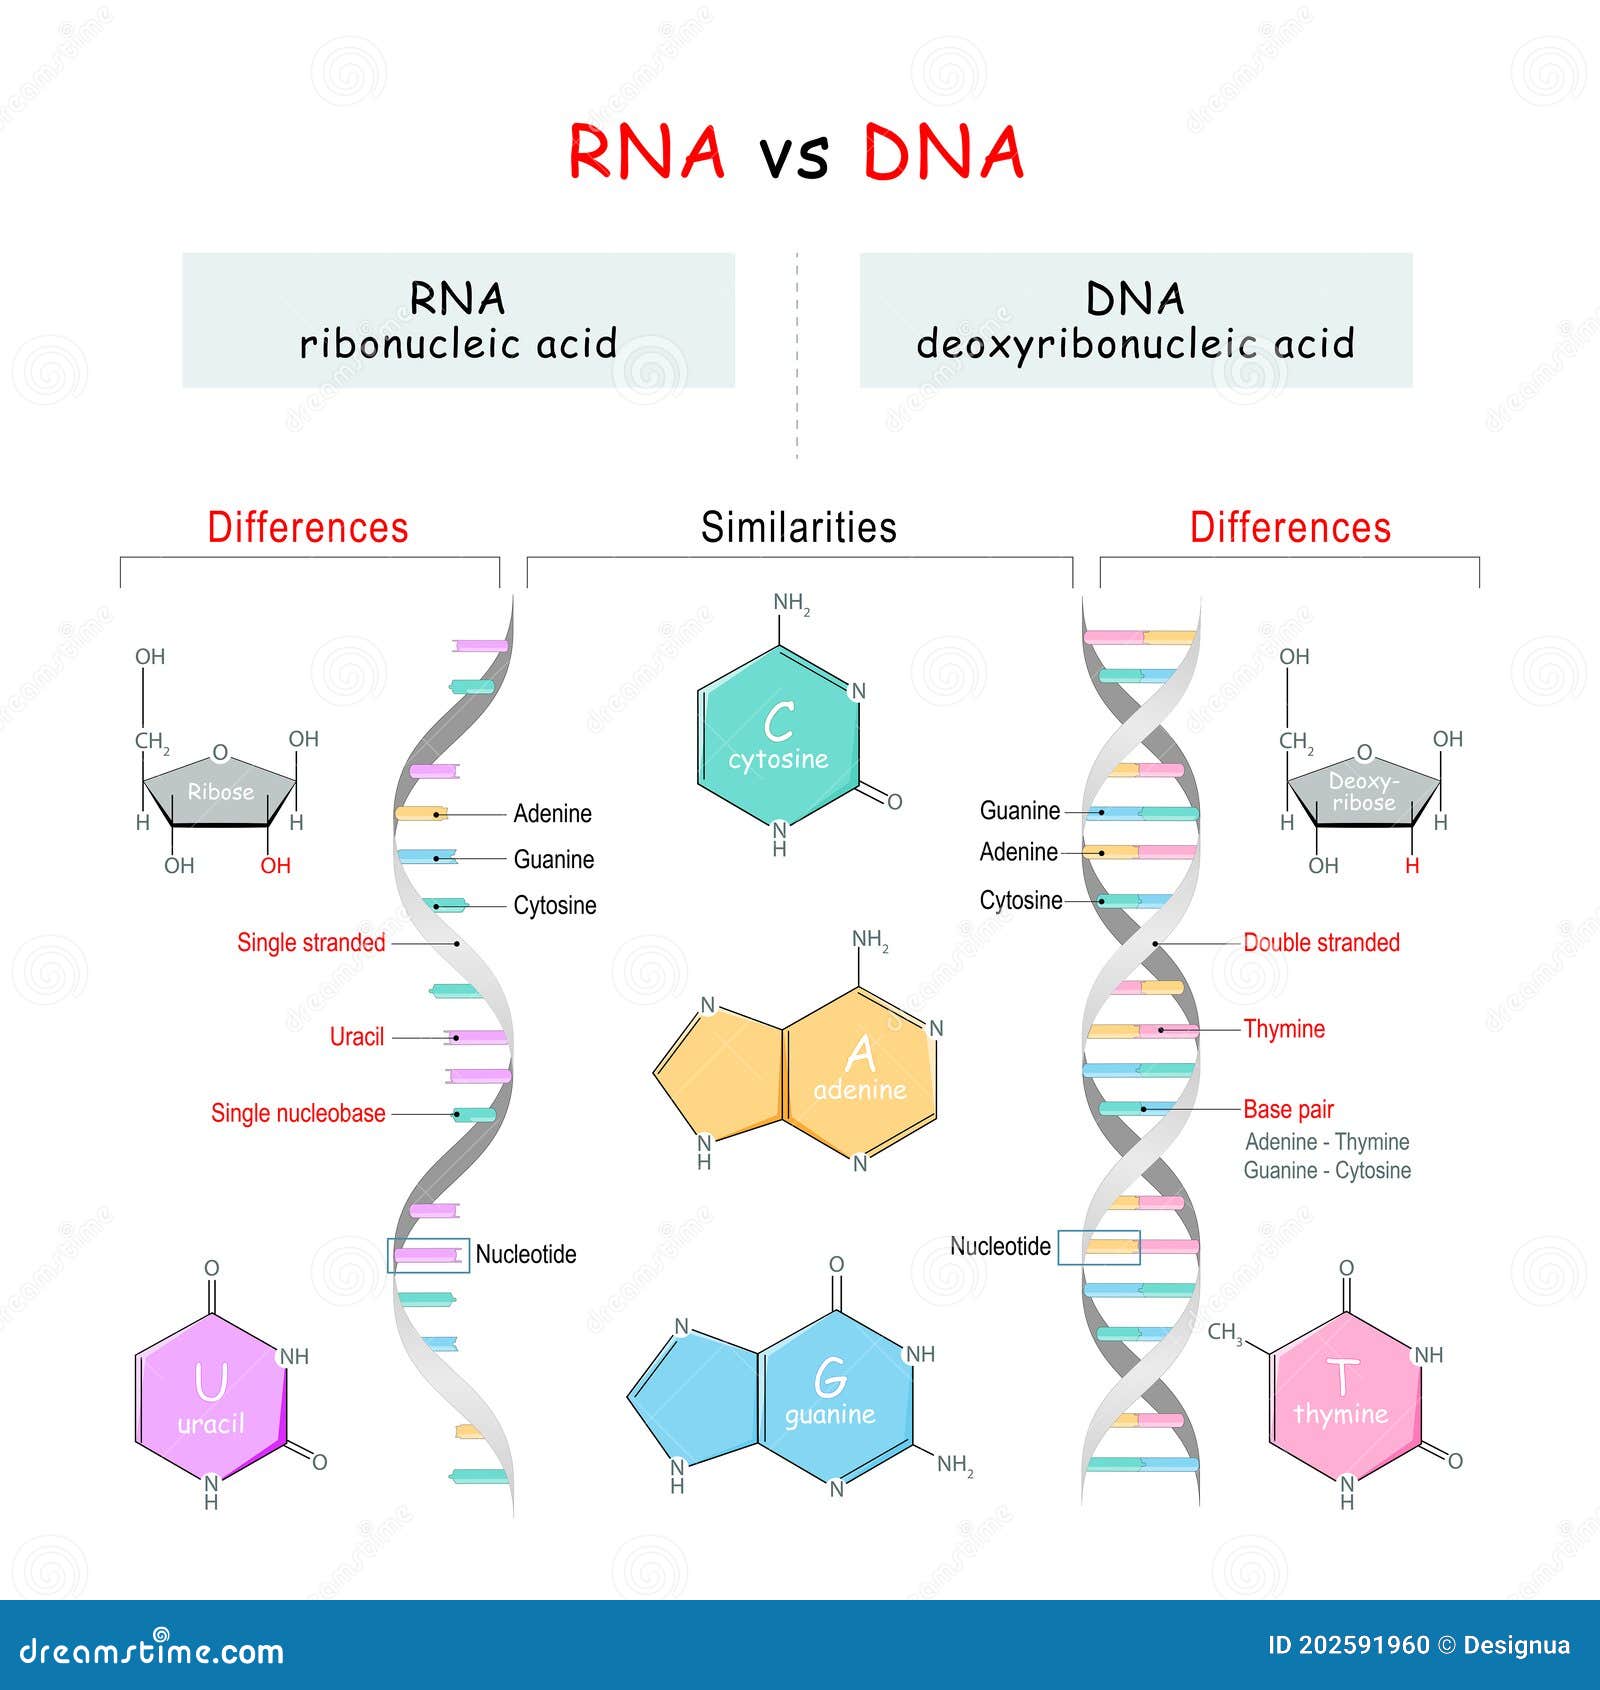
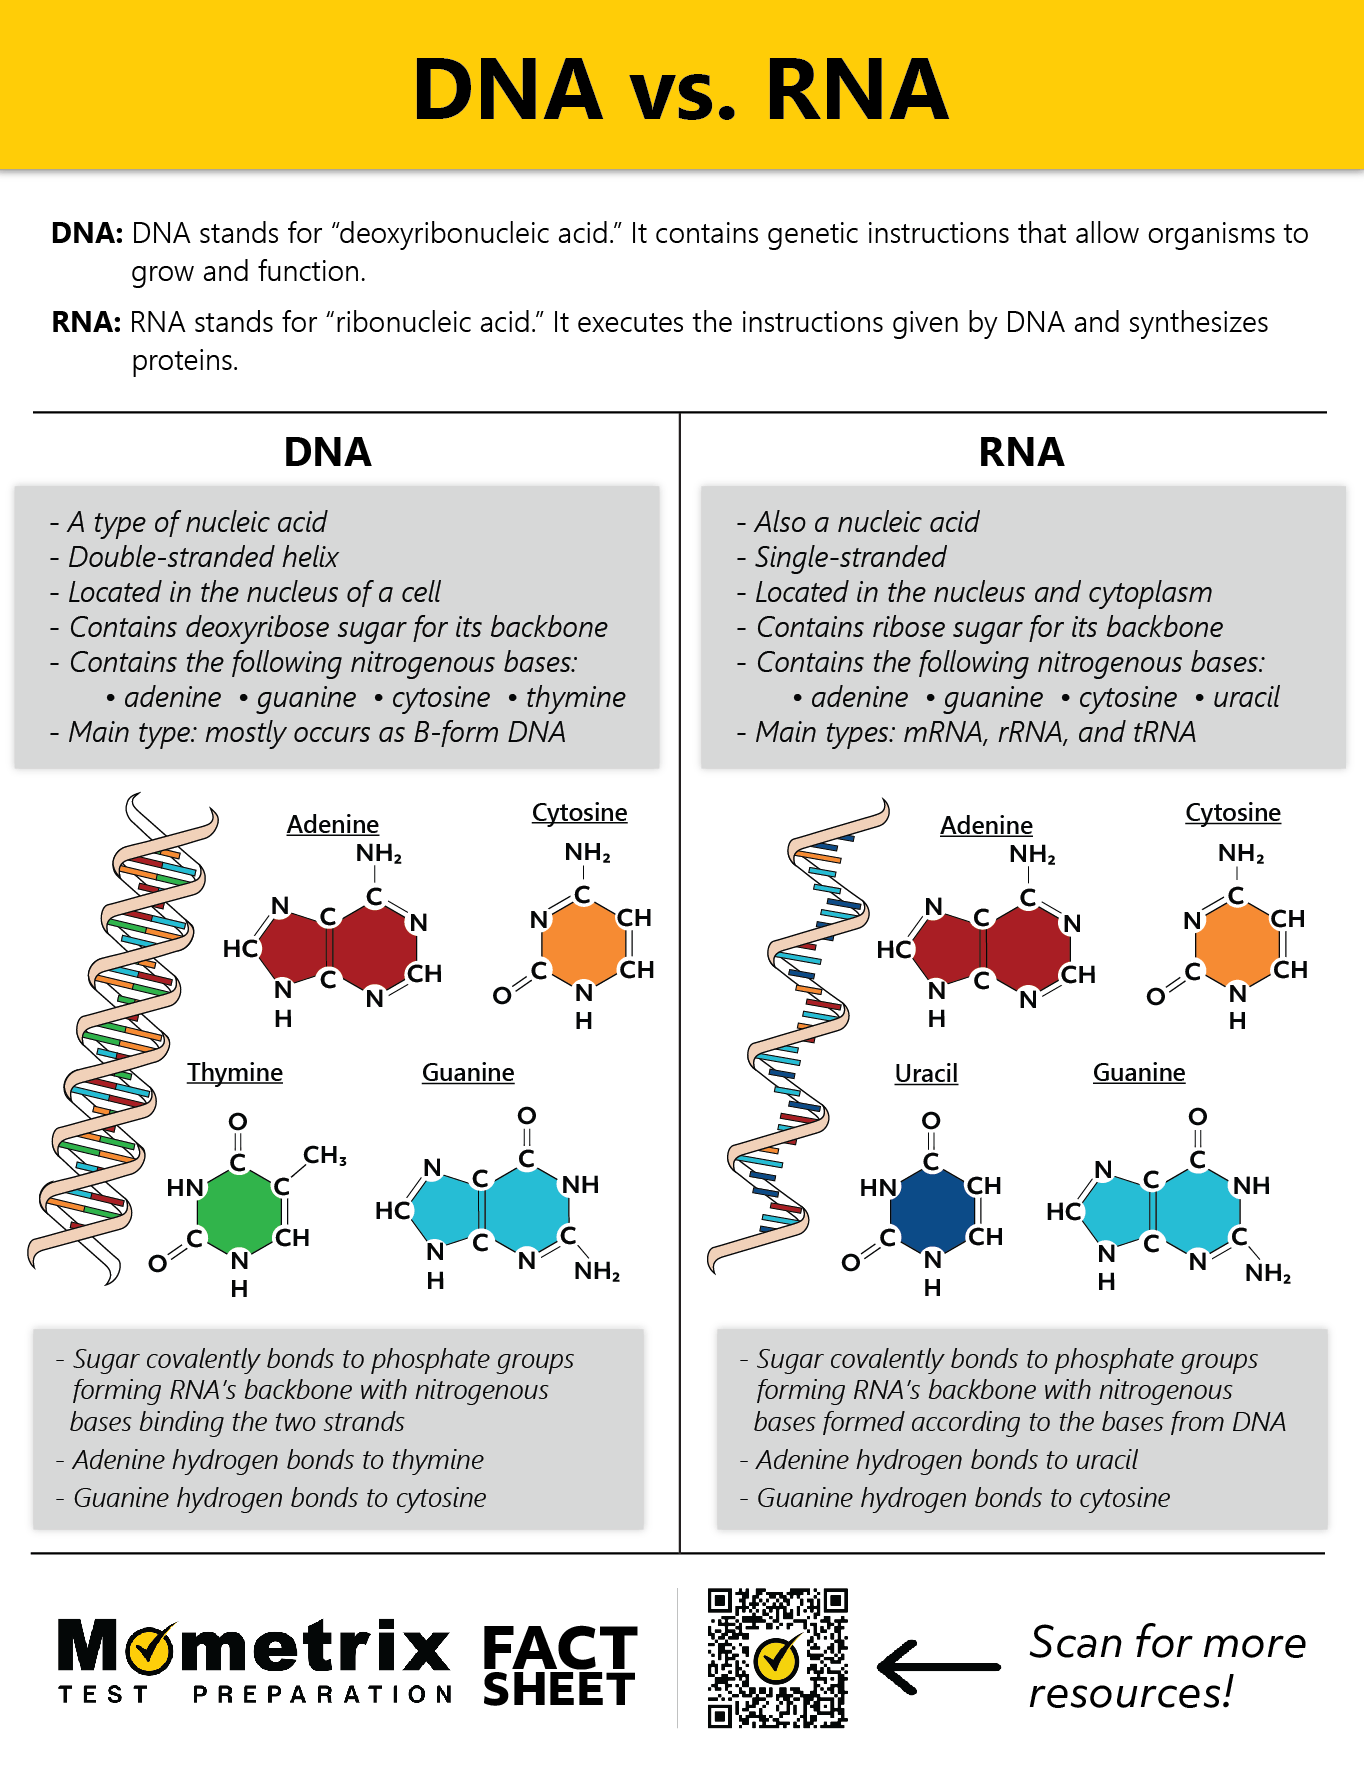
I spent way too many hours in a wet lab back when I was running RT-qPCR assays and got tripped up more than once by the fact that standard DNA sequencing primers would happily bind to your RNA samples if you weren't running DNase treatment. It's a small detail but it can ruin an entire experiment. The differences between Dna And Rna are not as abstract as most textbooks make them out to be. Once you understand what is physically different at the molecular level, a lot of the procedural headaches just go away. DNA is deoxyribonucleic acid and RNA is ribonucleic acid. That one word difference — deoxy vs ribo — is the entire root cause of everything else. The sugar in RNA has a hydroxyl group on the 2' carbon of the ribose ring. DNA has just a hydrogen there instead. That single oxygen atom makes RNA far more reactive and far less stable. In practice, RNA degrades much faster because that 2' OH can act as a nucleophile and cleave the phosphodiester backbone under basic conditions or even in certain cellular environments. DNA lacks that self-cleavage mechanism, which is one reason it ended up as the long-term storage molecule in almost all organisms. Both use adenine, guanine, and cytosine. The difference in bases is thymine versus uracil. DNA uses thymine, RNA uses uracil. Thymine is basically uracil with a methyl group slapped onto it. That methyl group costs energy to make, but it gives DNA an advantage: cytosine spontaneously deaminates to uracil all the time. If DNA used uracil natively, repair enzymes wouldn't be able to tell the difference between a legitimate uracil and one that showed up from deamination damage. With thymine in the mix, any uracil that pops up in DNA is immediately flagged as damage. RNA doesn't face that problem because it is already supposed to contain uracil and it has a short lifespan anyway.
Structure is another area where they diverge. DNA is almost always double-stranded and forms the familiar antiparallel double helix. RNA is typically single-stranded, but it folds back on itself into complex secondary and tertiary structures because complementary regions within the same strand can base pair. You get hairpins, stems, loops, pseudoknots, the whole thing. This structural diversity is why RNA can do catalytic work, which DNA essentially cannot do in a biological context.
Functional differences and why they matter for your protocols
DNA's job is storage. RNA's job is execution. That's the oversimplified version, but it holds up. DNA holds the genome, gets copied during replication, and generally stays out of the way. RNA comes in several flavors — mRNA, tRNA, rRNA, plus all the regulatory RNAs like microRNAs and siRNAs — and each one participates directly in gene expression or its regulation. The double helix of DNA provides redundancy. Each strand carries the information of the other, so if one strand gets damaged, the other can serve as a template for repair. Single-stranded RNA has no backup. That is another reason why RNA is inherently less stable and why RNA viruses tend to have higher mutation rates than DNA viruses. Their polymerases make more errors and the RNA genome itself is more susceptible to degradation. HIV is a classic example. Its RNA genome mutates fast enough that developing a vaccine has been exceptionally difficult. From a practical standpoint, if you are working with RNA in the lab, you need to treat it like something that wants to fall apart. RNases are everywhere — on your skin, in dust, on bench surfaces — and they are notoriously hard to destroy. Heat alone does not reliably inactivate them. I once ran a Northern blot where the control RNA looked perfectly intact and the experimental samples were completely degraded. Turned out the issue was a filter paper I had used to wipe down the bench earlier in the day. RNases had transferred to the surface and attacked my samples. I switched to treating the bench with RNase decontamination solution and baked my glassware instead, and the problem stopped. It sounds extreme but that is just how it is with RNA work.
Get the Full Details

Why base pairing rules apply differently in each molecule
Chargaff's rules — A pairs with T and G pairs with C — apply to double-stranded DNA. In RNA, base pairing is more flexible because of the structural complexity. You get wobble pairing in tRNA anticodons, non-canonical base pairs in ribozymes, and G-U wobble pairs that are surprisingly common in RNA secondary structures. A G and a U can form two hydrogen bonds instead of the usual three, and the RNA machinery generally accepts this without complaint. This is functionally important because it allows a single tRNA to recognize multiple codons, which is why the genetic code has degeneracy built into it. When you design primers or probes, keep in mind that DNA-DNA hybridization is more rigid and predictable than RNA-RNA or RNA-DNA hybridization. The melting temperature formulas you find online are calibrated for double-stranded DNA and they will give you approximate but not exact results for RNA duplexes. If you need precise Tm values for RNA oligos, you should use a nearest-neighbor model specifically parameterized for RNA rather than the simple Wallace rule most people reach for.
Reverse transcription is where the practical differences bite hardest
If you ever need to convert RNA into DNA — which is necessary for things like RT-qPCR, RNA-seq library prep, or cDNA cloning — you are relying on reverse transcriptase. This enzyme reads an RNA template and synthesizes a complementary DNA strand. The resulting molecule is called cDNA, and it looks chemically identical to regular DNA. That is the trick. Once you have cDNA, you can treat it like DNA for all downstream steps. One thing that catches people off guard is that reverse transcriptases vary significantly in their fidelity. Some, like M-MLV variants with point mutations, have reduced RNase H activity and give you longer full-length cDNA products. Others are higher fidelity but slower. If you are doing RNA-seq, read length and coverage uniformity depend a lot on which reverse transcriptase you choose. I found through trial and error that using a thermostable reverse transcriptase at higher temperatures (around 55°C) helped me get through secondary structures in GC-rich transcripts that otherwise caused premature termination. Standard M-MLV at 42°C left big gaps in my coverage until I switched protocols.
What about epigenetic differences?
DNA methylation is a major epigenetic mechanism. Methyl groups added to cytosine bases, usually in CpG contexts, can silence gene expression without changing the underlying sequence. RNA can also be chemically modified, and this is sometimes called the epitranscriptome. The most common RNA modification is N6-methyladenosine, or m6A, which affects mRNA stability, splicing, and translation efficiency. These modifications are dynamic and reversible, which is conceptually different from DNA methylation, where the marks are generally more stable across cell divisions. This is relevant if you are comparing gene expression data across experiments. RNA modifications can affect how efficiently a reverse transcriptase reads through a given transcript. Some m6A sites cause RT stop artifacts that show up as drops in sequencing coverage. It is easy to mistake a modification-driven coverage drop for a real biological signal if you do not know to look for it. I ran into this when analyzing RNA-seq data from a paper where the authors reported differential expression in a gene I knew was heavily modified. After cross-checking with m6A mapping data, the "differential expression" disappeared. It was just uneven reverse transcription.

Size and composition differences that matter for sequencing
DNA genomes vary enormously in size. A human genome is roughly 3 billion base pairs per haploid set. Bacterial genomes are on the order of a few million base pairs. RNA genomes are generally much smaller. The influenza virus genome is about 13,500 nucleotides split across eight segments. SARS-CoV-2 is around 30,000 nucleotides and is actually one of the largest known RNA virus genomes. There is a size ceiling for RNA genomes, probably because the error rate of RNA-dependent RNA polymerases scales with genome length, and beyond a certain point mutations accumulate faster than selection can remove them. This is known as the error threshold, and it is estimated to be somewhere around 30,000 to 100,000 nucleotides for most RNA viruses. For practical sequencing work, this means that RNA sequencing inherently involves an extra conversion step compared to DNA sequencing. You cannot load raw RNA directly onto most standard DNA sequencers. You have to make cDNA first, which adds time, cost, and potential bias. Long-read RNA sequencing technologies like PacBio's Iso-Seq or Oxford Nanopore's direct RNA sequencing attempt to bypass some of this, but direct RNA sequencing still has higher error rates than DNA sequencing and requires specialized library prep workflows that are not as widely adopted yet.
When DNA and RNA share more in common than you might think
Both are polyanions. Both have a sugar-phosphate backbone. Both use the same four-base coding system with minor variations. Both can form double helices under the right conditions. The differences are real but they are ultimately built on the same chemical foundation. That is why primers designed for DNA amplification can sometimes work in RT-PCR, and why many molecular biology reagents are compatible with both molecules. The main practical takeaway is that RNA requires more care but rewards you with information that DNA sequencing simply cannot give you. Gene expression levels, splice variants, non-coding RNA populations, RNA editing events — all of that is invisible to standard DNA sequencing. If you only sequence DNA, you are seeing the blueprint but not what the building is actually doing at any given moment. The tradeoff is that RNA work is fragile, temperamental, and demands clean technique throughout. If you respect that, it works fine. If you treat it like DNA, it will fall apart, literally.