How Chemical Bonds Actually Form
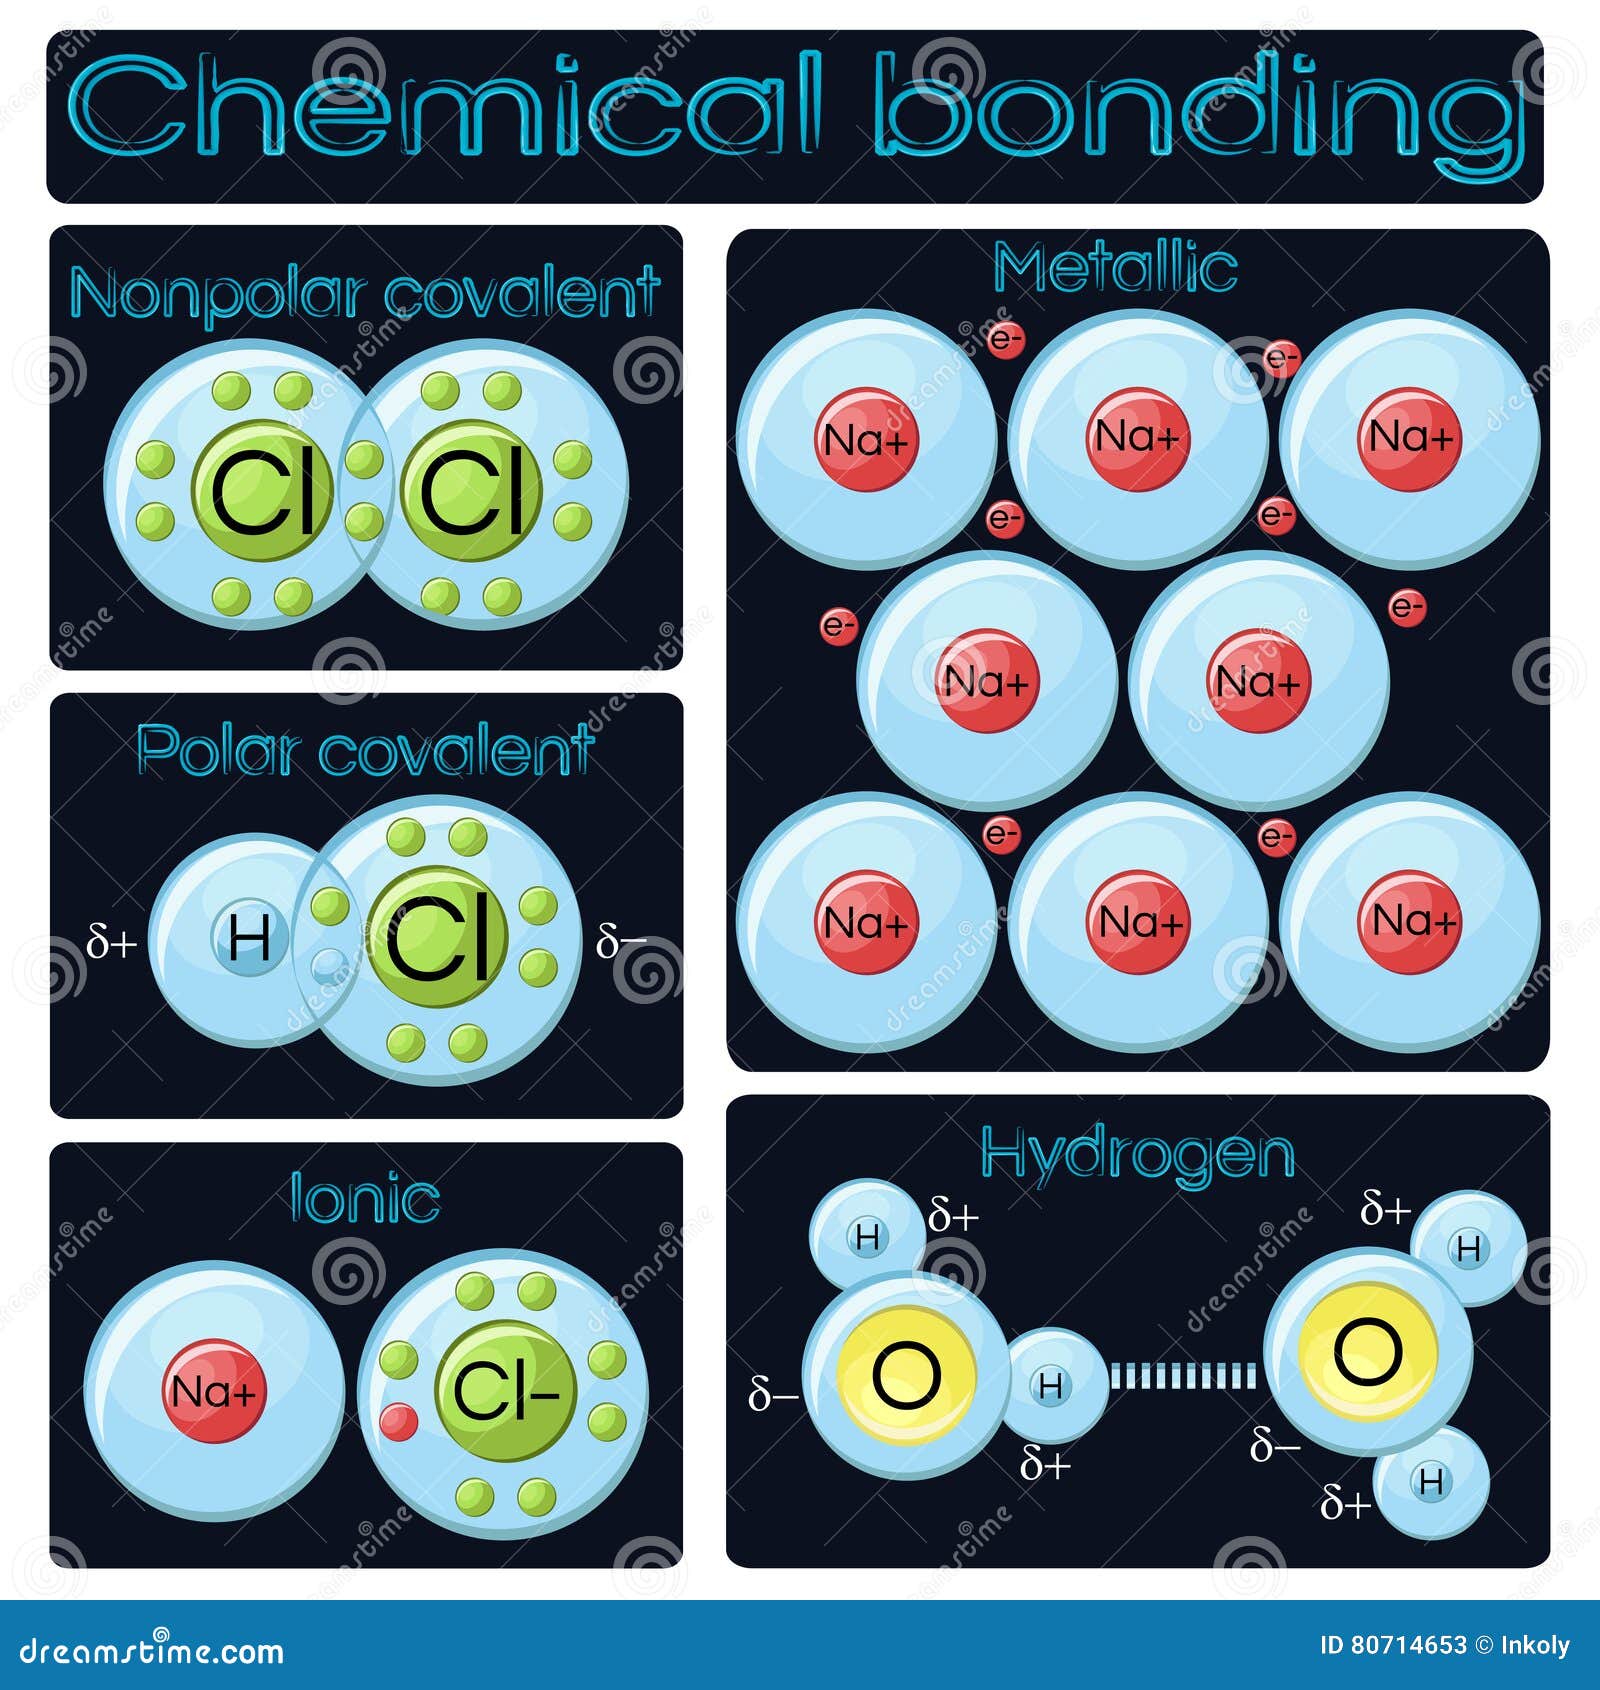
Most people learn about ionic, covalent, and metallic bonding in high school chemistry and think they've mastered the topic. They haven't. The reality is messier, more nuanced, and a lot more interesting than the three-category model suggests. When you step into actual lab work or materials science, the boundaries between bonding types blur constantly. That's where things get tricky. Let's start with what's actually practical rather than what's in the textbook. Ionic bonding happens when one atom hands off electrons to another, creating ions that stick together through electrostatic attraction. Sodium chloride is the classic example. But here's what instructors gloss over: purely ionic bonding barely exists. Even CsF, the most ionic compound you can name, has some covalent character. Pauling's electronegativity scale tells the story. The difference between cesium (0.79) and fluorine (3.98) is 3.19, which sits right around 91% ionic by most calculations. The rest is polarization effects, electron sharing creep, all of it. You need to know this because if you're working with supposedly ionic compounds and your measurements don't match theoretical lattice energies, this is why. Covalent bonding is where atoms share electron pairs. Simple enough. The complication comes in distinguishing sigma from pi bonds, understanding hybridization as a mathematical model rather than physical reality, and recognizing that bond order isn't always an integer. Benzene has a bond order of 1.5 for its carbon-carbon bonds because of resonance delocalization. Molecular orbital theory explains this cleanly, but even MO theory has limits when you get into transition metal complexes or excited states.
Metallic bonding is the hardest to pin down conceptually. You have a lattice of positive ions bathed in a sea of delocalized electrons. That's the model. In practice, the electrons aren't truly delocalized across the entire structure in most metals. You get band structure instead, with valence bands and conduction bands separated by varying energy gaps. That's why some metals conduct electricity better than others, and why the concept of "electron sea" breaks down when you're trying to predict actual material properties. There are also bonding types that sit between these categories and cause a lot of confusion. Hydrogen bonding isn't a true chemical bond in the same sense as the others. It's an electrostatic interaction between a hydrogen atom covalently bonded to an electronegative atom and another electronegative atom nearby. It's stronger than van der Waals forces but weaker than covalent bonds, typically in the 4 to 40 kJ/mol range. Water's anomalous properties, DNA base pairing, protein folding—all of it depends on hydrogen bonding. Don't underestimate how much of chemistry is just hydrogen bonds holding things together. Coordinate covalent bonds, or dative bonds, occur when one atom provides both electrons for the shared pair. This is central to coordination chemistry and Lewis acid-base theory. Ammonia donating its lone pair to a metal ion like copper in the formation of the tetraamminecopper(II) complex is a textbook example. These bonds are chemically indistinguishable from regular covalent bonds once formed. The distinction only matters for tracking electron movement in reaction mechanisms.
What I Learned the Hard Way
I spent a week troubleshooting a synthesis that kept producing unexpected products because I was treating a supposed ionic intermediate as fully ionic. The compound in question was an alkali metal complex with a large organic ligand. The literature described the metal-ligand interaction as ionic. X-ray crystallography later showed significant electron density between the metal and the coordinating atoms. It was a coordinate covalent bond disguised as ionic interaction by the simplicity of the structural model. I ended up having to re-evaluate the entire reaction mechanism once I recognized the partial covalent character. That cost me three days and a failed batch. Now I always check the electronegativity differences and consider polarization effects before assuming pure ionic behavior, especially with large organic anions that can distort electron clouds significantly. Another thing nobody warns you about: bond energies are averages. The C-H bond energy listed in tables is roughly 413 kJ/mol, but that value shifts depending on what else is attached to the carbon. A C-H bond next to a carbonyl group is weaker because the resulting radical is stabilized by resonance. If you're calculating reaction enthalpies using tabulated bond energies, expect errors in the range of 5 to 15 percent. For rough estimates that's fine. For precise work, you need formation enthalpies from calorimetry or high-level computational chemistry.
Get the Full Details

Pitfalls and Where the Models Fail
The biggest problem students and even practitioners run into is applying the wrong bonding model to a system where it doesn't fit. Take transition metal oxides. You might try to classify them as ionic, but many of them are actually covalent networks with significant metal-metal bonding. MgO is genuinely ionic. TiO2 is not. The difference comes down to charge density and polarizability. Small, highly charged cations like Ti4+ distort the electron clouds of nearby anions enough to create substantial covalent character. This is Fajans' rules in action, and it matters enormously if you're designing materials for catalysis or battery electrodes where bonding type determines electronic structure and reactivity. Another failure mode is assuming that bond length correlates linearly with bond strength across different bond types. A C-C single bond is about 154 pm. A C=C double bond is 134 pm. A CC triple bond is 120 pm. The trend looks straightforward. But compare that to N-N bonds. NN is incredibly short at 110 pm and one of the strongest bonds in chemistry at 945 kJ/mol. N-N single bonds in hydrazine are actually longer and weaker than you'd expect because of lone pair repulsion between the nitrogen atoms. The simple correlation between bond order and bond strength breaks down when you introduce lone pairs, steric effects, or ring strain. Van der Waals forces, including London dispersion forces, are often dismissed as negligible. They're not. In large organic molecules and supramolecular assemblies, dispersion interactions can contribute tens of kilojoules per mole to binding energy. This is why nonpolar hydrocarbon chains stack together in lipid bilayers and why fullerenes form crystalline solids despite having no permanent dipole moments. If you're modeling molecular recognition or self-assembly and ignoring dispersion, your results will be qualitatively wrong.
Practical Approach to Identifying Bonding Types
When you're actually working with a compound and need to determine its bonding character, start with electronegativity differences. A difference greater than about 1.7 suggests ionic character. Below 0.4 suggests nonpolar covalent. Between those values, you're in the polar covalent zone where things get complicated. But electronegativity alone won't save you. Look at the physical properties. Ionic compounds tend to be brittle solids with high melting points that conduct electricity when molten. Covalent compounds vary wildly—diamond is hard and insulating, graphite is soft and conductive, water is liquid at room temperature. Metallic compounds conduct electricity in the solid state and are malleable. Spectroscopic data is more reliable than any rule of thumb. X-ray photoelectron spectroscopy can tell you about oxidation states and bonding environments. Infrared spectroscopy reveals bond strengths through vibrational frequencies. A C=O stretch around 1700 cm¹ indicates a typical carbonyl, but if it shifts to 1650 cm¹, you've got conjugation or hydrogen bonding weakening the bond. Raman spectroscopy complements IR for symmetric vibrations that IR misses. If you have access to X-ray diffraction, bond lengths and angles from crystal structures give you direct evidence of bonding character. Computational chemistry has become the go-to tool for problems where experimental data is ambiguous. DFT calculations can map electron density distributions, showing you exactly where bonding electrons are concentrated. Is there electron density between two atoms? That's a bond. Is the density skewed toward one atom? That's polar bonding. Are the electrons delocalized over multiple atoms? You're looking at resonance or aromaticity. The calculations take time and require understanding of the underlying theory, but they remove a lot of the guesswork that used to dominate the field.
The key takeaway is that bonding exists on a spectrum, not in discrete categories. Your job is to figure out where a given interaction sits on that spectrum and use the appropriate model for whatever you're trying to do. The textbook categories are starting points, not conclusions.


