Setting Up the Grid
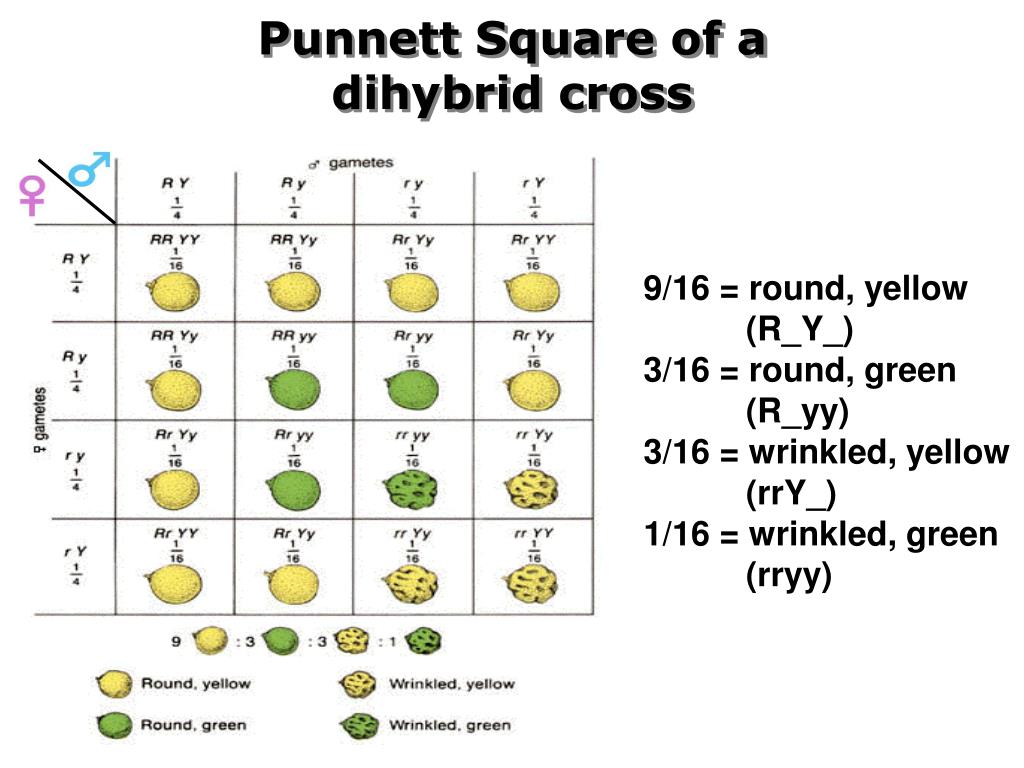
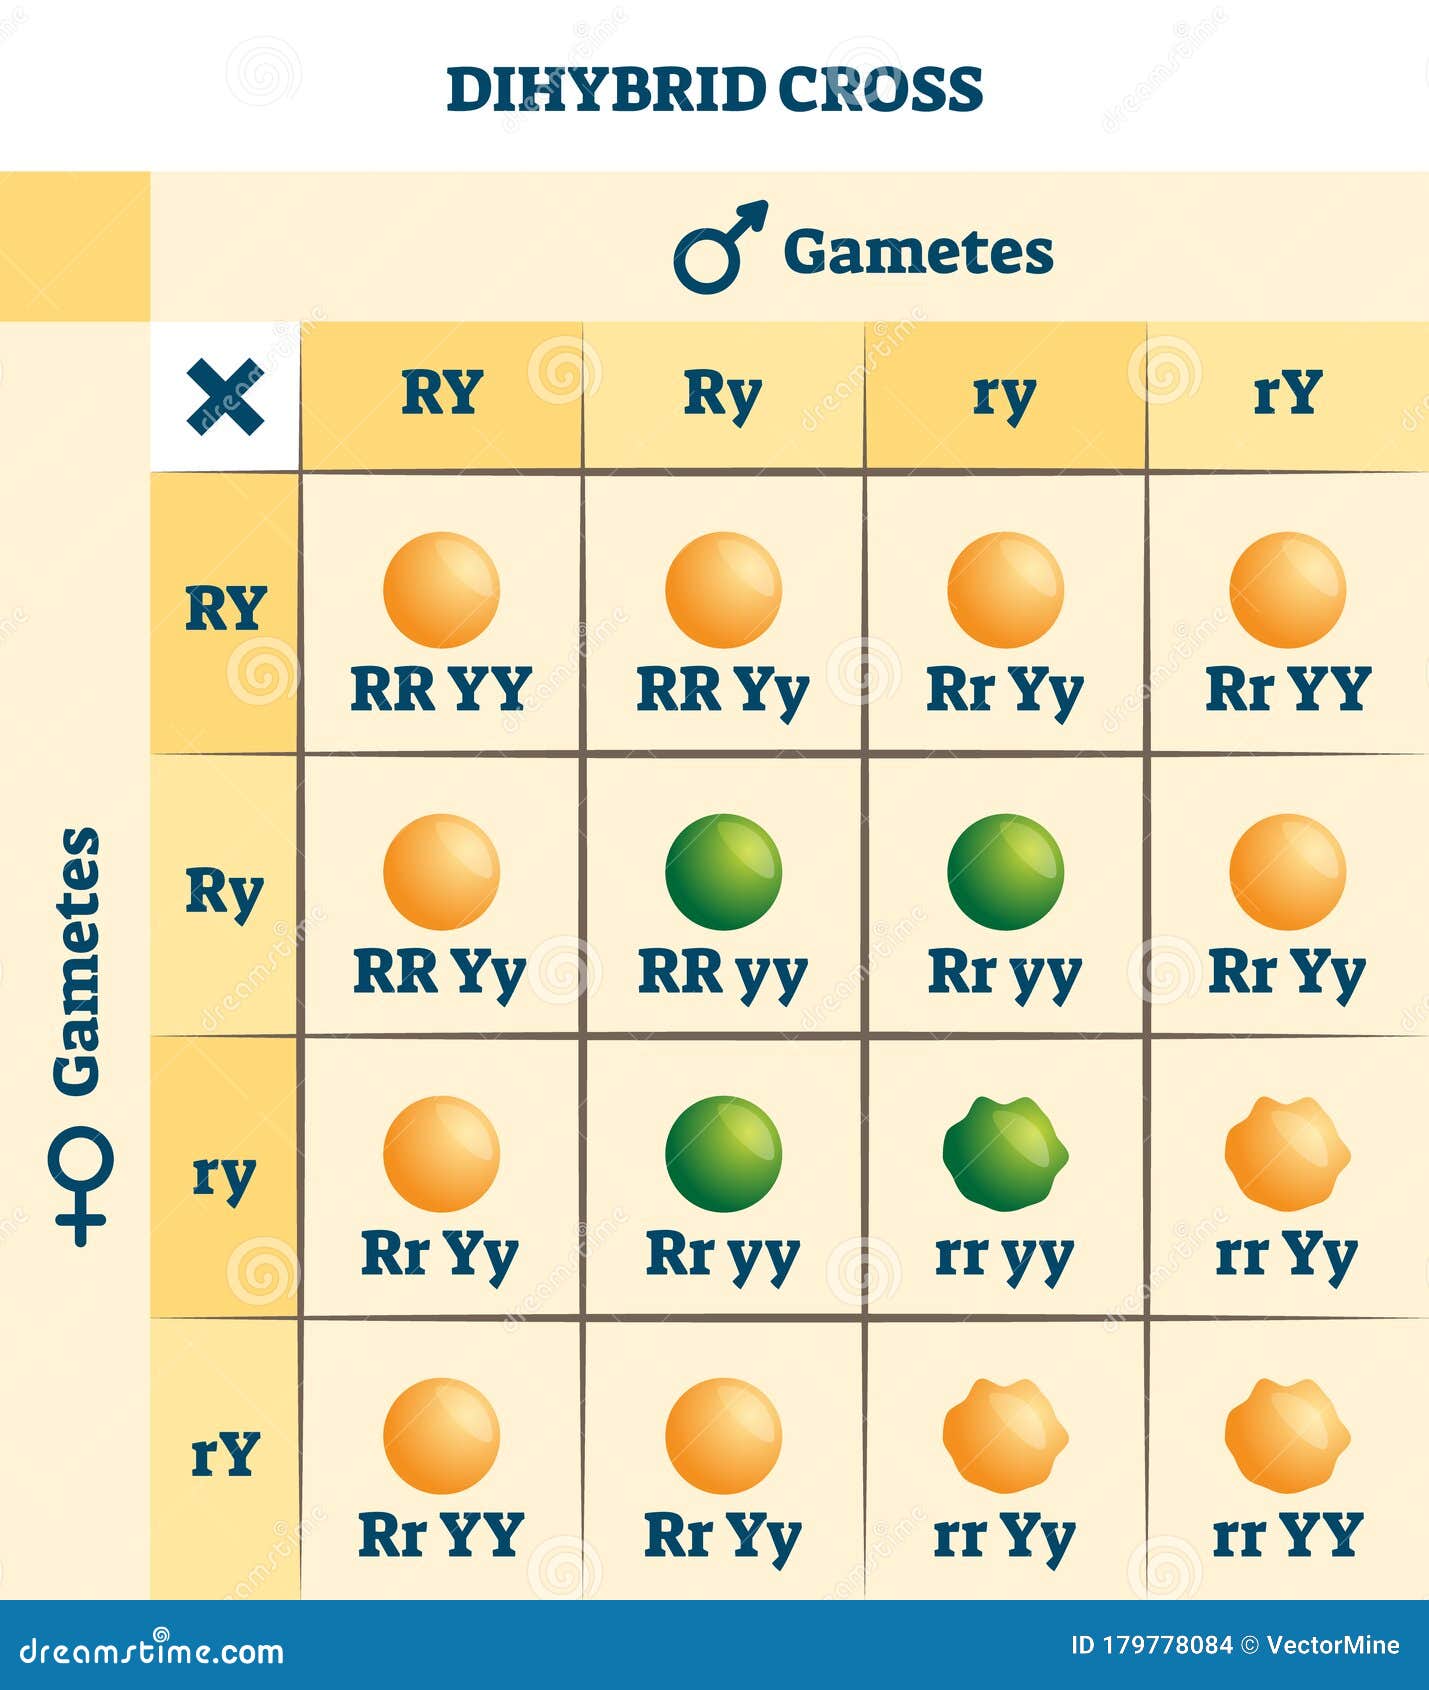
A dihybrid cross tracks two traits at once, which means each parent produces four types of gametes instead of two. You start by writing out those gamete combinations, then cross-reference them in a 4x4 grid that holds 16 boxes total. Take a cross between two heterozygotes for both traits—AaBb times AaBb. Each parent can produce AB, Ab, aB, and ab gametes. Those four go across the top and down the side, and each box in the middle shows one possible genotype outcome for an offspring. The result gives you that textbook 9:3:3:1 phenotypic ratio when both traits show complete dominance and the genes assort independently. Nine boxes will show both dominant phenotypes. Three will show the first dominant and the second recessive. Another three will flip that. And one box will show both recessive traits.
Dihybrid Cross Punnett Square Setup and Interpretation
Most people learn the grid layout before understanding what it actually represents. That's backwards. The grid is just a visual multiplication table for gamete combinations. Each box represents one fertilization event. The genotype inside that box combines whatever allele pair came from the top parent with whatever allele pair came from the left parent. If the top says AB and the side says ab, that box is AaBb. Simple, but it's easy to misread the alleles if you're not writing them carefully. I ran into a specific problem last year while grading labs. A student submitted a dihybrid cross where they had written the parental genotypes as AaBb but then only listed three gamete types instead of four. They had missed the Ab combination entirely. The resulting ratios were completely wrong, and the student couldn't figure out why. The issue wasn't a calculation error. It was a fundamental misunderstanding of how you derive gamete types from a heterozygous genotype. I had them redo it using a simple branching diagram before they attempted the full grid again. Once they could list the gametes correctly without the grid crutch, the rest clicked into place. The gamete derivation step is where most mistakes happen. Write the two gene pairs side by side—Aa and Bb. Then systematically combine every allele from the first gene with every allele from the second gene. A with B gives AB. A with b gives Ab. a with B gives aB. a with b gives ab. That ordering matters because rushing it is how people skip a combination or duplicate one by accident.
Reading the Results
Once the grid is filled, you read it in two layers. First, identify the genotype in each box. Second, convert those genotypes into phenotypes based on whatever dominance relationships exist. If both traits show complete dominance, any genotype with at least one dominant allele for that trait expresses the dominant phenotype. So AaBB, AABB, and AABb all look the same phenotypically because they carry at least one dominant A and at least one dominant B. Count up the phenotypes across all 16 boxes and you get 9 with both dominant traits, 3 with only the first dominant, 3 with only the second dominant, and 1 with both recessive. That ratio holds as long as the genes are on different chromosomes or far enough apart on the same chromosome that they assort independently. If you need the exact genotypic ratios instead of phenotypic ones, you can't just group them loosely. Each of the 16 boxes contains a unique or semi-unique genotype combination. AABB appears once. AABb appears twice. AaBB appears twice. AaBb appears four times. AAbb appears once. Aabb appears twice. aaBB appears once. aaBb appears twice. aabb appears once. So the genotypic ratio is 1:2:1:2:4:2:1:2:1. Most people don't need that level of detail, but it comes up when you're working with probability questions that specify exact genotypes.
Get the Full Details

When the Standard Method Falls Apart
The dihybrid cross Punnett Square assumes independent assortment. That means the allele a gamete receives for one gene tells you nothing about what it receives for the other gene. This assumption is false whenever the two genes are linked on the same chromosome. When genes are linked, parental combinations of alleles stay together more often than recombinant combinations. The 9:3:3:1 ratio breaks down completely. You get far more of the parental phenotype classes and far fewer recombinant types. I worked with a research lab a few years back that was studying a dihybrid cross in a species where two of their marker genes turned out to be only about 5 centimorgans apart. Their observed offspring ratios looked nothing like the expected 9:3:3:1. They spent two weeks troubleshooting before someone suggested checking the genetic map distance. Once they accounted for linkage and calculated the recombination frequency, the numbers made sense. The Punnett square wasn't wrong. The assumption behind it was wrong for their specific case. There are other situations where a standard Punnett square doesn't work well. Epistasis changes the phenotypic ratios entirely. If one gene masks the expression of another, that clean 9:3:3:1 can become something like 9:3:4 or 12:3:1 or 9:7. The grid still shows the right genotypes. The phenotype interpretation just gets more complicated. Sex-linked genes are another case where a basic dihybrid square needs adjustment because males and females inherit X-linked alleles differently. You have to split the cross by sex or use a modified grid setup.
A Faster Alternative for Independent Assortment
If you're confident the genes assort independently, you don't always need the full 16-box grid. The forked-line method or probability multiplication does the same work faster and with fewer places to make an error. Split the dihybrid cross into two separate monohybrid crosses. For AaBb times AaBb, that's Aa times Aa and Bb times Bb. The first cross gives you a 3:1 dominant-to-recessive ratio. The second does too. Multiply the probabilities: 3/4 times 3/4 gives you 9/16 for both dominant. 3/4 times 1/4 gives 3/16 for first dominant and second recessive. And so on. It takes about 30 seconds and produces the same phenotypic ratios. I recommend this approach for homework and exam situations where speed matters. The grid is useful for learning the mechanics and for cases where the assumptions need to be visualized explicitly. But in practice, most geneticists I know use the probability method for quick calculations and reserve the Punnett square for teaching or for crosses where linkage or other complications need to be laid out visually.
Common Mistakes to Watch For
The most frequent error is writing the wrong gametes. People sometimes write AABB as a gamete, which is impossible because a gamete carries only one allele per gene. Gametes are haploid. They carry one allele from each gene locus, never two copies of the same gene. This mistake usually comes from confusing the parental genotype with the gamete genotype. Parents are diploid. Gametes are haploid. Another common issue is misaligning the alleles when filling in the boxes. Write the alleles from the same gene together. AaBb, not ABab. The order doesn't change the biology, but inconsistent notation makes it easy to misread your own work later, especially when you're scanning 16 boxes under time pressure. People also forget to label which trait is which after filling in the grid. Without clear labels, distinguishing between the 3 boxes of first-dominant-second-recessive and the 3 boxes of first-recessive-second-dominant becomes guesswork. Write the trait names above the columns and to the left of the rows. It adds ten seconds to the setup and saves you from second-guessing your own counts later.

What This Method Cannot Tell You
The Punnett square is a tool for predicting outcomes under idealized conditions. It does not account for gene conversion, chromosomal nondisjunction, incomplete penetrance, variable expressivity, or environmental effects on phenotype. If any of those factors are at play, the actual offspring ratios will deviate from the predicted ones, and no amount of careful grid-filling will fix that. The square predicts probabilities, not certainties. A 9:3:3:1 ratio describes what you expect across a large number of offspring. With small sample sizes, random sampling variation can produce results that look wildly off even when the underlying genetics are perfectly normal. For linked genes, the square gives wrong predictions unless you manually adjust for recombination frequency. For epistatic interactions, it gives the right genotypes but the wrong phenotype ratios unless you apply the epistatic modification afterward. The method itself is neutral. It's the assumptions you bring to it that determine whether the output is meaningful for your particular cross.