Running DMA Without Losing Your Mind
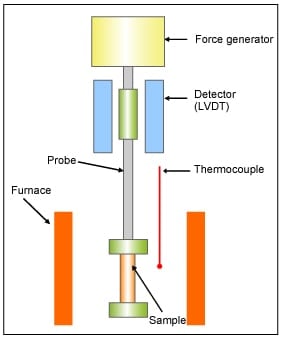
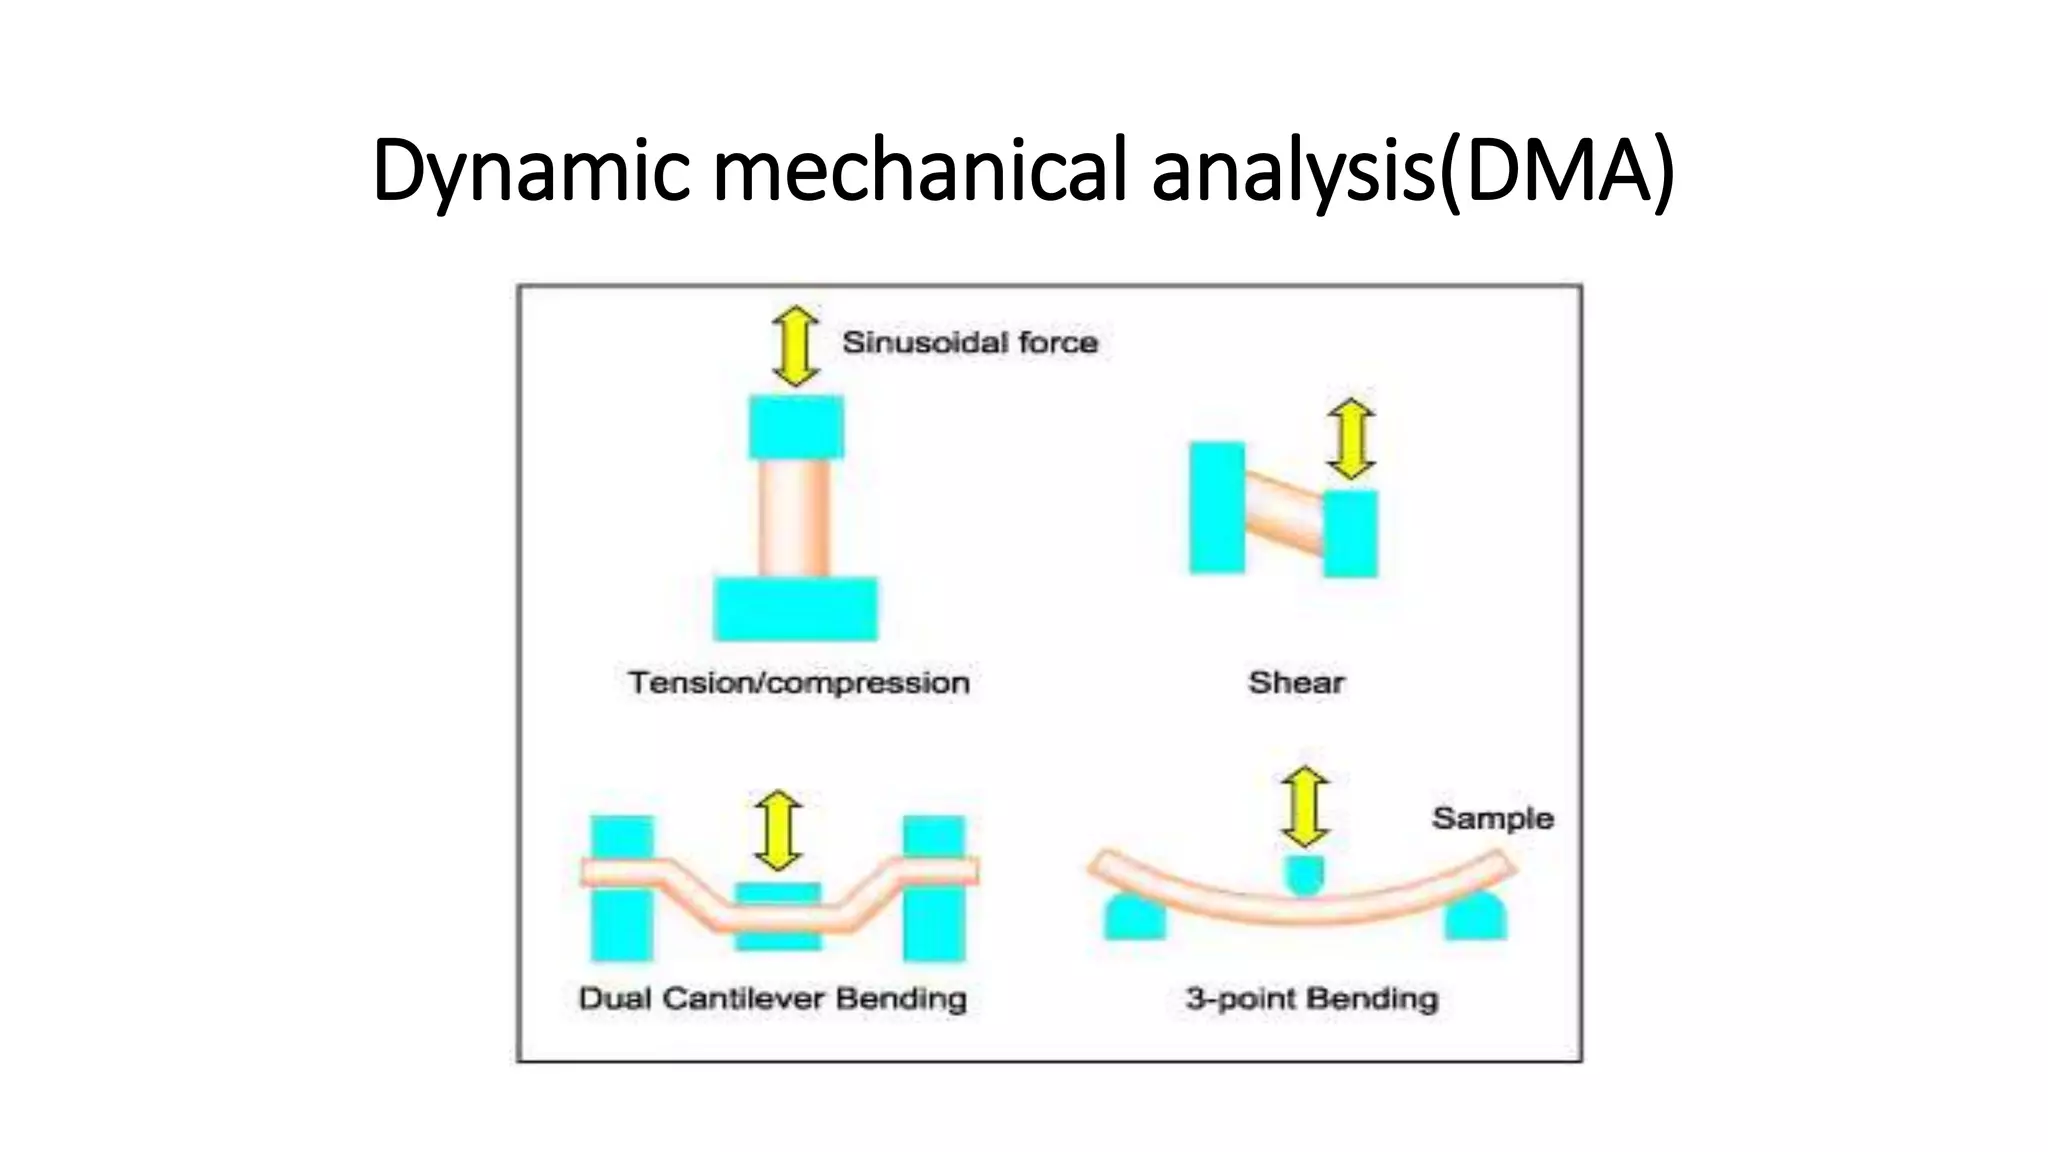
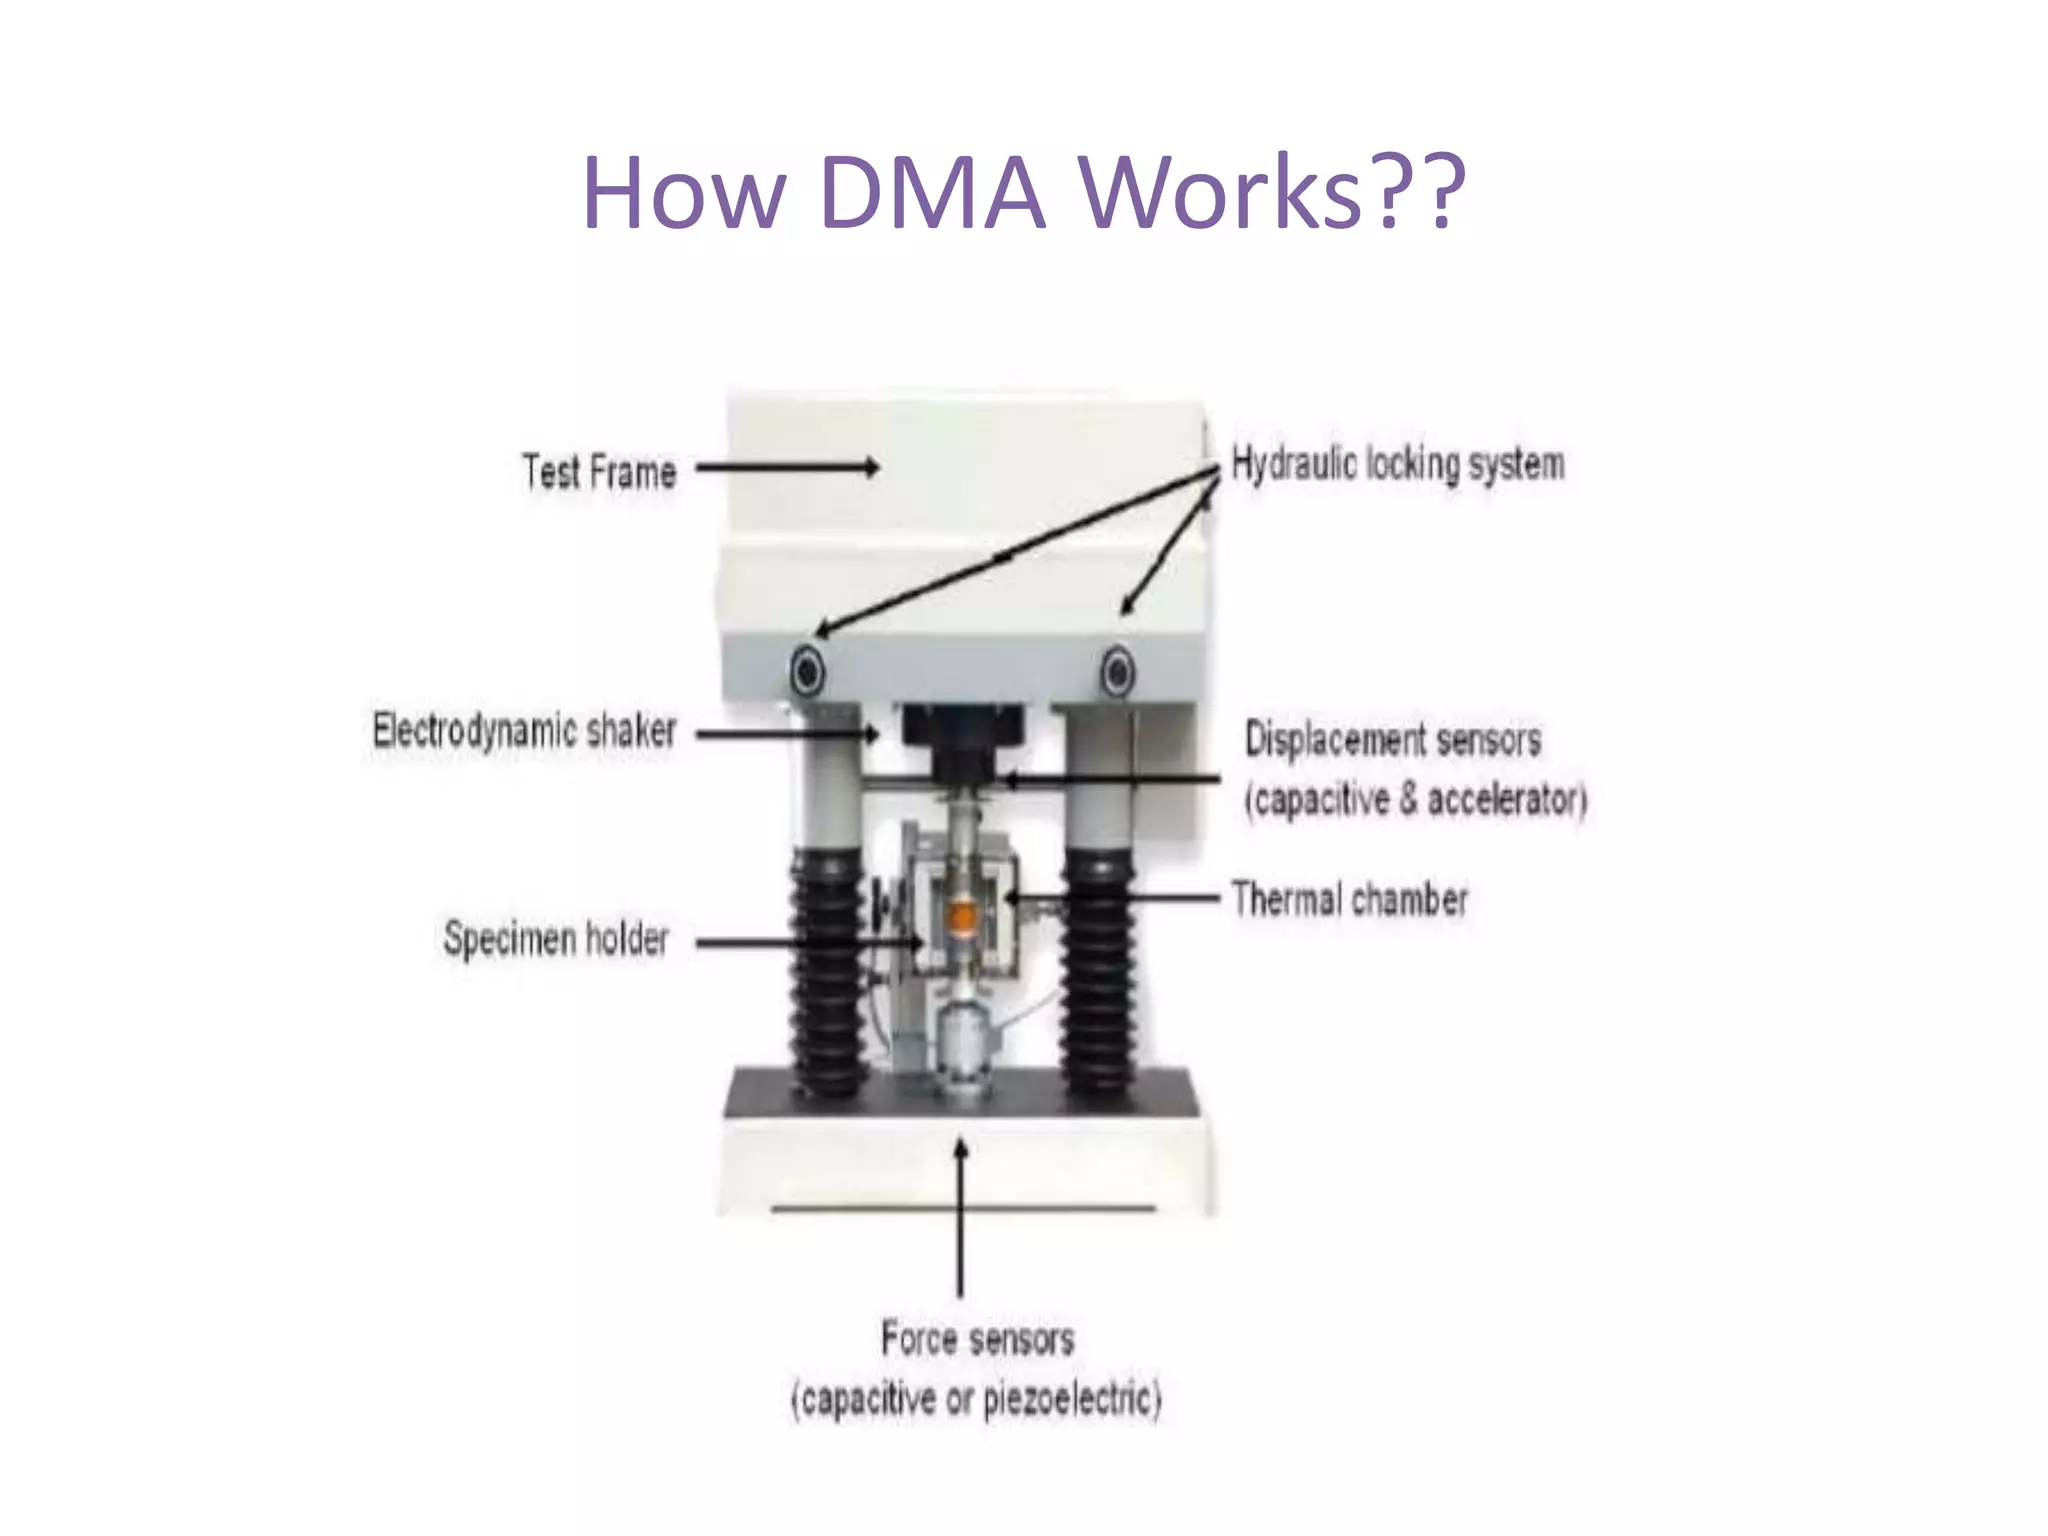
DMA is the best way to measure viscoelastic properties across temperatures, but it is also the most finicky technique you will encounter in polymer characterization. You put a sample in, run a temperature sweep, and get storage modulus, loss modulus, and tan delta curves. That is the easy part. Getting reproducible data is where everything falls apart. You apply a sinusoidal stress or strain and measure the resulting deformation. The phase lag between them tells you how much energy is stored versus dissipated. Storage modulus (E' or G') is the elastic response. Loss modulus (E'' or G'') is the viscous response. Tan delta is the ratio between the two and marks transitions like Tg most people look at. For tensile mode you clamp a dogbone or strip. For bending you use a three-point bend fixture. For compression you squish a cylinder. The mode matters because each one probes different deformation mechanics and the strain distribution through the sample thickness is not the same. This gets ignored constantly and then people wonder why their bending and tensile Tg values disagree by five degrees.
The Practical Setup
Start with geometry. The thickness and width need to be measured with calipers to at least 0.01 mm tolerance. A 0.1 mm error in thickness translates to roughly a 30 percent error in calculated modulus for bending mode because the calculation depends on the cube of the thickness. I have seen people report modulus values that were off by an order of magnitude from a simple measurement mistake. Fixture selection is not trivial. For thin films under 0.5 mm, tension mode is usually better because bending puts too much strain on the surface and the clamps can slip. For thick samples above 2 mm, three-point bending gives you a cleaner signal without needing special grip technology. Compression works for elastomers and soft materials but you need to watch for buckling if the sample is too tall relative to its diameter. Rule of thumb: keep the height to diameter ratio below two to one. Strain amplitude is where most people shoot themselves in the foot. You need to establish the linear viscoelastic region first. Run a strain sweep at a fixed temperature and frequency before you do anything else. Increase the strain from something like 0.01 percent up to maybe 1 percent and watch where E' starts to drop. That is your upper limit. If you run your temperature sweep at a strain level inside the LVR, your modulus values mean something. If you run above it, you are measuring nonlinear behavior and your Tg from tan delta will shift to a lower temperature. This is a very common source of inter-lab variability.
Running the Temperature Sweep
A typical protocol looks like this. Equilibrate at the starting temperature for five to ten minutes. Ramp at two to five degrees per minute. Apply a single frequency, usually one hertz for standard tests. Record data every degree or every half degree depending on how sharp your transitions are. Cooling rates and heating rates should match because thermal history affects your results, especially for semicrystalline polymers. Frequency selection matters more than people admit. A single frequency sweep will give you one snapshot. If you want to build a master curve, you need multiple frequencies across a temperature range. I usually run at 0.1, 0.5, 1, 5, and 10 hertz. That is nine to twelve samples minimum if you want clean data. The time investment is real. Each frequency at multiple temperatures can take two to four hours per sample depending on the range. For the time-temperature superposition to work, your shifts need to follow an Arrhenius or WLF relationship. If they do not, your material has multiple relaxation processes overlapping or your measurements are noisy. Check the shift factors before you spend hours trying to force a master curve.
Get the Full Details

A Real Problem I Ran Into
I was testing a filled epoxy system, something with glass beads at about forty percent loading. The tan delta peak for Tg was broadening uncontrollably as I approached the transition. My first instinct was to blame sample preparation. I remade the specimens three times. Same result. The peak was just wide and the modulus drop was gradual instead of sharp. The actual issue was thermal contact resistance at the clamp-sample interface. The filled material had a slightly rough surface from the mold release and the rigid clamps were not making uniform contact. As the material softened near Tg, the clamps were slipping microscopically during each cycle, introducing an artifact that looked like extra damping. I solved it by sanding the contact faces of the samples with 400 grit paper and applying a thin layer of silicone grease to the clamp faces. This improved the contact without adding significant compliance. The tan delta peak sharpened immediately and became reproducible across samples. The modulus values also shifted up by about eight percent because the slip was artificially lowering the apparent stiffness. This kind of problem does not show up in any manual. It takes a few bad runs to figure out.
Common Pitfalls That Ruin Data
Sample aging is one. Polyamides absorb moisture from the air quickly. If you run a dry sample and then a conditioned sample without controlling humidity, your Tg can shift by ten to twenty degrees. I always dry PA66 at eighty degrees in a desiccator for at least twelve hours before testing and run the DMA soon after. Storing samples in a desiccator between tests makes a measurable difference. Thermal expansion is another. As your sample heats up, it gets longer or shorter depending on the mode. In tension mode this changes the initial pre-load force. Most instruments auto-adjust the force, but the adjustment has limits. If your sample shrinks too much and loses contact with the grips, you will get nonsense data. I set a maximum force compliance threshold and watch the force trace during the run. If it drops to zero before the temperature range ends, the sample has lost clamping contact and the data is unusable past that point. For crosslinking studies, people often forget that the reaction itself generates heat. If you are doing an isothermal cure study at 150 degrees and the exotherm pushes the local sample temperature to 170 degrees, your kinetics data is wrong. Use a low heating rate or start at a lower isothermal temperature to minimize this effect.
When DMA Fails Completely
DMA does not work well for materials that are extremely brittle at test temperature. If your sample cracks during the run, you are done. I tested a highly filled thermoset once at room temperature and the specimen fractured halfway through the ramp. The modulus went to zero and the strain signal became erratic. There was no way to salvage it. Switching to compression mode did not help because the material was too brittle in all modes. The only solution was to heat the sample enough to make it ductile, run the test, and accept that you could not measure the glassy modulus below the transition. Porous or foam materials are another failure case. The standard calculations assume a homogeneous solid. When your sample is mostly air, the modulus values you get are structure-dependent rather than material-dependent. Two foams with the same polymer but different densities will give completely different DMA curves and there is no straightforward correction. If you need material properties from a foam, you have to either densify the sample or use a different technique like instrumented indentation. Heterogeneous blends where the phases have very different moduli can also produce confusing results. The measured response is a weighted average and deconvoluting the individual phase contributions from a single DMA curve is essentially impossible without additional assumptions or complementary techniques like DSC or microscopy.
Alternatives Worth Knowing
If your main question is just Tg, DSC is faster and easier. A standard DSC run takes fifteen minutes and gives you a clear glass transition step. DMA gives you Tg too, but also gives you modulus information that DSC cannot. If you need both, do DSC first for the quick answer and DMA for the mechanical properties. Running both on the same batch of material is standard practice in well-run labs. For rheological characterization of melts, oscillatory rheometry covers similar ground at higher temperatures and lower frequencies than most DMA instruments can reach. The sample geometry and loading are different, so direct comparison requires care, but the underlying viscoelastic principles are the same. There is no single best technique. DMA fills a specific niche between room temperature solids and elevated temperature behavior. It is worth the effort when you need modulus versus temperature data, but it is not the right tool for every problem. Know what question you are actually trying to answer before you load a sample.