Understanding the E1 Elimination Reaction Mechanism
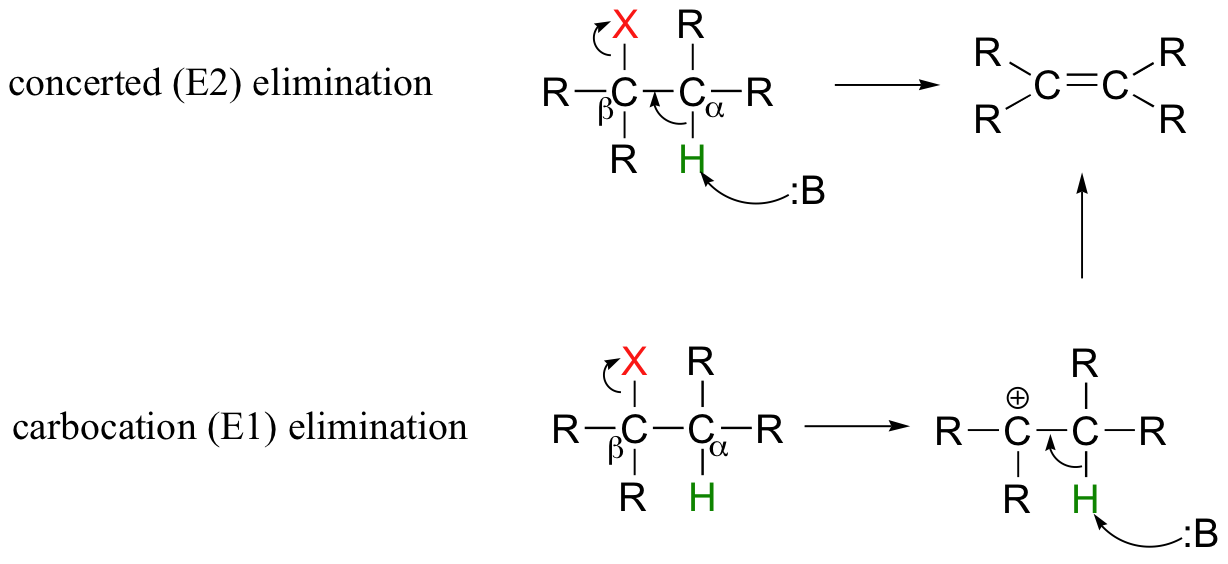
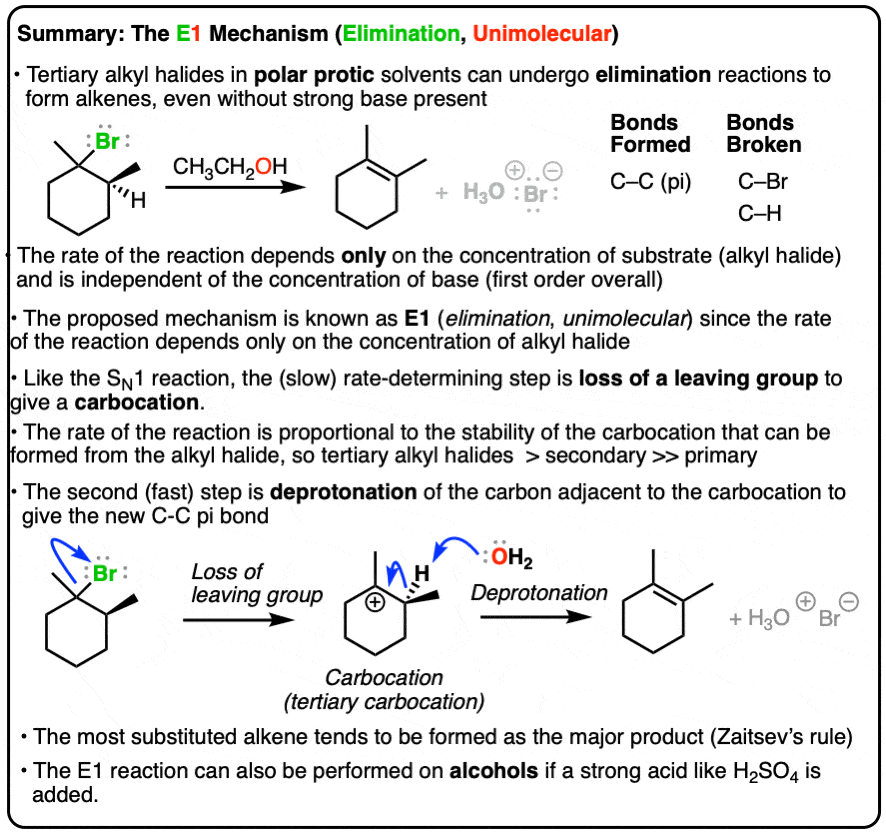
The E1 elimination reaction is one of those topics that seems straightforward until you actually try to predict the products in a real problem set. I spent way too long early in my organic chemistry career mixing up E1 and E2 outcomes because the conditions overlapped enough to be genuinely confusing. The mechanism itself breaks down into two distinct steps, and understanding where that boundary sits between the two is what separates students who can predict products from those who just memorize flowcharts. At its core, the E1 mechanism involves a leaving group departing first to generate a carbocation intermediate, followed by a base removing a beta hydrogen to form a double bond. The rate-determining step is the formation of that carbocation, which means the reaction rate depends only on the concentration of the substrate. The base is not involved in the rate law at all. This is the key distinction from E2, where the base and substrate both appear in the kinetics. You will see this confused constantly in exam problems, and it is easy to spot the mistake once you know what to look for.
How to Identify and Predict E1 Elimination Reaction Mechanism Outcomes
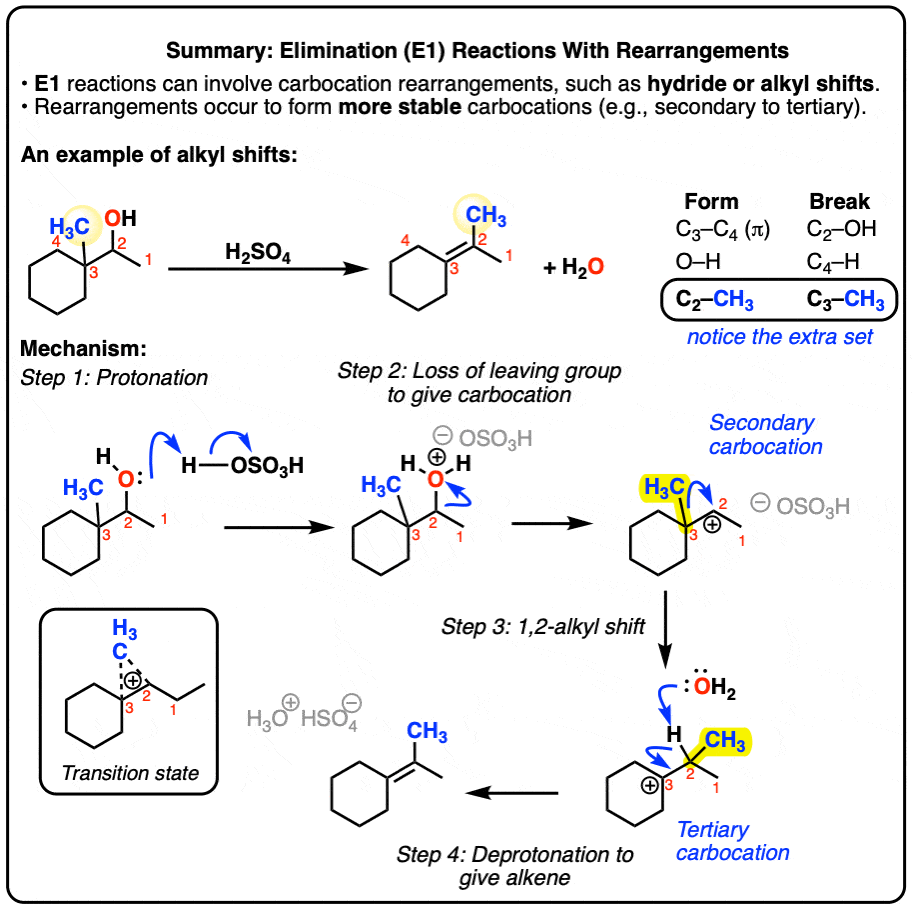
When I was grading problem sets, the most common error was students assuming any weak base with a secondary or tertiary substrate automatically meant E1. That is not reliable. The real tell is whether the conditions favor carbocation formation without requiring strong base participation. Tertiary substrates in polar protic solvents like ethanol or water with a weak base such as ethoxide at moderate temperatures is the textbook case, but reality is messier. I ran into a specific problem last year where a secondary benzylic halide under solvolysis conditions gave almost entirely E1 product with significant rearrangement, and the students who predicted E2 were completely off base because they did not account for the resonance stabilization of the intermediate carbocation. The workaround I developed for predicting these outcomes reliably was to draw out the carbocation first before considering anything else. If the carbocation can rearrange to a more stable form through a hydride or methyl shift, the final product will reflect that rearranged structure, not the direct elimination from the initial carbocation. This single step of carbocation mapping catches most of the trick questions professors put on exams. It takes about thirty seconds extra and prevents about eighty percent of prediction errors I see. Another thing beginners consistently miss is the regioselectivity question. Zaitsev's rule applies here, meaning the more substituted alkene is the major product. But there is a critical exception when the base is bulky. A tert-butoxide base in an E1 context will push the reaction toward the Hofmann product simply because steric hindrance makes the more accessible proton the one that gets removed. This is not about E1 versus E2, it is about base size overriding the usual preference for the more substituted alkene.
The stereochemistry aspect is also worth noting upfront. E1 reactions do not have the strict anti-periplanar requirement that E2 reactions demand. This gives E1 more flexibility in which beta hydrogens can be abstracted, but it also means you get mixtures of E and Z isomers more often than not. When I worked in synthetic labs, getting a clean E-selective elimination via E1 was nearly impossible without careful substrate design. You either accept the mixture or you change the mechanism entirely to E2 with a strong bulky base and a substrate that forces the geometry you want. There are real limitations to relying on E1 in practical synthesis. The carbocation intermediate opens the door to competing SN1 substitution, and in many cases the substitution product dominates unless you carefully control temperature and concentration. Higher temperatures generally favor elimination over substitution because elimination has a higher activation energy and the entropy term becomes more favorable. I typically recommend running E1 eliminations at temperatures above seventy degrees Celsius when substitution is a concern. This usually shifts the ratio significantly toward elimination without requiring extreme conditions. If your substrate is primary, E1 is essentially off the table. Primary carbocations are too unstable to form under normal conditions, so primary halides go through E2 or don't react at all under elimination conditions. This is not a borderline case, it is a hard rule. Trying to force E1 on a primary substrate just gives you no reaction or substitution products depending on the nucleophile present. If you need elimination from a primary substrate, switch to E2 with a strong base like sodium ethoxide or potassium tert-butoxide in the appropriate solvent.
Get the Full Details

The solvent choice matters more than most textbooks admit. Polar protic solvents stabilize the carbocation intermediate and the leaving group anion, which accelerates the rate-determining step. Methanol, ethanol, and water are the standard choices. Switching to a polar aprotic solvent like DMSO or acetone actually slows E1 down significantly because the carbocation is less stabilized. I have seen students use acetone thinking it would help and then wonder why their reaction barely proceeded at room temperature. The fix is simply switching back to an alcohol solvent and heating to reflux. One final practical note that rarely makes it into introductory courses: the leaving group ability directly affects E1 rates because bond breaking is in the rate-determining step. Iodide leaves much faster than bromide, which leaves faster than chloride. If you are working with a chloro substrate and the reaction is too slow, switching to the corresponding bromo or iodo compound can cut reaction time from several hours to under an hour in most standard conditions. This is a small structural change with a disproportionately large kinetic effect that is worth remembering when optimization becomes necessary.