How to Actually Calculate Effective Nuclear Charge Without Getting Confused
Understanding the Effective Nuclear Charge Periodic Trend
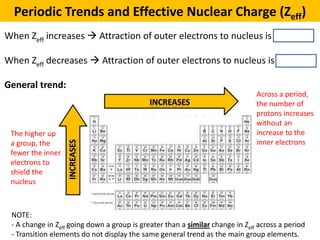
The first thing you need to understand is the calculation method, not some abstract definition. You find the effective nuclear charge by taking the actual nuclear charge — your proton count — and subtracting the shielding constant. For Slater's rules, that means grouping your electron configuration properly. (1s)(2s,2p)(3s,3p)(3d)(4s,4p)(4d)(4f)(5s,5p)... that grouping matters more than most people realize when you're working through problems. Here's how it actually works in practice. For a 3p electron in phosphorus, you have 1s2 2s2 2p6 3s2 3p3. The 3p electron you're examining gets shielded by the other four electrons in the n=3 shell — each contributing 0.35 — plus all eight electrons in the n=2 shell contributing 0.85 each, and the two electrons in the n=1 shell contributing 1.00 each. So that's 4 times 0.35, plus 8 times 0.85, plus 2 times 1.00, which equals 9.60. Your atomic number is 15, so Z_eff comes out to about 5.4. That's the number that actually matters for understanding why phosphorus holds onto its valence electrons the way it does. The periodic trend is straightforward in direction but messy in magnitude. Moving left to right across any period, effective nuclear charge climbs steadily because you're adding protons faster than you're adding shielding electrons. Across period 3, for example, you go from sodium at roughly 2.2 to chlorine at about 6.1. Each step adds one proton and one valence electron, and that valence electron only shields about 0.35 of the new nuclear charge. The net effect is a real, measurable increase in the pull the nucleus exerts on outer electrons.
Down a group, things get complicated quickly. A lot of textbooks say it stays roughly constant, and that's a useful approximation for introductory courses. But if you actually run the numbers with Slater's rules, you'll find it creeps upward. Going from lithium to sodium to potassium, each step adds a whole new principal shell, but the inner shells don't shield perfectly. The d and f electrons that appear further down the periodic table are particularly bad shielders, which is why the trend isn't as flat as you'd expect from a simplified model. I ran into a specific problem last semester that took me way too long to sort out. A student was trying to explain why gallium has a higher first ionization energy than aluminum, and they were using the simple textbook model that just looks at distance from the nucleus. That doesn't work here because gallium's valence electrons are in the 4p orbital, which is farther out, so the distance argument would suggest it should be easier to remove one. The real answer is the d-block contraction. Gallium has those ten 3d electrons sitting between the nucleus and the valence shell, and d electrons shield very poorly — they only contribute about 0.85 each instead of 1.00. So the effective nuclear charge on gallium's valence electrons jumps up significantly compared to what you'd predict without accounting for that. I had them recalculate using full Slater's rules, and the difference was about a full unit of Z_eff between aluminum and gallium. That explained the ionization energy inversion immediately. Another thing that trips people up regularly is how to handle transition metals. When you're calculating Z_eff for a 4s electron in, say, iron, you need to decide whether the 3d electrons count as shielding for it or not, and Slater's rules have a specific ordering that some students get wrong. The rule is that electrons in groups with higher n values don't shield electrons in lower n groups. So for a 4s electron, the 3d electrons do contribute to shielding, but at the 0.85 rate, not 1.00. A lot of people accidentally apply the 0.35 shielding between same-shell electrons incorrectly across the s and d orbitals, and that throws off their calculations by a noticeable margin.
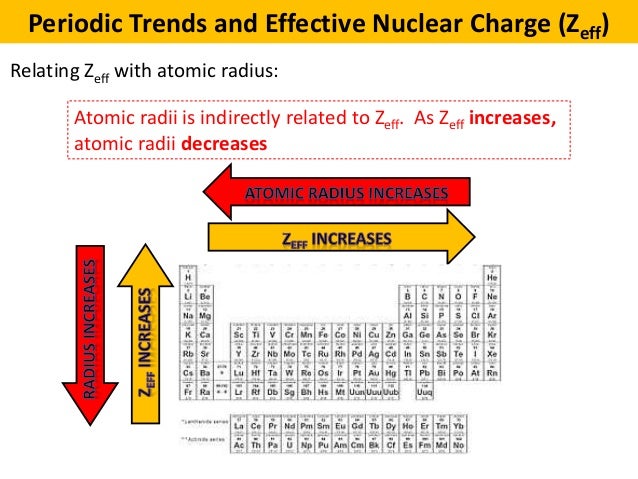
There's also a common misconception about Z_eff and atomic radius that I see in every class. Students will calculate that Z_eff increases across a period and then conclude that radius decreases, which is correct, but they often attribute it entirely to Z_eff. In reality, the decreasing radius across a period is also influenced by the fact that you're filling the same principal shell. The electrons aren't going into a new shell, so there's no compensating increase in average distance from the nucleus. When you move down a group, both Z_eff and the principal quantum number change, and the n effect dominates. That's why sodium is still much larger than lithium even though sodium's valence electron experiences a higher effective nuclear charge. The lanthanide contraction is probably the single most underappreciated consequence of imperfect shielding. After lanthanum, you start filling the 4f subshell, and f electrons are terrible at shielding — they contribute almost nothing by Slater's rules compared to what a full shell would contribute. This means elements following the lanthanides, like hafnium through gold, have significantly higher effective nuclear charges than their lighter congeners. The result is that hafnium and zirconium end up being almost the same size despite being in different periods. This matters for separations chemistry, catalysis, and anything where you'd normally rely on periodic trends to predict properties. One practical limitation you should know about: Slater's rules are an approximation, and a fairly rough one at that. They work reasonably well for main group elements, but they break down noticeably for heavier elements and for transition metals where relativistic effects start to matter. If you're working with elements past the sixth period, the calculated Z_eff values can drift from experimental observations by a significant amount. In those cases, more sophisticated calculations using Hartree-Fock or density functional theory give you results that are closer to reality, but they require computational tools rather than hand calculations. For most coursework and general chemical reasoning, Slater's rules are sufficient, but don't treat them as gospel when you're dealing with heavy elements or precision work.
Get the Full Details
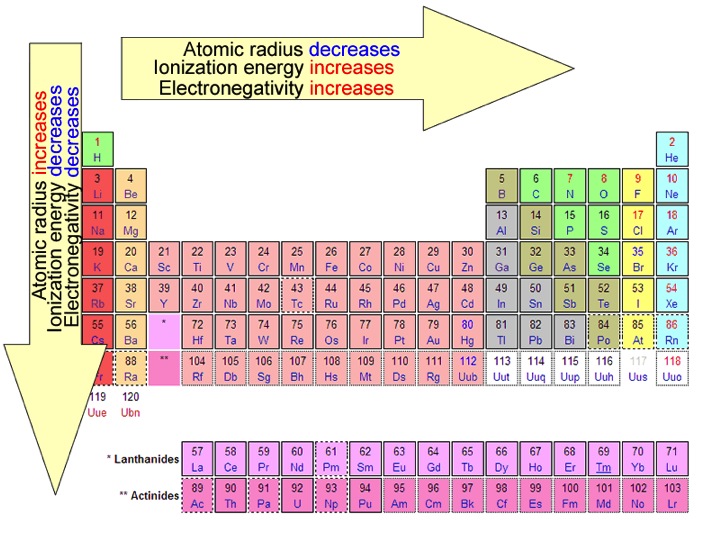
The other limitation I run into is that Z_eff alone doesn't tell the whole story for properties like electron affinity or electronegativity. Those depend on subtle quantum mechanical effects that the simple Z_eff model glosses over. For instance, the dip in electron affinity from nitrogen to oxygen isn't explained by a change in effective nuclear charge — it's about electron-electron repulsion in the newly occupied p orbital. Knowing Z_eff helps you understand trends in ionization energy and atomic radius reasonably well, but it's not a universal key for every periodic property.