Measuring Conductivity Without Losing Your Mind
The short version is that electrical conductivity of aqueous solutions measures how well ions move through water when you apply a voltage across it. The longer version involves cell constants, temperature compensation, and the fact that your meter will lie to you if you are not paying attention. I have spent years calibrating equipment and troubleshooting readings that made no sense at first glance, so here is the practical breakdown. Start with a clean cell. I cannot stress this enough. Most bad readings come from contamination on the electrodes, not from the solution itself. Rinse your conductivity cell with deionized water, then with a small amount of the sample you are about to measure. Do not wipe the electrodes with a cloth. Wiping creates static charge and scratches the plating. Just let the sample rinse through three times and move on. Calibrate using standards that bracket your expected reading. If you are testing something around 500 microsiemens per centimeter, calibrate with both 100 and 1000 µS/cm standards. A single-point calibration is acceptable only when you are doing quick field checks where approximate values are fine. For anything going into a report, two points minimum. The calibration process itself takes about five minutes if your meter handles it automatically. If you are doing it manually, factor in another ten minutes for preparation and documentation.
Temperature matters more than most people account for. The standard compensation reference is 25 degrees Celsius, and most meters apply a default coefficient of about 2 percent per degree Celsius. That default works for many common salt solutions but falls apart for others. Seawater, for example, has a temperature coefficient closer to 1.9 percent per degree. Organic-rich solutions can behave completely differently. If your lab deals with varied sample types, input the correct coefficient for each matrix. Looking it up takes thirty seconds and prevents systematic error that compounds quickly as temperature deviates from 25°C. Stir the sample gently during measurement. Standing solutions develop concentration gradients near the electrodes, especially at higher conductivities. A magnetic stir bar on low speed gives consistent readings. Do not stir so fast that you introduce air bubbles. Bubbles cling to the electrode surface and cause the reading to flutter. That fluttering is easy to misinterpret as a contaminated sample when it is just trapped air. Record the actual temperature along with the conductivity value. A compensated reading is only as good as the coefficient you fed into it. If you write down 500 µS/cm at 25°C but your sample was actually 30°C with a default coefficient applied, someone later trying to reproduce your work has no way to know the real uncompensated value was closer to 420 µS/cm. Keep both numbers.
I ran into a specific problem last year that illustrates why procedure matters. We were monitoring treated wastewater effluent from a facility and the conductivity readings jumped erratically between 800 and 1400 µS/cm on the same sample over a ten-minute window. The operator suspected the meter was failing. We swapped meters, checked the cell constant, recalibrated, and the problem persisted. The solution was apparently uniform. It turned out the sample line was drawing from a mixing tank where a chlorination step was cycling on and off. The chlorine was reacting with organic matter in the water, producing additional ions in situ. The conductivity was genuinely changing. Taking a grab sample and measuring it in the lab gave stable, reproducible results because the reaction had slowed dramatically outside the tank. The workaround was straightforward: we switched to continuous inline measurement at the point of interest and logged the data in real time instead of relying on grab samples. The inline probe showed the true variation pattern, and we could correlate it with the chlorination schedule. This is the kind of thing that looks like instrument failure until you actually investigate the process.
Get the Full Details

What Actually Determines Conductivity in Your Sample
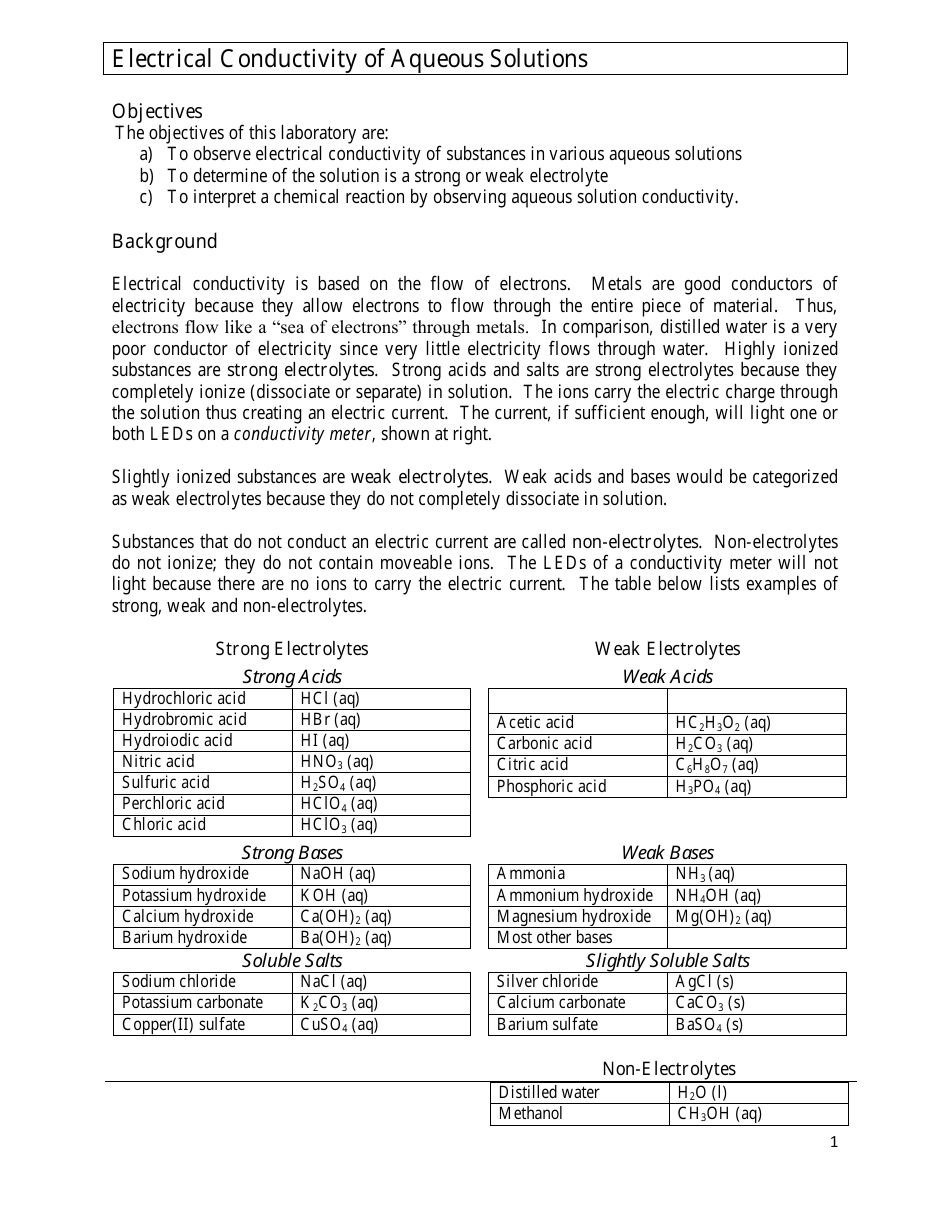
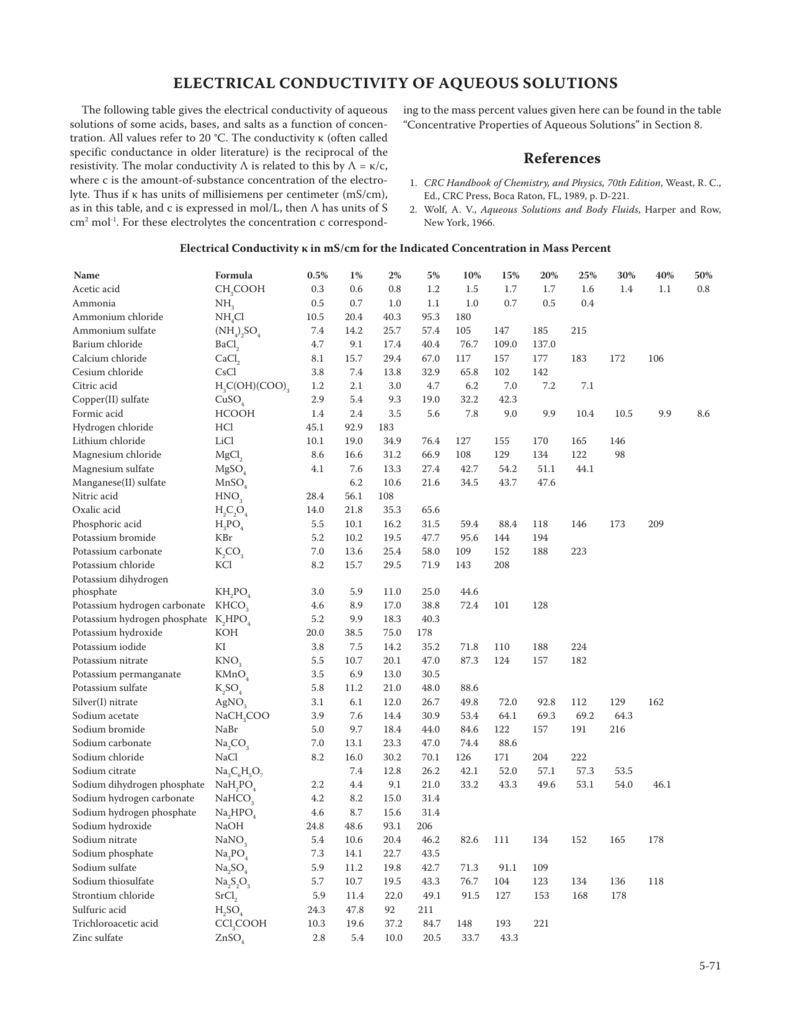
Ionic concentration is the primary driver, but the type of ion matters just as much. H+ and OH- ions conduct roughly five to seven times better than typical metal cations or halide anions due to the Grotthuss mechanism, which is the name for proton hopping through the water structure. That means a dilute acid or base solution will read significantly higher in conductivity than a neutral salt solution at the same molar concentration. Beginners sometimes miss this and assume a low reading means low contamination when the contaminant might actually be a strong acid or base at trace levels. Another counter-intuitive point is that conductivity does not increase linearly with concentration at higher ranges. Above about 0.01 M, ion-ion interactions become significant. The ions interfere with each other's movement through the solution, and the relationship curves downward. This is why high-purity water systems that are supposed to read near zero can show unexpected conductivity if even a tiny amount of salt gets introduced. At low concentrations, the relationship is essentially linear. At higher concentrations, you need to consult Kohlrausch's law and activity coefficients if you want quantitative accuracy. Most routine lab work stays in the linear range, but if you are analyzing concentrated brines or industrial process streams, that non-linearity will bite you. Non-ionic dissolved substances do not contribute to conductivity at all. Sugar, ethanol, most organic solutes. This is useful for diagnosing samples. If you have a solution that tastes sweet and reads high in total dissolved solids by evaporation residue but shows near-zero conductivity, you know the dissolved material is predominantly non-ionic. That distinction separates things like dissolved salts from things like dissolved organics, and it is something you can determine in under a minute with a conductivity meter if you pair it with a TDS measurement or a simple evaporation test.
Common Pitfalls That Waste Time
Using old calibration standards is the most frequent mistake. Conductivity standards degrade. CO2 from the air dissolves into open standards and increases their conductivity over time. A 1000 µS/cm standard that has been sitting open on the bench for a week might read 1050 or higher. Check the expiration date, seal standards tightly after use, and discard any standard that looks cloudy or has been exposed to contamination. Standards are relatively inexpensive compared to the cost of redoing a batch of samples because your calibration drifted. Another pitfall is ignoring the cell constant. Cells degrade over time. Graphite electrodes coat with deposits. Platinum black plating wears off. A cell rated at 1.0 cm^-1 might effectively become 1.1 or 0.9 after months of use. Verify the cell constant periodically by measuring a known standard and calculating the actual constant from your reading. If it has drifted more than 5 percent from the rated value, clean or replace the cell. This check takes two minutes and prevents hours of questionable data downstream. High conductivity samples can polarize the electrodes if your meter uses a DC voltage instead of AC. Polarization creates an opposing voltage at the electrode surface and drives your reading artificially low. All modern conductivity meters use AC excitation to avoid this, but if you are working with legacy equipment or very high-conductivity samples above 100 mS/cm, make sure your instrument is designed for that range. Some cheaper meters bottom out around 20 mS/cm and give increasingly inaccurate readings beyond that point without any warning.
There is also a practical limitation worth stating plainly: conductivity tells you the total ionic load but nothing about which ions are present. Two solutions can have identical conductivity and completely different chemistry. If you need to identify specific contaminants, conductivity alone will not get you there. You need ion chromatography, atomic absorption, or titration depending on what you are looking for. Conductivity is a screening and monitoring tool, not an identification tool. Treating it as anything more than that leads to wrong conclusions.

When Conductivity Measurement Fails Completely
Suspensions and colloids are a clear limitation. Particulate matter coats the electrodes and changes the effective cell geometry over time. The reading becomes unstable and meaningless. If your sample is turbid, filter it first or use a flow-through cell designed to minimize fouling. Even then, long-term monitoring in suspensions is unreliable. You are better off measuring the filtrate and accepting that you are characterizing the dissolved fraction only. Very low conductivity measurements below 1 µS/cm require special handling. Atmospheric CO2 dissolves into ultra-pure water and raises the conductivity within minutes. A freshly produced RO or DI water sample might read 0.05 µS/cm at the tap but climb to 0.5 or higher by the time it reaches your beaker. Measure inline if possible. If you must take a sample, use a sealed container, fill it completely to eliminate headspace, and measure immediately. Even then, your reading includes whatever CO2 the water absorbed during transfer. This is a known and accepted limitation in the industry, and anyone claiming otherwise is selling something. Organic solvents mixed with water also complicate things. The dielectric constant changes, ion pairing becomes more likely, and the temperature coefficient shifts unpredictably. Standard conductivity meters assume an aqueous medium. Mixing in methanol or ethanol can throw off readings by 20 to 40 percent depending on the proportion. If your application involves solvent mixtures, validate your method against known standards in that specific matrix before trusting the numbers.
Practical Workflow for Routine Measurements
A typical efficient workflow for a lab doing daily conductivity checks runs about fifteen minutes per sample once you are calibrated and warmed up. Rinse the cell. Calibrate with fresh standards if it has been more than four hours since the last calibration. Measure the sample in triplicate, recording each reading and the temperature. Clean the cell between samples if you are moving between very different matrices, or just rinse thoroughly if the samples are similar. Document everything. This process takes roughly one minute per sample once you are past the initial calibration, which makes conductivity one of the faster wet chemistry measurements available. That speed is why it remains a standard parameter in so many regulatory frameworks and process control applications despite being fundamentally simple in concept.