Understanding How Inductive Effects Actually Move Electrons
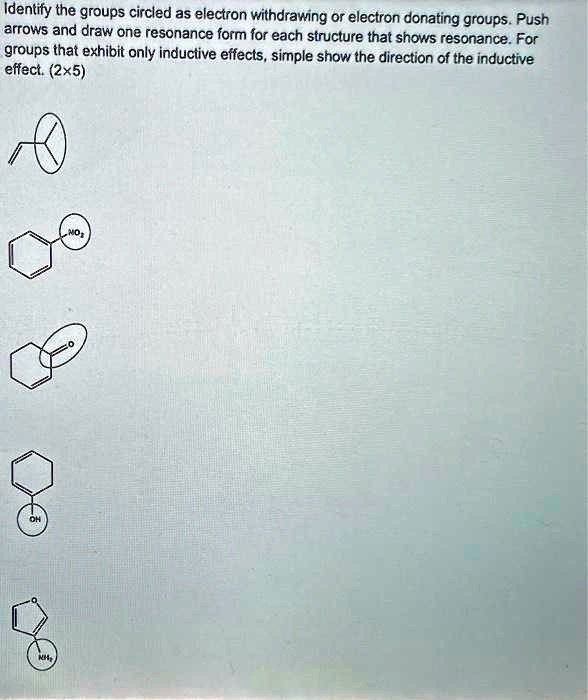
Most organic chemistry classes present electron withdrawing groups as a clean list you memorize for exams. The reality is messier. The inductive effect operates through sigma bonds, and its strength drops off dramatically with distance. Fluorine right next to a carboxylic acid changes everything. Fluorine two carbons away barely matters. This distance dependency is where students lose points and where actual synthetic work becomes tricky. I spent weeks troubleshooting a reaction where the expected regioselectivity completely flipped. We were working on a substituted cyclohexane ring with a trifluoromethyl group positioned meta to our reaction center. Literature predicted one outcome based on standard inductive withdrawal tables. The NMR didn't lie. The product was the opposite isomer. Turns out, at that distance on a rigid ring system, the inductive effect had attenuated so much that hyperconjugation from adjacent C-H bonds started competing. I ended up running a computational study just to confirm what was happening. The fix was switching from a CF3 group to a nitro group, which maintains strong inductive withdrawal even at meta positions due to its resonance contribution, not just pure induction.
Predicting Reactivity Using Electron Withdrawing Groups Via Induction
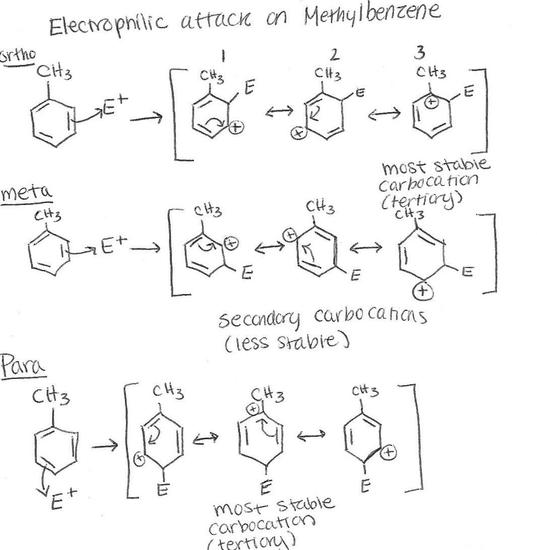
The practical approach starts with identifying which atoms or groups are more electronegative than carbon. Oxygen, nitrogen, halogens, and the nitro group top the list. Once you find them, you trace the sigma bond network from that group toward your site of interest. Each bond along the path weakens the effect roughly by half. This isn't a hard rule, but it's close enough for most ground-state predictions. Here is where people make mistakes. They treat induction and resonance as separate silos. In practice, they overlap constantly. Take a para-nitrophenol molecule. The nitro group withdraws electrons through both resonance and induction simultaneously. If you only account for induction, your pKa prediction will be off by about 2 units. That is the difference between predicting a compound is acidic enough to deprotonate with bicarbonate or thinking you need hydroxide. Always check whether the group has pi orbitals available for conjugation before attributing all electron withdrawal to pure induction. Another common oversight involves solvent effects. Inductive withdrawal is measured and discussed almost entirely in gas phase or nonpolar contexts in textbooks. In water or DMSO, solvation shells can reinforce or partially cancel the inductive pull. A cyano group on an aromatic ring behaves differently in acetonitrile than it does in ethanol. I learned this the hard way during a nucleophilic aromatic substitution where our rate data in different solvents didn't match the Hammett constants we pulled from the literature. The rho value shifted depending on solvent polarity. Rechecking with solvent-corrected tabulated values fixed the discrepancy. Don't skip this step when your experimental rates look wrong.
When you are evaluating a molecule for reactivity, start with the closest electron withdrawing group. Its influence dominates. Then move outward bond by bond. Write down the partial charges if it helps you visualize the polarization. The carbonyl carbon in an alpha-halo ketone carries a significantly higher partial positive charge than the one in a simple ketone. That extra charge density is what makes halogenated ketones more reactive toward nucleophiles and more prone to enolization. If you need quantitative data, the Taft equation gives you sigma* values for individual substituents. The standard Hammett chart works for aromatic systems. Neither handles aliphatic branching well, so take both with a grain of salt when your molecule gets complicated. The biggest limitation to keep in mind is that inductive effects are static models. They describe ground state electron distribution. They do not predict transition state behavior reliably on their own. When a reaction involves bond breaking or significant geometric rearrangement, field effects and steric factors can dominate over pure induction. I have seen experienced researchers rely too heavily on inductive reasoning and miss a steric block that completely shut down their proposed mechanism. Always run a quick steric check with models or CPK kits before committing to a synthetic route. The combination of induction, sterics, and electronics together gives you something closer to what actually happens in the flask.
Get the Full Details