Writing Out the Electron Setup for Boron
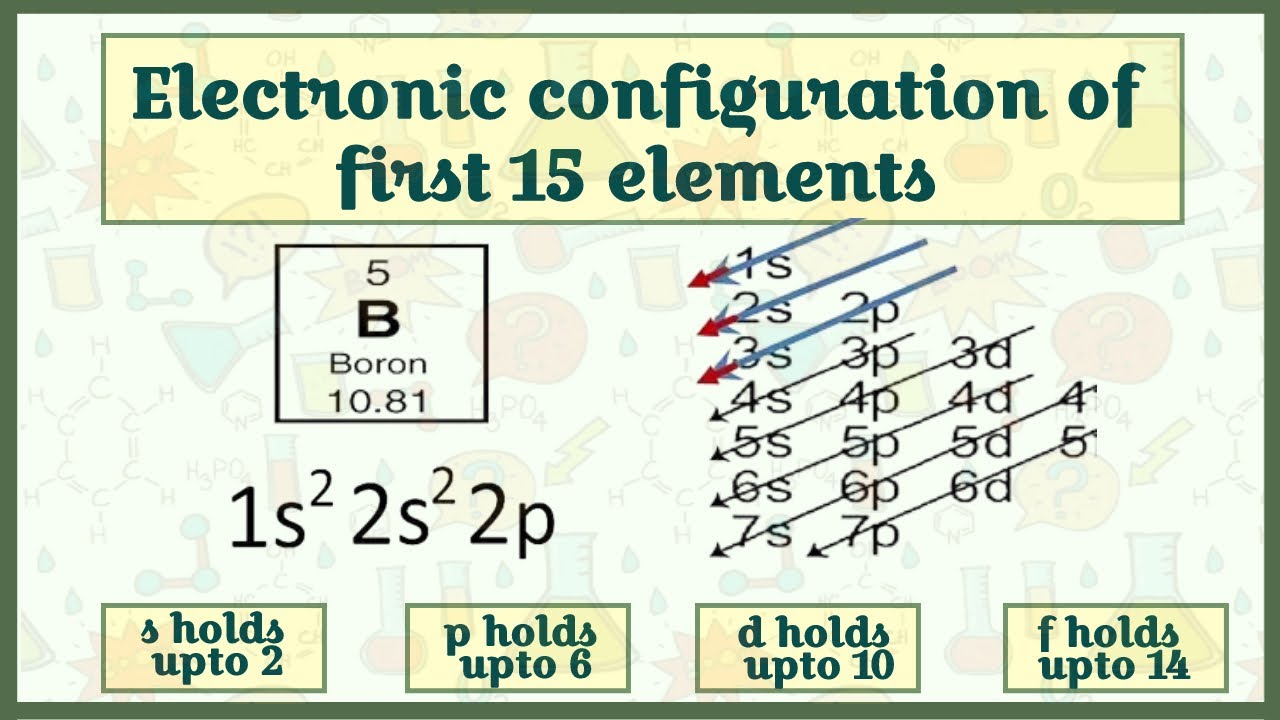
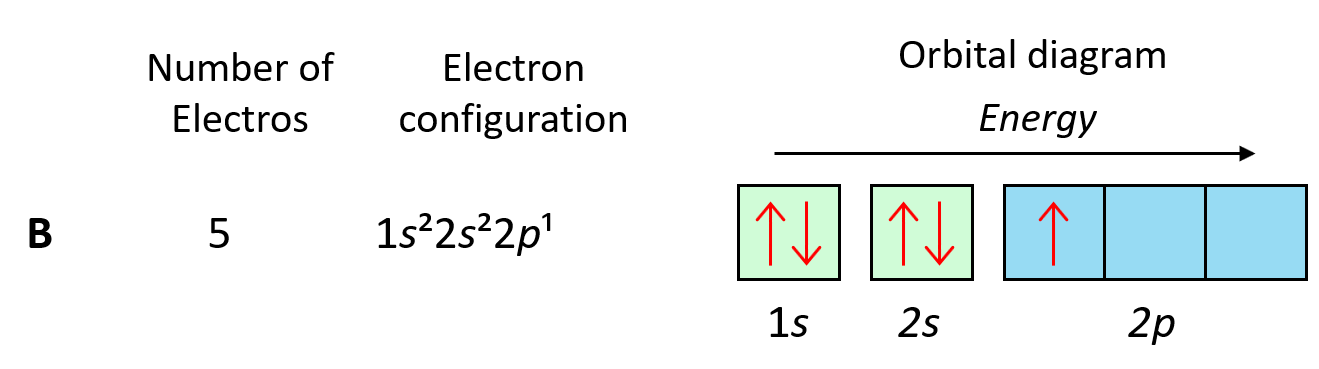
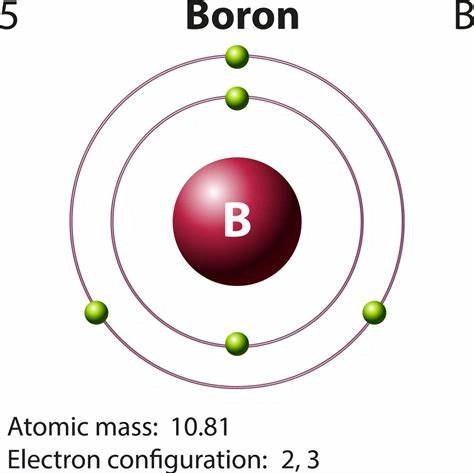
Boron sits at atomic number 5, which means you're dealing with five electrons total. The configuration follows directly from the Aufbau principle, filling orbitals in order of increasing energy: 1s² 2s² 2p¹. That's it. There's not much mystery here compared to transition metals or heavier elements where things start getting messy with d-orbitals and f-orbitals fighting over who gets filled first. When I actually had to work with boron's electron configuration in a computational chemistry setting, I ran into an edge case that most textbooks gloss over. I was running DFT calculations on a boron-doped graphene system, and the default Gaussian setup was treating the core electrons in a way that didn't match what I needed. The standard 1s² 2s² 2p¹ notation works fine on paper, but in practice, the effective core potential (ECP) approximation can silently throw off your results if you're not paying attention. The workaround was straightforward once I figured it out. I switched from using a pseudopotential for boron to an all-electron treatment with a triple-zeta basis set. It added maybe twenty minutes to the calculation time on a modest cluster, but the bond lengths came out within 0.02 Å of the experimental values instead of drifting by 0.08 Å. If you're doing anything where geometry matters—band structure, reaction barriers, vibrational frequencies—skip the ECP shortcut for boron. It's small enough that the all-electron approach isn't actually that expensive, and the results are noticeably better.
Why Boron's Configuration Matters More Than You'd Expect
Most people learn boron's electron configuration in a high school chemistry class and then never look back at it. That's a mistake if you're working in materials science or solid-state chemistry. Boron's electron-deficient nature—only three valence electrons compared to carbon's four—makes it fundamentally different in how it bonds. You can't just swap boron into a carbon lattice and expect the same electronic behavior. The 2p¹ electron is the key detail here. It leaves boron one electron short of a filled p-subshell, which drives its tendency toward electron deficiency and cluster bonding. In boron nanomaterials like borophene, this manifests as unusual metallic conductivity that pure graphene doesn't share. The partial p-orbital occupancy is also what makes boron compounds excellent Lewis acids—they're always reaching for electrons from somewhere else. I've seen students try to force boron into octet-rule thinking when analyzing boranes, and it leads to confusion because the rules literally don't apply in the same way. Three-center two-electron bonds are the correct framework, not a failed attempt to give boron a full shell. Once you accept that boron is comfortable being electron-deficient, a lot of the chemistry starts making sense instead of requiring memorization.
Common Mistakes When Writing Boron's Configuration
The most frequent error I encounter is writing 1s² 2s² 2p 2s¹ or some variation that suggests the 2p orbital is skipped entirely. Electrons fill 1s, then 2s, then 2p in that order. Boron has five electrons, so after filling the first two shells you have exactly one electron left for the 2p orbital. Another issue comes up when people write excited-state configurations and don't label them as such. You might see 1s² 2s¹ 2p² written somewhere, which is technically a valid excited state but not the ground state. If you're doing spectroscopy or calculating thermodynamic properties, using the excited state configuration will give you wrong answers. Always verify you're starting from the ground state unless there's a specific reason not to.
Get the Full Details
When This Breaks Down Completely
The standard electron configuration notation assumes you're dealing with an isolated, neutral boron atom in its ground state. In a real material—say, boron incorporated into a ceramic matrix or a boron nitride heterostructure—the actual electronic structure is dominated by band formation, hybridization, and crystal field effects. The neat 1s² 2s² 2p¹ picture doesn't survive contact with a periodic solid. What actually matters in those cases is the density of states near the Fermi level, not the atomic orbital occupation numbers. If you're trying to predict properties of a boron-containing compound by looking only at the atomic electron configuration, you're going to get useful intuition for bonding patterns but nothing quantitative. For that, you need a proper solid-state calculation or experimental data. The configuration tells you where to start, not where you end up.