Working With Reaction Enthalpy In Practice
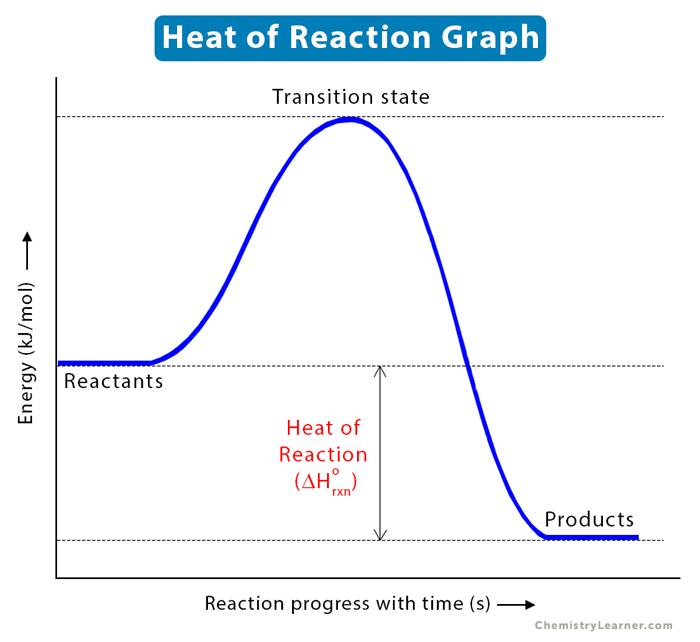
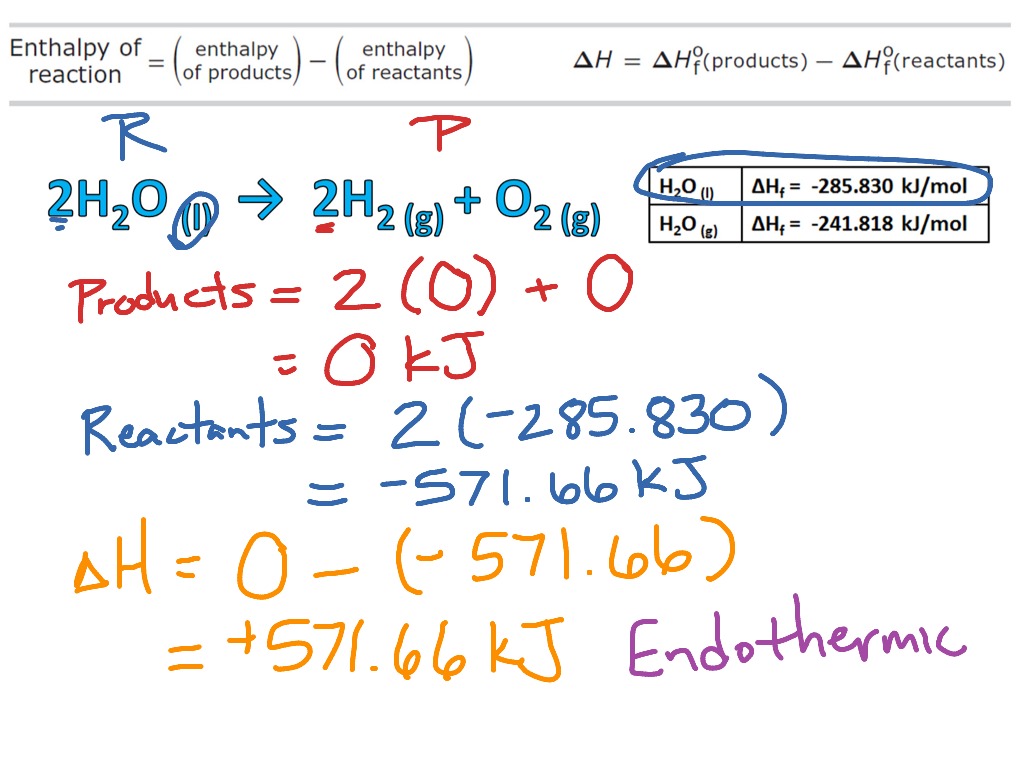
Most people learn that the enthalpy change of a reaction is just products minus reactants using standard formation values, and technically that is correct. The way you actually apply this in a lab or plant setting is considerably more annoying. I deal with reaction calorimetry and process heat integration, and the gap between textbook numbers and real measured values is where things break down. Here is how the calculation works when you are not in a controlled academic environment. Start with standard molar enthalpies of formation at 298.15 K and 1 bar. These are available in NIST Chemistry WebBook, the CRC Handbook, and several process simulation databases. The basic equation is: H°rxn = H°f(products) H°f(reactants)
The nu terms are stoichiometric coefficients, positive for products and negative for reactants. You multiply each coefficient by the corresponding formation enthalpy and sum. This gives you the standard reaction enthalpy at 25°C. It is straightforward until your reaction actually runs at a different temperature, which it always does. To adjust for temperature, you use Kirchhoff's law. The integrated form accounts for how heat capacity changes with temperature: H°rxn(T) = H°rxn(T) + [T to T] Cp dT
Where Cp is the difference between the heat capacity of products and reactants, each multiplied by their stoichiometric coefficients. Heat capacity is usually expressed as a polynomial in T: Cp = a + bT + cT² + dT². You integrate term by term. For quick hand calculations, assuming a constant average Cp over a small temperature range gives results within a few percent. Over a 200-degree swing, the error can reach 10 to 15 percent if Cp is large, which it often is. I once spent three hours tracking down a 40 kJ/mol discrepancy in a sulfonation reaction. The issue was that the literature value I was using assumed liquid water as a product, but my reaction conditions produced steam. The enthalpy of vaporization for water is 44 kJ/mol at 25°C, and we were generating two moles of water per mole of product. Subtracting that phase change enthalpy brought the calculated value in line with our calorimeter reading. This happens more often than you would expect. Always check the physical state assigned to each species in your data source.
Get the Full Details
Alternative Methods When Tabulated Data Is Unavailable
Bond enthalpy estimates exist and are faster, but they are rough. Average bond energies are derived from many different molecules, so applying them to a specific reaction introduces systematic error. For a hydrocarbon combustion, bond enthalpy methods typically land within 10 to 20 percent of the true value. For reactions involving heteroatoms or unusual bonding environments, the error grows. I use bond enthalpies only for sanity checks, never for design calculations. Combustion calorimetry gives empirical values. A bomb calorimeter measures the heat released when a compound is burned completely. From the measured temperature rise and the known heat capacity of the calorimeter assembly, you calculate the combustion enthalpy directly. This is the gold standard for validating formation enthalpies of organic compounds. The uncertainty is typically ±0.5 kJ/mol for well-behaved samples. The downside is that you need the sample, a calibrated instrument, and time. Most reactions of interest do not involve combustion at all, so you need to construct a thermochemical cycle using Hess's law to connect the combustion data to your target reaction. Computational chemistry can predict reaction enthalpies when experimental data does not exist. DFT methods with a decent basis set, such as B3LYP with 6-311+G(d,p), typically achieve errors of 5 to 15 kJ/mol for organic reactions. Coupled-cluster methods like CCSD(T) are more accurate but scale poorly with system size. I run Gaussian or ORCA calculations for novel intermediates where no database entry exists. The results are good enough to guide experimental design but not precise enough to replace measurement for safety-critical process parameters.
Common Pitfalls That Waste Time
The biggest mistake I see is ignoring the reference state. Standard formation enthalpies are defined for elements in their standard states at 1 bar. Carbon is graphite, not diamond. Oxygen is O gas, not atomic oxygen. If you accidentally use the formation enthalpy for O(g) instead of O(g), your result will be off by the O=O bond dissociation energy, roughly 498 kJ/mol. This is an easy typo and a catastrophic error. Another frequent issue is mixing solution enthalpies with formation enthalpies without accounting for the dissolution step. H°f values in most tables refer to the pure substance. If your reaction occurs in aqueous solution, you need to add the enthalpy of solution for each ionic species. The enthalpy of dilution is often non-negligible, especially for concentrated electrolyte solutions.Ignoring this can introduce errors of 10 to 30 kJ/mol for salt-forming reactions. Pressure effects are small for liquids and solids but matter for gases at high pressure. The correction involves the departure function, which requires an equation of state. For reactions at moderate pressures up to about 10 bar, the ideal gas assumption introduces less than 1 percent error in most cases. Above 50 bar, you should use a real gas model or measure the effect directly. I worked on a high-pressure hydrogenation process where neglecting the pressure correction on the reaction enthalpy led to an undersized heat exchanger. The reactor ran 8 degrees hotter than designed, which degraded the catalyst selectivity noticeably.
Tools And Data Sources
For routine calculations, the NIST WebBook is the most reliable free source. It provides curated thermochemical data with uncertainty estimates and references. The JANAF tables are the historical benchmark, though some of the original measurements are old. For process engineering work, the DIPPR 801 database is industry standard, covering thousands of compounds with temperature-dependent Cp and phase transition data. Aspen Plus and ChemCAD have built-in property methods that estimate missing enthalpies using group contribution techniques like Lydersen or Joback, but these estimates carry higher uncertainty, typically ±15 to ±25 kJ/mol. I keep a spreadsheet with the Kirchhoff integration pre-programmed for common reaction types. You input the polynomial coefficients for each Cp, the temperature bounds, and the standard reaction enthalpy, and it outputs the corrected value. It saves maybe twenty minutes per calculation compared to doing it by hand, which does not sound like much, but when you are evaluating ten different reaction pathways, it adds up. The spreadsheet also flags missing phase transition data, which catches errors before they propagate.
What This Approach Cannot Handle Well
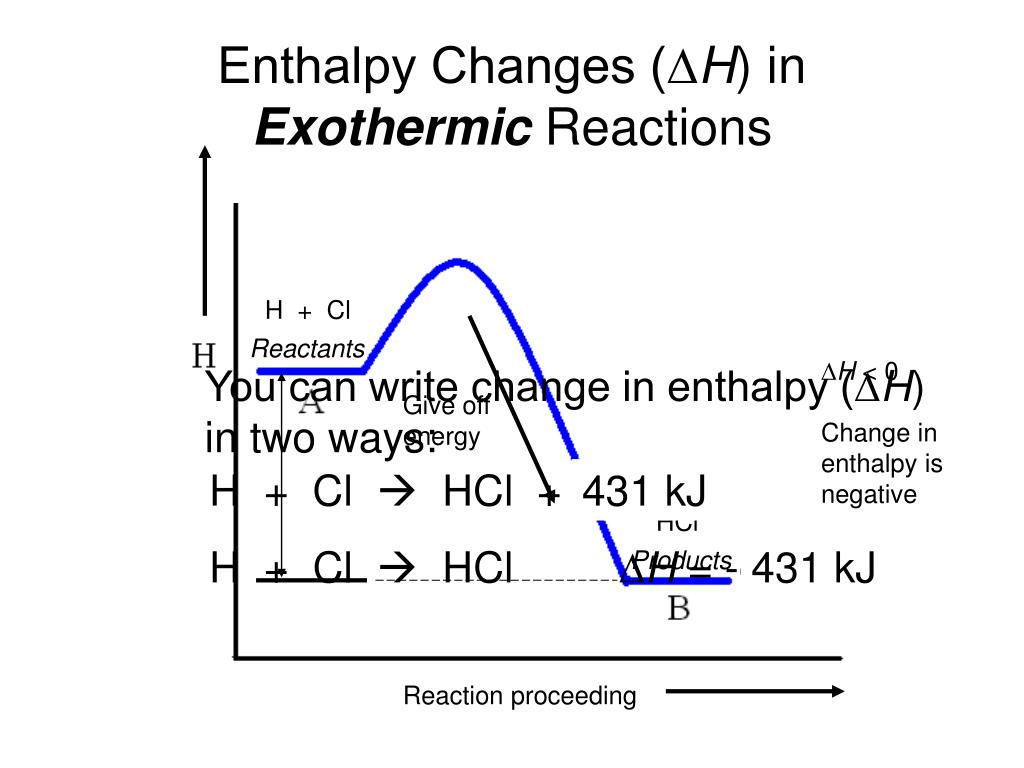
Reaction enthalpy calculations assume the reaction goes to completion or that you are calculating the standard enthalpy change for the stoichiometric equation as written. Equilibrium effects, partial conversion, and side reactions are separate problems. The enthalpy you calculate is for the idealized reaction, not for what actually happens in a reactor with incomplete conversion and competing pathways. If you need the actual heat duty, you must combine the reaction enthalpy with conversion data and selectivity data from your process simulation or pilot plant measurements. Non-ideal mixtures are another limitation. Activity coefficients affect the effective chemical potential, which indirectly influences the temperature dependence of the reaction enthalpy. For dilute solutions, this is negligible. For concentrated electrolytes or non-ideal organic mixtures, the deviation can be significant. I have seen deviations of 5 to 10 kJ/mol in highly concentrated sulfuric acid systems where ideal solution assumptions break down completely. If your reaction involves radicals, excited states, or photochemical pathways, standard thermochemical tables are not sufficient. The enthalpy of a radical species depends on its electronic state, and tabulated values often represent the ground state only. I encountered this with a free radical chlorination where the chain propagation steps involve Cl• atoms. Using molecular chlorine formation data without accounting for the Cl-Cl bond dissociation energy gave an answer that was 242 kJ/mol too high for the propagation step. You need formation enthalpies for the radical species themselves, which are available in specialized databases but require extra lookup steps.