Understanding Enthalpy of Formation Through Practice
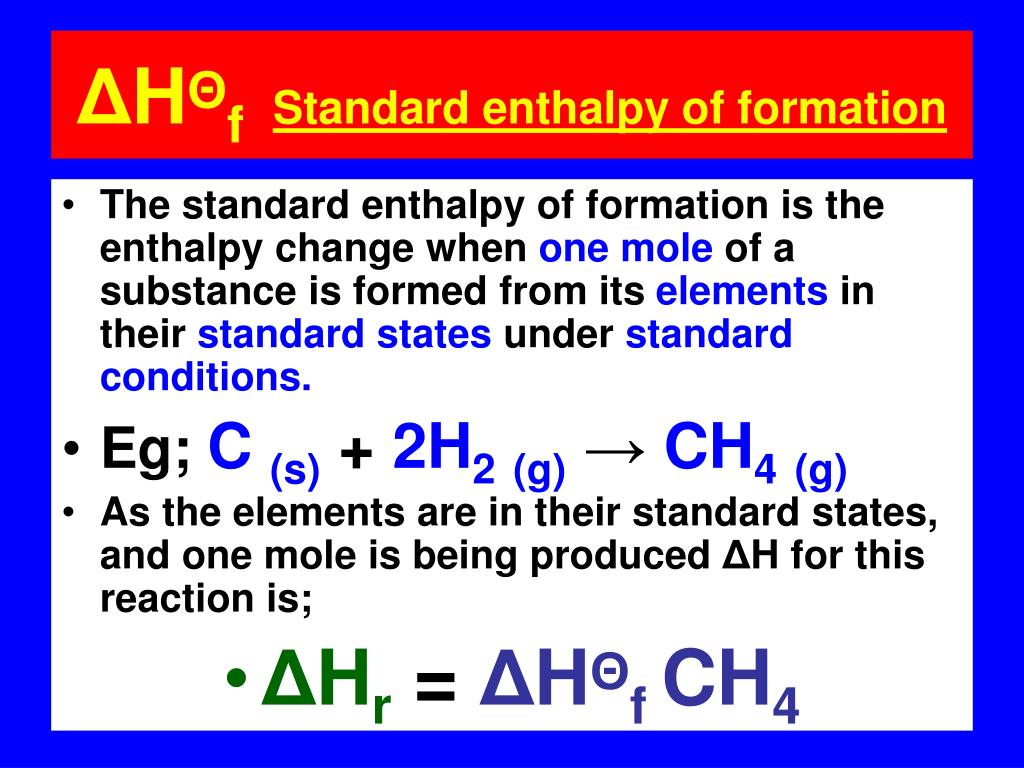
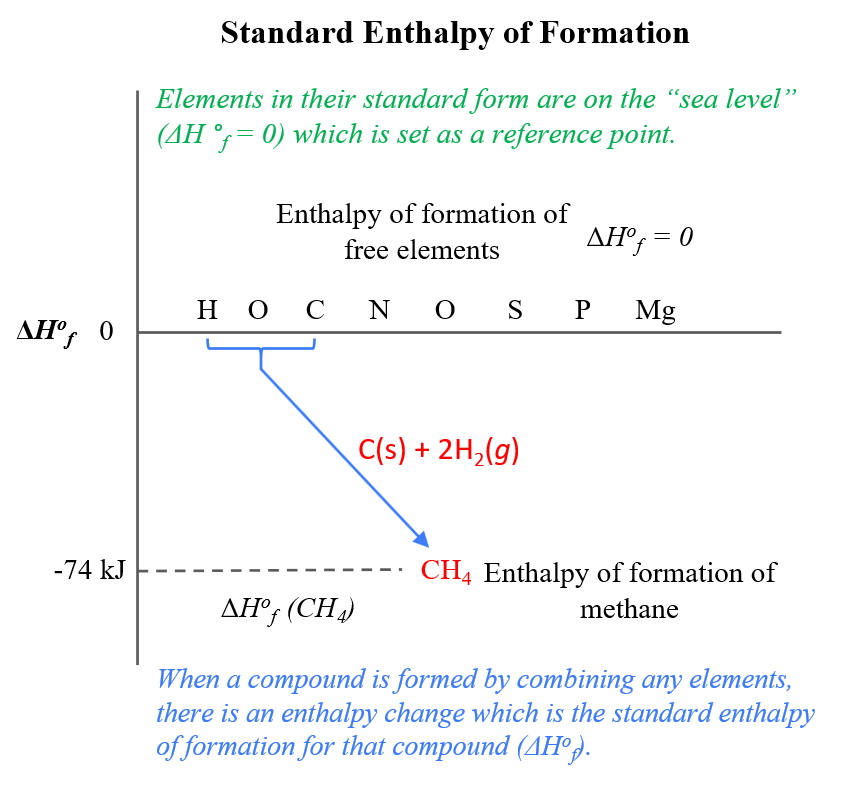
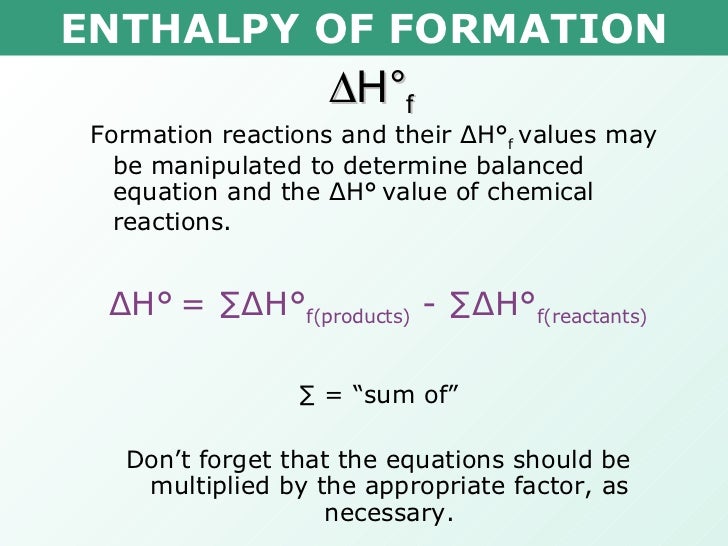
The first time I tried to calculate enthalpy of formation from combustion data, I got a result that was off by about 40 kilojoules per mole. The textbook didn't mention why. What I learned was that standard enthalpies of formation for organic compounds are often derived indirectly, and small rounding errors in the reference tables compound quickly when you're working with multi-step Hess's law problems. This is something you don't pick up from a quick review before an exam. The core equation you need is straightforward. Standard enthalpy change for a reaction equals the sum of standard enthalpies of formation of the products minus the sum of standard enthalpies of formation of the reactants, each multiplied by their stoichiometric coefficients. In symbols, that's H°rxn = nH°f(products) mH°f(reactants). The units are kilojoules per mole, and the standard state means 298.15 Kelvin and one bar pressure for gases, or one molar concentration for solutions. Here's what trips people up in practice. The H°f value for an element in its standard state is zero, but only for the standard state. Oxygen as O2 gas is zero, but ozone O3 is +142.7 kJ/mol. Nitrogen as N2 is zero, but atomic nitrogen is +472.7 kJ/mol. I once graded papers where students wrote H°f(N) = 0 and lost points because they confused the atomic form with the molecular standard state. This matters when you're setting up the equation correctly.
The method works best when you have a target reaction and you know the H°f values for all species involved. You look them up in a table, multiply by coefficients, and subtract. For example, if you want the enthalpy of formation of liquid benzene from its combustion data, you reverse the combustion reaction, use the known H°f values for CO2 and H2O, and solve for the unknown. The algebra is simple, but the sign conventions are where mistakes happen. Students frequently flip the sign when reversing a reaction instead of multiplying the entire H° by 1. A more practical problem I ran into involves phase changes. The standard enthalpy of formation for water vapor is 241.8 kJ/mol, but for liquid water it's 285.8 kJ/mol. That 44 kJ/mol difference is the enthalpy of vaporization at 298 K. If a problem gives you combustion data producing liquid water but asks for the enthalpy of formation relative to gaseous water, you need to account for that phase correction. I usually write down the phase of every species explicitly before plugging numbers in. It takes ten seconds and saves you from a systematic error. Another edge case is when the compound you're calculating doesn't exist as a stable substance under standard conditions. Some intermediates in combustion or atmospheric chemistry have positive H°f values, meaning they're thermodynamically unstable relative to their elements. The equation still works, but the physical interpretation changes. A positive enthalpy of formation tells you energy must be put into the system to create that compound from its elements. Benzene has a positive H°f of +49.0 kJ/mol, which explains why it's flammable—its formation from carbon and hydrogen is endothermic, so combustion releases that stored energy plus the energy from forming CO2 and H2O.
When working with ions in solution, you need a reference point. By convention, H°f(H+, aq) is defined as zero at all temperatures. This means all other ionic enthalpies of formation are measured relative to the proton. If you're calculating the enthalpy of neutralization for a strong acid and strong base, the net ionic equation is H+ + OH H2O, and the enthalpy change is simply the H°f of liquid water minus the H°f of hydroxide ion, since the proton contributes nothing by definition. The result is about 55.8 kJ/mol, which matches experimental calorimetry within a few percent. One limitation you should know: this approach assumes all reactants and products are in their standard states. If you're working at elevated temperatures or pressures, you need to apply heat capacity corrections using Kirchhoff's equation. The enthalpy of formation values in most tables are for 298 K only. At 500 K, the actual enthalpy change for a reaction can differ by tens of kilojoules per mole depending on the heat capacity difference between products and reactants. I usually flag this when students ask about high-temperature combustion without mentioning the temperature correction. Another practical issue is data availability. Not every compound has a measured H°f value. For some obscure organometallics or radical species, researchers estimate values using group additivity methods or compute them computationally. These estimates can have uncertainties of 10 to 30 kJ/mol, which is significant when you're trying to distinguish between two competing reaction pathways. I always check the source of the value and note the uncertainty if it's available in the table footnote.
Get the Full Details
For students working through problems, the quickest way to build fluency is to memorize the H°f values for a handful of common species: CO2(g) at 393.5 kJ/mol, H2O(l) at 285.8 kJ/mol, H2O(g) at 241.8 kJ/mol, NO2(g) at +33.2 kJ/mol, and NH3(g) at 46.1 kJ/mol. These appear in at least half of introductory thermodynamics problems. The rest you look up. Don't try to memorize the whole table—it's not efficient use of study time.
Common Pitfalls and How to Avoid Them
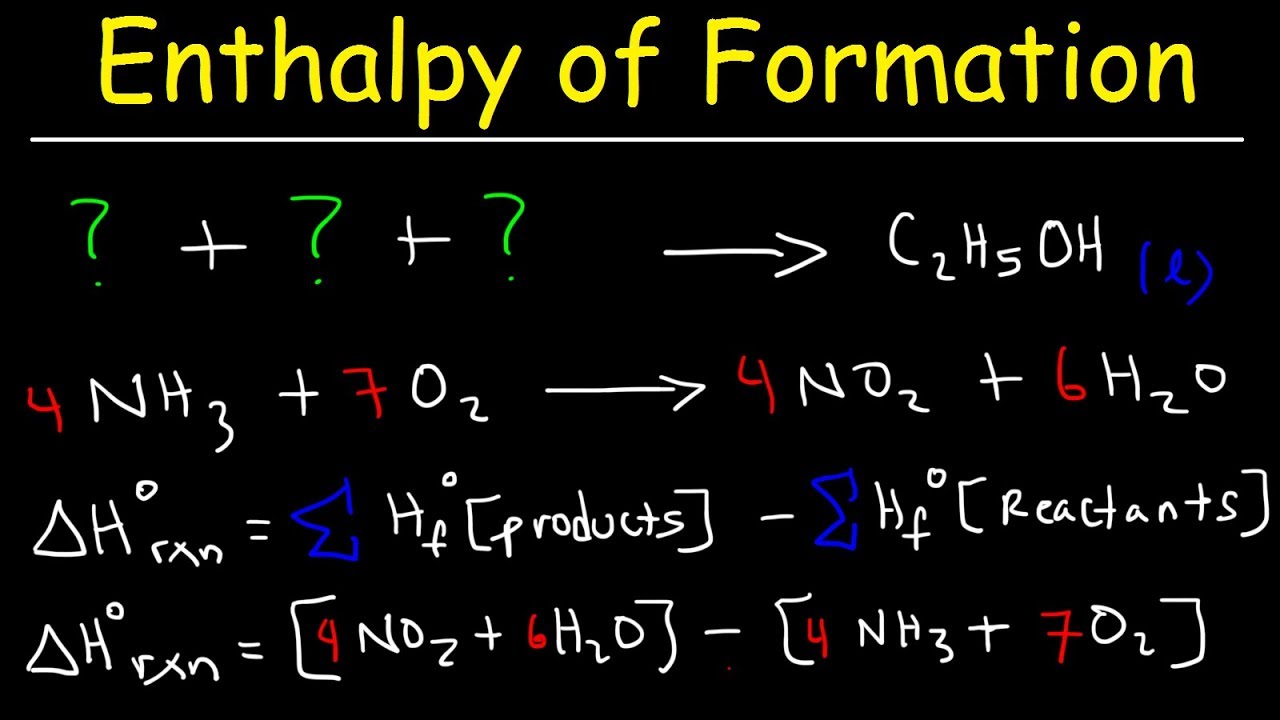
The biggest source of error is forgetting to multiply by stoichiometric coefficients. If your balanced equation has 2 moles of NH3, you must use 2 × H°f(NH3), not just H°f(NH3). I see this mistake in about one in four problem sets. Another frequent error is using the H°f for H2O(g) when the reaction produces liquid water, or vice versa. Always check the phase indicated in the problem statement or in the table entry you're using. A more subtle issue arises when dealing with reactions that don't go to completion. The enthalpy of formation equation assumes complete conversion according to the balanced equation. If you're working with equilibrium systems or partial conversion, you need to adjust your approach using extent of reaction or conversion fractions. This comes up more often in chemical engineering than in general chemistry, but it's worth knowing about if you're doing process design work later. If you need to look up values, the NIST Chemistry WebBook is the most reliable free resource. It lists standard enthalpies of formation with references to the original literature. Some university libraries also provide access to the JANAF Thermochemical Tables, which are the gold standard for high-precision work. For most coursework, the appendix values in your textbook are adequate, but they sometimes have outdated entries or typographical errors. I once caught a textbook listing H°f(SO2) as 296.8 when the correct value is 296.1 kJ/mol—a small difference that mattered for a precision calculation in my research.
The equation itself doesn't tell you whether a reaction is spontaneous. For that you need Gibbs free energy and entropy. Enthalpy is just one piece of the puzzle. A reaction can be endothermic and still proceed spontaneously if the entropy increase is large enough, like the dissolution of ammonium nitrate in water. Don't confuse exothermic with spontaneous—that's a separate misconception that shows up on exams regularly.