Calculating Osmotic Pressure Without Burning Three Hours
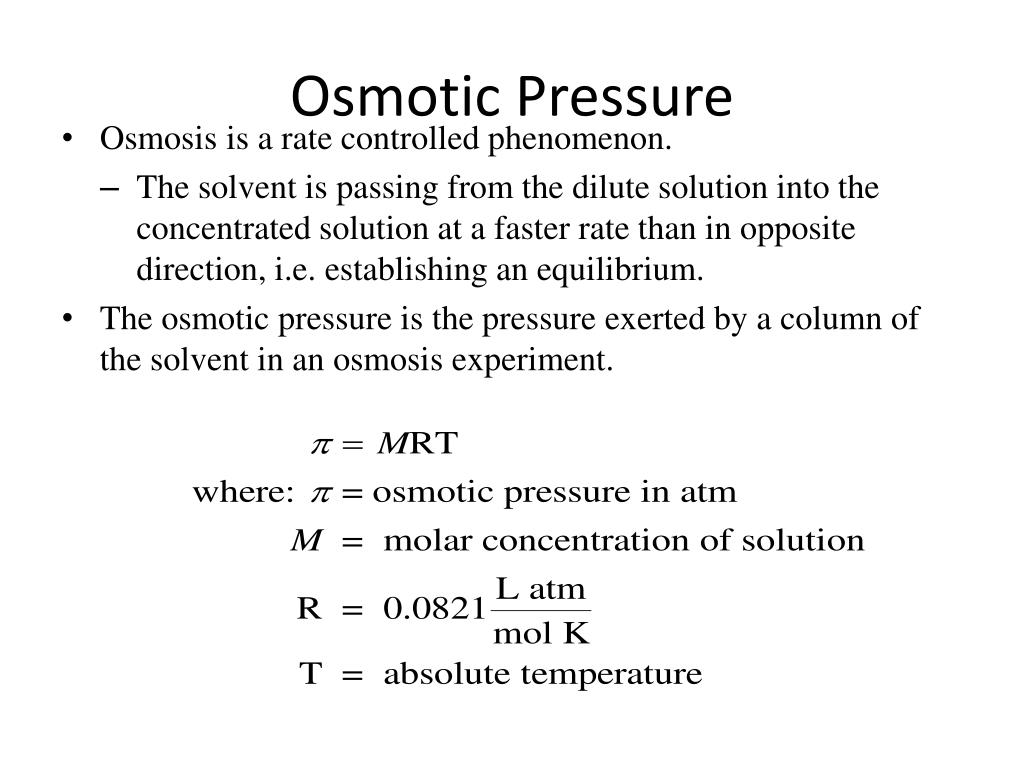
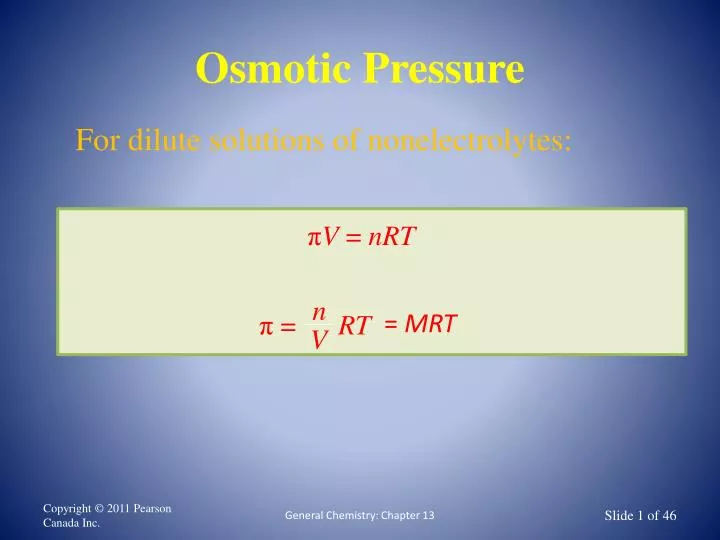
I used to treat osmotic pressure calculations like they were straightforward textbook problems, then spent a full afternoon recalibrating my spectrophotometer because I ignored the temperature dependency in my van 't Hoff factor work. That was before I learned to stop trusting standard conditions and start accounting for real solution behavior every single time. The Equation For Osmotic Pressure is essentially the van 't Hoff equation, and in its simplest form it looks deceptively tame: = iMRT. But that simplicity is exactly where people get tripped up, because the variables carry more weight than the formula suggests. represents osmotic pressure in atmospheres, i is the van 't Hoff factor accounting for solute dissociation, M is molarity, R is the ideal gas constant at 0.0821 L·atm/(mol·K), and T is absolute temperature in Kelvin. The derivation comes directly from comparing osmotic equilibrium to ideal gas law behavior, which is why the same constant R shows up here instead of some other coefficient.
Getting the Equation For Osmotic Pressure Right on the First Pass
Most people skip the unit conversion step and wonder why their answer lands somewhere between a atmospheric reading and a kilopascal value depending on which textbook they cross-reference. The biggest pitfall I see is using Celsius instead of Kelvin for T. A 25°C solution entered as 25 instead of 298.15 gives you an osmotic pressure roughly twelve times too low. That error doesn't announce itself. It just sits there quietly until someone tries to actually make something work at that pressure. Another thing nobody emphasizes enough is the difference between molarity and molality in osmotic pressure calculations. The van 't Hoff equation technically uses molarity, but molarity shifts with temperature because volume expands. If you're working at elevated or variable temperatures, converting to molality and then back, or simply using the molal osmotic coefficient approach, keeps your numbers honest. I switched to the molality-based method after watching a batch fail during a temperature swing from 20°C to 37°C, and the discrepancy was about 4% — small on paper, catastrophic at scale. For non-ideal solutions, which is most real-world samples, you need the osmotic coefficient multiplied into the equation, making it = iMRT. The osmotic coefficient corrects for ion pairing, activity deviations, and intermolecular forces that the ideal model ignores. At concentrations above 0.1 M for electrolytes, can deviate significantly from 1.0. Ignoring it means your calculated pressure will be wrong, sometimes by more than 10% for divalent salts.
Here is a practical walkthrough. Say you have a 0.5 M NaCl solution at 25°C. NaCl dissociates into two ions, so the van 't Hoff factor i is approximately 2, though experimentally it might sit closer to 1.9 due to ion pairing. The osmotic coefficient for 0.5 M NaCl is roughly 0.92. Plugging in: = 2 × 0.5 × 0.0821 × 298.15 × 0.92. That gives you about 22.5 atm. If you skipped the osmotic coefficient entirely and assumed ideal behavior, you would get 24.4 atm. The difference looks minor until you are designing a membrane system that operates right at that threshold.
Get the Full Details
Where This Equation Breaks Down and What to Use Instead
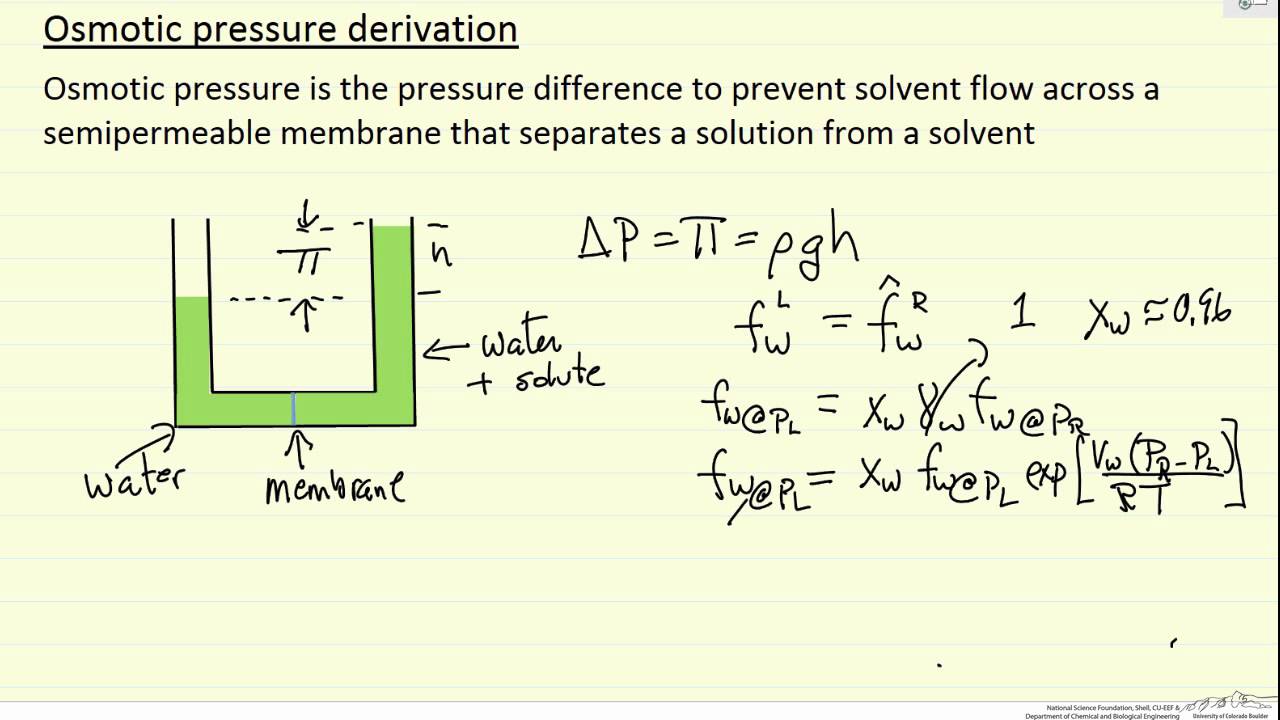
The van 't Hoff equation assumes dilute, ideal solutions. When you move into concentrated regimes, polyelectrolyte systems, or membranes with significant charge effects, it stops being useful. I ran into this directly while working with a PEG-based solution for a drug delivery application. The molecular weight distribution meant the osmotic pressure was nowhere near what the molar concentration predicted, and the standard equation gave readings that were completely off because PEG doesn't dissociate the way salts do and its effective particle count depends heavily on hydration shell behavior. In those cases, you need to shift to the virial expansion form: /RT = c/M + Bc² + Cc³, where c is concentration, M is molecular weight, and B and C are virial coefficients that capture non-ideal interactions. Getting those coefficients usually means measuring osmotic pressure at several concentrations and fitting the curve, or pulling them from literature for your specific polymer-solvent system. There is no shortcut around that. Another common failure mode is applying the equation across semipermeable membranes with significant solute rejection, not 100%. The theoretical osmotic pressure assumes perfect separation of solute and solvent. In practice, if your membrane allows even a small fraction of solute through, the effective osmotic pressure drops. I learned this the hard way during reverse osmosis testing where our calculated didn't match observed water flux, and the discrepancy traced back to a 3% solute passage rate through the membrane that nobody had measured.
For membrane design and reverse osmosis applications, the solution diffusion model or the modified van 't Hoff approach with a rejection coefficient incorporated tends to work better than raw osmotic pressure calculations. If you are working in biotech or pharmaceutical formulation, you will also want to account for Donnan equilibrium effects whenever charged species are present on one side of a membrane. That adds another layer of complexity the basic equation does not address at all. The bottom line is that the Equation For Osmotic Pressure is a starting point, not a destination. Get the basics right, watch your units, include the osmotic coefficient for anything beyond dilute solutions, and know when to abandon it entirely. Most errors I encounter come from people using the simple form well past its validity range and not realizing it until the numbers don't add up.