Working With Radioactive Decay Equations in Practice
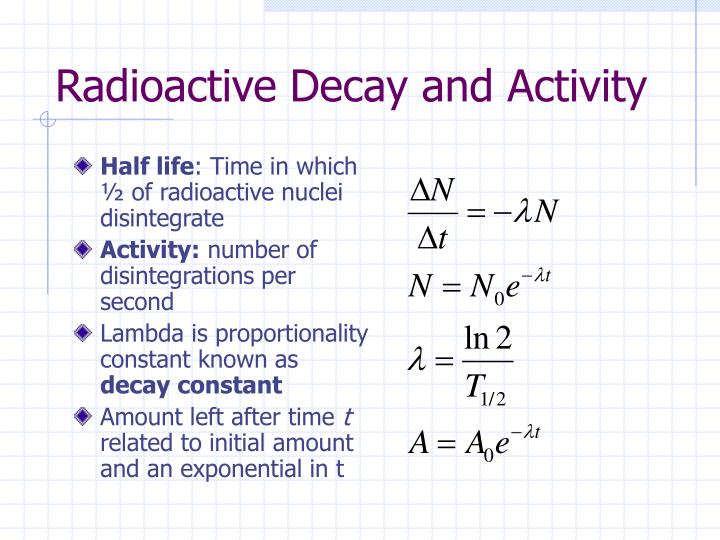
The Equation For Radioactive Decay is something most people learn as N(t) = Ne^(-t) in a first-year physics class and then never really think about again until they need it for actual work. When you actually use it, there are a few things that the textbook versions don't tell you, and I ran into them head-on a few years ago when I was recalibrating a gamma spectroscopy setup for environmental samples. Let me walk through how it actually works before we get into the messy parts.
Equation For Radioactive Decay and What Each Term Actually Means
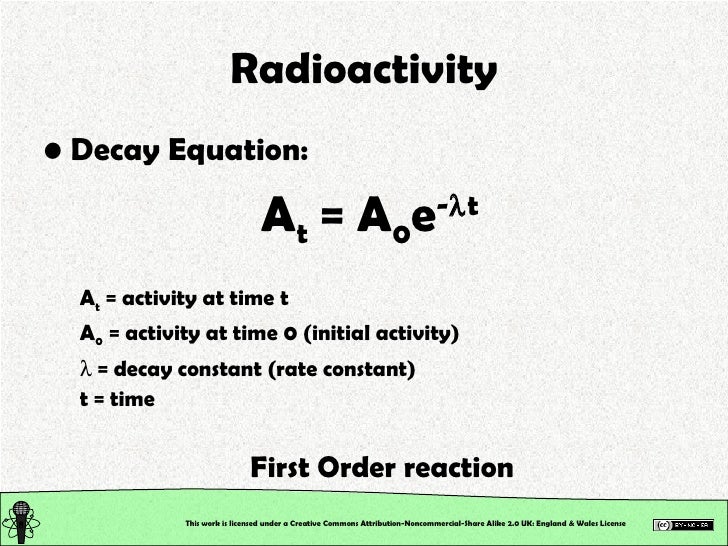
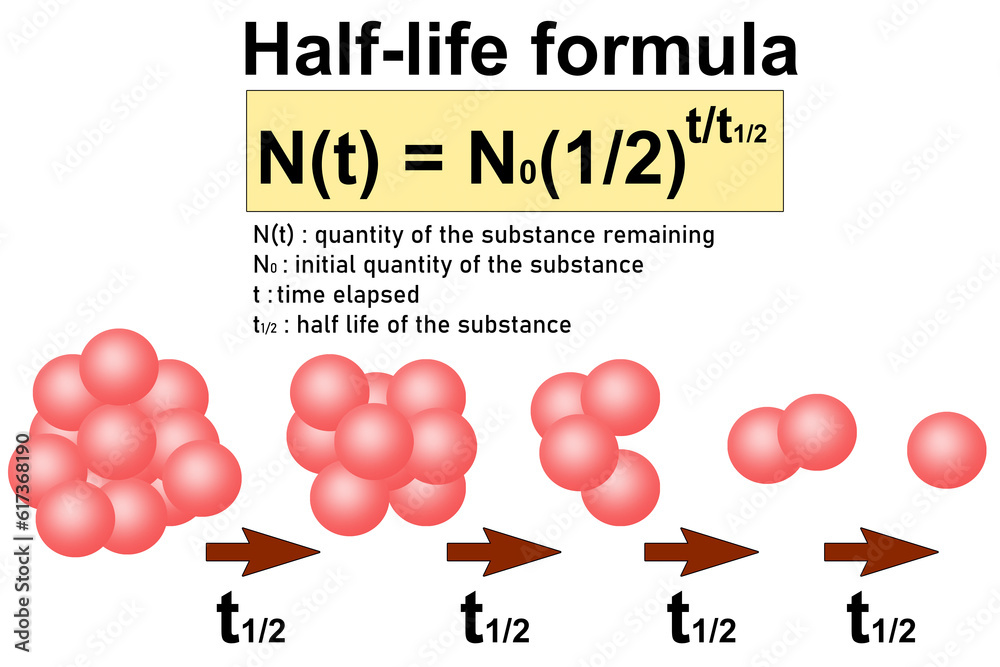
The basic form is: N(t) = N × e^(-t) N(t) is the number of atoms remaining at time t. N is the starting number of radioactive atoms. (lambda) is the decay constant, which is related to half-life by = ln(2)/t/. And t is the elapsed time. The activity form, which you'll see more often in lab work, is A(t) = A × e^(-t), where A is the measured count rate or disintegrations per second.
That's the straightforward part. The hard part comes when you start applying this to real measurements, because real measurements are never clean. I remember being handed a set of soil samples with Cs-137 contamination, and the lab had recorded the initial activity on a date that was, turns out, about three weeks earlier than the certificate of analysis stated. I didn't catch it at first. I plugged the activity straight into the decay equation without accounting for the actual elapsed time between the measurement date and the reporting date, and my calculated concentrations were off by roughly 2.1%. For trace-level environmental work, that's the difference between compliance and a failed audit. The fix was simple in hindsight — go back to the raw instrument logs, find the actual acquisition timestamp, and recalculate the decay correction using the true delta-t. But getting those logs required digging through old CSV files from a spectrometer that hadn't been used in two years.
Get the Full Details
When the Standard Equation Breaks Down
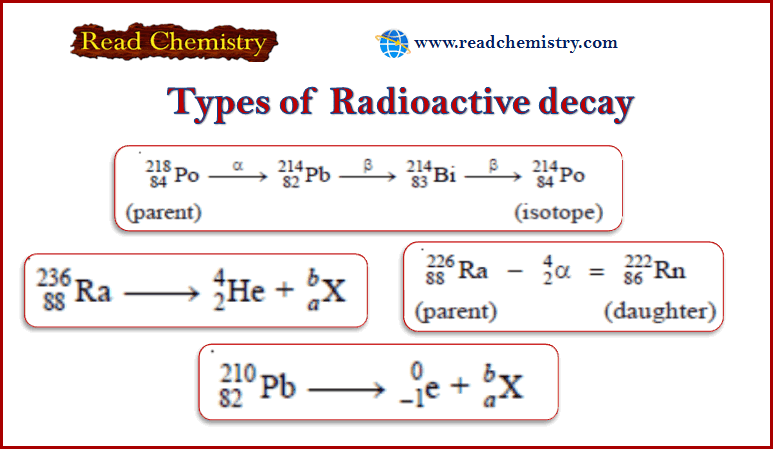
Here's what most guides skip: the single-isotope exponential decay equation assumes you're dealing with one nuclide decaying into something stable, or at least something you're not measuring. That assumption evaporates pretty quickly in anything beyond a textbook problem. When you have a decay chain — say, Ra-226 decaying through several daughters before landing on Pb-206 — the simple equation no longer applies. You need the Bateman equations. These account for the fact that daughter products are themselves radioactive and building up over time. I worked a case involving Po-210 in a dried beverage sample where the parent Pb-210 was still actively producing Po-210 through decay. If I had just used the standard exponential decay equation on the Po-210 activity without considering the ingrowth from Pb-210, the result would have been wildly wrong. The actual workflow involved solving the coupled differential equations numerically rather than analytically, which meant writing a short script in Python using scipy.integrate.odeint. That took maybe twenty minutes to set up once, and then it handled the chain correctly every time. Another thing nobody mentions is the problem of secular equilibrium. When a long-lived parent has a short-lived daughter, after about five half-lives of the daughter, the system reaches equilibrium and the activities become approximately equal. In practice, this means you can measure the daughter's gamma peaks and infer the parent's activity without having to separate anything chemically. It saved us weeks of lab work on a uranium series analysis because we just counted the Bi-214 and Pb-214 peaks and assumed equilibrium held. The catch is that equilibrium doesn't always hold. If the sample has been chemically processed — which environmental samples often have been, through leaching or precipitation — the daughter products can be fractionated away from the parent. You have to verify equilibrium independently, usually by checking multiple points in the chain, before you rely on this shortcut.
Practical Calculation Steps
For straightforward single-isotope work, here's the process I actually use instead of what the textbooks suggest: First, identify your isotope and look up the half-life from an authoritative source like the NNDC (National Nuclear Data Center) or the IAEA radionuclide database. Don't rely on memory or a random website — different sources sometimes list slightly different values, and for high-precision work that matters. For Cs-137, for instance, the accepted half-life is 30.05 years, but some older references say 30.17. That 0.4% difference adds up over long decay corrections. Second, calculate the decay constant. Lambda equals 0.693147 divided by the half-life, but make sure your time units match. If your half-life is in years and your elapsed time is in days, convert one of them. This sounds obvious, but it's the single most common error I see in lab reports. Someone will plug a half-life in seconds into an equation where everything else is in hours, and the result will be nonsense by several orders of magnitude.
Third, determine the elapsed time with precision. Not "roughly last month," but the exact number of seconds or days between the reference activity date and your current measurement date. I keep a running spreadsheet with a cell that auto-calculates days elapsed from a reference date using a simple datediff formula, so I never have to count manually. Fourth, apply the equation. A(t) = A × e^(-t). If you're working in Excel, the function is =A0*EXP(-lambda*t). In Python, it's A0 * math.exp(-lambda * t). Both give identical results to far more decimal places than your measurement precision justifies. Fifth, and this is where people cut corners, propagate the uncertainty. Your half-life has an uncertainty. Your initial activity has an uncertainty from counting statistics and calibration. Your elapsed time has uncertainty if the reference date isn't exact. The final uncertainty on A(t) comes from combining all of these. For exponential decay, the relative uncertainty in the result is roughly the sum of the relative uncertainties in A and t, though you should really do a proper error propagation calculation rather than just adding them. A Monte Carlo approach — sampling from the uncertainty distributions of each input parameter and running the equation thousands of times — gives you the most reliable uncertainty estimate, and it takes about thirty seconds to code.
Common Pitfalls That Waste Time
Branching decay is another one that trips people up. Some isotopes don't decay 100% through a single pathway. For example, N-16 decays primarily by beta emission to O-16, but a small branching ratio goes to an excited state that emits a different gamma energy. If you're quantifying using a specific gamma peak, you need the branching ratio — the gamma emission probability per decay — not just the raw decay equation. The IAEA publication "Gamma Ray Transition Intensities" is the standard reference for these values, and the numbers have been updated several times as measurement techniques improved. Self-absorption in your sample is another practical issue. The decay equation tells you how many atoms remain, but your detector doesn't see all of them equally. Dense or thick samples absorb their own gamma rays before they reach the crystal. I spent an afternoon characterizing this for a set of wet sediment samples by comparing measured activities against known standards prepared in identical matrices. The correction factor was about 12% for our heaviest samples, and applying it changed the final result enough to flip a regulatory decision. If you skip matrix-matched calibration, your numbers are just guesses with extra steps. There's also the problem of dead time in your detector. At high count rates, the electronics can't process every event, and you lose counts. The standard paralyzable or non-paralyzable dead time correction formulas exist, but they break down at very high rates — above about 10% dead time, the corrections become unreliable and the uncertainty explodes. The workaround is simple: dilute the sample or increase the source-to-detector distance. I once had a concentrated waste sample that was reading at 45% dead time. No amount of mathematical correction was going to save those numbers. We diluted it 1:10 and got clean data on the second pass.
Software Options
If you're doing this kind of work regularly, you probably don't want to calculate everything by hand or in a spreadsheet. A few tools I've actually used: SimplerLab is a decent entry-level tool for basic decay calculations and half-life lookups. It's free for non-commercial use and handles single-isotope decay well. The interface is functional if a bit dated, but it gets the job done for routine work. You can find it at simplerslab.com. For decay chains and more complex scenarios, RADAR from the National Center for Radiological Standards is more capable, though it's a Windows-only application and the interface feels like it was last updated in 2003. It handles Bateman equations internally, so you don't need to code anything.
Many labs just write their own scripts. Python with the built-in math module handles single-isotope decay in three lines. For chains, the phits package or even a custom NumPy implementation covers most practical needs. I keep a reusable Python module with functions for decay correction, ingrowth calculation, and uncertainty propagation. It took me about an afternoon to build, and it's saved me countless hours since.

When to Question Your Results
The decay equation itself is not approximate — it's exact for a single isolated nuclide. But every input to that equation comes with uncertainty, and some of those uncertainties are systematic rather than statistical. Calibration drift in your detector, changes in geometry between measurements, variations in sample density, and even temperature effects on the detector response can all introduce bias that dwarfs the statistical uncertainty from counting. I've seen cases where the calculated decay correction was smaller than the uncorrected calibration drift, meaning the decay math was the least significant source of error in the whole measurement chain. If your decay-corrected results show inconsistencies — say, duplicate samples give widely different activities after correction, or results contradict known physical constraints — the problem is almost never the equation. It's somewhere in the measurement chain. Start by checking the calibration, then the geometry, then the sample preparation, and only then revisit your decay correction assumptions. The equation itself is straightforward. The work is in making sure everything around it is right.