Understanding Chemical Equilibrium Without the Textbook Gimmicks
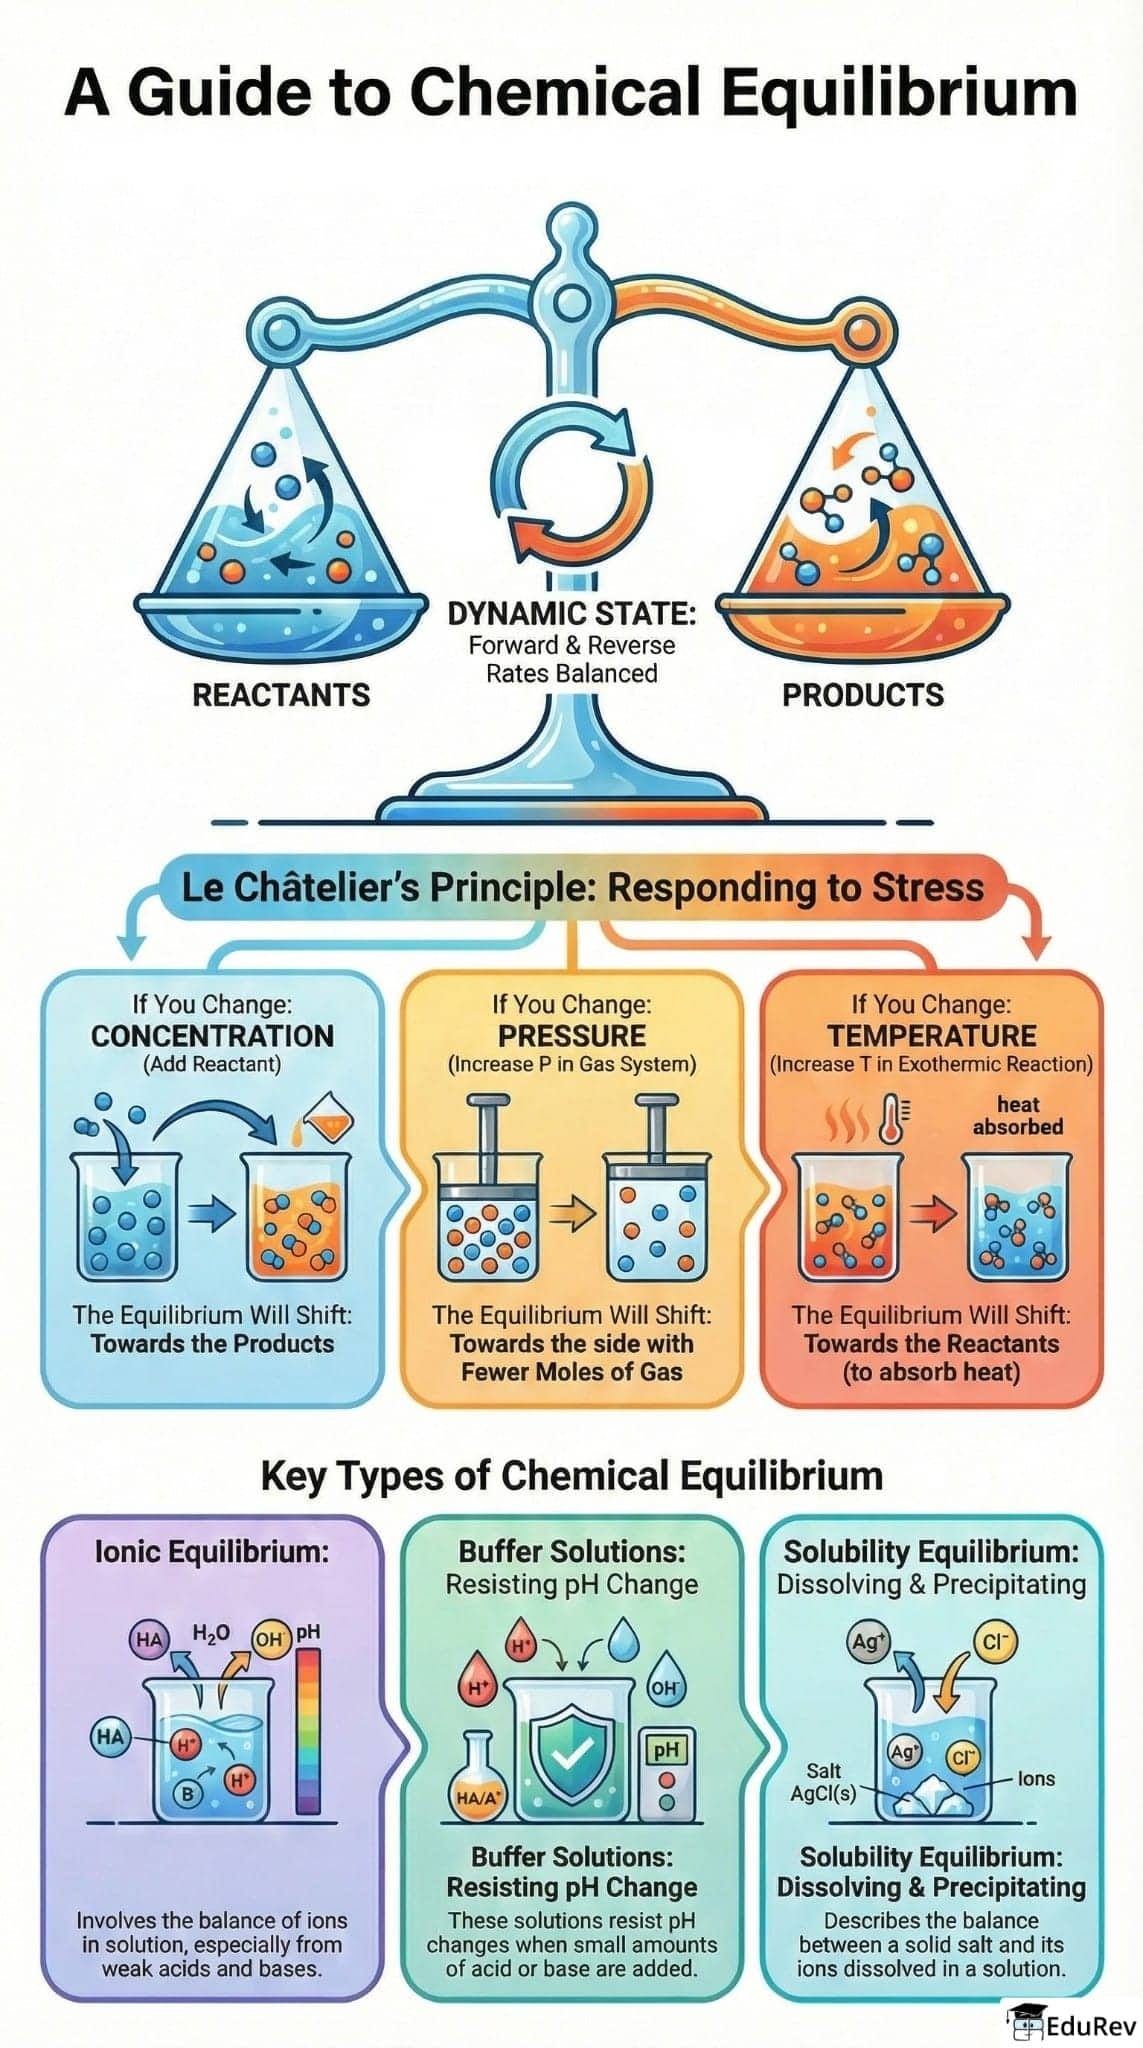
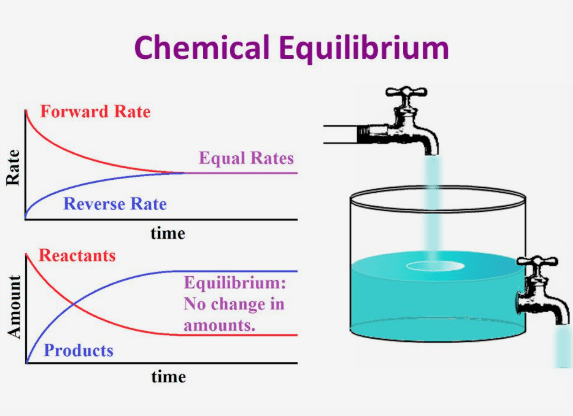
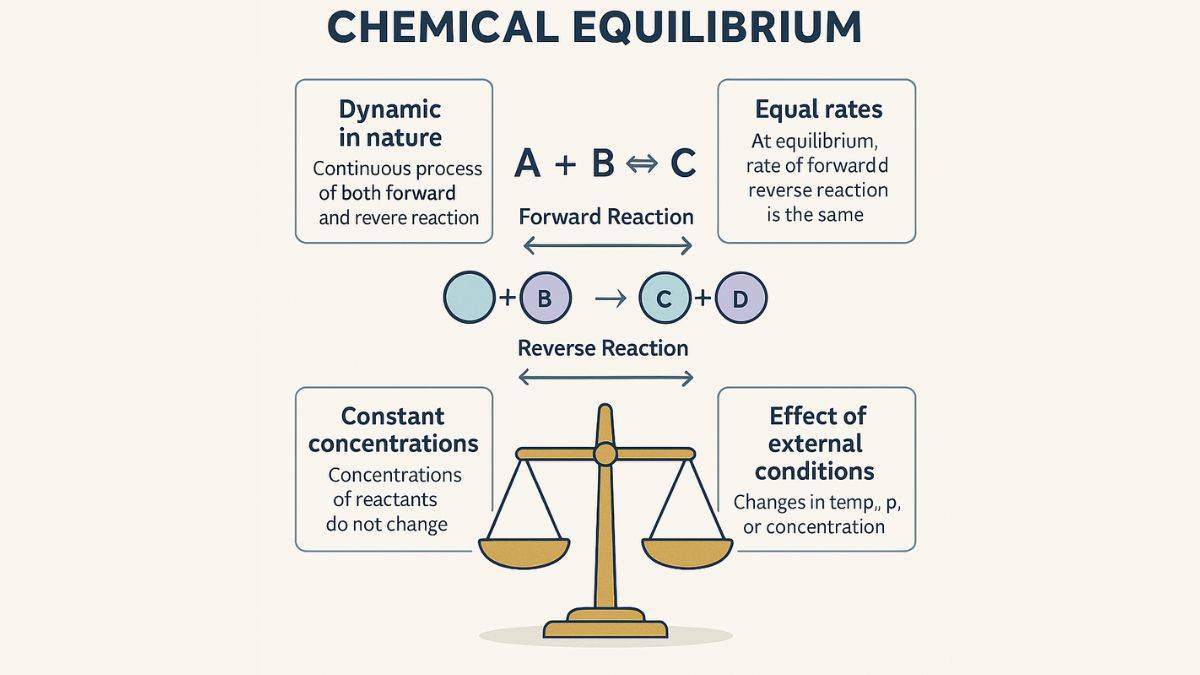
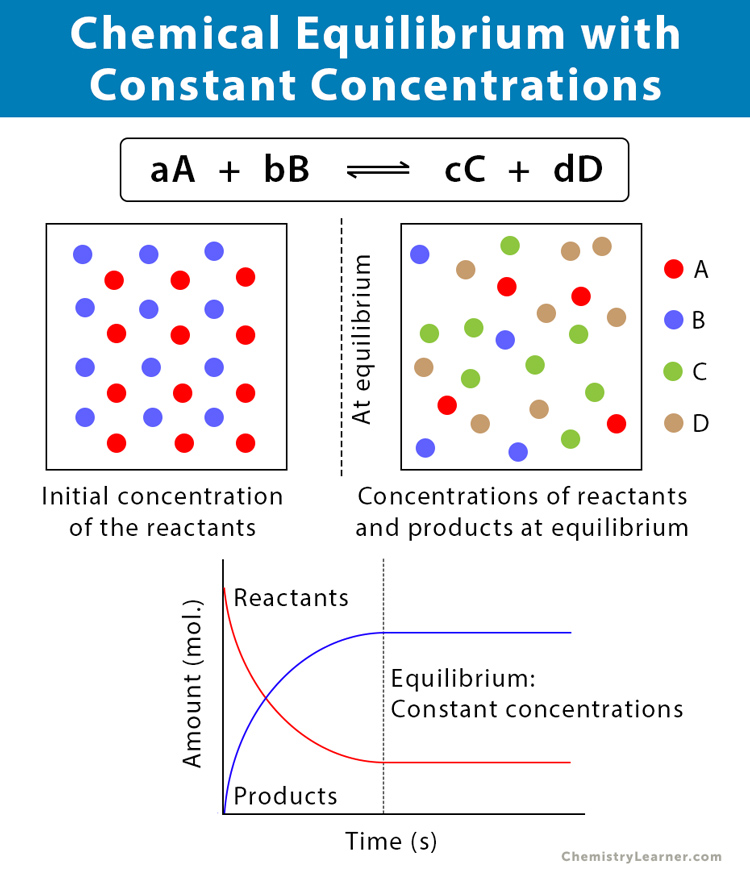
I spent three years in analytical chemistry labs before I actually understood what equilibrium means instead of memorizing Le Chatelier's principle for exams. Here's the thing nobody tells you: equilibrium doesn't mean "nothing is happening." It means everything is happening at the same rate in both directions, and that distinction changes how you approach every calculation. At its core, chemical equilibrium describes a state where the forward and reverse reaction rates are equal, resulting in no net change in concentrations of reactants and products over time. The equilibrium constant K tells you the ratio of product concentrations to reactant concentrations at that steady state, each raised to their stoichiometric coefficients. But K is temperature-dependent, which matters more than most students realize. The practical application starts with writing the equilibrium expression correctly. Take a simple reaction: N(g) + 3H(g) 2NH(g). The equilibrium expression becomes K = [NH]² / ([N][H]³). You need to be careful about states of matter here. Solids and pure liquids don't appear in the expression. I had a student once include water as a liquid in her K expression for an aqueous reaction, which threw off her entire calculation. She was frustrated for weeks before she caught it.
The ICE Table Method: Your Most Reliable Tool
ICE stands for Initial, Change, Equilibrium. It's not glamorous but it works when other methods fail. You set up a table tracking concentrations through the reaction progress, solve for the unknown using your equilibrium expression, and usually end up with a quadratic equation or something worse. Here's a real example from my teaching experience. Consider the decomposition of PCl at 250°C: PCl(g) PCl(g) + Cl(g), with K = 1.80. If you start with 0.100 M PCl, your ICE table looks like this: PCl: 0.100 -x 0.100 - x
PCl: 0 +x x
Cl: 0 +x x
Setting up the equation: K = x²/(0.100 - x) = 1.80. This gives you x² + 1.80x - 0.180 = 0. Using the quadratic formula, x 0.0935 M. The equilibrium concentration of PCl is approximately 0.0065 M, meaning about 93.5% decomposition. That seems high but it's consistent with the large K value. One common mistake: students often assume x is small and skip the quadratic. That approximation only works when K is very small (usually less than 10) relative to your initial concentration. For the Haber process calculations I regularly do in industrial chemistry contexts, this assumption fails almost immediately.
Get the Full Details
When Equilibrium Calculations Break Down
I wish someone had told me earlier about activity coefficients. The standard equilibrium expressions assume ideal behavior, meaning activities equal concentrations. Real solutions, especially at higher ionic strengths, deviate significantly. In my experience with concentrated electrolyte systems, using activities instead of concentrations can shift calculated equilibrium positions by 20-40%. The Debye-Hückel equation helps estimate activity coefficients, though it has its own limitations at high concentrations. Another issue that trips people up: equilibrium only applies to closed systems at constant temperature. Open systems, flowing reactors, and biological systems with continuous input and output aren't at equilibrium. I worked on a project involving enzymatic reactions where the "steady state" approximation looked superficially like equilibrium but was fundamentally different. The product was being consumed as fast as it formed, creating a false impression of equilibrium conditions. Heterogeneous equilibria present their own challenges. When solids participate, only the gaseous or dissolved species appear in K expressions. But surface area affects reaction rates, not equilibrium positions. I've seen multiple students confuse kinetics with thermodynamics on this point. A finely divided solid reacts faster than a single large piece, but both reach the same equilibrium composition given sufficient time.
Multiple Equilibria and Coupled Reactions
Real systems rarely involve just one equilibrium. Acid-base chemistry in blood involves carbonate, phosphate, and protein buffers all interacting simultaneously. I found that writing mass balance equations alongside charge balance equations provides the most systematic approach to solving these problems. For a practical case involving solubility equilibria, consider AgCl in the presence of ammonia. The silver ions form a complex with ammonia, shifting the solubility equilibrium. You need to solve two equilibrium expressions simultaneously: Ksp for AgCl and Kf for the complex formation. This is where systematic methods like the algebraic approach or numerical iteration become necessary. Spreadsheet solving or even simple graphical methods work well for these problems. My go-to workaround when facing coupled equilibria that resist analytical solution: define all relevant equilibria, write the mass balance equations, identify the dominant species at approximate concentrations, and iterate. This usually converges within two or three cycles for typical laboratory conditions. For extreme cases with multiple competing equilibria, specialized software packages like PHREEQC or MINTEQ handle the matrix algebra efficiently.
Le Chatelier's Principle: Proper Application
Everyone learns Le Chatelier's principle, but few apply it correctly. The principle predicts how a system at equilibrium responds to disturbances: pressure changes, concentration changes, or temperature changes. However, it's easy to misapply it. Consider adding an inert gas at constant volume versus constant pressure. At constant volume, total pressure increases but partial pressures remain unchanged, so equilibrium position doesn't shift. At constant pressure, the system volume expands, partial pressures decrease, and equilibrium shifts toward more moles of gas. I've seen this distinction tested repeatedly because it catches students who memorize without understanding. Temperature effects require knowing whether the reaction is exothermic or endothermic. For the Haber process, H = -92 kJ/mol, making it exothermic. Increasing temperature shifts equilibrium toward reactants, reducing ammonia yield. This is why industrial ammonia synthesis operates at elevated temperatures despite the equilibrium penalty: kinetics demand higher temperatures, and the actual yield is managed through recycling unreacted gases.
The practical compromise in industrial equilibrium processes usually involves balancing yield against rate and economic factors. High pressure favors ammonia production but requires expensive equipment. Moderate temperature provides acceptable rates. Catalysts speed both forward and reverse reactions equally, improving rate but not equilibrium yield. Understanding these tradeoffs separates competent engineers from those who just follow textbooks.
Measuring Equilibrium Constants Experimentally
Direct measurement requires reaching equilibrium before taking readings, which can take hours or days depending on the system. Spectrophotometry works well for colored species, allowing continuous monitoring without disturbing the system. I prefer this method over sampling approaches because it eliminates perturbation errors. For gas-phase equilibria, manometric measurements of pressure changes work straightforwardly. The key is ensuring temperature stability, since K varies with temperature. Even 0.1°C fluctuations can introduce measurable errors in precise work. Solving equilibrium problems becomes intuitive with practice. Start with simple single-reaction systems, master the ICE table approach, then progress to coupled equilibria and non-ideal systems. The mathematical complexity increases, but the fundamental principles remain consistent across all scenarios.
Equilibrium concepts underpin countless applications beyond textbook problems: drug delivery systems, environmental chemistry, materials processing, and biological energetics. Understanding what equilibrium means beyond the definition helps you recognize these connections and apply the principles flexibly across different contexts.