Understanding Lewis Bases Through Practical Use
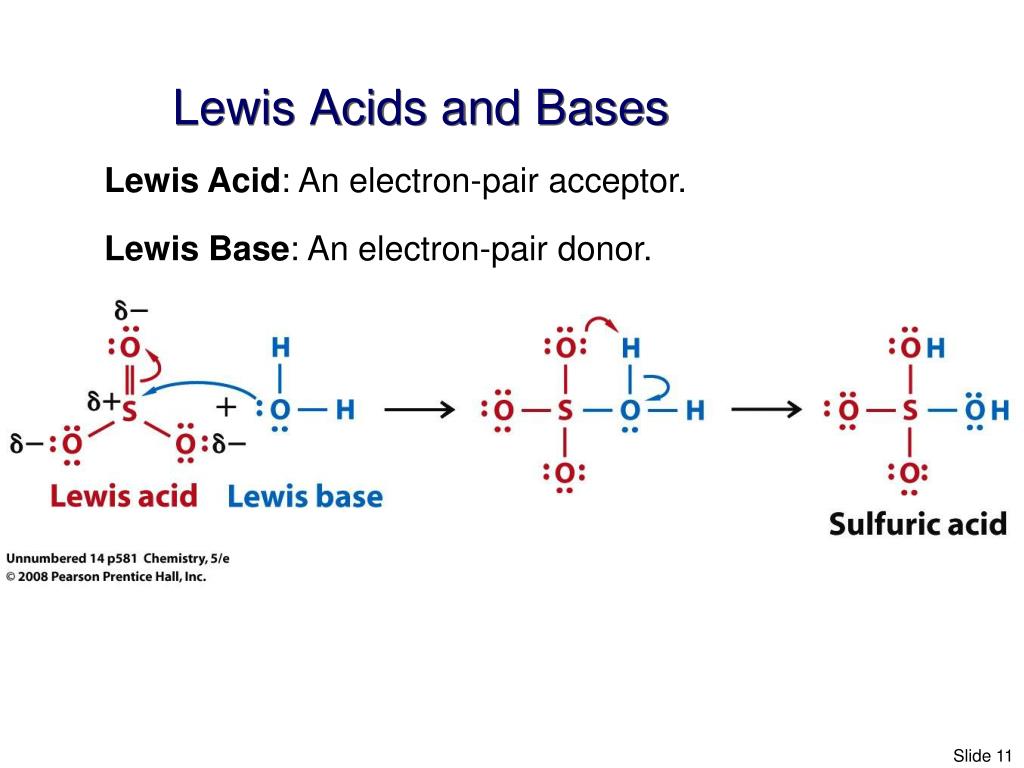
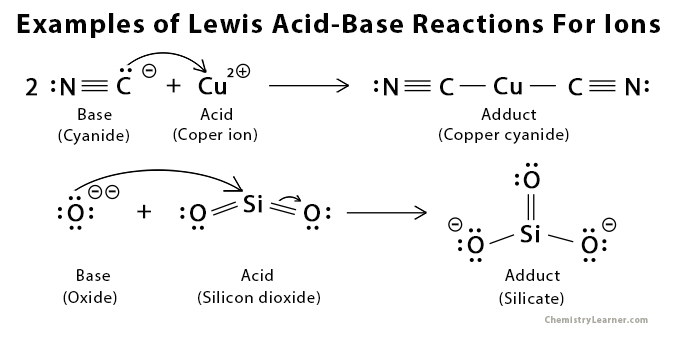
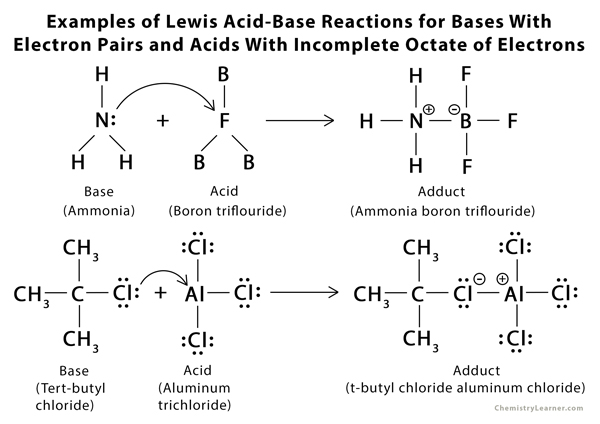
A Lewis base is simply a species that can donate a pair of electrons to form a coordinate covalent bond. That's it. No drama, no fancy language needed. The definition comes from Gilbert Lewis back in 1923, and it's actually more general than the Brønsted-Lowry idea because it doesn't require anything to do with protons at all. Any molecule or ion with a lone pair that it's willing to share qualifies, and that range is surprisingly wide. Ammonia (NH3) is the textbook answer most people learn first, and for good reason. The nitrogen has one lone pair sitting right there on the surface of the molecule, ready to donate it to any Lewis acid that comes along. When you mix ammonia with boron trifluoride (BF3), that lone pair attacks the electron-deficient boron and forms NH3-BF3. Clean, straightforward reaction. Water works the same way with its two lone pairs on oxygen, and hydroxide (OH-) is even more reactive because the negative charge makes that electron density much easier to push toward an acceptor. Halide ions like Cl- and Br- are also classic Lewis bases, though they're weaker than ammonia or hydroxide because the electron density is spread over a larger, more diffuse orbital. The tricky part isn't memorizing the examples. It's figuring out which lone pairs are actually available and how strongly they'll bind in a given situation. Not every lone pair on every molecule participates equally. Consider a molecule like dimethyl sulfoxide (DMSO). The oxygen has two lone pairs, but one is held in an sp2-type orbital that points away from the sulfur, and the other is more tightly bound in a different orientation. In practice, DMSO coordinates through oxygen in most reactions, but there are edge cases where the sulfur itself can act as the donor center under special conditions, especially with very soft metal centers. I ran into this when I was trying to coordinate a late-transition-metal complex and kept getting unexpected product distributions. The solvent wasn't just sitting there inert — DMSO was slowly displacing the intended ligand through the sulfur rather than the oxygen, and it took me three failed runs before I caught it by running a 1H NMR on the crude mixture and noticing the methyl peaks had shifted in a way that only made sense if the coordination geometry around sulfur had changed.
The workaround was switching to acetonitrile as the solvent, which is a much harder donor and doesn't compete nearly as aggressively with the ligands I actually wanted bound to the metal. That costs about an extra day of work per project because you have to optimize everything again in a different solvent system, but it saved me from spending weeks chasing artifacts.
How To Identify And Work With Lewis Bases in Practice
Start by looking at the structure and finding lone pairs. Atoms like nitrogen, oxygen, phosphorus, and sulfur are your usual suspects. But here's what most people miss: the strength of a Lewis base depends heavily on the context, not just the presence of a lone pair. Hardness and softness matter a lot. A hard Lewis base like fluoride or ammonia prefers to bind to a hard Lewis acid like boron or aluminum. A soft Lewis base like triphenylphosphine or iodide will happily bind to a soft acid like palladium or platinum instead. If you mix hard with soft, the bond tends to be weak and the equilibrium doesn't favor the adduct. This is the HSAB principle, and it's something you need to keep in mind constantly when designing reactions or choosing ligands for catalysis. Another thing beginners regularly get wrong is assuming that a negatively charged species is always a stronger Lewis base than its neutral counterpart. That's true most of the time — hydroxide is a stronger donor than water — but it breaks down when steric hindrance gets involved. Tris(tert-butyl)phosphine is a ridiculous molecule to look at, and despite having a lone pair on phosphorus, it's an exceptionally poor Lewis base in practice because nothing can get close enough to that lone pair for effective donation. I once tried to use it as a ligand precursor in a Pd-catalyzed cross-coupling reaction and spent two days wondering why the catalyst just wouldn't activate. The phosphine was too bulky to coordinate properly to the metal center, so the active species never formed. Switching to tri-tert-butylphosphine with shorter methyl branches instead fixed the problem immediately, though the yield was still lower than with less bulky analogues because the steric protection now working against product release. When you're actually running a reaction, think about what you're trying to achieve. Are you stabilizing a reactive intermediate? Then pick a Lewis base that binds strongly enough to protect it but weakly enough to let it react when needed. Tertiary amines like triethylamine are commonly used as bases in organic synthesis because they're good at deprotonating without coordinating too tightly to anything else in the mixture. If you need something stronger, you might go with an amide base like LDA, but then you're dealing with air and moisture sensitivity that adds real time to your workflow.
Get the Full Details
The counterintuitive part about Lewis basicity is that solvent effects can completely flip the order. In gas phase, ammonia is a much stronger base than water, which matches what you'd predict from basicity tables. But in aqueous solution, the picture changes because solvation stabilizes the charged species differently. A small ion like F- gets heavily solvated by water, which pulls electron density away and makes it less available for donation. A larger ion like I- is poorly solvated and its lone pairs remain more accessible, which is why iodide behaves as a better nucleophile and Lewis base in polar protic solvents even though it's the conjugate base of a strong acid. This is why your pKa tables won't always predict reactivity correctly — they measure proton affinity, not electron-pair donation to a general Lewis acid. If you're working with organometallics or main-group chemistry, you'll also run into the issue that some Lewis bases can bridge between multiple metal centers, acting as mu-2 or mu-3 ligands. Alkoxides and amides do this frequently, and it changes the stoichiometry and structure of your product entirely. One equivalent of base might tie up two metal centers instead of one, which throws off your calculations if you haven't accounted for it. I've seen this ruin a lot of reactions because people treat alkoxides as simple mono-dentate bases when the metal center they're targeting is electron-deficient enough to pull the oxygen into bridging mode. The bottom line is that identifying a Lewis base is easy. Predicting how it will behave in a real reaction requires knowing the acid it's paired with, the solvent environment, the steric constraints, and whether there are competing coordination sites. Write down the structures of everything involved, check the hardness and softness, and think about what each lone pair is actually doing before you start mixing things together. It saves a lot of wasted reagents and failed reactions.