How to Tell Them Apart Without Overcomplicating It
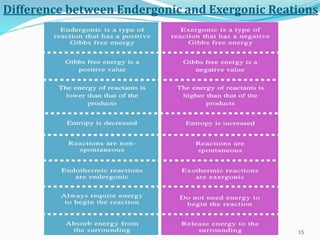
You run into this in intro biochemistry and then again when you're actually trying to figure out whether a pathway will work in a test tube. The textbook version is clean. Real lab work is not. An exergonic reaction releases free energy. Delta G is negative. The products sit at a lower energy level than the reactants and the system can do work on its surroundings as it goes. An endergonic reaction absorbs free energy. Delta G is positive. It needs an external source of energy coupled into it or it just won't proceed at a meaningful rate. The Gibbs free energy equation, G = H TS, is what actually determines the direction. People forget the entropy term and blame everything on enthalpy. That's where the confusion starts. A reaction can be endothermic and still be exergonic if the entropy increase is large enough and the temperature is high enough. I once spent two days troubleshooting why a supposedly spontaneous decarboxylation wasn't happening at room temperature. Turned out the entropy gain was real but TS wasn't big enough at 298 K to overcome the enthalpy cost. Bumped the temperature to 340 K and it ran clean.
Here's the part most guides skip: exergonic does not mean fast. A reaction can have a huge negative G and still sit there doing nothing because the activation energy barrier is massive. Diamond to graphite is exergonic. Your jewelry is fine. Endergonic reactions are everywhere in metabolism precisely because they're coupled to exergonic ones. ATP hydrolysis is the standard coupling currency, but it's not the only one. NADH, proton gradients, thioester bonds in acetyl-CoA — different systems use different energy currencies depending on the pathway and the organism. When you're designing a synthetic route, you need to pick the right coupling partner, not just assume ATP solves everything. I ran into a case last year where we were trying to drive a unfavorable phosphorylation in a cell-free system. Standard ATP coupling gave maybe 12% yield. The problem was product inhibition — the ADP built up and pushed the reaction back toward reactants. Switching to a creatine phosphate regeneration system bumped it to about 67% because the coupled reaction pulled the equilibrium forward harder and the byproduct didn't interfere. Takes five minutes to set up compared to buying ATP every time, though you do need the creatine kinase enzyme.
The common mistake beginners make is treating G as a fixed number. It's not. G depends on actual concentrations, not standard state conditions. G°' is useful for comparing reactions on paper. The real G in your flask could be completely different if you're running at millimolar concentrations instead of the standard 1 M. I've seen people design pathways based on tabulated G°' values that looked favorable on paper and then get nothing when they actually mixed the reagents. Calculate the actual G using the reaction quotient. It takes thirty seconds and saves hours of failed experiments. Another thing that trips people up: recognizing that coupling doesn't change G of the individual reactions. It changes the overall G of the combined process. The endergonic step still has a positive G. The exergonic step still has a negative G. What you're doing is making the sum negative. Enzymes don't change G either. They only lower the activation energy. If someone tells you an enzyme makes an endergonic reaction spontaneous, they don't understand what's happening. For practical purposes, if you're working with biochemical pathways, learn to spot which steps are near-equilibrium versus which are far from equilibrium. The near-equilibrium ones are reversible and respond to substrate and product concentrations. The far-from-equilibrium steps are the regulated ones — usually the ones with large negative G values. Those are your control points. In glycolysis, for example, phosphofructokinase-1 has a G well outside equilibrium and is the main regulatory step. Hexokinase and pyruvate kinase are similar. The rest of the pathway can flip direction if concentrations change enough.
Get the Full Details

When you're reading papers that report kinetic data without giving G values, don't take the rate constants as evidence of thermodynamic favorability. A fast reaction isn't necessarily exergonic and a slow one isn't necessarily endergonic. They're measuring different things. I've encountered a few papers where the authors conflated the two and drew incorrect conclusions about pathway feasibility. It happens more often than it should. For lab work, the easiest way to confirm whether a reaction is proceeding in the direction you think is to measure the actual concentrations of reactants and products at equilibrium, not just at time zero. If your product yield plateaus at 5%, that's your equilibrium position telling you something. You can either change conditions to shift it or accept that the reaction isn't viable under those conditions. Chasing a thermodynamically unfavorable reaction with more enzyme or longer time is just wasting resources.
Edge Cases Where the Simple Model Breaks Down
Multiphase systems. Membrane transport. Non-aqueous solvents. Enzyme-catalyzed reactions in crowded cellular environments. The standard G equations assume ideal dilute aqueous solution, which is never what you actually have. Activity coefficients matter. Ionic strength matters. Macromolecular crowding changes effective concentrations in ways that aren't trivial to calculate. If you're doing computational work on reaction pathways, don't trust standard G°' tables alone. Run a quick check with actual concentrations using the Nernst-like relationship for Gibbs energy. The difference between G°' and G can be 10 to 20 kJ/mol in typical cellular conditions, which is the difference between "spontaneous" and "not happening." I once modeled a synthetic biology pathway where three steps looked fine on paper using standard values. After accounting for actual intracellular concentrations and pH, two of those steps had positive G and would stall the whole thing. We redesigned the pathway with different enzymes and shifted the operating pH slightly. Worked on the third attempt. The first two attempts were pure dead ends because we hadn't calculated real G.
The takeaway is straightforward. Know your delta G. Know whether it's standard or actual. Know that kinetics and thermodynamics are separate problems. And when something isn't working, check which one you've been confusing for the other.