What You Actually Need to Do Before Titration
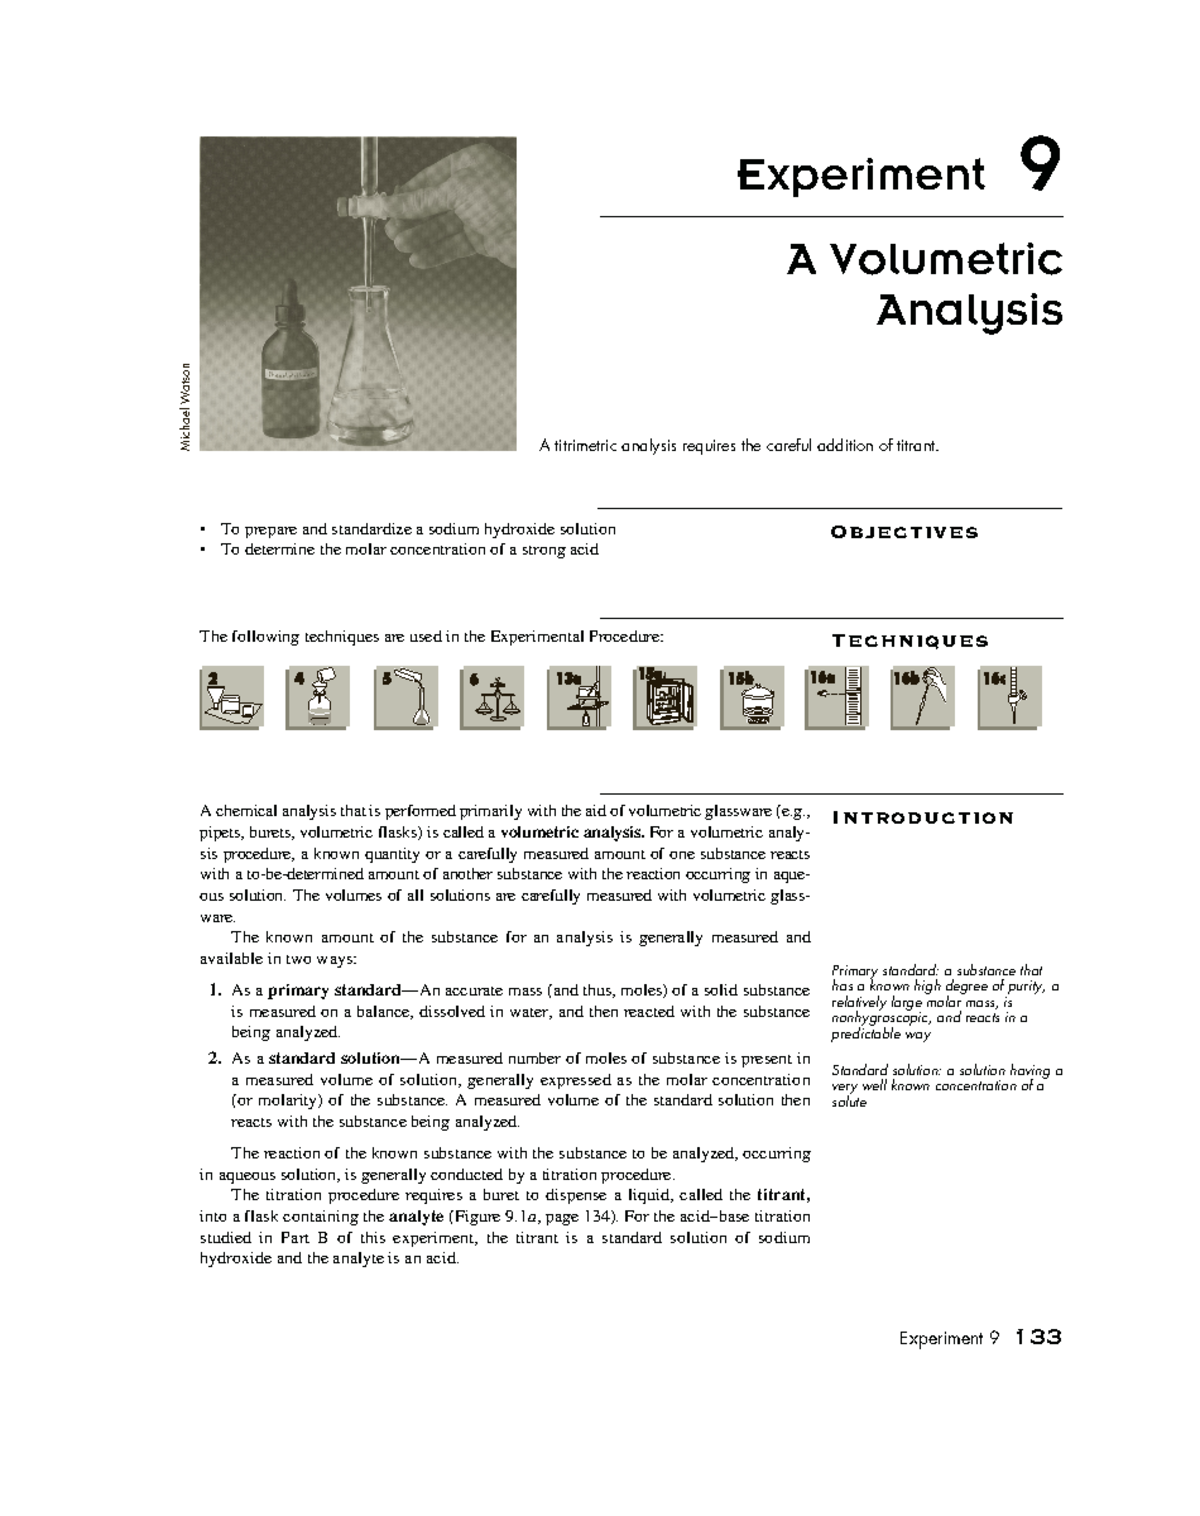
Most students treat the prelab as a paperwork exercise. They copy the procedure, plug numbers into a few equations, and submit. That approach gets you a decent grade on the worksheet and a miserable time when you actually stand at the bench. The prelab is where you figure out the problems before they cost you data points. I have run this experiment dozens of times across different sections. The one thing that consistently trips people up is not the math. It is the preparation. You need to understand what the titration is actually measuring, how much of each reagent you will need, and what could go wrong before you ever touch the burette.Experiment 9 A Volumetric Analysis Prelab
This experiment is a standard acid-base titration, usually involving an unknown concentration of an acid or base and a standardized solution. You will determine the unknown concentration by finding the equivalence point. The prelab work sets up the stoichiometric calculations, the expected volume ranges, and the procedural checks you need to run. Here is the practical sequence I go through every time, not the theoretical sequence your lab manual gives you. First, write out the balanced chemical equation. Not from memory. Look it up if you have to. Get the stoichiometric ratio right because every calculation after this depends on it. One wrong coefficient and your final concentration is off by a factor of two, and you will not catch it until the TA looks at your data sheet.
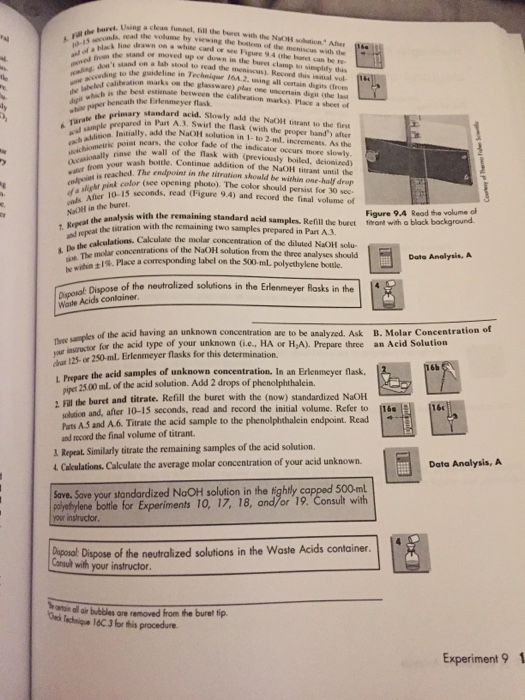
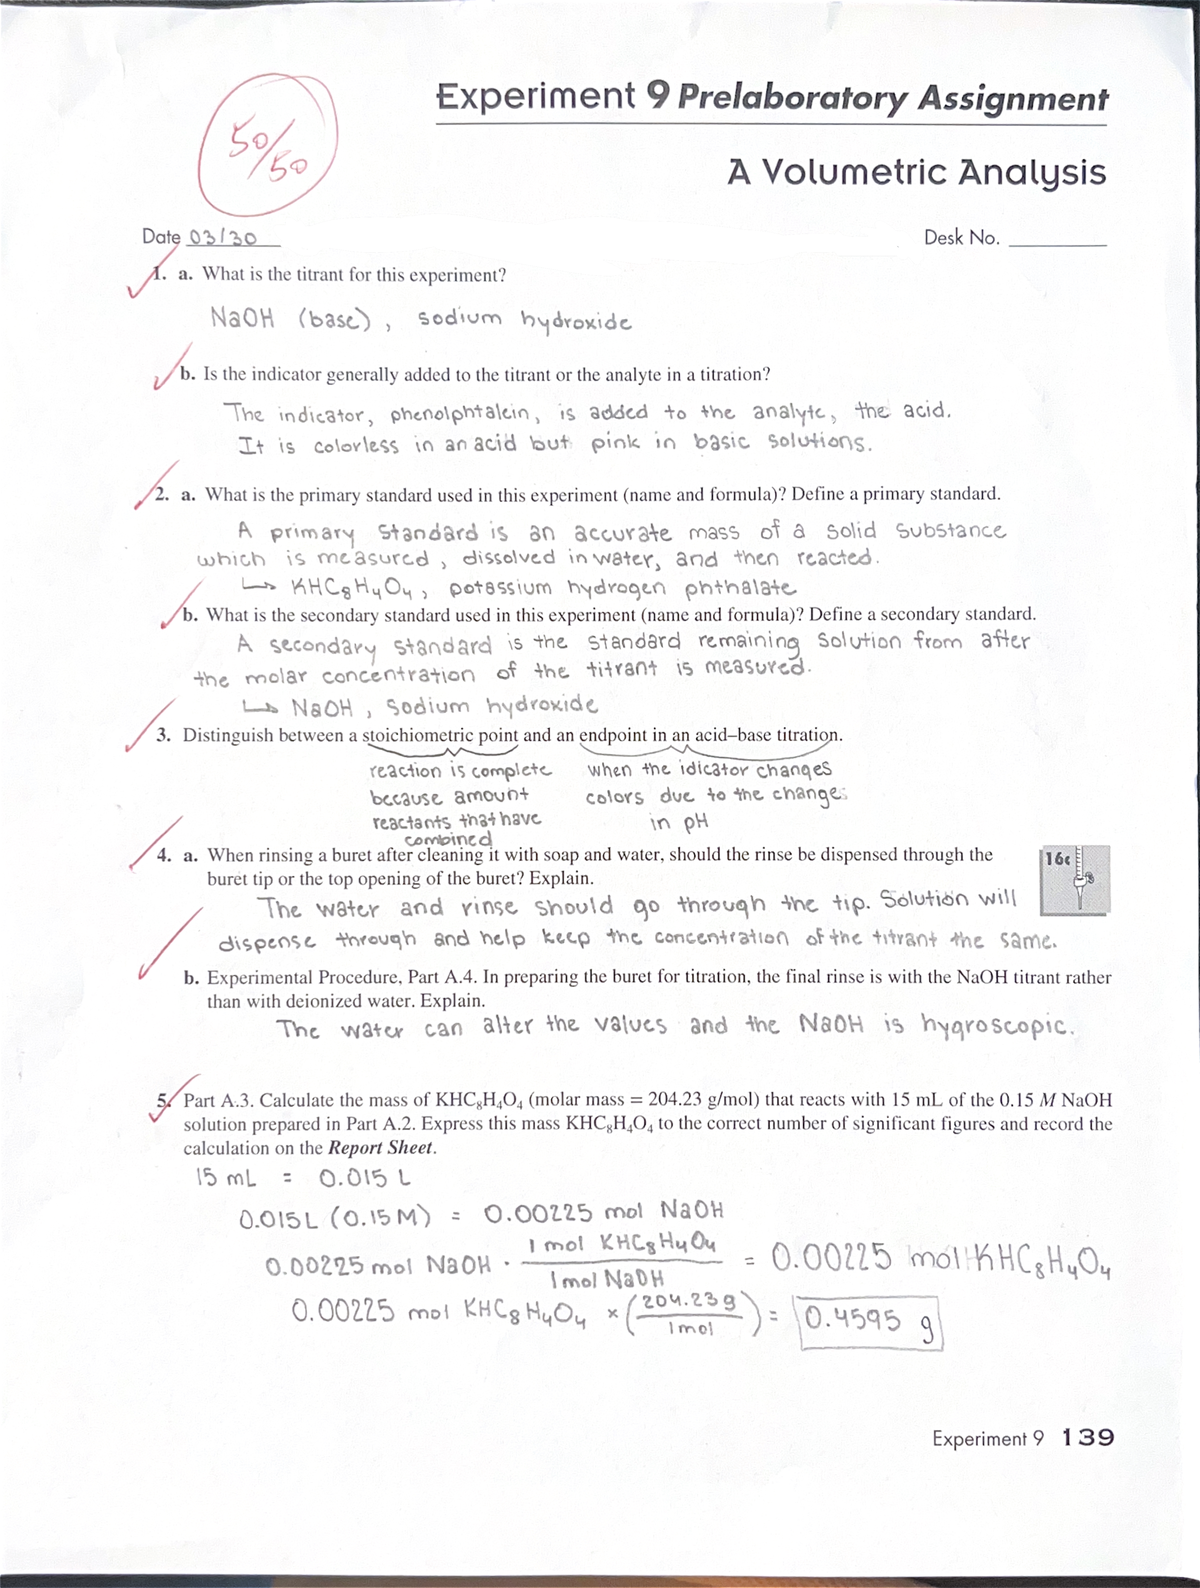
Next, calculate the expected volume of titrant. Take the approximate concentration of your unknown, assume you are pipetting a standard aliquot like 25.00 mL, and solve for the volume of titrant needed to reach the endpoint. I always calculate for both the low and high ends of the unknown concentration range if the manual gives one. This tells you how many burette fillings you will need. Running out of titrant halfway through a trial is a real problem, and it wastes about twenty minutes while you clean and refill everything. Then, determine how much primary standard you need if you are standardizing your own titrant. This is where students make the worst mistakes. If you are standardizing NaOH with KHP, you need between 0.4 and 0.6 grams per trial for a reasonable volume around 20 to 30 mL of titrant. Weighing less than 0.3 grams amplifies the balance uncertainty to the point where your standardization error balloons. Weighing more than 0.7 grams pushes your titrant volume past 35 mL and forces a refill before you finish your three trials. I learned this the hard way in my second semester when I weighed 0.21 grams of KHP and spent an hour watching my burette readings hover uselessly near the 10 mL mark while the uncertainty in my concentration estimate sat at nearly four percent. After the calculations, go through the glassware checklist. Burette, pipette, Erlenmeyer flasks, funnel, wash bottle. Inspect each piece before you start. Check the burette for chips near the stopcock. A chipped tip causes irregular droplet formation and makes endpoint detection unreliable. Rinse the burette with distilled water first, then condition it with the titrant solution. I do two 5 mL rinses through the burette, letting the liquid contact the entire inner surface including the tip. Skipping the condition rinse dilutes your titrant by a variable amount depending on how wet the glass is from the water rinse. That variable dilution is exactly the kind of error that shows up as scatter in your replicate trials.
The pipette needs the same conditioning treatment. If you are using a volumetric pipette, never deliver the last drop by blowing it out unless the pipette is explicitly marked as a blow-out type. Most volumetric pipettes are calibrated to drain freely with a small retention drop. Blowing it out adds roughly 0.02 to 0.05 mL extra volume, which compounds across your trials and shifts your calculated concentration consistently in one direction. For the indicator, phenolphthalein is standard for strong acid-strong base work. Add two to three drops, not a splash. Some students pour in a generous amount thinking more color change is better. Extra indicator introduces additional weak acid or base species into your system, which shifts the endpoint slightly. The shift is small, maybe a milliliter or two in extreme cases, but it is enough to matter when you are aiming for precision in the sub-percent range. One edge case that is worth planning for: carbon dioxide absorption. If your NaOH titrant has been sitting open for more than a few hours, it has absorbed CO2 from the air and formed carbonate. This changes the effective concentration and can shift your endpoint by making it less sharp. I keep my NaOH in a bottle with an soda lime trap and rarely worry about this, but if your stock solution looks old or the endpoint seems fuzzy, prepare a fresh batch or switch to an indicator system less sensitive to carbonate interference, like using a mixed indicator or running a blank correction.
Get the Full Details
Calculation Framework
Your prelab should include a sample calculation section. Do not just write final answers. Show the full dimensional analysis so anyone grading it can follow your logic. Start with the mass of KHP or the volume and concentration of your known solution, convert to moles using the molar mass, apply the stoichiometric ratio from your balanced equation, then divide by the volume of titrant used. Express your final concentration in mol/L with the correct number of significant figures based on your measurements. I also calculate the relative average deviation for my trials if I have preliminary data. This gives you a quick check on whether your technique is producing consistent results before you finish the full experiment. A relative average deviation above two percent in a standard acid-base titration usually signals a technique problem rather than random error. In my experience, it is most often an endpoint overshoot or inconsistent titrant delivery speed near the finish. One thing the manuals do not emphasize enough: temperature matters. Molar concentrations change with temperature because volume changes. If you are doing high-precision work and the lab is significantly warmer or cooler than the calibration temperature of your glassware, typically 20°C, you should note the ambient temperature and apply a correction if your institution requires it. For routine undergraduate labs, this is rarely a hard requirement, but it is worth knowing that your 25.00 mL pipette delivers slightly more volume at 25°C than at 20°C, and the difference is real even if it is small.
Common Problems and What to Do About Them
Endpoint overshoot is the most frequent issue. The color jumps from pale pink to deep magenta in a fraction of a second, and you have already added too much titrant. When this happens, do not discard the trial and start over immediately. Record the volume you added, note the overshoot, and consider back-titrating with your unknown solution if the procedure allows it. This saves you the cleaning and setup time. If your lab manual does not permit back-titration, just accept the trial as a learning point and move on. Your TA will notice the notation and usually accounts for it. Another frequent problem is air bubbles in the burette tip. A bubble that escapes during titration registers as liquid volume delivered but is actually empty space. Your recorded volume will be higher than the true volume, and your calculated concentration will be wrong. Check the tip before you start, during the initial zeroing, and occasionally during the run. Tap the burette gently or open the stopcock fully for a second to dislodge any trapped air. I check this at least three times per trial, and I have caught enough bubbles this way to prevent several bad data sets over the years. Meniscus reading error is a quiet source of inconsistency. Different students read the bottom of the meniscus at slightly different angles, introducing parallax error. Always position your eye level with the meniscus and use a white card behind the burette to make the curve clear. This simple habit reduces reading variance noticeably between trials.
The procedure works well for strong acid-strong base systems. It breaks down for very dilute solutions below about 0.01 M because the pH change at the equivalence point becomes gradual rather than sharp, making endpoint detection unreliable. It also struggles with weak acid-weak base combinations where the equivalence point pH is near neutral and no common indicator gives a clear color change. If you encounter one of these scenarios, you need a potentiometric endpoint determination instead of an indicator-based method. No amount of prelab preparation will fix a fundamentally unsuitable method. This prelab format is straightforward if you treat it as a planning document rather than a compliance task. Work through the calculations, check the glassware, anticipate the failures, and you will spend less time troubleshooting at the bench and more time getting clean data.