Electron behavior isn't something you memorize. It's something you learn to predict by watching where the electrons actually go.
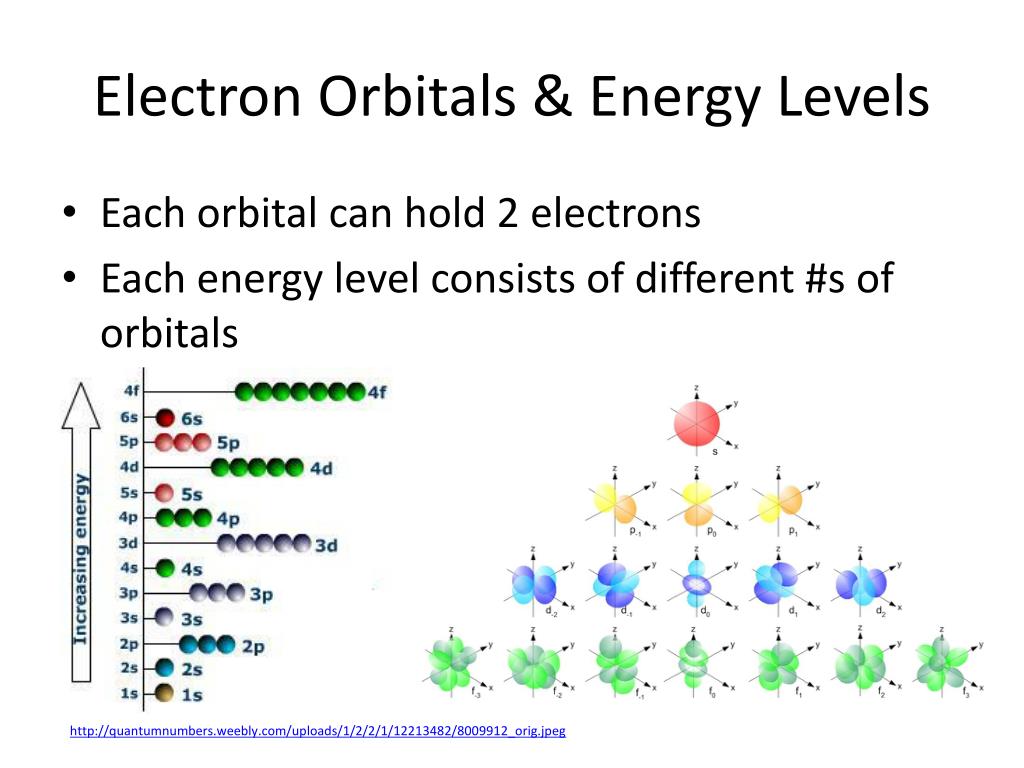
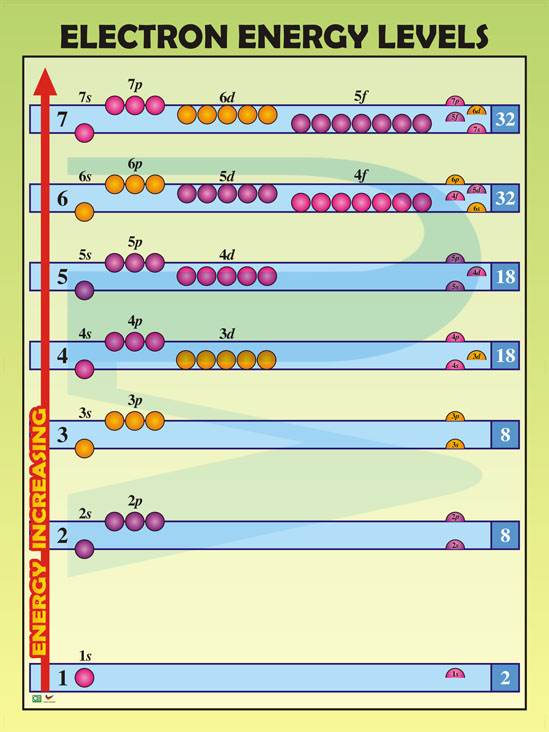
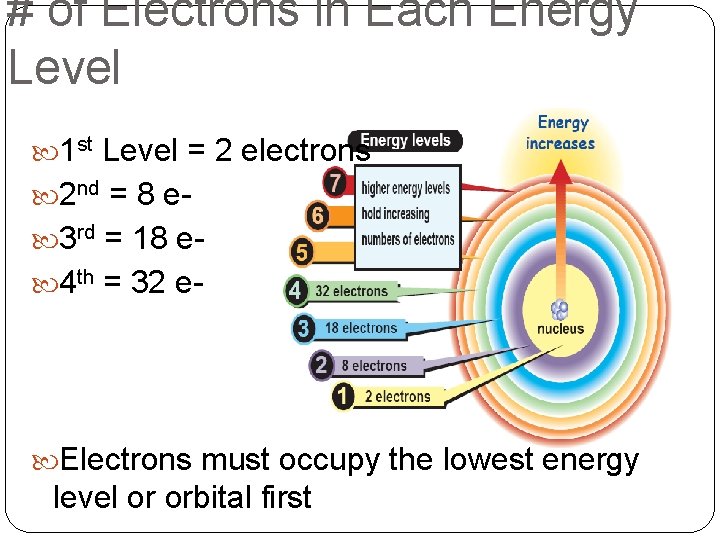
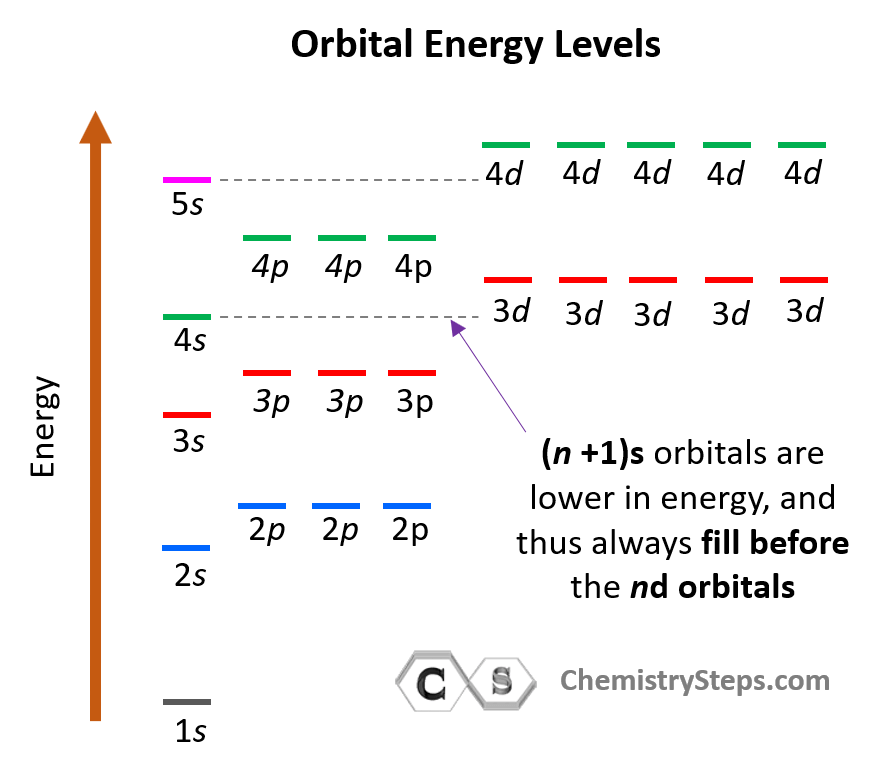
When I first started working with quantum chemistry software and spectroscopy data, I made the same mistake everyone makes. I treated energy levels like steps on a ladder. Each rung holds a fixed number of electrons, they go up, they come down, end of story. That framework gets you through introductory chemistry. It breaks down the moment you actually have to model something or interpret a real spectrum. Energy levels are really just solutions to the Schrödinger equation for a given system. The quantum numbers (n, l, ml, ms) label those solutions. The principal quantum number n determines the general size and energy of the orbital. The angular momentum quantum number l determines the shape. But here is what the textbooks gloss over: the energy of an orbital depends heavily on the electron configuration already present in the atom. This is why the 4s orbital fills before the 3d in potassium and calcium, but once the 3d starts filling, the energy ordering flips and 4s electrons ionize first. Every time I tell someone this, they look genuinely confused, because it contradicts the simple Aufbau diagram they learned. The diagram is an approximation for neutral atoms in their ground state. It is not a universal law.
Explain How Energy Levels Relate To Electron Behavior
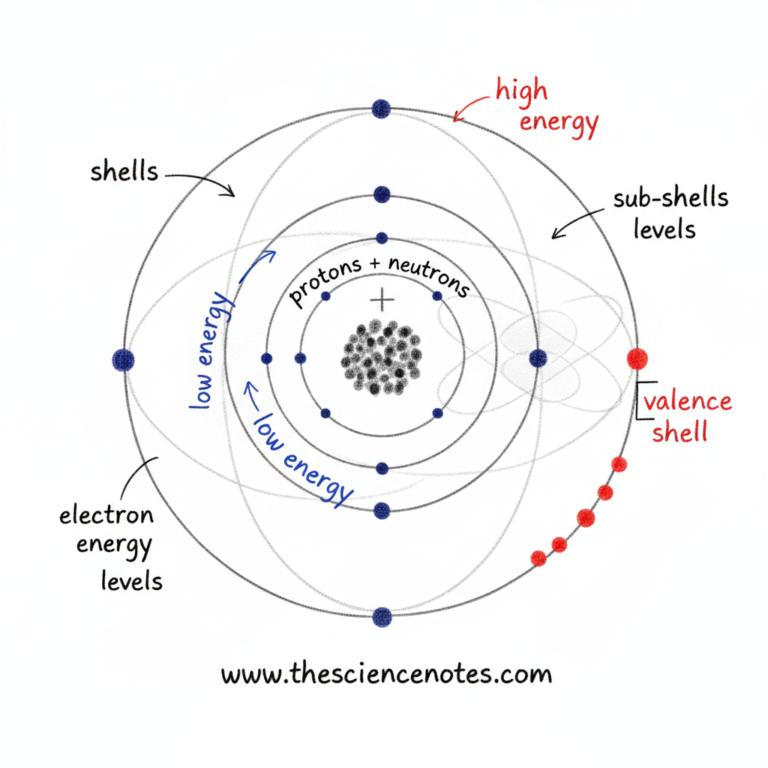
The core relationship is straightforward. Electrons occupy the lowest available energy states first, that's the Pauli exclusion principle in action. Each orbital holds two electrons with opposite spins. When an electron absorbs energy, it moves to a higher energy level. When it drops back down, it emits a photon with energy equal to the difference between the two levels. This is why we get line spectra instead of continuous ones. Each element has a unique fingerprint because the spacing between energy levels is unique to that element's nuclear charge and electron count. But the practical reality is messier. Multi-electron atoms introduce electron-electron repulsion that shifts energy levels significantly. You can't solve the Schrödinger equation exactly for anything with more than one electron. We use approximations like Hartree-Fock or density functional theory, and even those have known failure modes. I spent about three weeks last year debugging a computational chemistry project where the predicted excitation energies from a standard DFT functional were off by nearly two electron volts for a transition metal complex. The problem wasn't the code. The problem was that the functional couldn't handle the near-degenerate d-orbital states properly. Switching to a range-separated hybrid functional fixed it, but it doubled the computation time. That's a tradeoff you need to be aware of before you start. There is also the issue of excited states. A ground-state calculation tells you about the energy level structure, but it doesn't tell you how electrons behave when they are perturbed. Time-dependent DFT or configuration interaction methods are needed for that. These methods are computationally expensive and still approximate. If you're working with larger systems, say molecules beyond about 100 atoms, you're often forced to use semi-empirical methods that sacrifice accuracy for speed. The results can be qualitatively reasonable but quantitatively unreliable, especially for properties that depend on subtle energy differences like reaction barriers or charge transfer excitations.
One thing that catches people off guard is the role of spin. Hund's rule says that electrons fill degenerate orbitals singly before pairing up. This isn't just a rule. It's a consequence of exchange energy. Parallel spins reduce electron-electron repulsion through the Pauli principle, which lowers the overall energy. In practice, this means that the total spin state of a system matters enormously for its magnetic and optical properties. Transition metal complexes with unpaired electrons behave very differently from closed-shell systems, and standard calculation protocols sometimes miss this if you don't specify the right multiplicity at the start. Another counter-intuitive point is that energy levels aren't static. They shift with the chemical environment. Solvent effects, nearby charged groups, and even crystal packing can change the energy gap between orbitals by fractions of an electron volt, which translates to noticeable shifts in absorption and emission wavelengths. When I was analyzing UV-Vis spectra for a series of organic dyes, the calculated gas-phase energy gaps didn't match the experimental solution-phase data at all. Adding a polarizable continuum model to account for solvent effects brought the calculations into agreement, but only after I calibrated the model against a few known reference compounds. You can't just slap on a solvent model and expect accurate results without some empirical grounding. The degeneracy of energy levels is another area where intuition fails. In hydrogen, all orbitals with the same n are degenerate regardless of l. In multi-electron atoms, this degeneracy is lifted by shielding and penetration effects. An s orbital penetrates closer to the nucleus than a p orbital of the same n, so it experiences less shielding and has lower energy. This is why the energy ordering goes s, then p, then d, then f within each shell. But even this ordering has exceptions. Chromium and copper, for example, have ground state configurations that deviate from the expected pattern because half-filled and fully-filled d subshells gain extra stability from exchange energy and symmetry.
Get the Full Details
When you move to molecules, energy levels become molecular orbitals formed from linear combinations of atomic orbitals. The number of molecular orbitals equals the number of atomic orbitals you combine. Bonding orbitals are lower in energy than the constituent atomic orbitals. Antibonding orbitals are higher. Nonbonding orbitals stay roughly at the same energy. The gap between the highest occupied molecular orbital and the lowest unoccupied molecular orbital, the HOMO-LUMO gap, correlates roughly with optical absorption energy, but it's not a direct measurement. TDDFT and other excited state methods give better estimates, but they come with their own assumptions and limitations. If you're trying to work with this practically, start by understanding what approximation you're using and what it's good for. Don't assume a ground-state DFT calculation gives you accurate excitation energies. Don't assume the Aufbau principle tells you the actual ionization order for a transition metal. Don't assume gas-phase calculations match solution-phase experiments. And when your results don't match expectations, check the multiplicity, the convergence criteria, and the basis set size before you start questioning your interpretation of the physics. The biggest bottleneck I've encountered is the balance between accuracy and computational cost. For small systems, high-level wavefunction methods like CCSD(T) can give excellent results, but they scale as N7 with system size. That means a molecule twice as large takes roughly 128 times longer to compute. For medium-sized systems, DFT is the workhorse, but you need to choose the right functional. B3LYP is popular but notoriously unreliable for dispersion interactions and reaction barriers. WBF2 or M06-2X might be better depending on what you're studying, but they're more expensive. There's no universal best choice, and the literature is full of case studies where a functional that works great for one class of molecules fails miserably for another.
Spectroscopic interpretation is another area where the theory-practice gap shows up clearly. The energy level diagram you see in a textbook assumes isolated atoms or simple molecules. Real spectra show broadening from vibrational coupling, rotational structure, solvent interactions, and instrumental resolution. A sharp theoretical transition energy becomes a band. Assigning peaks in a real spectrum requires simulating the full vibrational structure, not just the electronic transition. Gaussian and ORCA can do this, but the simulations take time and careful setup. Missing a single imaginary frequency in a transition state optimization can invalidate an entire vibrational analysis. For anyone actually working in this space rather than just studying it, the lesson is practical. Learn to read the output files carefully. Check the SCF convergence. Verify that your geometry is a true minimum with no imaginary frequencies unless you're looking at a transition state. Run a basis set sensitivity test. Compare multiple functionals if the property you care about is sensitive to electronic structure details. And when something goes wrong, which it will, systematic debugging beats guessing every time.