Getting the Layers to Actually Separate
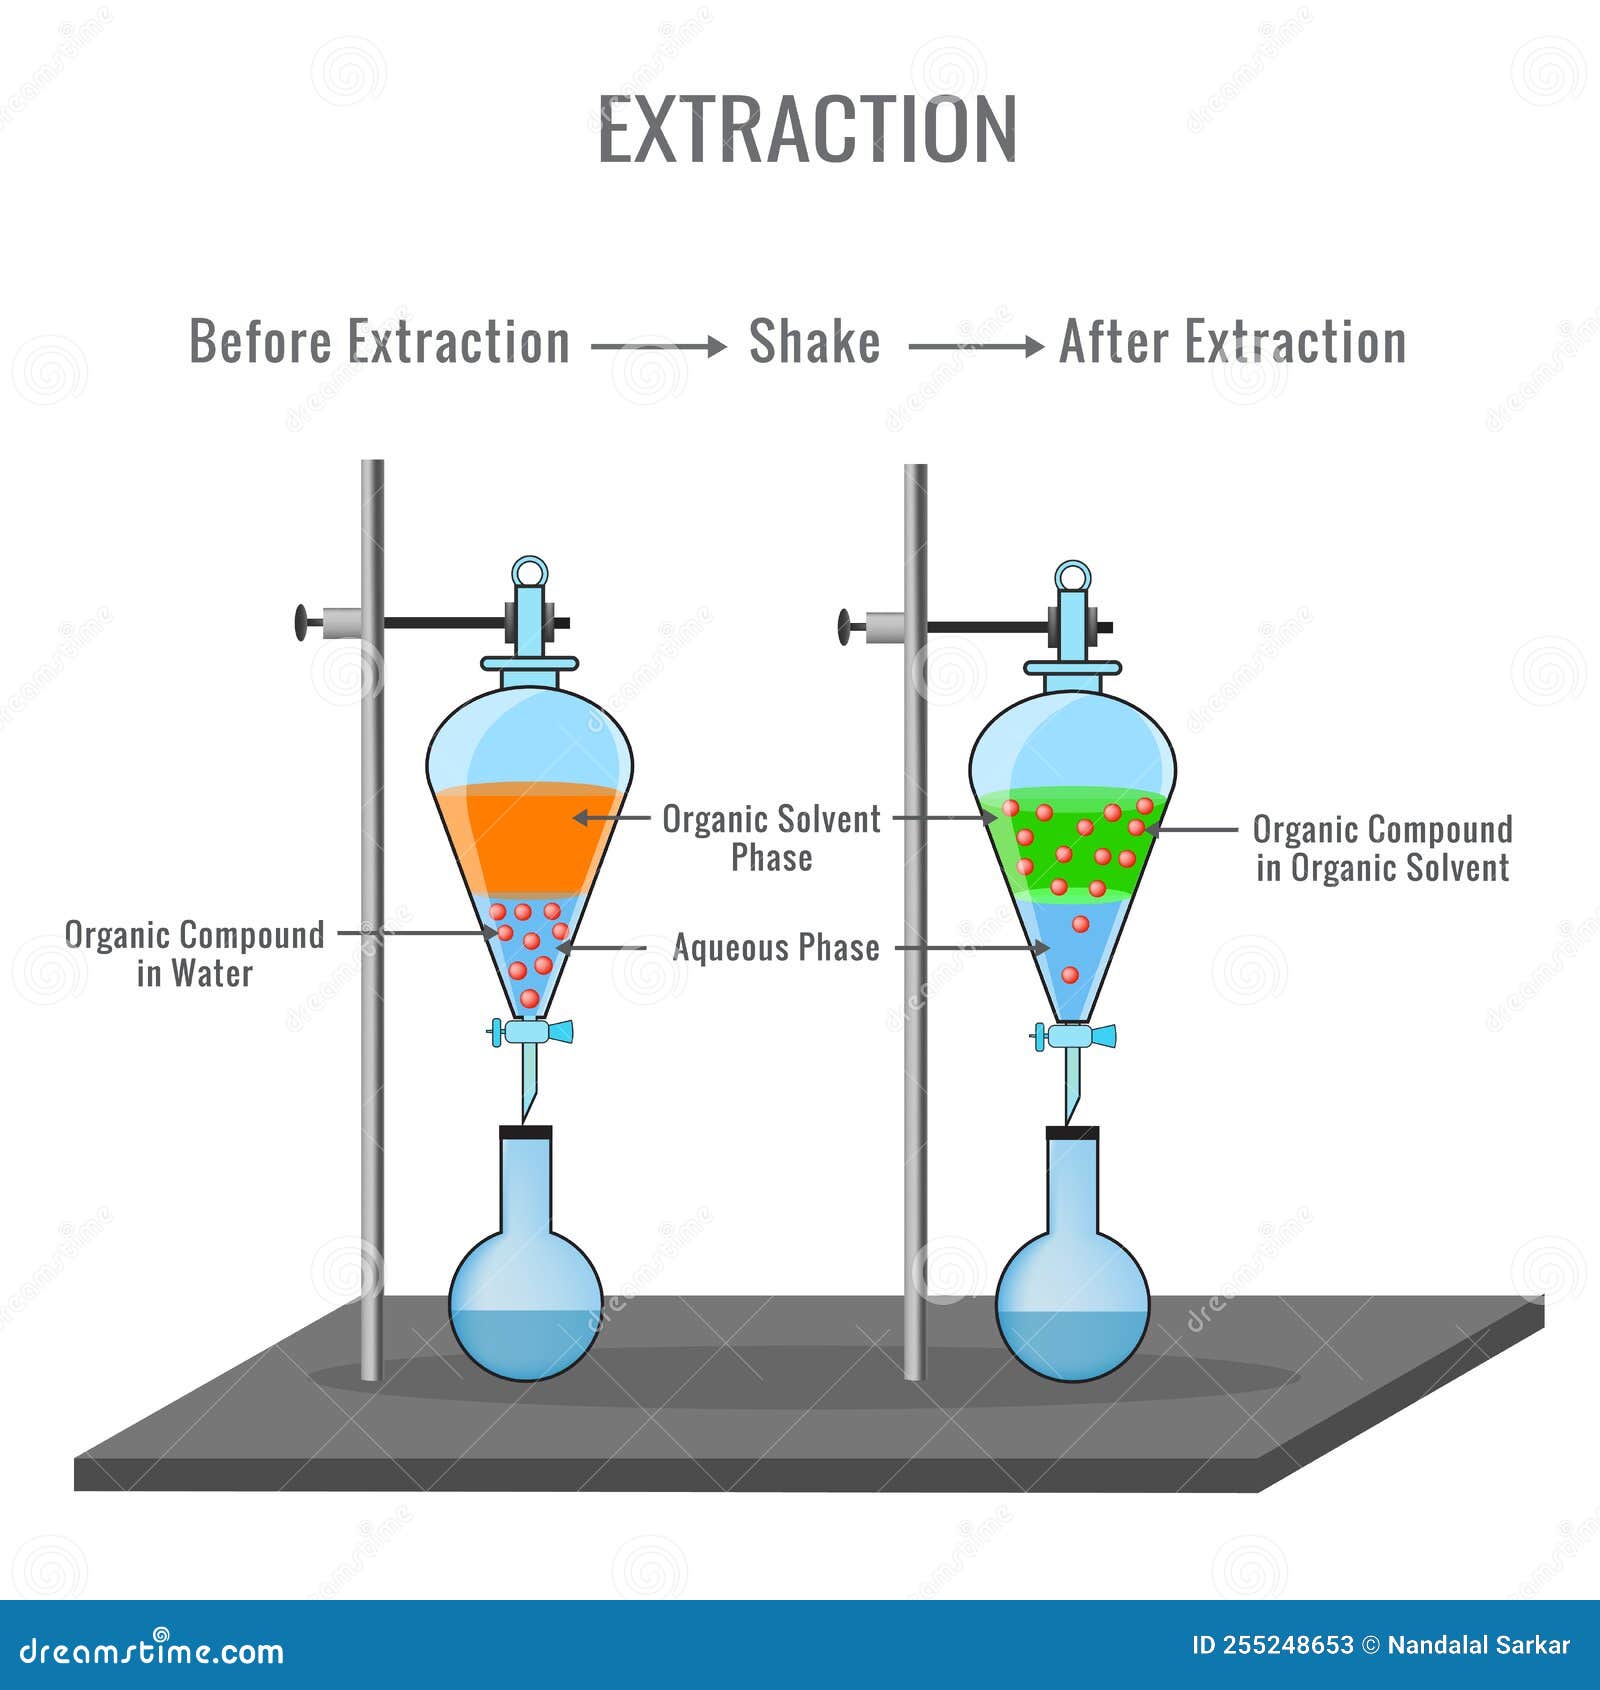
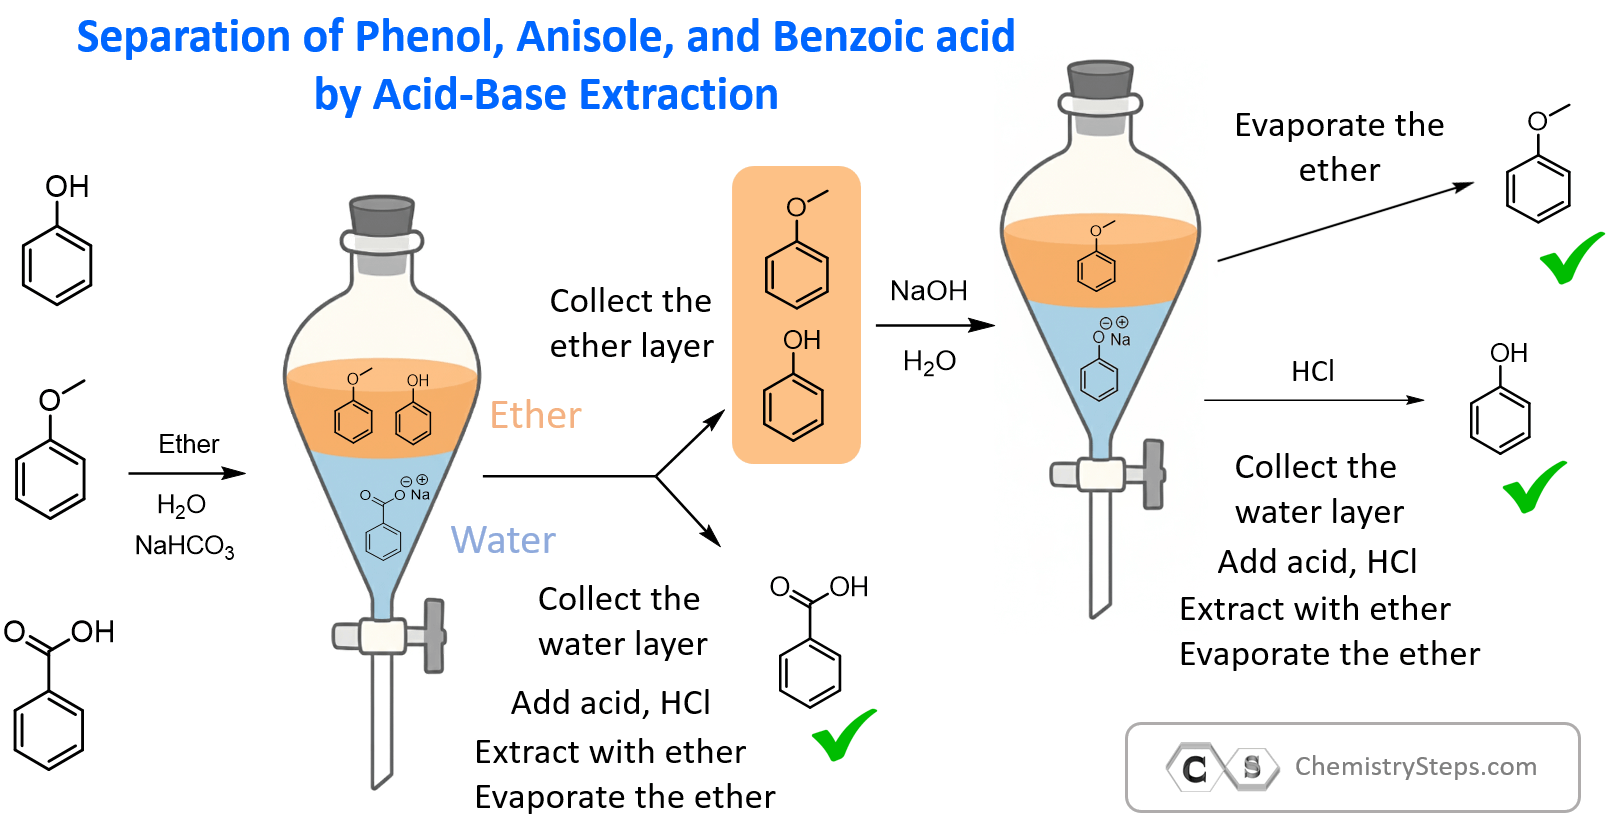
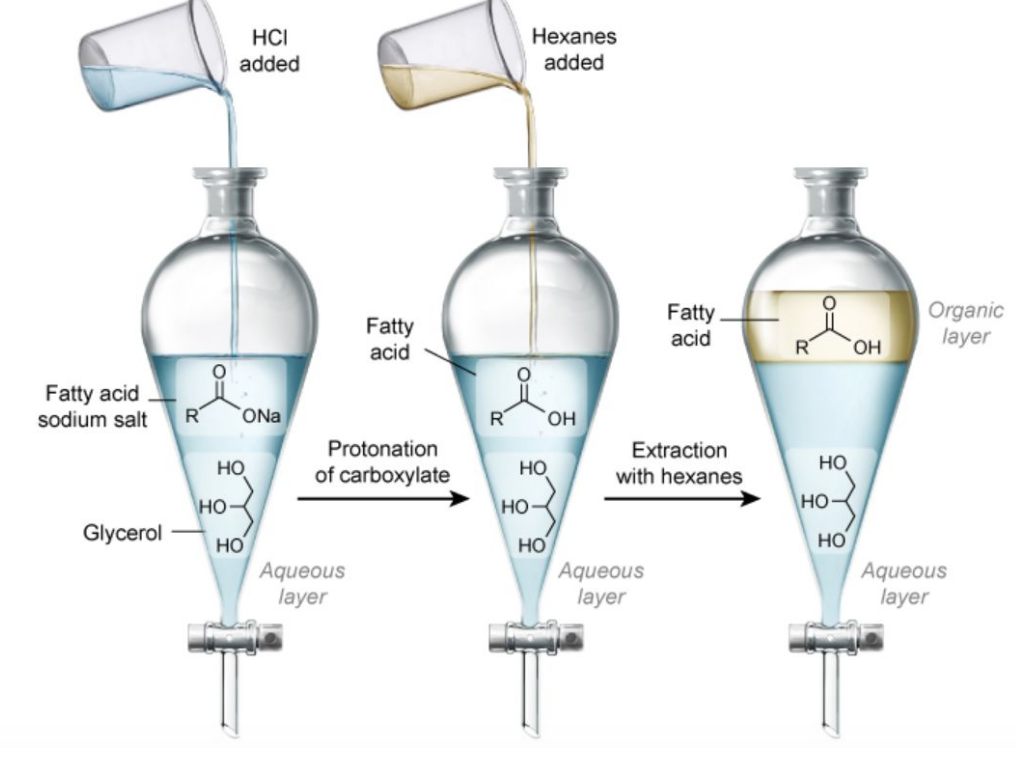
Most people learn liquid-liquid extraction in undergrad and then never do it properly again until something goes wrong at 11pm on a Thursday. The technique itself is mechanically simple, but the details are where you lose product or waste three hours chasing an emulsion. Let me walk through how I actually run these things in the lab now, not how the textbook describes them. At its core, you're using two immiscible solvents to move a target compound from one phase into the other based on its solubility preference. That preference is quantified by the partition coefficient, Kd, which is just the ratio of concentrations at equilibrium. The extraction efficiency depends on that coefficient, the volume ratio of your solvents, and how many discrete extraction steps you perform. Three extractions with a small volume each are almost always better than one giant extraction with the full volume. That's basic math, but people still pour everything into one funnel and wonder why their yield is mediocre. I ran into a particularly annoying case last year involving a moderately polar product dissolved in ethyl acetate that I needed to wash free of some acidic byproduct. Standard procedure would be a bicarbonate wash. The problem was that my reaction mixture also contained a significant amount of unreacted starting material that had similar solubility characteristics in both layers. After the first wash, I checked the aqueous layer by TLC and there it was, co-extracting right along with my product because the pH wasn't selective enough. The workaround was straightforward: I acidified the aqueous layer back down with dilute HCl, re-extracted with fresh ethyl acetate, and then carefully basified a separate aliquot only after confirming via pH paper where my product actually partitioned. It added maybe twenty minutes to the workup but saved me from running a column on contaminated material.
Choosing the Right Solvent Pair
The classic water and organic solvent combo works for most things, but the choice of organic solvent matters more than students realize. Ethyl acetate and water are a common pair because they're relatively non-toxic and form clean separations. Dichloromethane and water give faster separation since DCM is denser than water, so it settles to the bottom. That seems convenient until you're trying to collect the bottom layer and you accidentally pull solution from the flask instead of just the DCM layer because you misread which side was which. I've made that mistake twice. Now I always label the layers before I open the stopcock. For very polar compounds that won't stay in an organic layer, you can switch to a water-saturated butanol system or even try a biphasic PEG-water system if you're dealing with something truly stubborn. These alternative systems are slower and require more careful handling, but they exist for a reason. Not every extraction has to follow the standard textbook recipe.
Practical Details That Make the Difference
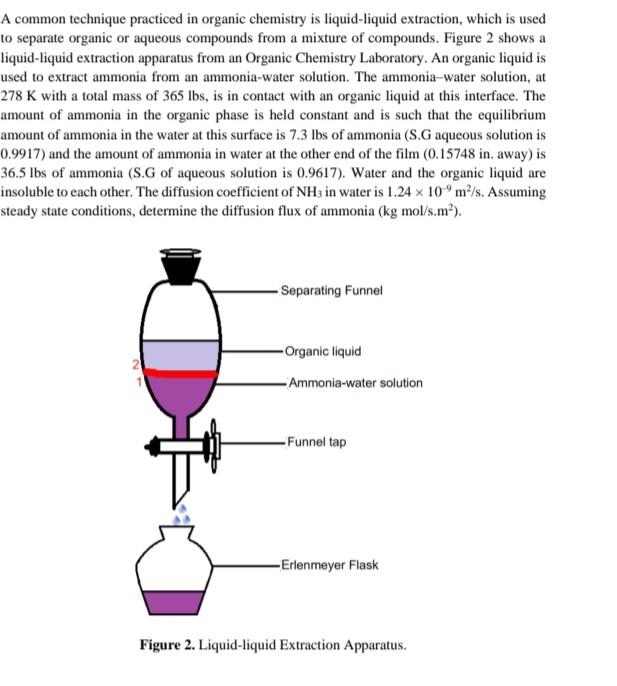
Here's what actually happens when you're at the fume hood with a separatory funnel in your hand. You load your mixture, add the second solvent, close the stopper, and invert. Then you immediately vent by opening the stopcock while keeping the funnel tilted upward. If you don't vent promptly, pressure builds up and something explodes out of the top of the funnel. This isn't theoretical. I watched a grad student get a face full of ether because he forgot to vent during a diethyl ether extraction of a volatile product. He was fine physically, but he lost the contents of his funnel and had to start over from scratch. After shaking, you let the layers separate. This takes longer than you think, especially if there are surfactant-like impurities present or if you're working with viscous reaction mixtures. I typically wait at least five minutes between shakes and check for a clear interface before proceeding. Rushing this step is how you end up with cloudy layers and no idea where your product went. When draining, open the stopcock slowly and watch the interface closely. Stop draining the moment you see the boundary approaching the stopcock. You want to leave a small amount of the bottom layer behind rather than accidentally pulling over some of the top layer. Transfer each layer to a separate, clearly labeled flask. Never assume you know which is which based on memory alone.
Get the Full Details
Dealing with Emulsions
Emulsions are the bane of extraction work. They happen when you have fine droplets of one solvent suspended in the other, creating a cloudy, stable mixture that refuses to separate. They're more likely when you're working with concentrated solutions, when there are solid particulates present, or when your compound has amphiphilic character. I've seen emulsions persist for hours in flasks that were left overnight on the bench. The simplest approach is usually just to let it sit longer. Sometimes a gentle swirl or a brief sonication can help coalesce the droplets. If that doesn't work, adding a small amount of saturated brine can break the emulsion by increasing the ionic strength of the aqueous phase and reducing the solubility of the organic solvent in water. A drop or two of a surfactant-breaking agent like hexane or isopropanol can also help in some cases. For really stubborn emulsions, I'll filter the mixture through a small plug of Celite, which physically traps the interfacial material and lets the layers separate cleanly on the other side.
Drying the Organic Layer
Once you've isolated your organic layer, you need to remove residual water before you concentrate. Anhydrous sodium sulfate is the workhorse drying agent for most extractions. It's cheap, works fast, and doesn't adsorb much product. Magnesium sulfate is faster but more expensive. I use magnesium sulfate when I'm in a hurry and sodium sulfate when I'm processing multiple samples and cost matters. The trick with drying agents is knowing when you've added enough. Add a spatula tip, swirl, and watch. If the solid flows freely like sand, you need more. If it clumps together, you've reached the saturation point. I typically add slightly more than I think I need and let it sit for ten minutes. Then I filter or decant the dried solution before concentrating under reduced pressure. Skipping this step or doing it poorly means you'll end up with wet product that refuses to crystallize or gives you messy NMR spectra with water peaks everywhere.
When Extraction Fails You
No single extraction technique handles every situation well. If your product is extremely hydrophilic, repeated extractions may still leave significant amounts in the aqueous layer. In those cases, ion pairing reagents can help move the compound into the organic phase, but they introduce new purification challenges. If your product is thermally sensitive, concentrating under high vacuum at elevated temperature can degrade it. I've lost material this way by not paying attention to the bath temperature during rotary evaporation. Keeping the water bath below thirty degrees Celsius and using a proper vacuum pump with a cold trap has prevented most of those losses going forward. For compounds that are difficult to extract cleanly, chromatography is often the more practical route despite the extra time and solvent consumption. Extraction excels at crude purification and bulk separation, not at achieving analytical purity. Knowing the boundary between those two regimes is part of what makes this technique useful rather than frustrating. The real skill in Extraction Technique In Organic Chemistry comes from recognizing when your particular system is behaving unexpectedly and having the patience to troubleshoot it step by step rather than forcing it through a standard procedure that wasn't designed for it. Most problems have simple solutions if you take the time to understand what's actually happening in the funnel.