Working with First Order Reactions in the Lab
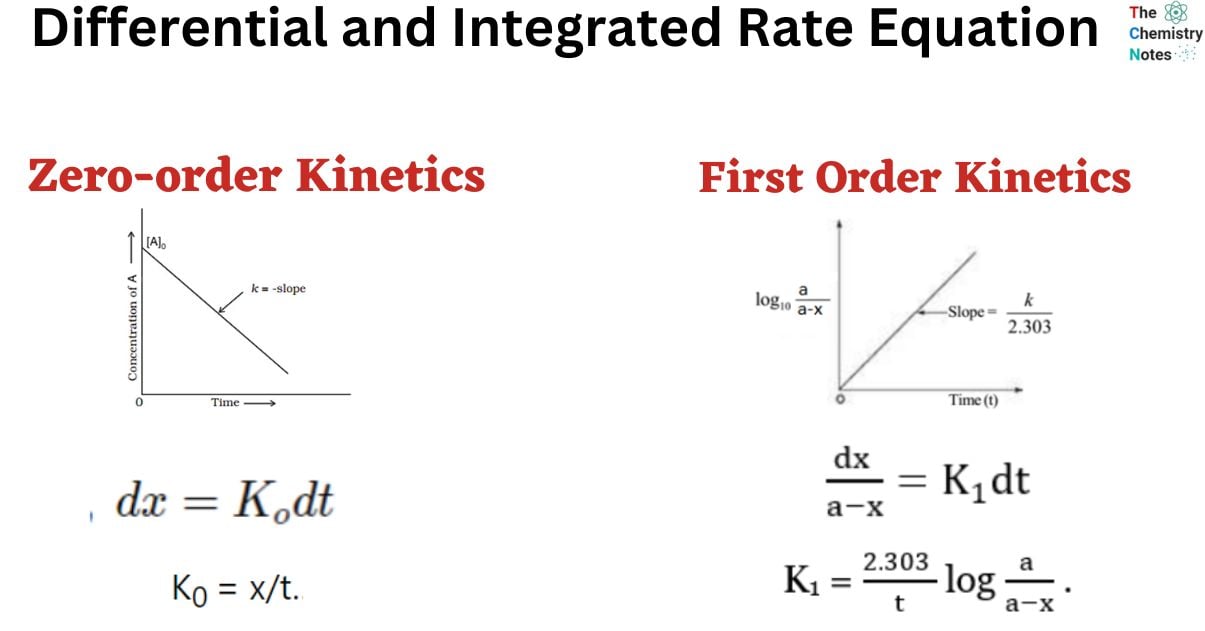
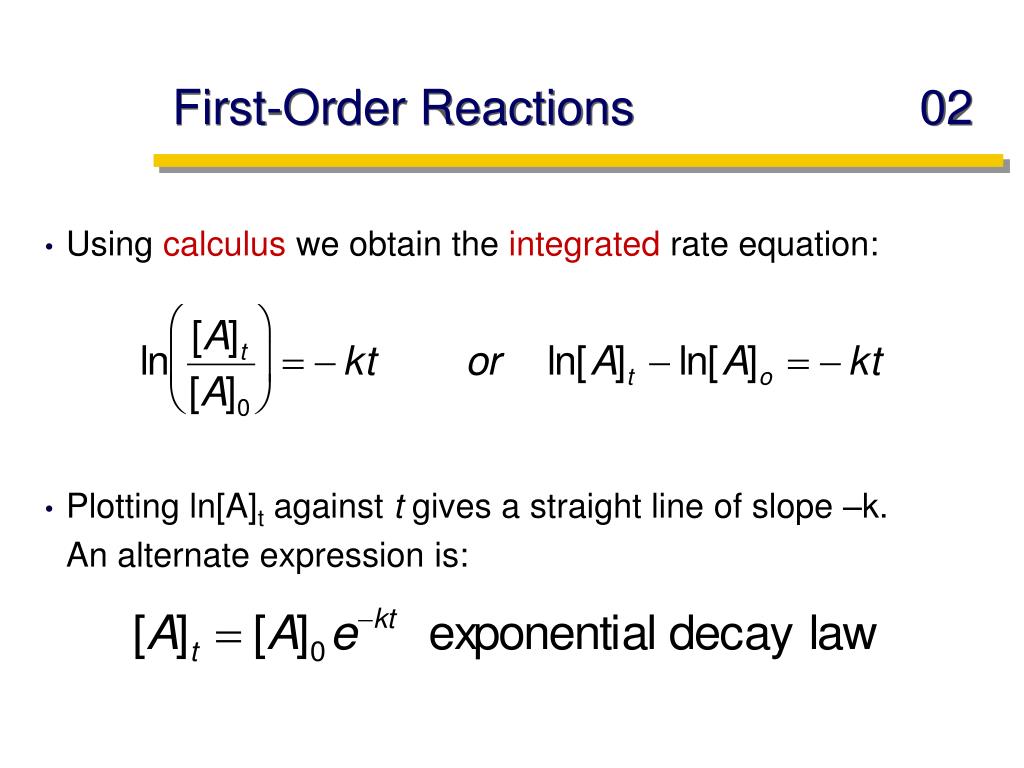
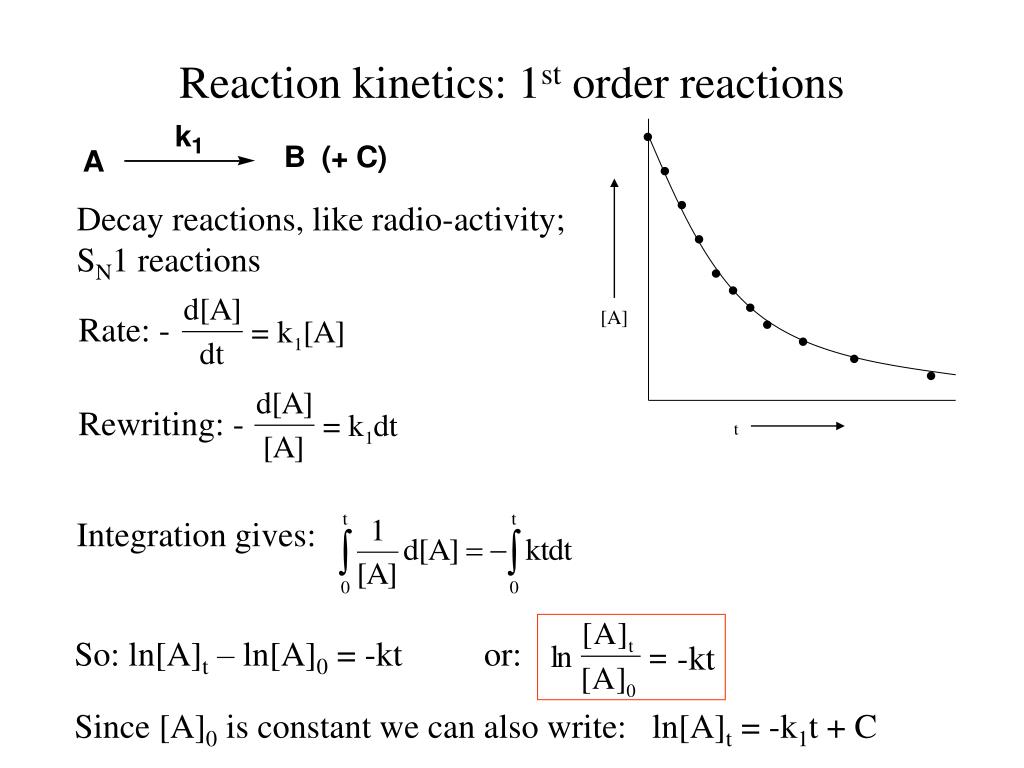
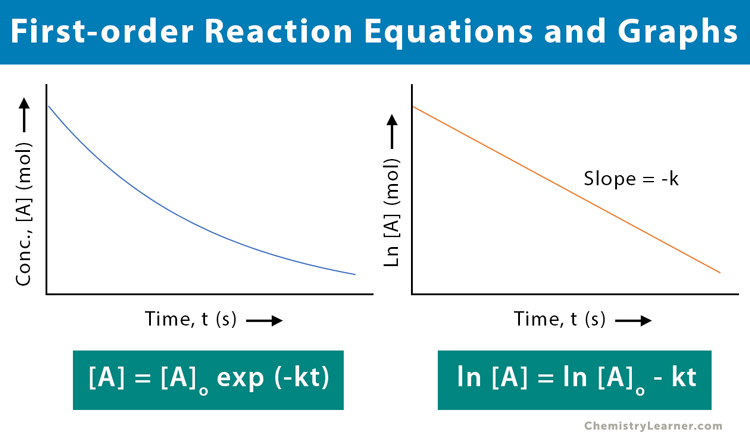
The First Order Kinetics Equation is one of those things you learn in your second semester of physical chemistry and then forget until a real problem forces you to remember it. It shows up everywhere once you actually need it: drug half-lives in pharmacokinetics, atmospheric decay of pollutants, radioactive contamination cleanup, even aging of organic materials in conservation labs. The equation itself is simple, which is probably why people assume they understand it. They don't. Here's what it looks like when you're actually using it: ln([A]t / [A]0) = -kt
Or the more common form most textbooks give you: [A]t = [A]0 * e^(-kt) Where [A]t is the concentration at time t, [A]0 is the initial concentration, k is the rate constant, and t is time. If you're working with half-lives instead of raw concentrations, you can use t_1/2 = ln(2) / k. That's the whole toolbox for first order systems.
First Order Kinetics Equation — What You Actually Need to Know
I spent about three days wrestling with a degradation study last year where the numbers refused to behave. We were tracking how a certain active pharmaceutical ingredient broke down under elevated temperature conditions. The plot of ln(concentration) versus time should have been a straight line if the reaction was truly first order. It wasn't. It curved upward slightly at the tail end. The first thing you'd be told to do is check your instrument calibration. So we did that. HPLC was fine. Samples were fine. Then I noticed something nobody had mentioned in any of the guidance documents: the impurity that formed near the end of the degradation profile was absorbing at the same wavelength we were monitoring for the parent compound. We thought we were measuring leftover API. We were actually measuring API plus a degradation product. The "upward curve" was artificial concentration reading caused by spectral overlap. The workaround was switching to a different detector wavelength where the impurity had negligible absorbance. Once we did that, the plot went perfectly linear and the calculated half-life matched the literature value within five percent. If you're seeing non-linearity in a first order plot, don't just keep recalculating. Check whether your analytical method is actually measuring what you think it's measuring.
Get the Full Details
Another thing that trips people up: the rate constant k has units of inverse time. Not inverse concentration. Inverse time. That's the single biggest difference between first order and second order kinetics, and it's the reason your dimensional analysis keeps failing when you try to force a second order model onto first order data. The units don't lie. If k doesn't come out in 1/time, something is wrong with your model or your data. There's also a common misconception about half-life. For first order reactions, the half-life is constant. It does not depend on the starting concentration. This feels unintuitive because in everyday life, if you have twice as much of something, it takes longer to use it up. But in first order kinetics, the rate is always proportional to however much is currently there. So if you start with twice the concentration, you also have twice the rate. They cancel out. The half-life stays the same whether you begin at 0.1 M or 10 M. This property is why first order kinetics is so useful in pharmacology. A drug with first order elimination will always take the same amount of time to reduce its concentration by half, regardless of dose. That's not true for zero order or second order systems.
When I'm doing these calculations manually now, I skip the natural log transformation and just use an online solver or a quick Python script. The math hasn't changed but the time savings are real. What used to take me twenty minutes of manual plotting and regression takes about ninety seconds with a script that also gives me confidence intervals on the rate constant. If you're doing this work regularly, automate it early. There are cases where first order kinetics simply doesn't apply and you need to accept that. Enzyme-catalyzed reactions at high substrate concentrations shift toward zero order because the enzyme is saturated. Reactions between two molecules where the rate depends on both concentrations are second order. Some degradation pathways show autocatalytic behavior where the product accelerates its own formation, giving you sigmoidal curves that no amount of logarithmic transformation will fix. If your ln(C) vs. time plot has an R-squared of 0.94 or lower, you probably shouldn't be calling it first order. That's not a borderline case. That's a signal that the mechanism is more complex than a simple unimolecular decay. I've seen people report first order rate constants from data that clearly wasn't first order because they needed a number for a regulatory submission. Don't do that. It comes back to haunt you during audit.
The integrated rate law approach also breaks down if you're dealing with reversible reactions. Once the reverse reaction becomes significant, the simple exponential decay equation no longer describes the system. You need to solve the full differential equation with both forward and reverse rate constants. In practice, this usually means fitting to a different model or working in a regime where the reverse reaction is negligible. The latter is often possible in the early to middle stages of a reaction before equilibrium approaches become relevant. Temperature dependence is another layer you can't ignore. The Arrhenius equation ties the rate constant to temperature: k = A * e^(-Ea/RT). If you're determining a half-life at one temperature and need to predict it at another, you need the activation energy. Without it, you're guessing. Accelerated stability studies are built on this principle, and they're only as reliable as the activation energy you plug in. I've seen activation energies pulled from similar compounds in the literature used as placeholders. That works okay if the chemistry is close. It fails hard if the degradation mechanism changes with temperature, which happens more often than you'd expect. Bottom line: the equation is straightforward. The application is where things get messy. Make sure your data actually fits the model before you trust the numbers it produces.