Working with First-Order Kinetics in the Lab
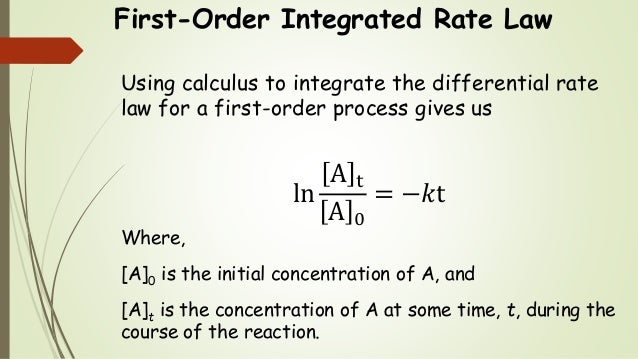
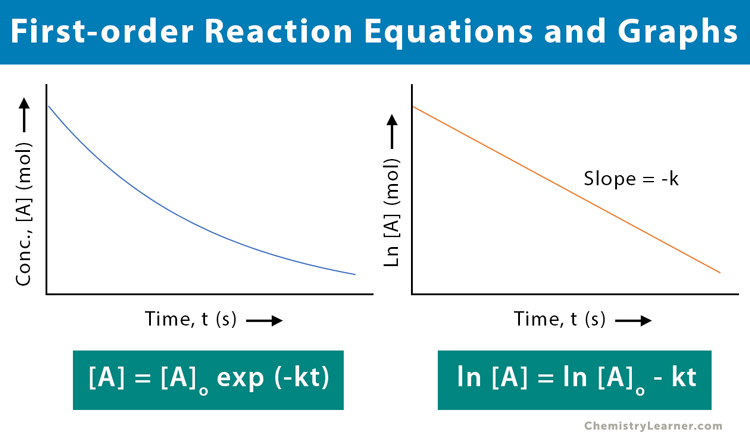
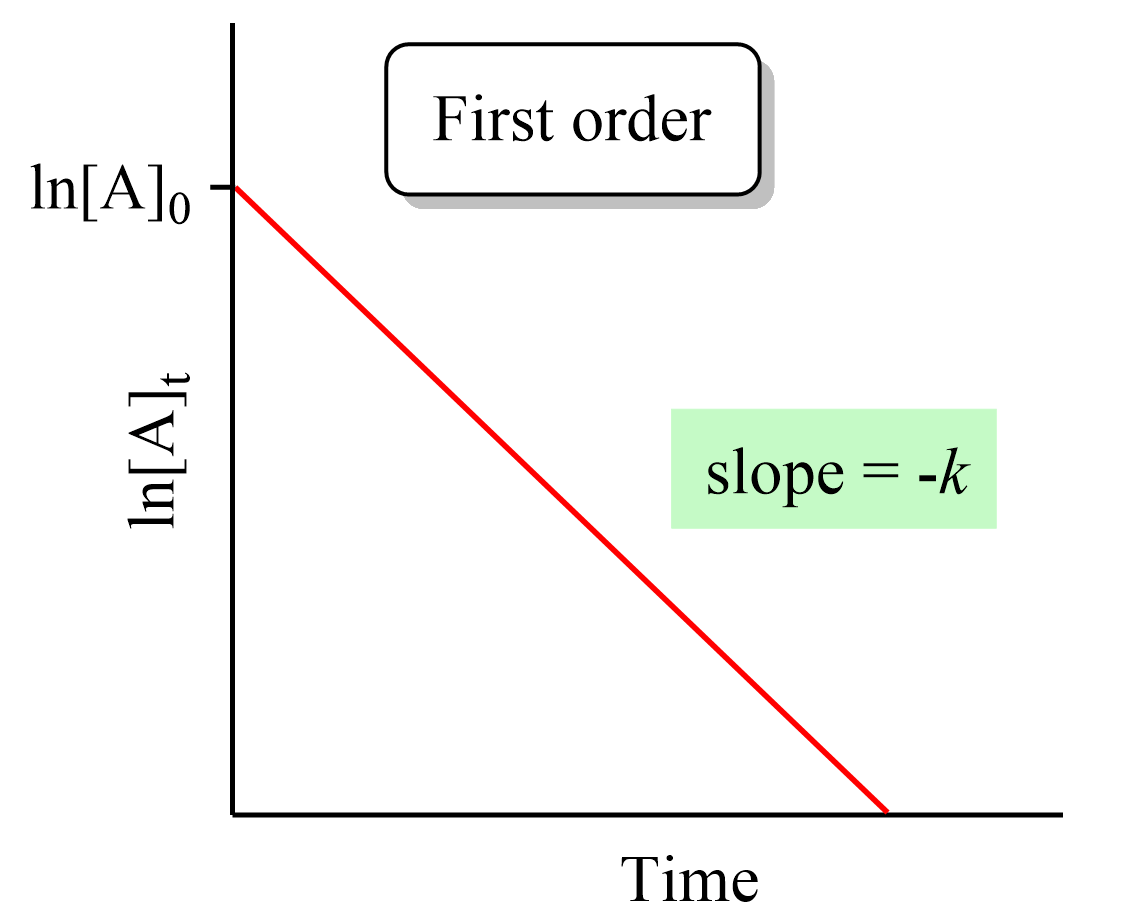
Most people learn the integrated rate law for first-order reactions in their second semester of chemistry and then never really think about it again until something goes wrong during an actual experiment. The equation itself is straightforward enough — concentration decays exponentially over time, and the half-life stays constant regardless of how much stuff you started with. The problem is that everything in practice looks first-order until you actually try to fit data to it. The standard form is ln[A] = -kt + ln[A], where [A] is the concentration at time t, [A] is the initial concentration, and k is the rate constant. Some textbooks also write it as the differential form, d[A]/dt = -k[A], which is the more fundamental statement. Both describe the same thing, but they're used differently. The integrated form is what you'll plug concentrations into when doing regression. The differential form is what matters when you're trying to understand why your reaction isn't behaving the way you expect it to. I remember running a degradation study on a pharmaceutical compound a few years back — something I thought would be a clean first-order process based on the literature. The plots of ln(concentration) versus time looked fine for the first twenty minutes, then the points curved away from the line. Not a little bit. Enough that the R² value dropped from 0.997 down to 0.89 by the end. Everyone on the project wanted to force-fit a line through the whole dataset and call it acceptable. I spent three days digging into what was happening and found that the compound was undergoing a secondary parallel reaction at higher conversion, producing a degradation product that absorbed at the same wavelength we were monitoring. The first phase was genuinely first-order, but the analytical method couldn't distinguish between the parent and the byproduct once the byproduct built up past about fifteen percent. Switching to HPLC with UV detection at a different wavelength fixed it completely. The real rate constant was right there in those first twenty minutes — I just had to stop trying to make the whole dataset fit the model.
Here's something beginners consistently get wrong: a constant half-life does not prove first-order kinetics. It's necessary but not sufficient. I've seen people confirm "first-order" behavior simply because the half-life appeared roughly constant across three data points measured at different starting concentrations. That's not proof. A zero-order reaction in a closed system with limited reagent can show an apparently constant half-life over a narrow concentration range, and so can some second-order processes if you're only looking at a small window of the curve. The way to actually verify the order is to vary the initial concentration deliberately and check whether the half-life changes. For true first-order, t/ = ln(2)/k, independent of [A]. If your half-life shifts even slightly when you change the starting concentration, the reaction isn't first-order, no matter how good your linear fit looks. Another thing nobody warns you about is the problem of baseline drift in spectrophotometric measurements. When you're tracking absorbance over hours or days for a first-order decay, the instrument baseline can shift by a few thousandths of an absorbance unit. That sounds negligible, but when you take the natural log of the readings, even small additive errors become multiplicative distortions that curve your plot. I usually run a blank reference cell alongside the sample and subtract the blank drift before converting to concentration. This takes maybe ten extra minutes per run but prevents you from spending hours wondering why your residuals show a systematic pattern. The main limitation of assuming first-order kinetics is that it only applies when one reactant's concentration is effectively constant or when the reaction genuinely has a molecularity of one. In enzymatic reactions, for example, you'll see pseudo-first-order behavior only when the substrate concentration is well below the Michaelis constant. Push the substrate concentration up and the whole model breaks down — you're now in mixed-order territory and the half-life will depend on how much substrate you started with. I've seen this come up repeatedly in formulation work where someone assumes first-order stability degradation across the entire shelf-life period, then gets surprised when the product fails accelerated stability testing because the degradation mechanism shifted at higher temperatures.
If you're working with real data and need to determine the rate constant, don't rely on two-point calculations using the integrated equation. Take at least six to eight time points spread across two or three half-lives, plot ln[A] versus time, and let the regression do the work. Report the confidence interval on the slope — that's your uncertainty on k. If the confidence interval is wider than twenty percent of the estimated k value, you don't have enough data or your system isn't actually first-order. Both are equally common.
Get the Full Details