What you actually need to know
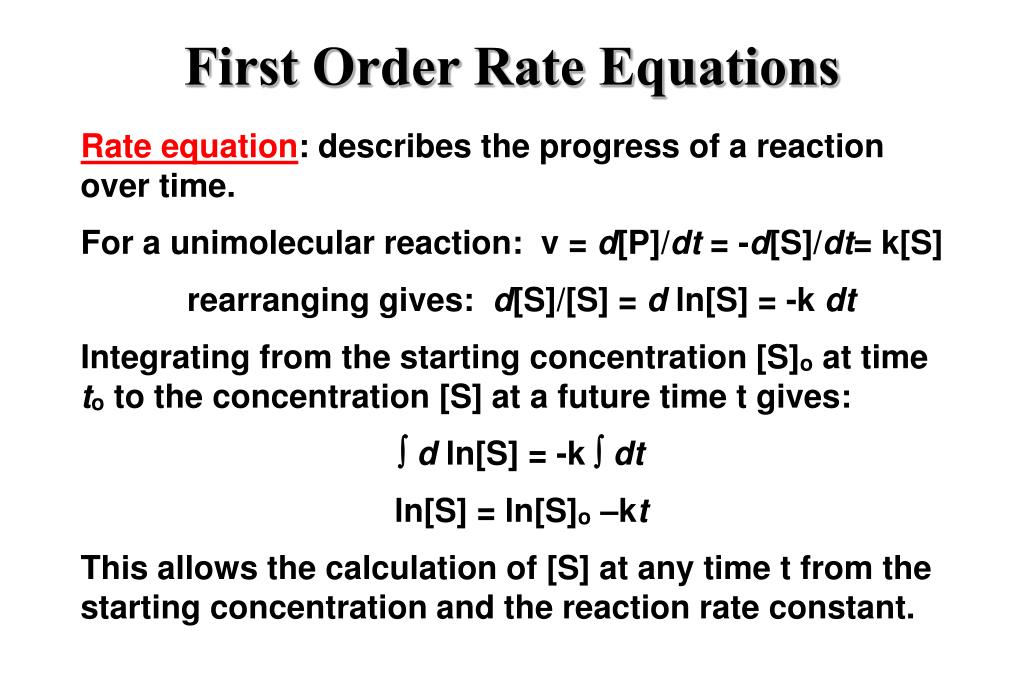
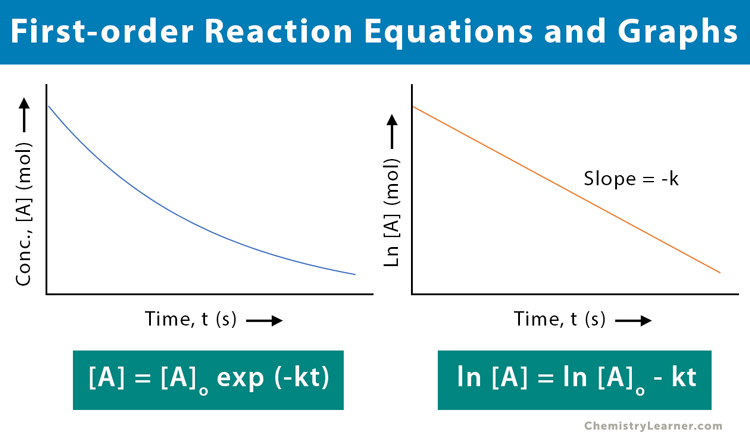
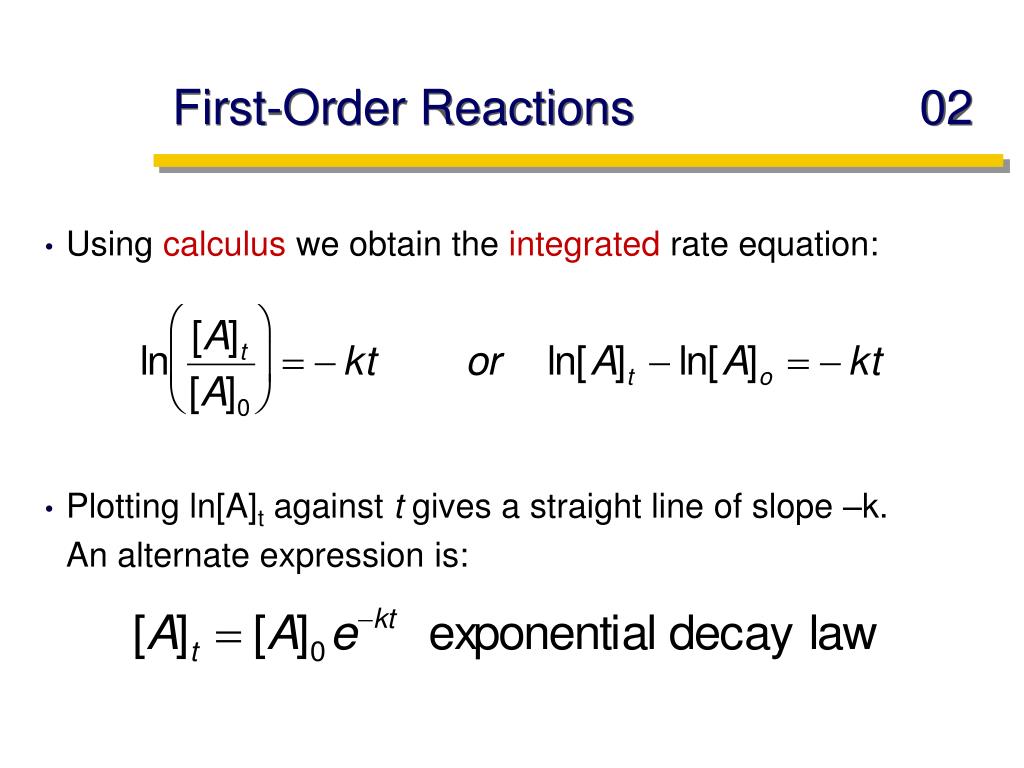
The First Order Reaction Equation describes how the concentration of a reactant changes over time when the rate depends on just that one species raised to the first power. It shows up everywhere in kinetics — radioactive decay, drug elimination, pollutant breakdown — because first-order behavior is simpler than it looks and comes up constantly in real lab work. There are two forms you will use, and they are equivalent depending on whether you start with an integrated expression or need a differential rate. The integrated form is: ln[A] = -kt + ln[A]
Or rearranged: [A] = [A] · e^(-kt) Where [A] is concentration at time t, [A] is the initial concentration, and k is the rate constant with units of inverse time (s¹, min¹, hr¹). The differential form is simply rate = k[A].
I spend most of my time working with the integrated form because experimental data comes as concentration measured at various time points, not as a rate. You plot ln[A] versus t and the slope gives you -k directly. A straight line through those points means the reaction is first order over the range you measured. That is the standard check. One practical detail people mess up: the natural log. Make sure your calculator or software is using ln and not log base 10. I once fed log values into the linear fit and got a rate constant that was off by a factor of 2.303. It took me twenty minutes to realize I had forgotten which log function the instrument software was returning. The fix was to just switch to ln in the spreadsheet and refit. That 2.303 factor is the conversion between log bases and it keeps catching people who are moving fast. Here is a quick walkthrough with actual numbers. Say you start with 0.500 M of a compound and after 40 minutes the concentration is 0.320 M. You want k.
Get the Full Details
ln(0.320) = -k(40) + ln(0.500) ln(0.320) = -1.1394 ln(0.500) = -0.6931
-1.1394 = -40k - 0.6931 -0.4463 = -40k k = 0.01116 min¹
Half-life from that is ln(2)/k = 62.1 minutes. The half-life is constant for a first-order process regardless of starting concentration. That is the thing that makes first-order clean and why it is easy to spot in data — every successive halving takes the same amount of time. There are edge cases where the equation behaves badly. I ran into this with a degradation study of a pharmaceutical intermediate. The reaction looked first order for the first three hours, then the rate slowed down noticeably. I thought I had introduced systematic error, so I ran a second batch under identical conditions and got the same curvature. The real cause was product inhibition. The accumulating degradation product was binding to the remaining reactant in a way that reduced the apparent rate. The integrated first-order equation still worked fine for the initial window, but if you try to fit the entire dataset you get garbage. My workaround was to restrict the linear regression to the first 60 percent of conversion and report k only for that range. I also added a note in the methods section explaining the deviation past that point. Hiding it doesn't help, and neither does pretending the model covers everything. Another counter-intuitive thing: temperature changes do not change the order. They change k, not the form of the equation. If you measure at 25°C and 40°C, both datasets should still be linear on a ln[A] vs t plot if the mechanism stays the same. What changes is the slope. You can use two temperature points to estimate activation energy through the Arrhenius equation, ln(k/k) = (Ea/R)(1/T - 1/T), but that is a separate calculation. The First Order Reaction Equation itself is temperature-agnostic in its structure.

The biggest limitation of this whole approach is assumption. You are assuming the reaction is truly first order before you even look at the data. Many reactions are only approximately first order under restricted conditions — usually when one reactant is in large excess or when the mechanism simplifies at low conversion. If you do not verify linearity across the full range, you are fitting noise and calling it a rate constant. A single correlation coefficient is not proof. Look at residuals. Systematic curvature in the residual plot means the model is wrong, not that your data is bad. A common pitfall is ignoring the units of k. If your time axis is in seconds, k is in s¹. If you switch to minutes somewhere in the middle of a calculation, your half-life will be wrong by a factor of 60. This happens more often than it should, especially when people copy rate constants from papers without checking the original time units. Always write the units next to k and double-check them before computing t/ or predicting concentrations at later times. If the reaction is not first order — and sometimes it simply is not — the differential method can still help you figure out the true order. You measure initial rates at several starting concentrations and plot log(rate) versus log([A]). The slope is the order. For a strict first-order reaction the slope is 1. If it comes out as 0.85 or 1.15 you have a partial-order system or experimental scatter, and you need to decide which interpretation fits the chemistry before committing to the integrated equation.
For practical purposes, the integrated first-order equation is reliable, fast, and straightforward when the assumptions hold. It is not a universal tool, and it breaks down without warning if the mechanism changes mid-reaction or if side reactions consume or produce the species you are tracking. Keep the data range reasonable, check the residuals, and do not force the model where it does not belong.