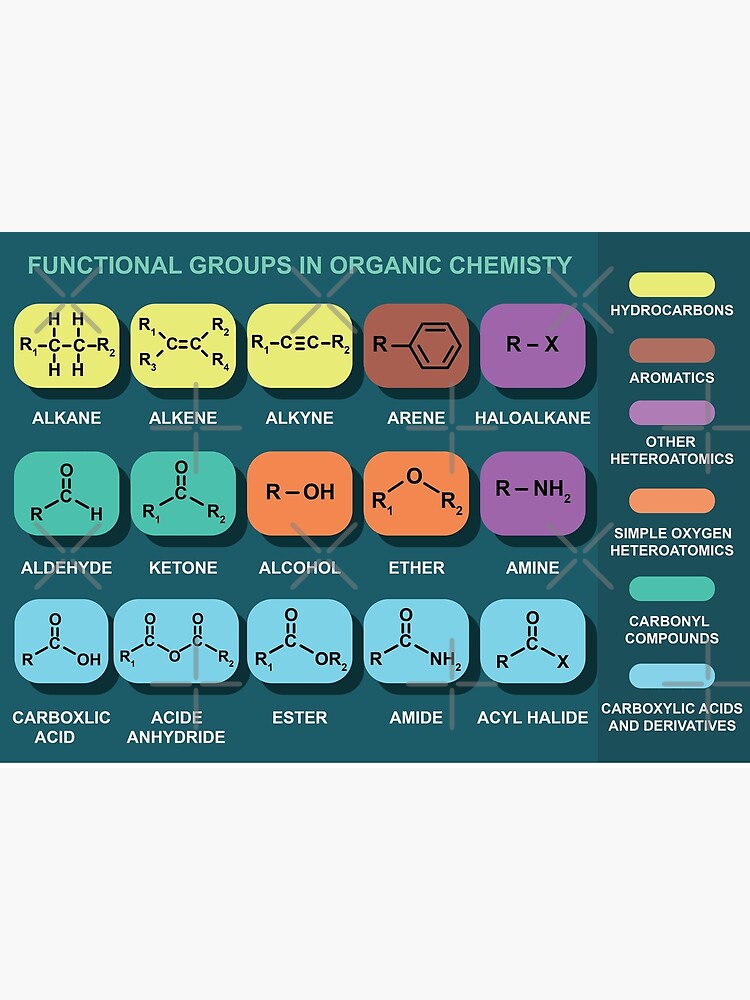
Why Functional Groups Actually Matter Outside the Textbook
When you're actually working in a lab, functional groups are the things that determine whether your reaction proceeds, stalls out, or explodes in your face. The textbook definition is straightforward enough, but the practical side is where people trip up. I'm going to walk through how I approach this, the mistakes I see constantly, and one edge case that cost me two weeks once. Start by memorizing the common ones, sure. Hydroxyl, carboxyl, amine, carbonyl, ester, ether, thiol, phosphate. That's table stakes. But the real skill is recognizing how they behave together in a molecule, not just what each one does in isolation. I always tell beginners to think about electron density and steric access first. Everything else follows from that. Here's the practical method I use when I'm given an unfamiliar compound and need to figure out what it'll do. First, I draw out the full structure with all lone pairs visible. Not a mental sketch, actually draw it on paper. Then I identify every functional group present, label them, and rate each one as electron-donating or electron-withdrawing relative to its neighbors. This takes about five minutes for a moderately complex molecule. From there, I map out the most likely reactive sites. The site with the highest electron density is usually your nucleophile, and the site adjacent to an electron-withdrawing group is your electrophile. Most reactions I run are just one of those two attacking the other.
Let me give you a concrete example. I had a synthesis project where the target molecule had both a secondary alcohol and a primary amine on the same carbon chain. I needed to selectively protect the amine without touching the alcohol. The textbook answer would say "use a Boc group" and move on. In practice, I found that standard Boc anhydride conditions also began acylating the alcohol at room temperature, which ruined the selectivity. The workaround was to cool the reaction mixture to 0 degrees Celsius and add the Boc reagent dropwise over twenty minutes while monitoring by TLC. At that temperature, the amine reacted cleanly and the alcohol stayed put. It added maybe twenty minutes to the procedure but saved me from purifying a mixture that would have been a nightmare.
The Part Nobody Teaches You
Functional groups don't exist in vacuum. A hydroxyl group next to a carbonyl behaves completely differently than one three carbons away. The proximity matters enormously, and this is where most students lose points on exams and waste time in the lab. When a hydroxyl is directly adjacent to a carbonyl, you get intramolecular hydrogen bonding that can stabilize certain conformations and make that particular OH less nucleophilic than it would otherwise be. I've seen people miss this and then wonder why their esterification didn't proceed as expected. Another thing that comes up constantly: protecting groups. People treat protecting groups like optional extras. They're not. If your molecule has multiple reactive functional groups and you only want one to react, you need to protect the others or the selectivity will fall apart. The most common mistake I see is choosing a protecting group that's incompatible with your reaction conditions. You pick a benzyl ether because it's robust, but then your reduction step uses palladium on carbon with hydrogen gas, and poof, your protecting group is gone along with your desired product. Always check compatibility before you commit to a protection strategy. It saves hours of troubleshooting later. There's also the matter of pKa values and what they actually tell you in practice. A carboxylic acid has a pKa around 4 to 5, which means at physiological pH it's mostly deprotonated. But in an organic solvent with no water around, that same carboxylic acid might sit there protonated and completely unreactive toward a base-catalyzed mechanism you were counting on. Solvent effects are real and they're easy to overlook. I once ran a reaction in dry DCM where I expected a carboxylate to form and act as a nucleophile. Nothing happened for six hours. Switched to DMF, which stabilizes the ionic intermediate, and the reaction proceeded in forty minutes. Same reagents, same temperatures, different solvent behavior.
Get the Full Details

Common Pitfalls and What to Do Instead
Pitfall number one: assuming that two molecules with the same functional groups will behave the same way. They won't. Molecular context changes everything. Ethanol and phenol both have hydroxyl groups, but phenol is roughly a million times more acidic because the resulting conjugate base is resonance stabilized. If you treat them the same in a reaction design, you'll end up confused about why your phenol required a stronger base than you calculated. Pitfall number two: ignoring the solvent's functional group. THF is an ether, which is generally inert, but under strongly acidic conditions it can open up and form peroxides. If you're running an acidic reaction in old THF that's been sitting around, you're introducing an uncontrolled variable. Fresh solvent, distilled if possible, and definitely test for peroxides if the bottle isn't brand new. This isn't theoretical, it's happened to me and it's not fun to clean up. The bigger limitation of relying solely on functional group recognition is that it breaks down when you get into sterically crowded molecules or conjugated systems where electronic effects cancel each other out. In those cases, computational modeling or at least a careful literature search for analogous compounds is necessary. There's no shortcut around that. Functional group analysis gets you to the starting line, but it won't tell you the entire race course.
Quick Reference for the Most Common Groups
Haloalkanes: C-X bond, generally electrophilic at the carbon, good substrates for substitution and elimination reactions. Reactivity order is I > Br > Cl > F in most cases. Alcohols: C-OH, can act as nucleophiles when deprotonated or as leaving groups when protonated. Oxidation state depends on whether it's primary, secondary, or tertiary. Primary goes to aldehyde then carboxylic acid. Secondary goes to ketone. Tertiary generally doesn't oxidize under normal conditions. Aldehydes and ketones: C=O with hydrogen or carbon substituents respectively. Always the most reactive carbonyl in a molecule unless something stronger is present. Nucleophilic addition is the standard reaction pattern.
Carboxylic acids and derivatives: The carbonyl carbon here is less electrophilic than in aldehydes and ketones because the oxygen lone pair donates back into the carbon. Esters, amides, and acid chlorides follow from this core structure, with acid chlorides being the most reactive and amides the least. This reactivity ladder matters when you're planning a synthesis and need to control which carbonyl reacts. Amines: C-NH2 or substituted versions. Basic and nucleophilic. Can be acylated, alkylated, or converted to diazonium salts depending on the application. Primary and secondary amines form imines with aldehydes and ketones. Tertiary amines don't have the hydrogen needed for that pathway.