Working With Gas Dissolved In A Liquid
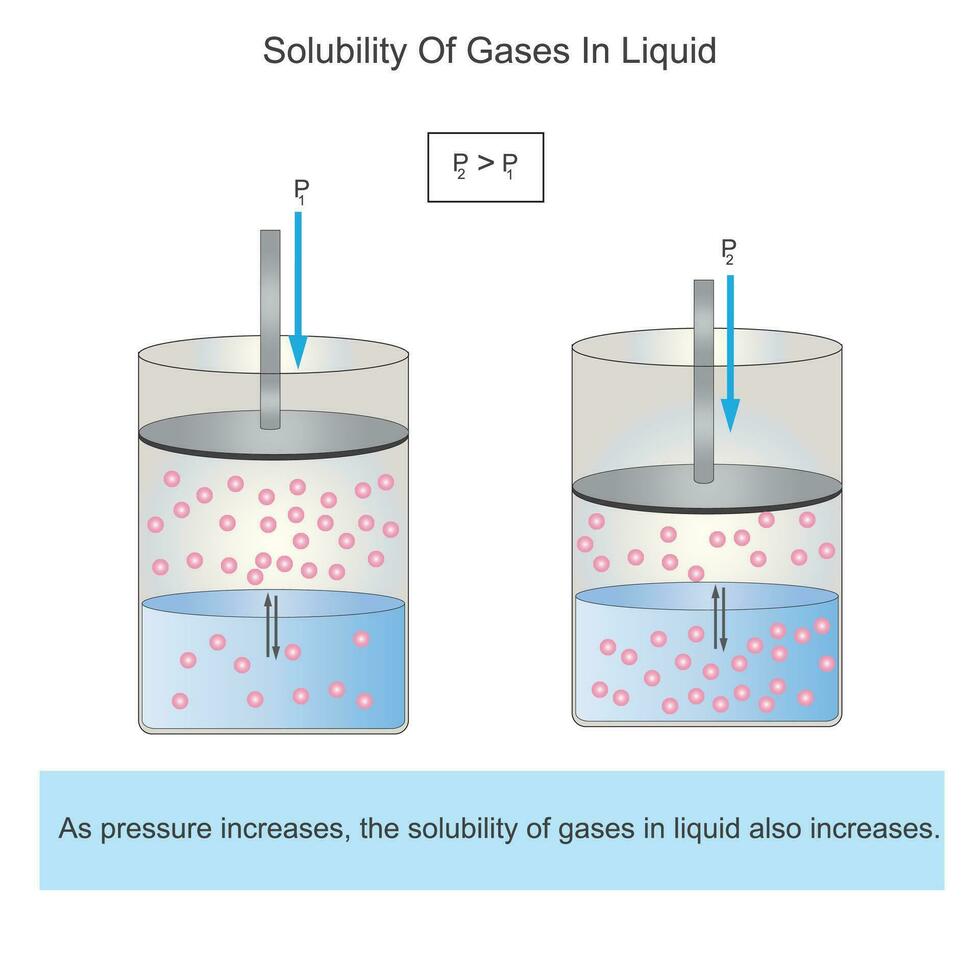
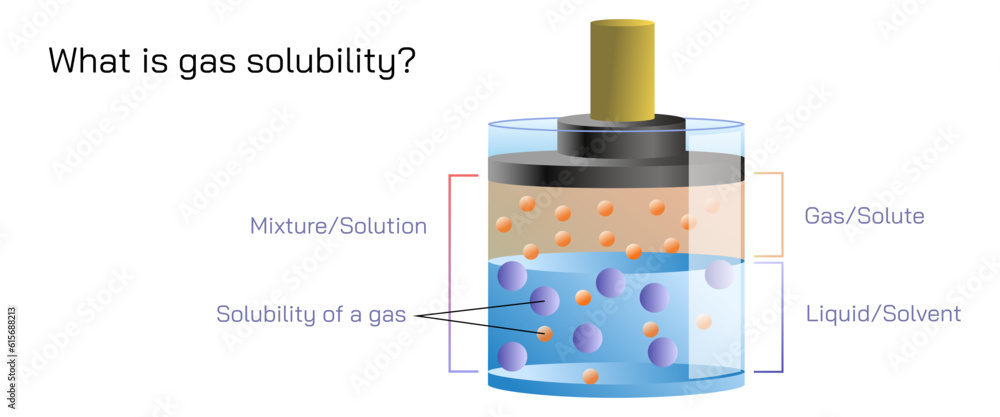
When a gas sits above a liquid, molecules constantly cross the boundary between phases. Some dissolve into the liquid. Others escape back out. At equilibrium, the rates match and the concentration stays steady. That concentration depends on three things: the partial pressure of the gas above the liquid, the temperature, and what else is already in the liquid. That's the whole picture in one sentence. Henry's Law is the equation people actually use. It says the concentration of a dissolved gas equals the Henry's law constant multiplied by the partial pressure of that gas. The constant changes with temperature and solvent. For oxygen in water at 25 degrees Celsius, the constant works out to roughly 1.3 times ten to the negative three moles per liter per atmosphere. That means at one atmosphere of pure oxygen, you get about 1.3 millimolar dissolved oxygen. Air is only 21 percent oxygen, so in practice you get around 0.27 millimolar. That number is why most freshwater fish can survive but only just barely when the water sits stagnant on a hot day.
Why Gas Dissolved In A Liquid Matters In Practice
I spent a few years running HPLC systems and the degassing step always came up as a nuisance nobody wanted to deal with. The mobile phase has air bubbled into it during mixing and storage. When that pressurized mobile phase hits the detector flow cell, the pressure drop causes the gas to come out of solution as microbubbles. Those bubbles scatter light and make your baseline go wild. Peaks get distorted. Retention times drift. You spend half your morning troubleshooting something that isn't actually broken. The standard workaround is a membrane degasser. Helium is sparged through the solvent lines and then vacuumed away, or you just run the lines through a heated vacuum chamber. The membrane approach is cleaner for modern systems. The solvent flows on one side of a gas-permeable tube and a vacuum pulls gases through the polymer wall without letting any liquid pass. It takes about twenty minutes to degas a liter of mobile phase, and once it's done, the baseline stays stable for hours. If you skip this step entirely, expect a ten to fifteen percent increase in noise and maybe an hour wasted re-running samples before you figure out what's going on. Another edge case that cost me a day once involved preparing standard solutions for dissolved oxygen calibration. I was making zero-oxygen standards by sparging nitrogen through deionized water. The problem was that my nitrogen line had a small leak downstream of the regulator. Atmospheric oxygen was creeping in through that tiny leak at maybe two percent of the total flow. Over four hours of sparging, the water never dropped below 0.3 parts per million dissolved oxygen instead of hitting the near-zero target I needed for the calibration curve. The fix was installing a downstream oxygen sensor in-line and watching the reading stabilize before trusting it. You'd think bubbling an inert gas through water for hours would do the job, but a small leak changes everything.
The Temperature Problem Nobody Talks About
Most people learn that gas solubility decreases as temperature rises. That's true for the vast majority of common gases in water. But the relationship isn't linear and the magnitude varies wildly between gases. Carbon dioxide drops off sharply from zero to sixty degrees Celsius. Oxygen follows a similar pattern but with a smaller absolute change. Hydrogen and the noble gases behave differently at elevated temperatures, sometimes showing a solubility minimum before rising again. If you're working with systems that heat up significantly, looking up a single Henry's law constant at room temperature and applying it everywhere will introduce errors. I've seen people miss a fifteen percent shift in dissolved gas concentration between twenty degrees and forty degrees because they assumed the constant stayed fixed. The salinity effect matters too. Seawater holds about twenty percent less oxygen than fresh water at the same temperature and pressure. The dissolved ions compete for hydration shells and effectively push gas molecules out. If you're modeling gas exchange in coastal systems or running experiments with brine, using freshwater constants will systematically overestimate your dissolved concentrations. The practical adjustment is to multiply your Henry's law result by a salinity correction factor, which you can find in standard tables or approximate with a simple linear reduction per part per thousand of salinity.
Get the Full Details

Pressure Changes And Real-World Consequences
Increasing pressure forces more gas into solution. That's straightforward. The reverse is where things get interesting and occasionally dangerous. Decompression disease in divers comes from dissolved nitrogen coming out of solution in blood and tissue when pressure drops too quickly. The physiology is complex, but the underlying principle is pure physical chemistry. At depth, elevated pressure drives nitrogen into the blood at concentrations that are harmless while submerged. Ascend fast and the nitrogen precipitates as bubbles. The rule of thumb for recreational diving is to ascend no faster than nine meters per minute and do decompression stops. That's not arbitrary. It gives the nitrogen time to safely diffuse back into the bloodstream and be exhaled rather than forming bubbles in the joints and bloodstream. In industrial settings, pressurized water treatment systems face a similar issue on a smaller scale. When water is held under pressure to dissolve oxygen for boiler treatment and then suddenly depressurized, the excess dissolved oxygen flashes out as microbubbles. This can cause pitting corrosion in downstream equipment. The workaround is to keep the system pressurized until the water reaches the point of use, or to install a vent upstream to let the gas escape slowly rather than forming damaging bubbles inside pipes and pumps.
Gas Dissolved In A Liquid And What Breaks The Model
Henry's Law works well at low to moderate pressures and for gases that don't react with the solvent. It breaks down when the gas chemically interacts with the liquid. Ammonia in water is a classic example. It doesn't just dissolve. It reacts to form ammonium and hydroxide ions. The apparent solubility is orders of magnitude higher than Henry's Law would predict because the reaction continuously removes dissolved ammonia molecules, pulling more gas in. Carbon dioxide in alkaline solution shows the same behavior. If you're modeling CO absorption into a caustic scrubbing solution, applying Henry's Law directly will give you answers that are completely wrong. You need to account for the chemical equilibrium alongside the physical solubility. Another limitation people run into is at very high pressures. Above roughly ten atmospheres, the linear relationship between partial pressure and concentration starts to deviate. The Henry's law constant itself becomes pressure-dependent. For most laboratory and industrial work this doesn't matter because you're operating well below that threshold. But if you're working with supercritical fluids or high-pressure reactor systems, you'll need to use a different model like the van 't Hoff equation adjusted for pressure or consult empirical solubility data for your specific gas-solvent pair. The most common practical mistake I see is assuming that stirring or agitating a liquid increases how much gas it can hold. It doesn't. Agitation only speeds up how quickly equilibrium is reached. The final concentration is set by pressure, temperature, and the Henry's law constant. A still beaker of water exposed to air at sea level will absorb oxygen at the same eventual concentration as a vigorously stirred one. The stirred beaker just gets there in minutes instead of days. If your dissolved gas readings are inconsistent, check your pressure and temperature control before you blame the mixing.
There's also the issue of what gas is actually above the liquid. In an open container, the liquid equilibrates with the partial pressure of that gas in the atmosphere. That's fine for oxygen and nitrogen because their atmospheric concentrations are stable. But for volatile organic compounds or trace gases, the partial pressure above the liquid may not reflect what you think it is. A solvent like acetone sitting in an open beaker will establish its own vapor pressure above the liquid, which dominates the gas phase composition. Trying to measure dissolved air in that setup gives you garbage data because the headspace isn't air anymore. Seal the system and control the headspace composition if you need accurate results. Temperature control during measurement is another subtle source of error. Henry's law constants change by roughly two to three percent per degree Celsius for most gases in water. If your sample warms by just two degrees between preparation and measurement, your calculated concentration will be off by that much. I use a thermostatted bath at twenty-five degrees for all my calibration work and let samples equilibrate for at least thirty minutes before measuring. It adds time to the protocol but eliminates one of the more frustrating sources of variability.