Getting Reliable Ethanol Readings by Gas Chromatography
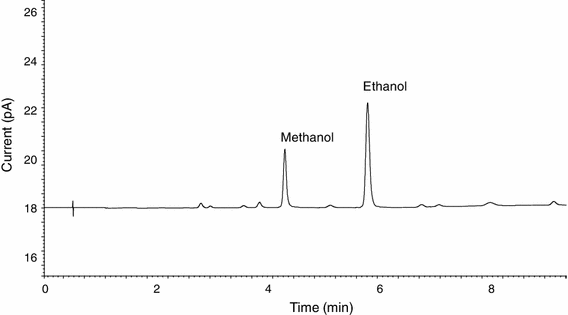
The standard approach uses a HP-PLOT Q or similar porous polymer column. Set the injector to split mode around 20:1, keep the column at 45°C for three minutes, ramp to 80°C at 10 degrees per minute, and run the detector at 150°C. I run ethanol samples this way every week. The peak comes through clean between 1.8 and 2.2 minutes on a typical 30 meter column. Water is the real problem here. If your sample is aqueous, you need to handle it differently than a neat spirit. Here is what actually works in a working lab, not what the manufacturer brochure says. I use an internal standard of n-propanol at 0.5% v/v. You add it to every sample and calibration standard. Without it, you are chasing temperature drifts and injection volume inconsistencies. My calibrators run from 1% to 60% ethanol. I prepare them gravimetrically with Class A flasks. Weight is more accurate than volume for liquids at this level. The real edge case that bites people is acetaldehyde interference. It elutes almost exactly where ethanol does on some column chemistries. I encountered this when analyzing a fermented beverage matrix. My ethanol peak looked normal until I checked the mass spectrum. The abundance ratio for m/z 45 to m/z 47 was off. Turns out the acetaldehyde was co-eluting and inflating my readings by about 3%. The fix was switching to a Zebron ZB-WAXplus column with a thicker film. That added roughly 30 seconds to the run but pulled the peaks apart cleanly. It also meant I could drop the oven start temperature to 35°C and hold it longer.
Another thing nobody warns you about: ethanol is hygroscopic. If your standards sit out even briefly they absorb water from the air and your concentration readings drift. I keep my stock solutions in sealed ampoules. Once opened I use them within two weeks and recheck the concentration against a fresh primary standard. Your calibration curve should not be older than 30 days regardless of what the method documentation claims. Detector choice matters more than the column for quantitation accuracy. FID is the default and it works fine for most applications. But if you are working below 0.1% ethanol, like in some residual solvent testing, you need a different approach. I use headspace sampling with a flame ionization detector in those cases. The headspace equilibrium gives you consistent partitioning and you avoid injecting liquid altogether. Run time goes up to about 12 minutes per sample but you gain precision at the low end. Data processing is where most errors creep in. Set your integration window to capture the full ethanol peak plus a small baseline region before and after. Automatic integration tends to cut the tail off ethanol peaks on polymer columns. That tail can account for 8 to 12% of the total area if you are not careful. I manually verify every integration point during method validation and then lock the parameters so the software does not second guess itself on subsequent runs.
If you need a reference method document, the official USP chapter 467 covers residual solvents including ethanol and provides acceptable GC parameters. The AOAC Official Method 985.16 is the food and beverage standard. Both are available through their respective organizations for download. The parameters vary slightly between them. USP runs a higher oven temperature and uses a different column dimension. AOAC specifies a longer equilibration time. Pick the one that matches your matrix and stick with it.
Get the Full Details