How transcription actually works once you stop thinking about eukaryotic noise
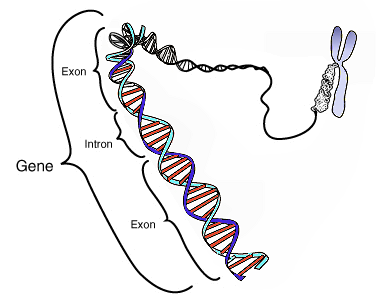
Most people learn about gene expression through the lens of what happens in a nucleus. You have promoters, enhancers, splicing machinery, chromatin remodeling, all of that. Prokaryotes don't have any of that overhead. When you look at Gene Expression In Prokaryotes, you're basically looking at a direct pipeline from DNA to RNA to protein with minimal intermediary steps. That's the simplification. The reality is a bit messier and more interesting. A prokaryotic promoter has two conserved regions that RNA polymerase recognizes — the 35 box and the 10 box. The 10 region is usually something like TATAAT, called the Pribnow box. The 35 region is typically TTGACA. Sigma factors mediate the initial binding, and the best-known example is in E. coli. Once the polymerase docks, it melts about 14 base pairs around the 10 region to form an open complex. There's no need for ATP-dependent chromatin remodeling because there's no nucleosome to deal with. The transcript comes out as a continuous strand. No introns. No 5' capping. No polyadenylation in the eukaryotic sense, though some bacterial mRNAs do get short poly(A) tails that actually target them for degradation rather than stabilizing them. That's a detail most textbooks skip.
Operons and the polycistronic shortcut
One of the things that trips people up is the operon concept. A single promoter drives transcription of multiple coding sequences in one mRNA molecule. The lac operon is the textbook example, but it's worth noting that not all genes in an operon are co-transcribed at equal levels. Position effects matter. A gene closer to the promoter tends to be expressed more simply because RNA polymerase falls off the template before reaching downstream sequences. This is called transcriptional polarity and it's a real practical concern when you're building synthetic circuits. I spent a week trying to get uniform expression from a three-gene operon in E. coli and couldn't figure out why the third gene was essentially silent. Turns out the ribosome binding sites weren't spaced properly, and the downstream gene was getting translated but the transcript was degrading before the ribosome could even load. I added a RBS variant with stronger Shine-Dalgarno complementarity and a ribosome-protection sequence between the second and third cistrons. Expression balanced out after that. There's a paper from the Collins lab that covers this kind of positional effect pretty thoroughly if you want to dig into it.
Regulation happens fast and it's combinatorial
Prokaryotic regulation isn't just on and off switches. You have attenuation, where transcription terminates prematurely based on translation coupling. The trp operon is the classic case — the ribosome stalls on the leader peptide when tryptophan is scarce, and that changes which hairpins form in the mRNA, keeping the terminator hairpin from closing. When tryptophan is abundant, the ribosome moves quickly, the terminator forms, and transcription stops before the structural genes are reached. Repressors and activators work differently than you might expect from eukaryotic systems. A repressor like LacI doesn't block elongation — it prevents the polymerase from even initiating by sitting on the operator sequence. Activation often involves DNA looping. The AraC protein in the arabinose system is a good example. It binds to two distant sites and loops the intervening DNA, which changes the promoter's accessibility depending on whether arabinose is present. Here's a counter-intuitive point: not all regulation is negative. Some of the most important control mechanisms in prokaryotes involve positive regulation — proteins that help RNA polymerase bind more effectively. Without these activators, basal expression levels are often too low to be biologically useful. CRP-cAMP is one of those systems. When glucose is low, cAMP rises, CRP binds cAMP, and the complex docks near certain promoters to boost transcription. This is how the cell senses carbon source quality and adjusts its metabolic gene expression accordingly.
Get the Full Details
Termination isn't always clean
There are two main termination mechanisms. Rho-dependent termination uses the Rho helicase, which loads onto the nascent RNA at rut sites and chases down to the polymerase, unwinding the RNA-DNA hybrid. Rho-independent termination relies on a GC-rich hairpin followed by a run of U residues. The hairpin destabilizes the elongation complex, and the weak rA-dT bonds in the hybrid let the RNA fall off. The thing people miss is that termination efficiency is rarely 100%. Some polymerases read through both types of terminators, producing longer transcripts that can interfere with downstream genes or generate antisense RNA. In synthetic biology applications, this readthrough is often a problem you have to engineer around. Adding a T7 terminator downstream of your construct or using a transcriptional insulator sequence usually helps, but it adds complexity to the design.
Translation and transcription are coupled
This is probably the single most important difference from eukaryotic systems. In prokaryotes, ribosomes load onto the mRNA while it's still being synthesized. Translation initiation doesn't require the transcript to be fully processed or exported — there's no export step because there's no compartment boundary. The ribosome binds the Shine-Dalgarno sequence, which base-pairs with the 16S rRNA, and translation begins almost immediately. This coupling has consequences. Ribosomes physically block premature termination signals. If transcription produces a rho-dependent terminator hairpin but a ribosome is already translating through that region, the terminator won't form and the polymerase continues. This is how attenuation works in the trp system, and it's also a design principle you exploit when building synthetic genetic circuits. Put a ribosome binding site upstream of a terminator and you can make the terminator conditional on translation.
Practical considerations when you're working with this
If you're cloning expression constructs for prokaryotic systems, the choice of promoter matters a lot. T7 promoters give very high expression but can be toxic if the gene product interferes with cell growth before induction. The pET system is the standard here, but you need a strain that expresses T7 RNA polymerase, like BL21(DE3). The DE3 prophage carries the T7 polymerase gene under lacUV5 control, so you induce with IPTG and the polymerase comes on. Bassler quorum sensing systems in Vibrio harveyi show that prokaryotic gene expression isn't just about internal metabolism. Cells sense population density through autoinducer molecules and coordinate behavior at the population level. Bioluminescence, virulence factor production, biofilm formation — all regulated through cell-to-cell signaling. This is fundamentally different from the inducer-repressor model you learn in introductory courses. One thing I've found useful when troubleshooting expression problems is checking the codon composition of your insert. E. coli prefers certain codons over others, and rare codons can cause ribosome stalling, premature termination, or frameshifting. The codon adaptation index gives you a quick metric, but it's not perfect. I've seen cases where a gene with a reasonable CAI still expressed poorly because of mRNA secondary structure near the start codon. Running an RNA folding prediction on the 5' end of your transcript and mutagenizing the problematic structure often resolves it.
Another practical tip: prokaryotic mRNA half-lives are short, usually on the order of a few minutes. This means your expression levels can change rapidly when you shift conditions. If you're doing a time course experiment, the timing matters more than you might expect. A sample taken 30 seconds late could look completely different.
When prokaryotic expression falls apart
Not every gene expresses well in a bacterial host. Membrane proteins, proteins requiring eukaryotic post-translational modifications, and genes that produce toxic products are common failure cases. E. coli doesn't have glycosylation machinery, disulfide bond formation in the cytoplasm is inefficient due to the reducing environment, and proteolysis can chew up misfolded proteins before they accumulate to useful levels. If you need disulfide bonds, the periplasm is a better place to target your protein because it's oxidizing. Signal sequences like PelB or OmpA can route your protein there. For glycosylation, you'd need to move to a different expression system entirely — yeast, insect cells, or mammalian cells. There's no workaround that makes E. coli do eukaryotic modification faithfully. Protein aggregation into inclusion bodies is another frequent issue. The protein folds too fast, hydrophobic regions interact before the structure is complete, and you end up with insoluble precipitate. This isn't always a loss. Inclusion bodies are relatively easy to purify — centrifuge the cells, resuspend the pellet, spin again. Then you can solubilize with urea or guanidine and refold by dilution or dialysis. The refolding step is where the real work happens, and it's often where people give up. Screening denaturant gradients, redox couples, and additives like arginine or glycerol can improve yields, but there's no universal protocol.
What you should take away
Prokaryotic gene expression is simpler than eukaryotic systems but it's not trivial. The coupling of transcription and translation, the variety of regulatory mechanisms beyond simple repression, the practical challenges of expression and folding — all of these matter when you're actually working in a lab. The conceptual framework is straightforward: promoter, coding sequence, terminator. The execution is where the details accumulate. If you're approaching this from a computational angle, tools like RegPrecise and DBTBS can help you predict promoter sites and regulatory motifs. For experimental work, the key is understanding that prokaryotic cells are fast and responsive. They adapt to environmental changes in minutes, and your experimental conditions determine what you observe. A culture grown at 37°C behaves differently from one grown at 30°C. Induction timing, IPTG concentration, and culture density all matter more than you might assume from reading a protocol. The field has moved beyond just studying model organisms like E. coli and Bacillus subtilis. Metagenomic data has revealed diverse regulatory networks in environmental bacteria, and synthetic biology is using prokaryotic expression systems for everything from metabolic engineering to protein therapeutics. The basics haven't changed, but the applications keep expanding.