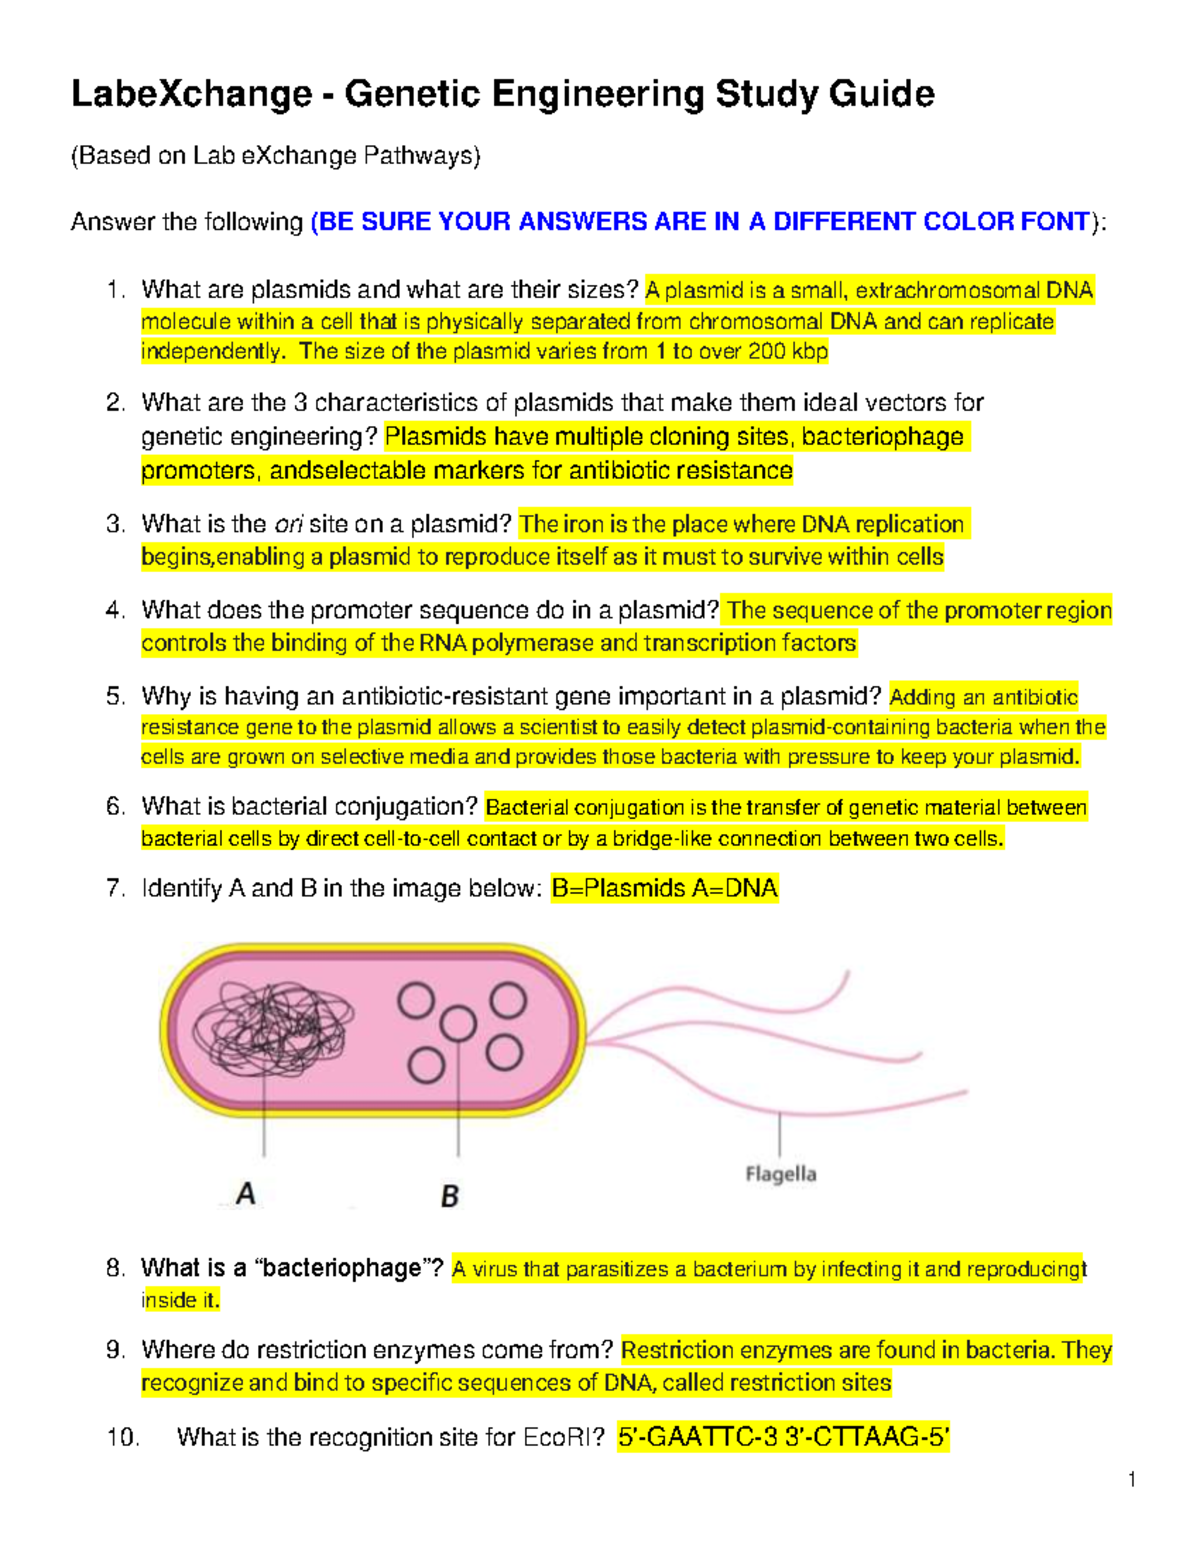
Building a Practical Genetic Engineering Modern Biology Study Guide
I spent too many semesters trying to cram everything about genetic engineering into one document. The best study guide I ever made wasn't organized by topic. It was organized by method, because that's how the work actually gets done in a lab. If you're trying to study this material for a course or just need to understand how it all connects, here's what actually works. A solid guide needs to cover the tools first, then the logic behind why you'd pick one over another. Start with restriction enzymes and the different types—Type II is what you'll use in undergrad labs, but you should know Type IIS exists for Golden Gate assembly. Move into vectors, and don't just list plasmid features. Explain why you'd choose a fosmid over a BAC for a particular project. Most students skip that distinction and then get confused during cloning exercises. The second major section should be transformation and selection. Antibiotic resistance markers are basic, but you need to understand blue-white screening, counter-selection with sacB, and when to use chemical competence versus electroporation. I had a student once try to transform electrocompetent cells using calcium chloride heat shock and spend three hours wondering why the efficiency was terrible. The cells were fine. The method was wrong.
After that comes CRISPR-Cas systems. This is where the field has moved the fastest, and your study guide needs to reflect that. Don't just explain Cas9. Cover Cas12, Cas13, and the difference between HDR and NHEJ repair pathways. The repair mechanism you get determines whether you're doing knock-in or knock-out work, and confusing the two will wreck an entire experiment.
The Core Methods You Need to Know Cold
PCR is the foundation, but your guide should go beyond the basic cycle. Include hot-start PCR, touch-down protocols, and when you'd use a high-fidelity polymerase versus a standard one. Taq gives you A-tailing for TA cloning, but if you're doing site-directed mutagenesis or cloning into a vector where frame matters, Q5 or Phusion saves you from dealing with extra steps later. Gel electrophoresis seems simple, but there are practical details that matter. Agarose concentration determines resolution for different fragment sizes. A 1% gel resolves 500 bp to 10 kb fragments well. If you're running small oligos or primers, that same gel won't separate them properly. Use a polyacrylamide gel or a high-percentage agarose instead. Students rarely learn this until they've already run the wrong gel and complained about smearing. Sanger sequencing and next-generation sequencing belong in the same section, but they serve different purposes. Sanger is for confirming a single clone or verifying a construct. NGS is for validating genome edits at scale, checking off-target effects, or doing RNA-seq. If your study guide treats them as interchangeable, it's not accurate to how labs actually use these technologies.
Get the Full Details

Common Pitfalls That Cost Time and Samples
One thing nobody warns students about is the degradation problem with CRISPR reagents. Once you resuspend your guide RNA, it degrades faster than people expect, especially if the buffer isn't properly protected. I remember working through a cloning project where we kept getting zero positives after transformation. We rechecked every primer, every enzyme, every vector preparation. The issue turned out to be that our gRNA stock had been freeze-thawn five times over three weeks, and the efficiency had dropped to nearly zero. Aliquoting into single-use volumes fixed it immediately. Another issue is incomplete restriction digests leading to self-ligation. You set up a double digest, run the gel, and see your band at the right size. But if the enzyme wasn't fully active—wrong buffer, insufficient time, star activity from glycerol carryover—the vector might recircularize without inserting your fragment. Always phosphatase-treat the vector when you're cloning anything difficult, and always include a no-insert control to measure background. When working with larger constructs above 10 kb, recombination in E. coli becomes a real problem. The bacteria will rearrange repetitive sequences inside your plasmid. Using recA-deficient strains like Stbl3 or TOP10 helps, but it doesn't eliminate the risk. If you're assembling something large, consider doing it in yeast using Gateway or Gibson assembly rather than relying on standard cloning in regular DH5.
How to Structure Your Actual Study Sessions
Don't read through chapters linearly. Work backward from problems. Start with a protocol you need to perform, look up what each step does, then fill in the theory. For example, if you need to do a site-directed mutagenesis, figure out what primer design rules matter, then learn why those rules exist. The DpnI digestion step makes more sense once you understand methylation patterns in dam+ strains versus dam- strains. Make flashcards for enzyme recognition sequences, but also make them for conditions. What buffer works with both EcoRI and HindIII? At what temperature does Taq polymerase work optimally? Which antibiotic survives autoclaving? These details show up on practical exams more often than anyone expects. For the molecular biology calculations, practice molarity conversions and dilution problems until they're automatic. You'll need to calculate how much DNA to add to a 50 microliter reaction, how much primer to resuspend to get a 100 micromolar stock, and how to convert nanograms of plasmid into moles for ligation ratios. Get comfortable with the formula nanograms times Avogadro's number divided by length in base pairs times 650 gives you femtomoles. It takes five seconds once you've done it enough times.
Resources Worth Using
Sambrook and Russell's Molecular Cloning remains the reference most people cite, even though it's dense. The current edition covers CRISPR applications and newer cloning methods, but it's not a quick read. For something faster, Addgene's protocols page is reliable and frequently updated. Their papers on CRISPR delivery methods and vector designs are actually useful for someone building a study guide. NCBI's Gene database and HomoloGene are worth knowing for comparative genetics sections of your guide. When you're studying gene function across species, having a quick reference for orthologs saves hours of searching. The Ensembl genome browser is better for visualizing genomic context and regulatory elements. If you want a downloadable study guide formatted for actual use, the one I ended up relying on most was compiled from lab notebooks and course materials rather than any single textbook. It had protocol flowsheets on one side and troubleshooting notes on the other. That format forced you to think about what goes wrong, not just what should go right. You can find similar resources on university lab websites, or you can build your own by taking apart the protocols you actually perform and annotating each step with the reasoning behind it.