Working With Molecular Geometry in Practice
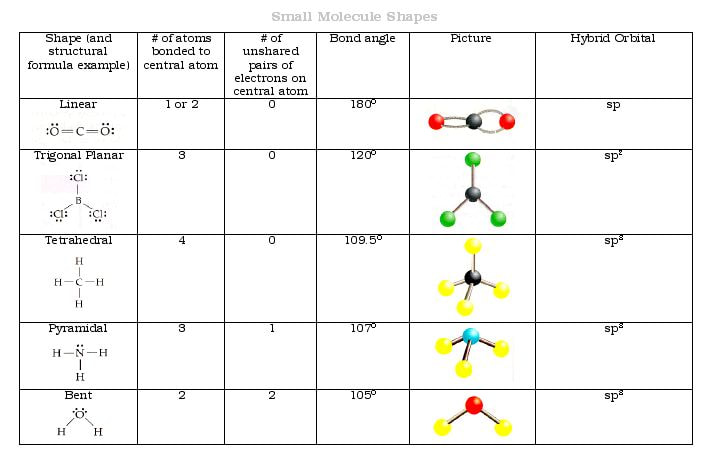
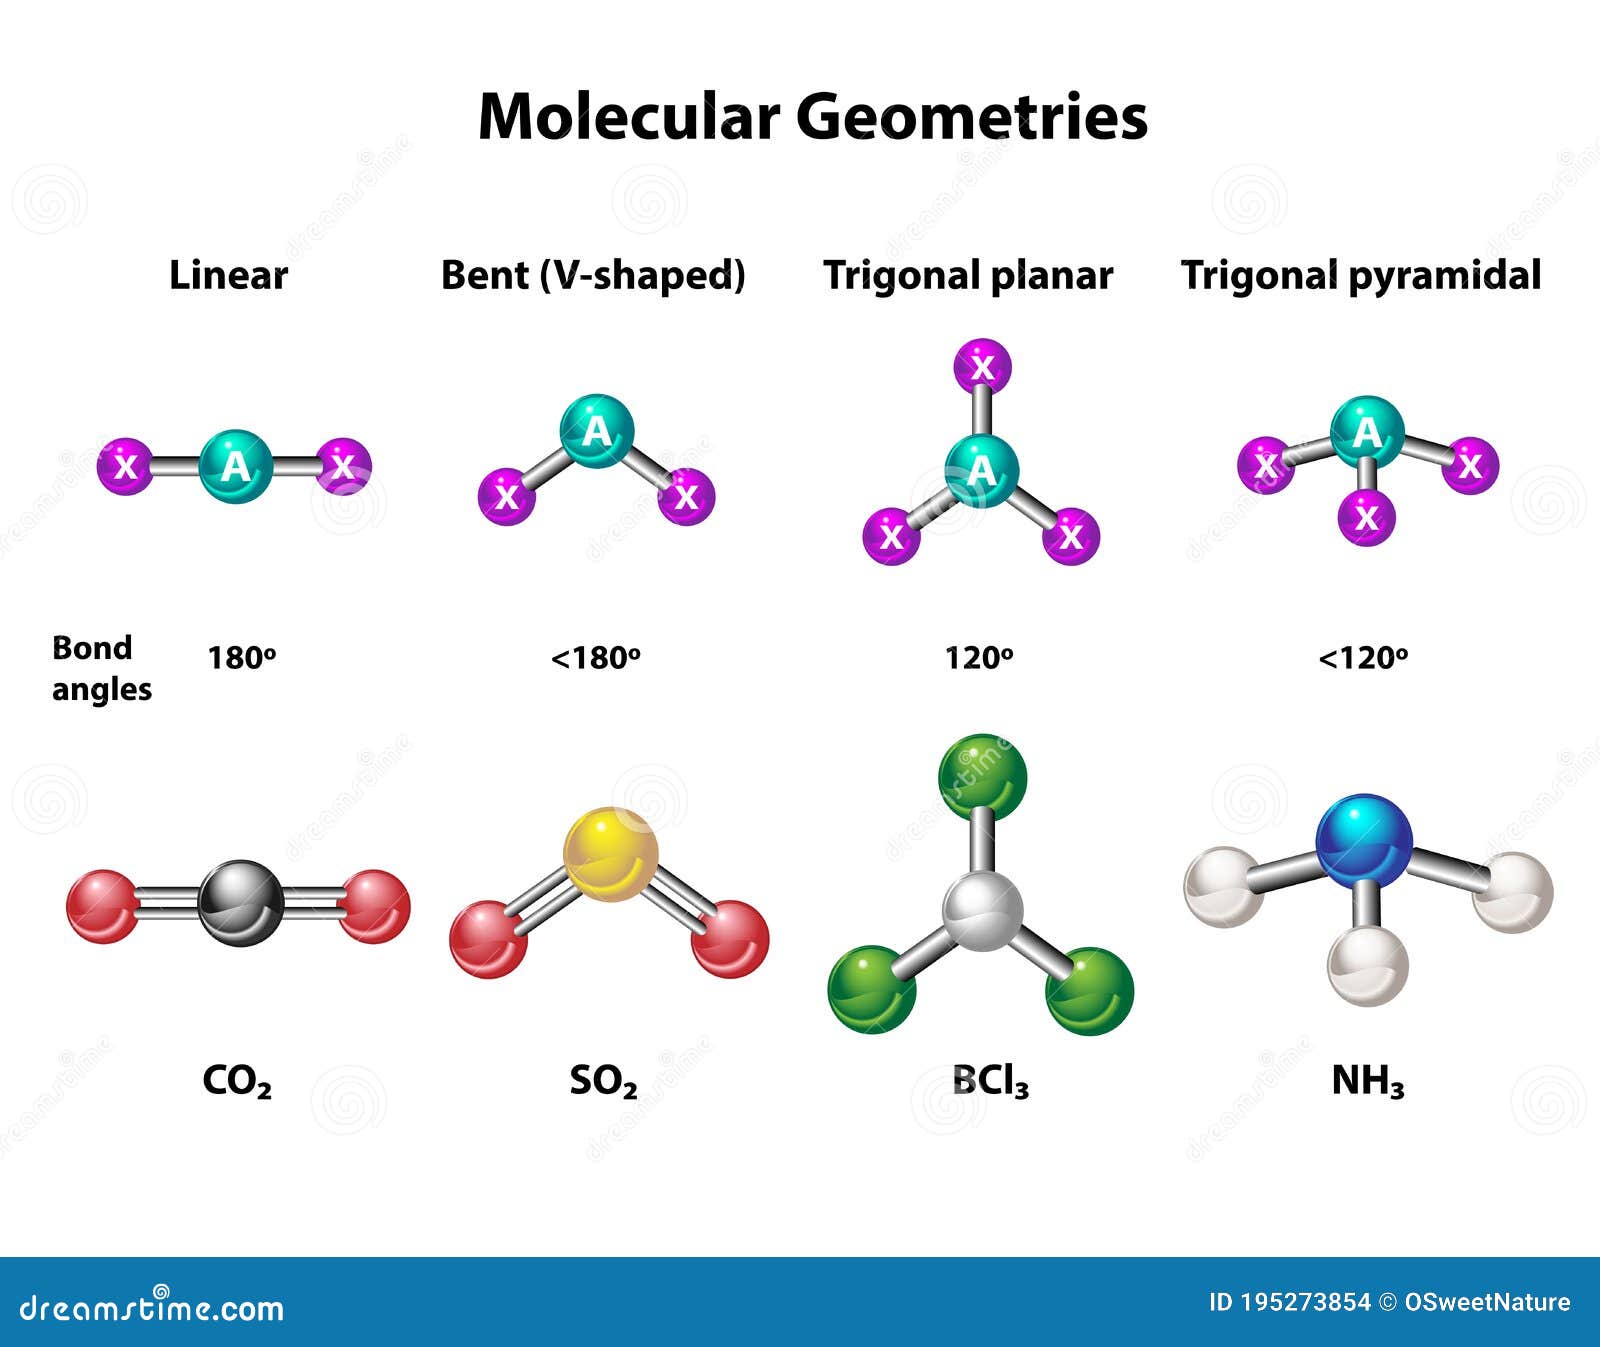
The first thing most people get wrong is assuming VSEPR theory is enough. It gets you through introductory chemistry, and then you hit a wall when you actually need to model something that doesn't fit a simple AXn pattern. I spent months struggling with transition metal complexes because I kept reaching for a model that was never designed for d-orbital complications. At its core, molecular geometry describes the three-dimensional arrangement of atoms around a central atom. You're mapping where every nucleus sits in space relative to every other nucleus. Bond angles, bond lengths, stereochemistry — it's all coordinates on a 3D grid. The reason this matters practically is that geometry dictates reactivity, polarity, and how molecules interact with enzymes, catalysts, and sensors. For small organic molecules, VSEPR gives you decent predictions. Water is bent at roughly 104.5 degrees. Methane is tetrahedral at 109.5. These are straightforward cases with minimal computational overhead. But as soon as you introduce lone pairs that distort angles, or atoms from the third period and beyond, the predictions start drifting from reality.
I learned this the hard way trying to model a sulfur hexafluoride analog for a research project. VSEPR says octahedral, perfect symmetry. The actual molecule has subtle distortions that VSEPR completely misses because it treats all electron domains as equivalent point charges. What you actually need is a quantum mechanical approach to get reliable geometry predictions beyond simple cases.
The Computational Approach
Most professionals use computational chemistry packages to determine molecular geometry accurately. The standard workflow involves building an initial structure, running an energy minimization, and then refining with increasingly sophisticated methods until the geometry converges. The tool I use regularly is Gaussian, though ORCA is more accessible if you don't have a licensed copy and need something free. Here's how a typical workflow goes: you start with a rough structure, either from crystallographic data or built manually in a program like Avogadro or GaussView. You specify a method and basis set, then the software iteratively adjusts atomic positions to find a local energy minimum. The output gives you bond lengths, angles, dihedrals, and a fully optimized 3D structure. For routine organic molecules, DFT with a basis set like B3LYP and 6-31G* gives results that usually match experimental crystal structures within 0.02 angstroms for bond lengths and a few degrees for angles. That's good enough for most purposes. If you need higher precision, switching to a triple-zeta basis set like 6-311+G(d,p) or def2-TZVP will tighten those numbers further.
Get the Full Details

The real problem comes with larger systems. A molecule with fifty or more atoms can take hours or even days on a standard workstation, depending on the method. I had a project where I needed the geometry of a coordination complex with a ligand framework containing about 80 heavy atoms. Running B3LYP with a decent basis set would have taken roughly two weeks on my setup. I switched to semi-empirical methods like PM7, which got the geometry in about 45 minutes, then used those coordinates as the starting point for a targeted DFT refinement that added another two hours. Total time dropped from two weeks to under three hours with acceptable accuracy.
Common Pitfalls
The biggest mistake I see is using an inappropriate initial geometry. Optimization algorithms find the nearest local minimum, not necessarily the global minimum. If you start with a strained conformation, the algorithm will relax it to some local minimum — which might be completely wrong for what you're studying. Always check multiple starting conformations, especially for flexible molecules. Another frequent issue is ignoring solvent effects. Gas-phase calculations often produce geometries that differ noticeably from solution-phase structures. For charged species or molecules with strong dipoles, this difference can be significant. Using an implicit solvent model like SMD or PCM usually adds maybe 20 to 30 percent to computation time but brings your results much closer to experimental conditions. I once submitted optimized geometries for a paper without checking for imaginary frequencies. The reviewer caught it. Two of my structures had imaginary frequencies, meaning they weren't true minima — they were transition states or saddle points. I wasted three days going back, adjusting the constraints, and reoptimizing. Always run a frequency calculation after geometry optimization. If there are no imaginary frequencies, you have a valid minimum. If there's exactly one, you might have a transition state. If there are two or more, your structure hasn't converged properly.
When Geometry Calculations Fail
Not every molecule cooperates. Open-shell systems with unpaired electrons, transition metals with near-degenerate states, and highly excited electronic configurations can cause convergence failures that no amount of tweaking parameters will reliably fix. Standard DFT functionals struggle with these cases, and even high-level wavefunction methods like CCSD(T) can become impractical for anything beyond small systems. For transition metal complexes specifically, the choice of functional matters enormously. B3LYP sometimes over-stabilizes certain spin states relative to others, leading to incorrect ground-state geometries. I've found that wB97X-D or M06-2X tend to perform better for organometallic systems, though no single functional works universally. If accuracy is critical, benchmarking against known experimental structures of similar complexes is worth the effort. There's also the matter of dispersion forces. Standard DFT functionals without empirical dispersion corrections can give poor geometries for systems where weak intermolecular interactions matter — large aromatic systems, supramolecular assemblies, proteins. Adding a dispersion correction like Grimme's D3 or D4 usually takes negligible extra computational cost and improves results noticeably for these cases.

If you're working with very large biomolecules where full quantum mechanical treatment is impossible, molecular mechanics force fields like AMBER or CHARMM are the fallback. They're fast but approximate. Bond lengths and angles are parameterized for specific atom types, so they work well for proteins and nucleic acids but fall apart for unusual chemistries or non-standard residues. The trade-off is speed versus accuracy, and you need to know which side of that line your problem falls on. For download links and software, Gaussian requires a commercial license. ORCA is freely available for academic use at orca.nikitin.info. Avogadro is open source at avogadro.cc. These cover the main needs for geometry determination across most common applications.