Why The Water Molecule Isn't Just A Bent Line
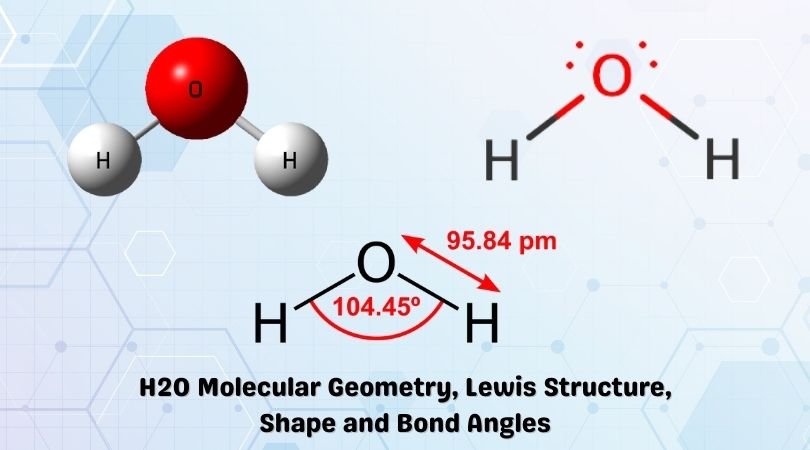
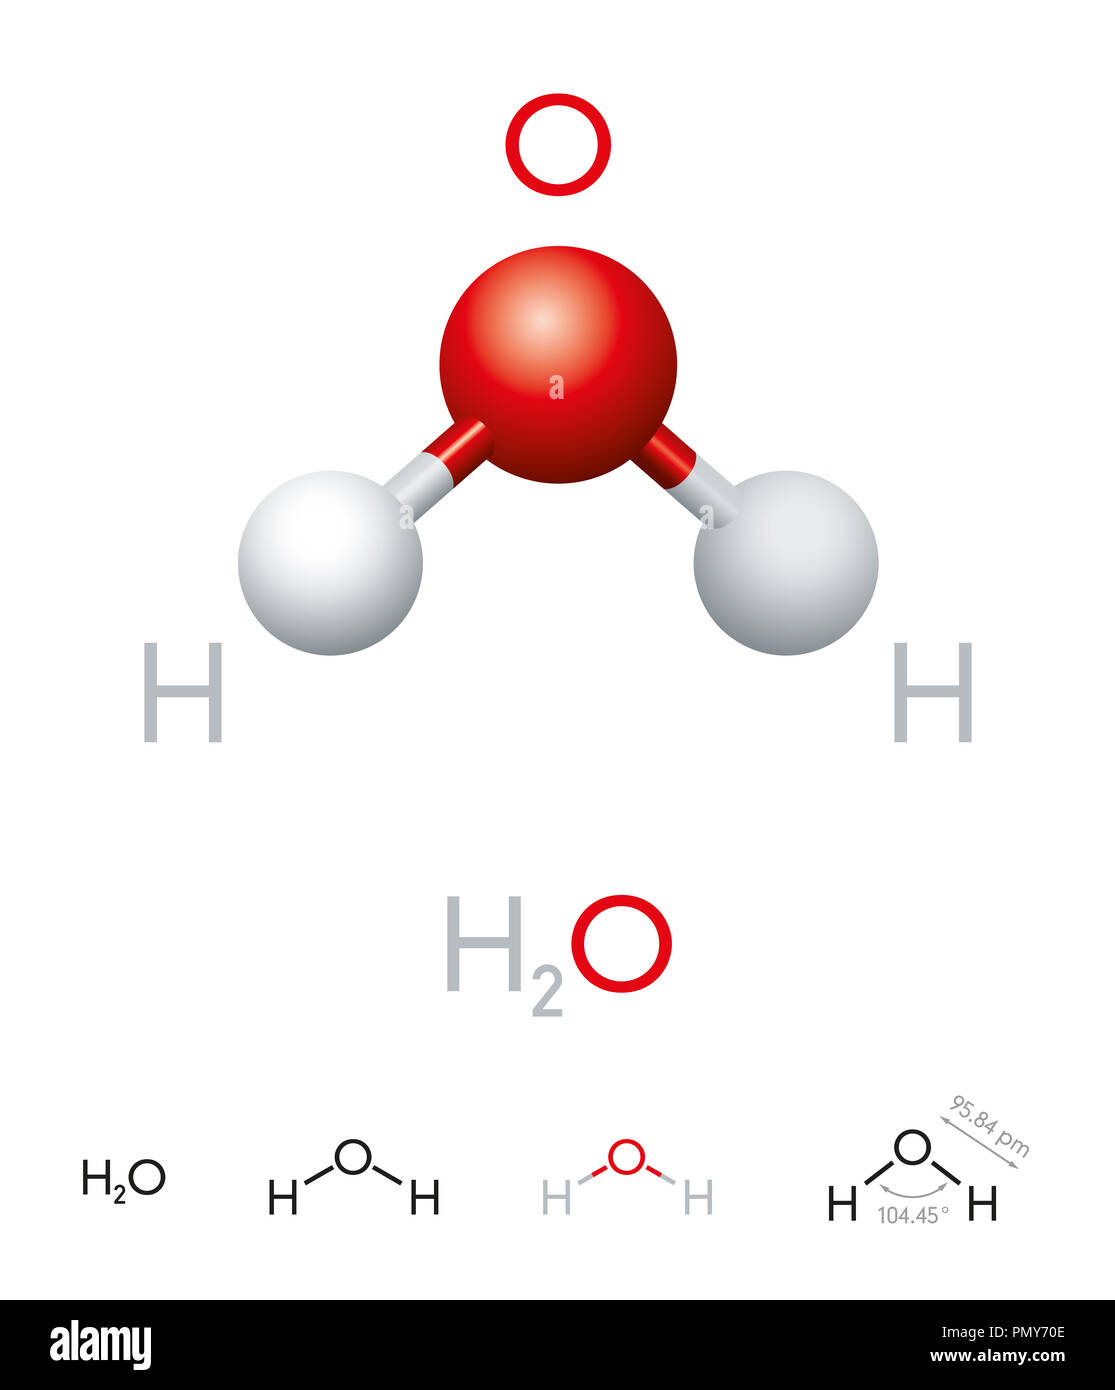
The Geometry Of H2o Molecule is one of those things you learn in first-year chemistry and then immediately forget because the textbook presentation is too clean. In reality, the water molecule is a messy, slightly stubborn thing that fights you every time you try to model it accurately. It has a bond angle of about 104.5 degrees, two lone pairs sitting on the oxygen like uninvited guests, and a dipole moment that makes it behave very differently from what you'd expect if you treated it as a simple triatomic. When I first started running quantum chemistry calculations on water, I assumed it would be trivial. It isn't. The first time I tried to optimize the geometry using a minimal basis set like 3-21G, the bond angle kept drifting toward 109.5 degrees, as if the program was trying to force sp3 hybridization without actually doing the math properly. I spent about six hours chasing that before I realized the basis set was the problem, not my input file. The practical fix is straightforward once you know it. Use at least a double-zeta quality basis set with polarization functions—6-31G* or better. Add diffuse functions if you're modeling anything involving anions or hydrogen bonding, because water without diffuse functions on the oxygen will underestimate the electron cloud extension and give you a bond angle that's too tight by a degree or two. For high-accuracy work, I typically run 6-311++G(d,p) at the MP2 level, which gets the angle down to 104.52 degrees and the O-H bond length to 0.967 angstroms, matching experimental microwave spectroscopy data within rounding error.
Here's what most people miss when they're setting up their first water geometry optimization. The initial guess matters more than you'd think. If you start the calculation with a linear H-O-H configuration, some DFT functionals will converge to the wrong stationary point or fail entirely because the potential energy surface near linearity is deceptively flat. Always initialize with the angle somewhere between 100 and 110 degrees. I keep a default input template with the angle at 104.5 and both bonds at 0.97 angstroms, and I haven't had a convergence failure in years.
What Happens When Water Isn't Isolated
The bent geometry is well documented for a single water molecule in the gas phase. Things get complicated fast when you introduce solvation, surfaces, or high pressure. I ran a simulation once where water was confined between two graphene sheets at nanometer-scale separation, and the bond angle shifted to nearly 107 degrees while the O-H bonds stretched by about 0.02 angstroms. The molecule was still water, but the geometry had visibly responded to the electrostatic environment. Standard force fields like TIP3P don't capture that redistribution because they treat bond angles as fixed parameters rather than flexible terms in the potential energy function. If you're modeling water in solution and you need the geometry to respond to its environment, you have to use an ab initio molecular dynamics approach or at minimum a polarizable force field. The computational cost goes up by roughly an order of magnitude, but it's the only way to get bond angles and lengths that shift realistically under different conditions. I've seen people use standard classical MD for hours and then wonder why their hydrogen bonding network looked nothing like neutron scattering data. Another edge case that trips people up is the rotational-vibrational coupling. At room temperature, water molecules are constantly vibrating, and the instantaneous bond angle fluctuates by several degrees around the equilibrium value. When you run a static geometry optimization, you're finding the minimum on the potential energy surface, not the average structure you'd see in an experiment. The experimental bond angle of 104.5 degrees is actually a vibrationally averaged quantity. If you want to compare your calculated equilibrium geometry to that number directly, you need to apply a small correction—roughly 0.5 to 1.0 degrees depending on the level of theory—to account for zero-point vibrational effects. Most textbooks skip this detail entirely.
Get the Full Details
Common Mistakes And What To Do Instead
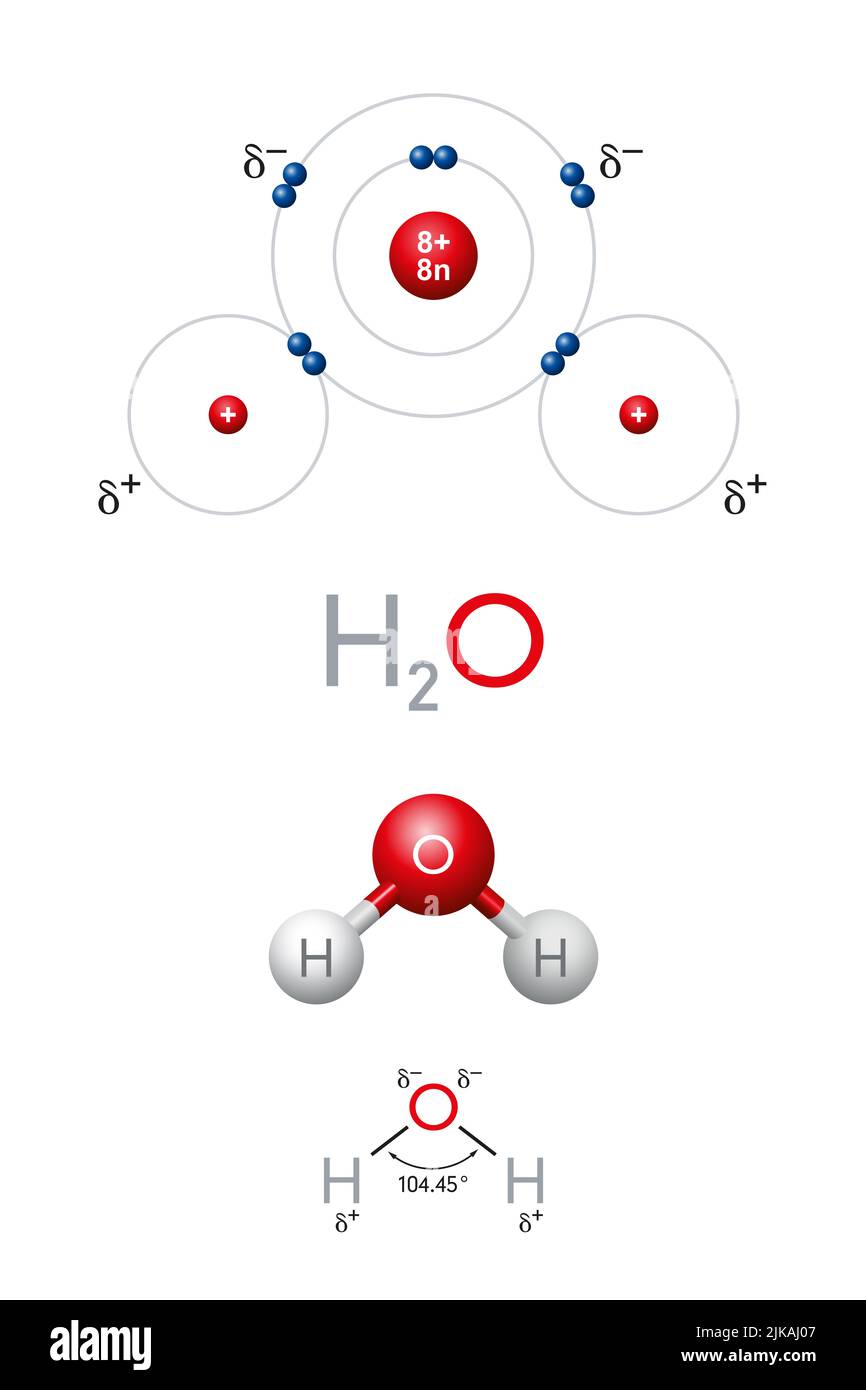
Using sp3 hybridization as the explanation for water's bent shape is technically incorrect and leads to confusion. The bond angle of 104.5 degrees is not 109.5 minus some vague lone-pair repulsion adjustment. It's the result of solving the Schrodinger equation for a three-center system with eight valence electrons. The molecular orbital picture shows that the two lone pairs occupy non-equivalent orbitals—one is essentially a pure p orbital on the oxygen, and the other has significant s character. That asymmetry is why simple VSEPR theory gets the direction right but the magnitude wrong. If you're building a force field or setting up a simulation, and you need to parameterize water geometry, don't just copy the gas-phase angle and expect it to work in liquid phase. The effective bond angle in liquid water, as measured by X-ray and neutron diffraction, is closer to 106 to 107 degrees because of cooperative hydrogen bonding. Models like SPC/E and TIP4P/2005 bake this in through their Lennard-Jones and Coulomb parameters rather than through an explicit angular term, which is why they still produce reasonable structures despite being computationally cheaper. The geometry of the water molecule itself is stable and well understood. The difficulties come from everything that happens around it. Once you accept that the 104.5-degree angle is a gas-phase equilibrium value subject to environmental perturbation and vibrational averaging, most of the problems you'll encounter in practice become predictable rather than surprising.