Working With Gibbs Free Energy in Real Lab Conditions
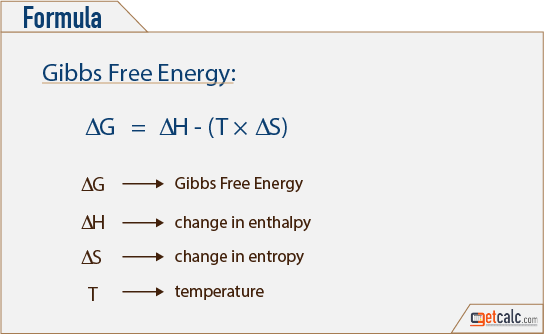
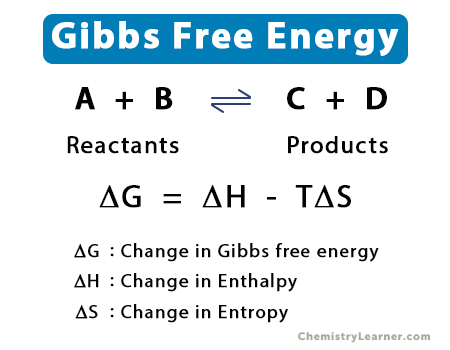
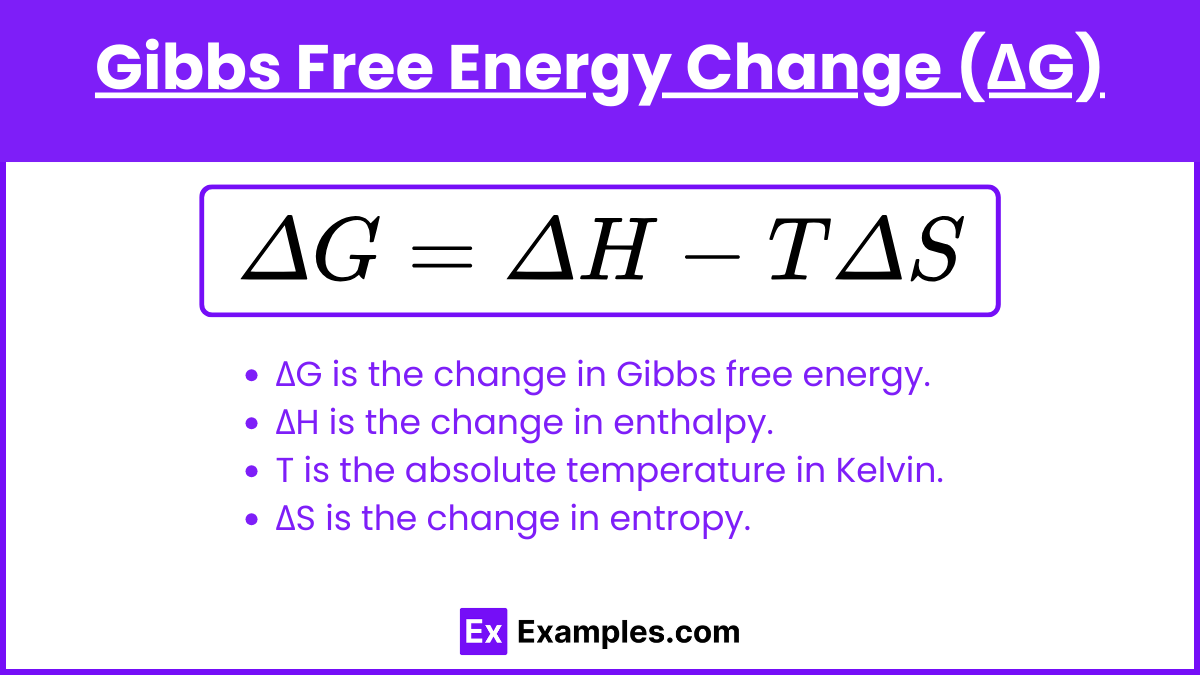
The Gibbs Free Energy Equation is G = H - TS, where G is Gibbs free energy, H is enthalpy, T is temperature in kelvin, and S is entropy. It tells you whether a process will happen spontaneously at constant temperature and pressure. That's the textbook definition. The reality of using it is messier than that. I spent three years running reaction optimization in a pharma lab, and the first time I actually used the Gibbs Free Energy Equation by hand for a real project, I nearly cost the company two weeks of batch time. The problem wasn't the equation itself. It was the data around it.
Why the Gibbs Free Energy Equation Fails Before You Even Start
Beginners treat G as a fixed number. It isn't. It changes with temperature, pressure, concentration, and solvent. The standard Gibbs Free Energy Equation assumes everything is at standard state — 1 M concentrations, 1 atm pressure, pure substances. Biological systems and industrial reactors rarely respect those conditions. Here's a specific example from my experience. We were evaluating an esterification reaction for a drug intermediate. The literature value for H was negative, meaning exothermic, and S was slightly positive. At room temperature, the Gibbs Free Energy Equation gave a clearly negative G. The reaction should proceed spontaneously. It didn't. Not really. The issue was water accumulation. The esterification produces water as a byproduct, and as water built up in the reaction mixture, the equilibrium shifted backward. The standard G didn't account for the actual reaction quotient Q. I should have been using G = G° + RT ln Q the entire time. Instead, I was looking at a textbook number and trusting it. That cost us about 11 days before someone caught it. Now I calculate Q before I calculate anything else.
Temperature Dependence Is Where People Get Tripped Up
The Gibbs Free Energy Equation shows that G decreases as T increases if S is positive, and increases as T increases if S is negative. This sounds straightforward until you try to apply it across a wide temperature range. The assumption that H and S are constant with temperature breaks down fairly quickly. In practice, they shift because heat capacities of reactants and products differ. If you need accuracy over more than a 50-degree range, you have to integrate the heat capacity term. The full expression becomes: G(T) = H(T_ref) - TS(T_ref) + [Cp dT] - T[(Cp/T) dT]
Get the Full Details

This is where most people stop doing the math and just plug in room-temperature values regardless of their actual operating temperature. If your process runs at 350 K and your H and S are measured at 298 K, you're probably off by a meaningful margin. For rough screening it's fine. For something you're going to scale, it's negligence.
Entropy Changes Are Harder to Measure Than Enthalpy Changes
Differential scanning calorimetry gives you H pretty directly. Entropy is trickier. You usually have to integrate Cp/T over a temperature range, or extract it from equilibrium constant measurements at multiple temperatures using the van 't Hoff approach. Both methods introduce error. The van 't Hoff method assumes H is constant over your temperature range, which brings us back to the same heat capacity problem. A practical shortcut I learned the hard way: if you can measure the equilibrium constant at just two temperatures, you can estimate both H and S simultaneously. Two data points. Two unknowns. It's not as accurate as five or six points with a proper regression, but in early-stage work when you're trying to decide whether a reaction is even worth pursuing, it cuts the characterization time from several days to a couple of hours.
Phase Transitions Break the Standard Equation
At a phase transition — melting point, boiling point, solid-solid transition — G equals zero by definition. The Gibbs Free Energy Equation still works, but you have to account for the discontinuity in entropy. The entropy change at a phase transition is simply H_transition / T_transition. If you're modeling a system that crosses a phase boundary, you can't use a single set of H and S values across that boundary. You need separate parameters for each phase. I ran into this with a crystallization step where the solvent was supersaturated and the product was precipitating out. The aqueous phase G calculation looked favorable, but the solid phase formation wasn't captured because I was treating the system as homogeneous. Once I added the solubility product term and recalculated G for the precipitation step, the picture changed completely. The reaction I thought was spontaneous in solution was actually being driven by the removal of product from solution via precipitation. Different driving force, same result, wrong mental model.

When the Gibbs Free Energy Equation Gives a False Positive
A negative G means the reaction is thermodynamically favorable. It does not mean the reaction will happen at an observable rate. Kinetics and thermodynamics are separate questions. I've seen people dismiss a catalyst study because G was positive, only to later discover the reaction was actually occurring through a different mechanistic pathway that the simple Gibbs Free Energy Equation didn't capture. Or they push a reaction aggressively because G is very negative and get surprising side products because they never considered activation barriers. The converse is equally common. A reaction with positive G under standard conditions can proceed if you continuously remove a product. Le Chatelier's principle is essentially a kinetic trick on thermodynamic data. The Gibbs Free Energy Equation tells you where equilibrium lies, not how to manipulate it. That's an important distinction that separates people who use this equation correctly from people who use it incorrectly.
A Quick Reference for Common Mistakes
Mistake one: Using G° without adjusting for actual concentrations or partial pressures. Always calculate Q first. Mistake two: Treating H and S as temperature-independent over wide ranges. They aren't. Check your Cp values. Mistake three: Confusing G with reaction rate. A favorable G says nothing about how fast anything happens.
Mistake four: Applying standard-state values to non-aqueous or mixed solvent systems. Activity coefficients change everything in organic solvents. Mistake five: Ignoring pressure effects on condensed phases. For liquids and solids, pressure dependence is usually negligible below 100 atm. For gases, it matters a lot. Use fugacity instead of pressure if you're working above 10 atm.
What I Actually Do Before Running a Reaction
First, I look up or measure H and S at a known temperature. Second, I estimate the heat capacity difference and decide whether temperature correction matters for my range. Third, I calculate Q based on my actual starting concentrations and product expectations. Fourth, I compute G using the full equation, not just the standard value. Fifth, I check whether any phase boundaries or solubility limits could shift the equilibrium unexpectedly. This takes about 20 minutes for a straightforward system. For a complex multicomponent reaction, it can take longer. The alternative is running the reaction, watching it fail, and then spending three weeks figuring out why the thermodynamics didn't predict what actually happened. I prefer the 20-minute version.