Why Your Electrolyzer Isn't Hitting Theoretical Efficiency
Most people treat the 1.23 V number like it's a target. It isn't. It's a lower bound that exists in idealized textbook conditions and doesn't account for anything happening inside an actual cell. When I first started working with proton exchange membrane (PEM) electrolyzers, I was frustrated that my lab setup required nearly double that voltage at reasonable current densities. The gap between 1.23 V and what actually showed up on the power supply wasn't a measurement error. It was the overpotential doing exactly what it's supposed to do, which is waste energy as heat. The thermoneutral voltage sits at about 1.48 V, and that's where the story gets interesting. Below 1.48 V, the reaction is endothermic and draws heat from the surroundings. Above 1.48 V, you're dumping excess energy into the system as waste heat. At 1.8 V, which is a common operating point for commercial PEM stacks, the electrical efficiency based on the higher heating value drops to somewhere around 65 percent. That's not a flaw in the design. That's thermodynamics writing the check that kinetics forces you to cash.
Gibbs Free Energy For Water Electrolysis
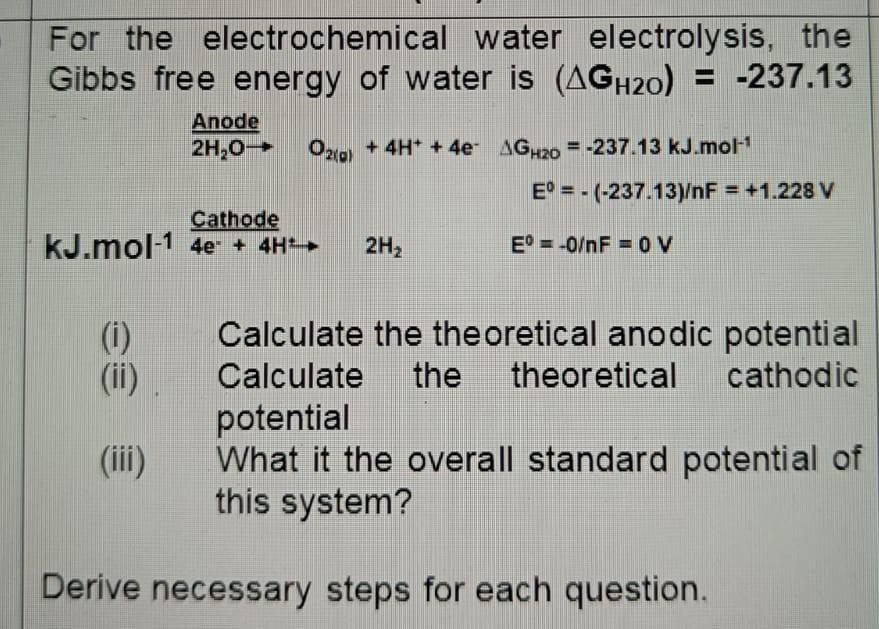
The core equation is straightforward. G equals negative nF times E, where n is 2 for the overall water splitting reaction, F is Faraday's constant at 96,485 coulombs per mole, and E is the reversible cell potential. At standard conditions, that gives you roughly 237.1 kilojoules per mole of hydrogen produced. Divide by the charge passed, and you get 1.23 volts. The enthalpy change H is 285.8 kilojoules per mole, which corresponds to 1.48 volts. The difference, TS, is about 48.7 kJ/mol and represents the thermal energy term that either needs to be supplied or gets released depending on your operating voltage. Here's what most guides skip: the Gibbs free energy changes significantly with temperature. At 80 degrees Celsius, G drops to approximately 228 kJ/mol, which means the theoretical minimum voltage falls to about 1.18 V. This isn't a minor correction. Running a solid oxide electrolyzer at 800 degrees C pushes the reversible voltage down to roughly 0.95 V because you're harvesting thermal energy from the high-temperature reservoir instead of supplying it all as electricity. That's why high-temperature electrolysis looks so attractive on paper. The catch is materials degradation at those temperatures, and the balance of plant complexity that comes with managing thermal cycling in a stack that's also handling steam at elevated pressures. I learned this the hard way during a project where we were comparing PEM and alkaline cells side by side at different temperatures. I had calculated the expected voltage using the standard G value at 25°C and then tried to apply it directly to our 60°C alkaline operation. The numbers were off by nearly 40 millivolts, which sounds small until you're trying to optimize stack efficiency at the margin. The workaround was straightforward but tedious: I pulled the temperature-dependent Gibbs energy values from NIST thermodynamic tables and recalculated the reversible potential for each operating point. That corrected the baseline by a meaningful amount. More importantly, it reminded me that tabulating standard values at 298 K and forgetting about temperature dependence is one of the most common mistakes in this space.
The Nernst equation also matters here, especially when you're not running at 1 atmosphere of pressure. Pressurizing the product gases shifts the reversible potential upward. At 30 bar, you're looking at roughly a 50 mV increase in the theoretical voltage requirement. That's because you're doing extra work compressing the products against pressure. Some people see pressurized electrolysis and immediately think it's a free lunch because you skip the downstream compression step. The cell voltage penalty is real, though it's often smaller than the system-level savings from avoiding a separate compressor. Whether it's worth it depends entirely on your application and the rest of your process integration. Another thing that trips people up is the difference between the thermodynamic limit and the actual decomposition voltage you measure. A fresh PEM cell at open circuit will sit near 1.23 V, but the moment you draw current, the voltage jumps. The anode oxygen evolution reaction is the main culprit. It has sluggish kinetics on most catalysts, and the overpotential scales roughly logarithmically with current density. At 1 A/cm², you might be adding 300 to 400 mV of overpotential at the anode alone. The cathode hydrogen evolution reaction is comparatively easy, usually contributing less than 50 mV at the same current density. That asymmetry is why anode catalyst development dominates the research literature. Kinetic overpotentials aren't the only loss mechanism. Ohmic resistance through the membrane and the electrode layers adds another voltage penalty that scales linearly with current. In a well-designed PEM cell, membrane resistance might contribute 100 to 150 mV at 1 A/cm². Contact resistances at the bipolar plate interfaces and gas diffusion layer boundaries are smaller but not negligible, especially as cells age and compression changes over time. Concentration overpotentials at high current densities are the last piece, and they tend to show up as sharp voltage increases near the limiting current region where mass transport can't keep up with the reaction rate.
Get the Full Details

One practical detail that matters more than people realize: the state of water in the reaction equation changes everything. The standard G value of 237.1 kJ/mol assumes liquid water as the reactant. If you're running high-temperature steam electrolysis, the reactant is gaseous water, and the thermodynamics shift accordingly. Using the gas-phase value changes the reversible voltage calculation enough that plugging the liquid-phase number into a steam electrolyzer model will give you systematically wrong results. It sounds obvious, but I've seen it happen in peer-reviewed work more often than I'd like to admit. The bottom line for anyone building or modeling an electrolysis system is that Gibbs free energy gives you the thermodynamic floor, not the operating point. Your actual cell voltage will always be higher, and the gap between theory and practice is where engineering decisions get made. If you're designing a system, start with the correct thermodynamic values for your actual operating conditions, then budget realistically for overpotentials and ohmic losses. Assuming you can operate near 1.23 V is the fastest way to produce a design that doesn't work in the lab.