What GMP Actually Looks Like on the Floor
Most people think Good Manufacturing Practice is a checklist you tick off before an inspector walks in. That is not how it works. It is a living documentation system that covers everything from raw material intake to final product release, and the moment you treat it as paperwork instead of a workflow, things fall apart quickly. I spent years dealing with GMP in pharmaceutical and food production environments, and the harsh reality is that the audits themselves are rarely the problem. The problem is what happens three months after the auditor leaves when everyone goes back to their old habits because the documentation burden felt arbitrary and imposed from outside.
The Real Mechanics of Good Manufacturing Practice
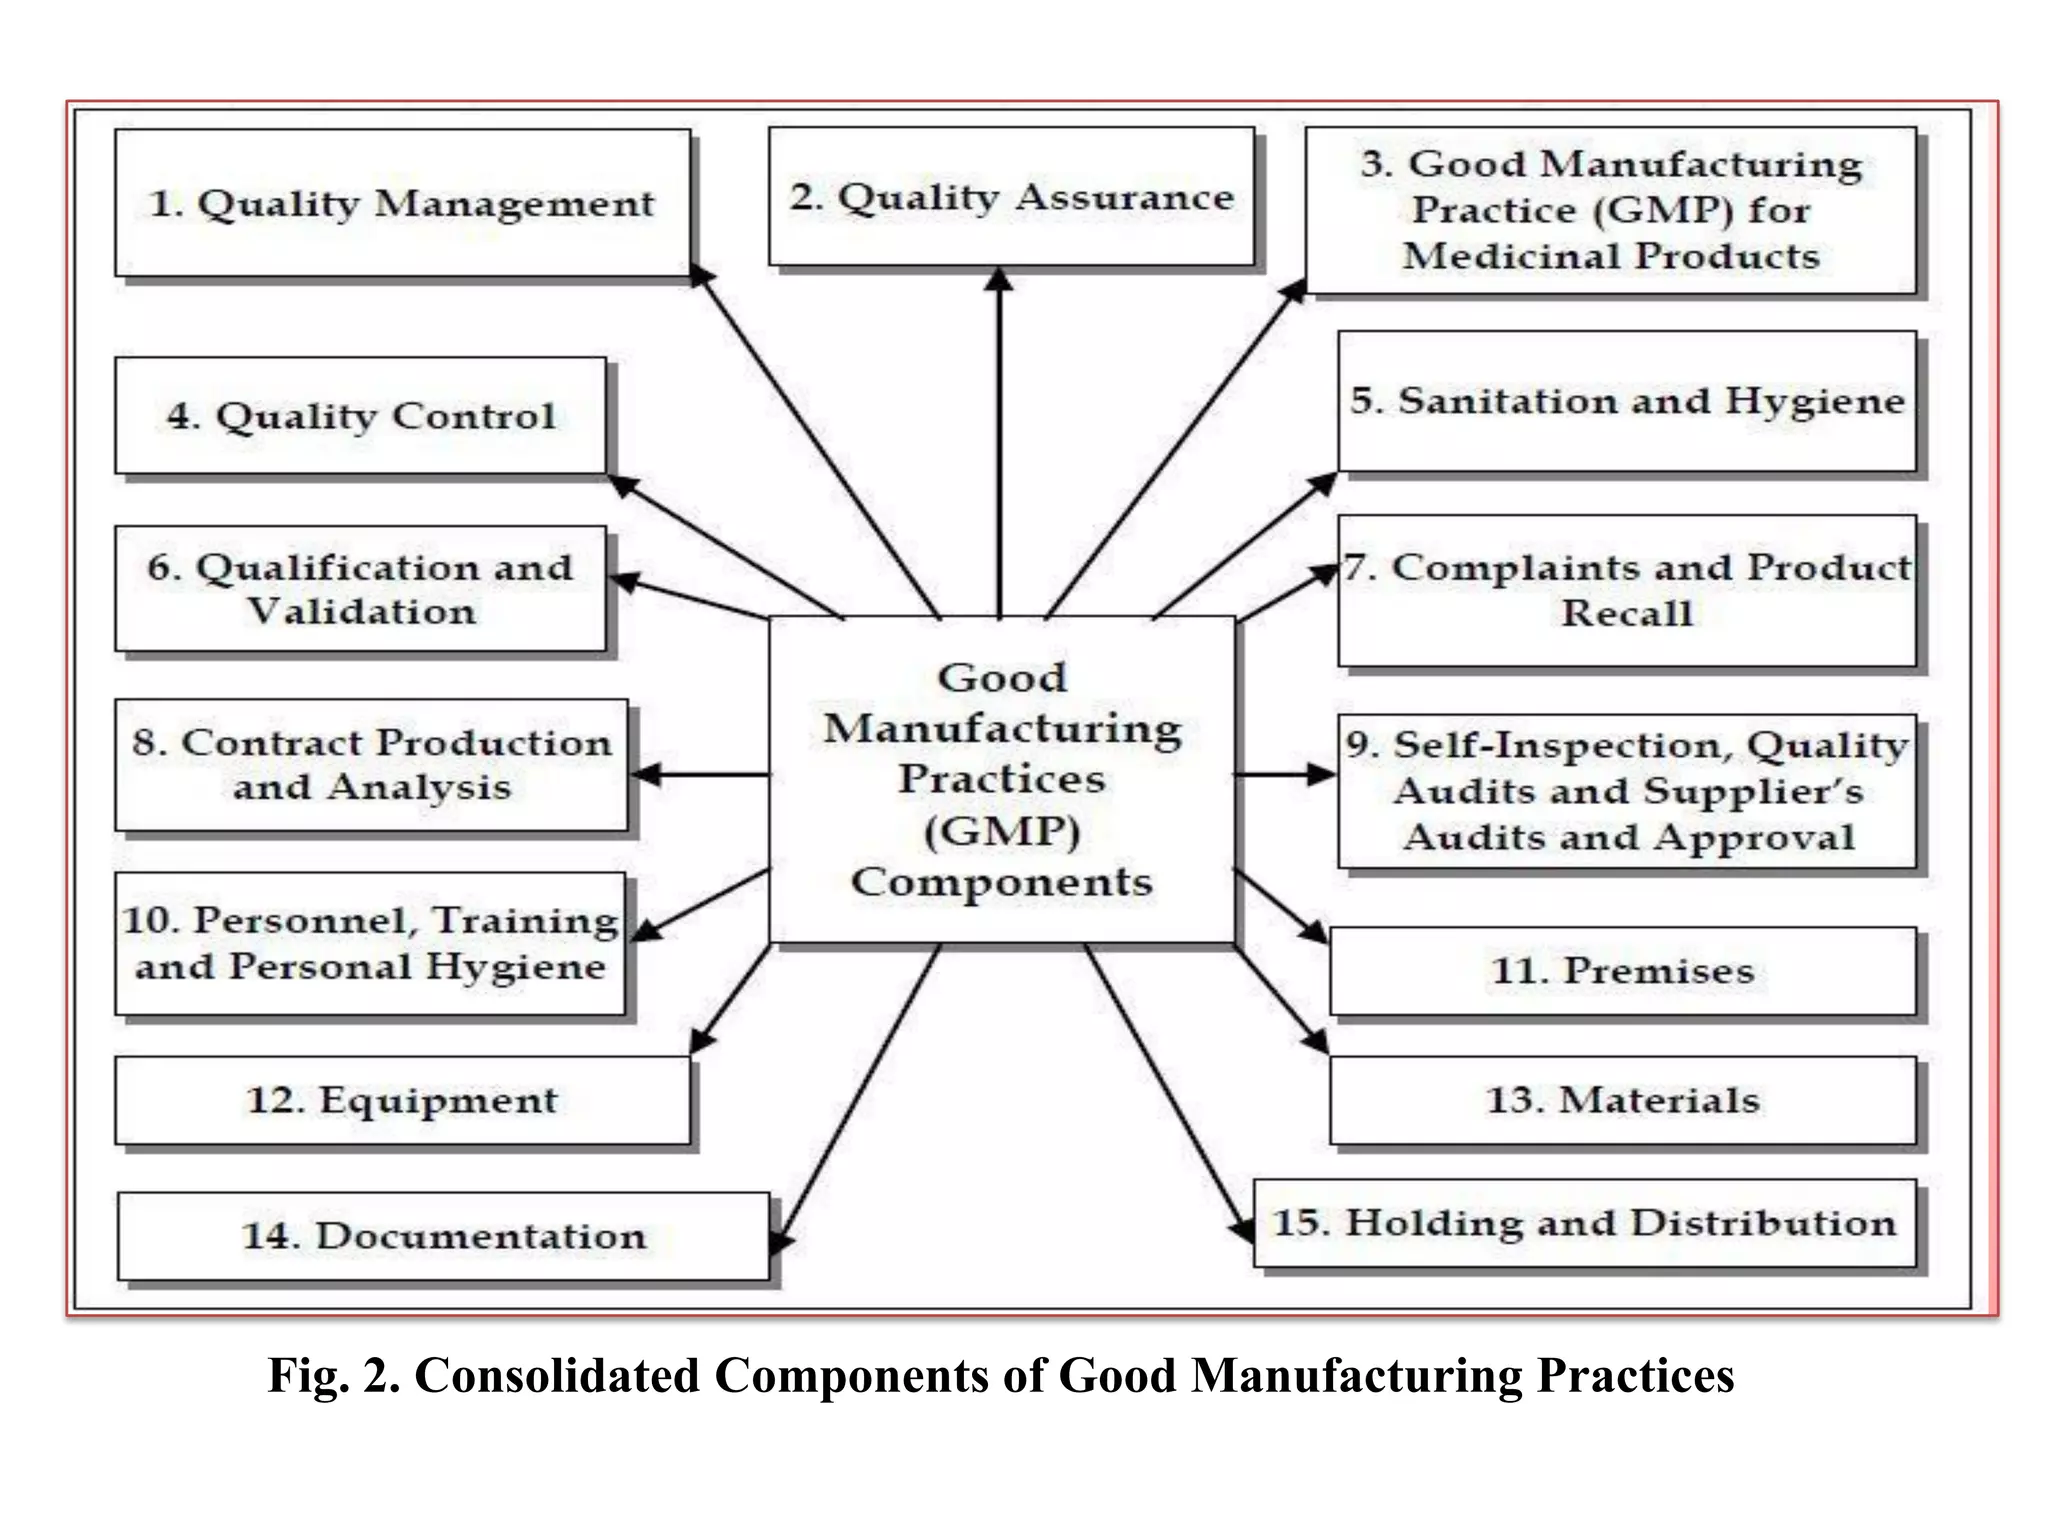
GMP is built on a few core pillars that every quality professional deals with daily: preventing contamination, ensuring consistent quality, maintaining full traceability, and validating that every process actually does what it claims to do. The regulations come from bodies like the FDA, EMA, and WHO, but they are not interchangeable. A process that satisfies FDA 21 CFR Part 210 and 211 will not automatically satisfy EU GMP Annexes without mapping the gaps. Here is something most beginners miss. Cross-contamination control is not primarily about physical barriers. It is about airflow management and the sequence of operations. I once had a facility where we were failing particle count tests in a Grade C cleanroom, and everyone assumed it was the HEPA filter integrity. After two weeks of investigation, we found the real culprit was the personnel door opening sequence. The air was being pulled backward through the gowning area because the pressure differential chart on the wall didn't match what the sensors were actually reporting. Fixing the calibration schedule dropped our rejection rate from about 12% to under 2% within a month. Documentation under GMP follows a simple principle but gets extremely complicated in practice. You write the procedure, you follow the procedure, and you record that you followed the procedure. Any deviation requires a documented explanation and a risk assessment. This is not bureaucracy for its own sake. When a batch fails downstream and you need to determine whether it was a raw material issue, a process error, or equipment contamination, your documentation is the only thing standing between a full recall and a targeted corrective action.
Key Areas That Actually Matter
Personnel and training. Every person who enters a controlled manufacturing environment needs documented training for their specific role. Not generic safety training, but role-specific procedures that they have demonstrated competence in. I have seen companies pass audits with minimal effort because their training records showed signatures but no evidence of competency assessment. That changes nothing about whether the operator actually knew what to do during a real deviation. Facilities and equipment. Your building design, HVAC system, and cleaning protocols are all part of GMP. Equipment qualification follows a strict sequence: Installation Qualification, Operational Qualification, and Performance Qualification. Skipping the IQ because you are confident the vendor installed it correctly is one of the most common and most expensive mistakes I have seen. An unqualified piece of equipment can invalidate an entire production batch if an audit uncovers that gap. Raw material control. Every incoming material needs a verified supply chain, proper identification testing, and storage conditions that maintain its specifications throughout its shelf life. I once dealt with a situation where a co-manufacturer swapped a lubricant grade on a filling line without updating the documentation. The change was functionally minor but violated the approved Bill of Materials. The product was already in distribution before we caught it during a routine material reconciliation. We had to pull three batches retroactively at a cost of over 200,000 euros.
Get the Full Details
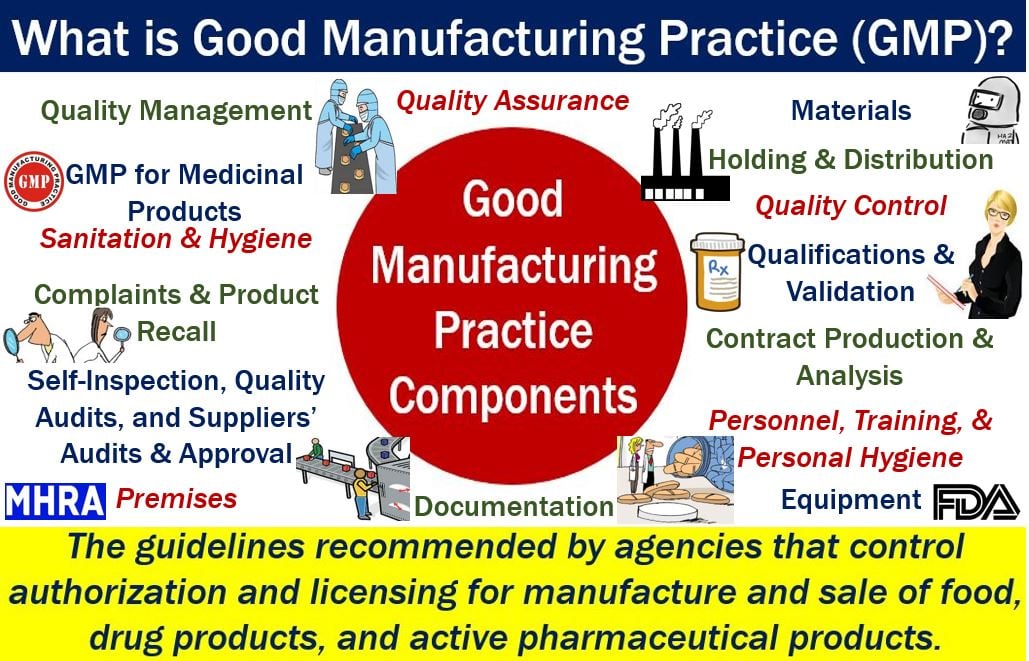
Validation and quality control. Process validation proves that your method consistently produces a product meeting its specifications. Analytical method validation proves your tests actually measure what they claim to measure. Stability testing proves the product remains acceptable throughout its labeled shelf life. These are not optional exercises. They are legal requirements, and the data must be generated, reviewed, and retained exactly as the regulations specify.
Common Pitfalls That Break Compliant Operations
The biggest blind spot I see is the assumption that deviation handling is about blame. It is not. It is about understanding cause and preventing recurrence. When operators hide deviations because they fear punishment, the quality system loses its primary learning mechanism. I implemented a no-blame deviation reporting culture in one facility and saw our documented deviations triple in the first quarter. That sounded terrible until we analyzed the data and realized we were catching problems we would have otherwise shipped out undetected. Another pitfall is the document revision dance. Every change to a GMP document should go through a controlled revision process with version history, effective dates, and supersession of old versions. I have seen multiple sites where the current procedure and the archived version were the same document sitting in different folders with no version control. An inspector will spot that in five minutes. Data integrity is the current hot topic across all GMP-regulated industries, and for good reason. ALCOA+ principles — Attributable, Legible, Contemporaneous, Original, Accurate plus Complete, Consistent, Enduring, and Available — are the standard framework. But the practical challenge is that most legacy systems were not designed with these principles in mind. Migrating paper-based records into electronic systems creates validation burdens that many companies underestimate. A properly validated electronic system with audit trails, access controls, and backup procedures usually costs between 50,000 and 200,000 euros depending on scope, and it takes six to eighteen months from project start to go-live.
What GMP Does Not Solve
Good Manufacturing Practice will not make your product better. It will make it consistently the same as what you intended to make. If your formulation is flawed or your process design is unsound, GMP compliance will only ensure that every batch is equally flawed. The system prevents variability and contamination, but it does not replace sound product development and process engineering. There are also contexts where GMP frameworks create real bottlenecks. For small-scale manufacturers and biotech startups, the documentation and validation overhead can consume more resources than the actual production work. I know several companies that found it more efficient to use CDMOs for regulated manufacturing while keeping their own operations leaner and less formally structured. That trade-off is real and worth evaluating honestly. Regulatory expectations also evolve. The FDA has shifted toward more risk-based inspection models, and the EU is increasingly focusing on data integrity during inspections. What passed review in 2019 might not pass in 2026. Staying compliant is not a one-time project. It is a continuous cycle of monitoring, training, and adaptation.
Practical Steps to Strengthen Your GMP System
Start by mapping your current quality system against the specific regulations that apply to your product category and target markets. The gaps will tell you where to focus. Then prioritize the documentation that has the highest impact on product quality and patient safety, not just the paperwork that sounds most impressive during an audit. Invest in competency-based training programs rather than completion-based ones. Run mock audits quarterly, not just when you suspect an official one is coming. Treat deviations as data points, not failures. And build your change control process so that any modification, no matter how small, flows through a documented review that considers quality, regulatory, and operational impact. The companies that get it right treat GMP as an operating system, not an inspection checklist. It is tedious, it is expensive, and it will frustrate you constantly. But it is also the difference between a product that stays on the market and one that ends up as a case study in regulatory enforcement.