How the Henderson-Hasselbalch Equation Actually Works in the Lab
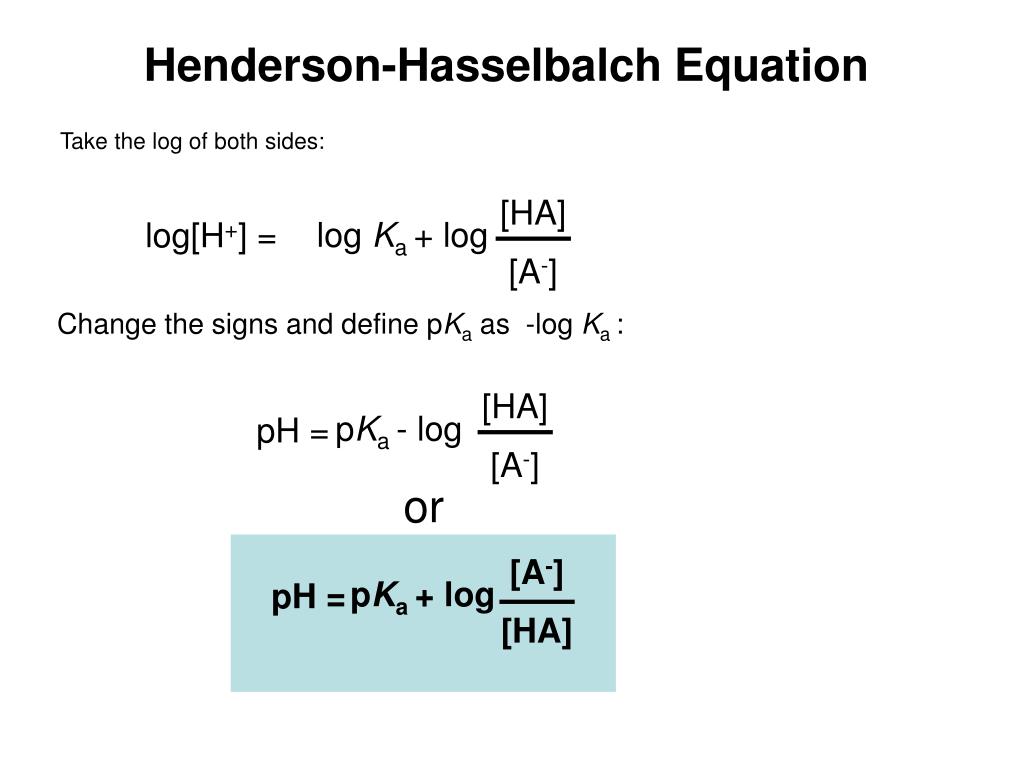
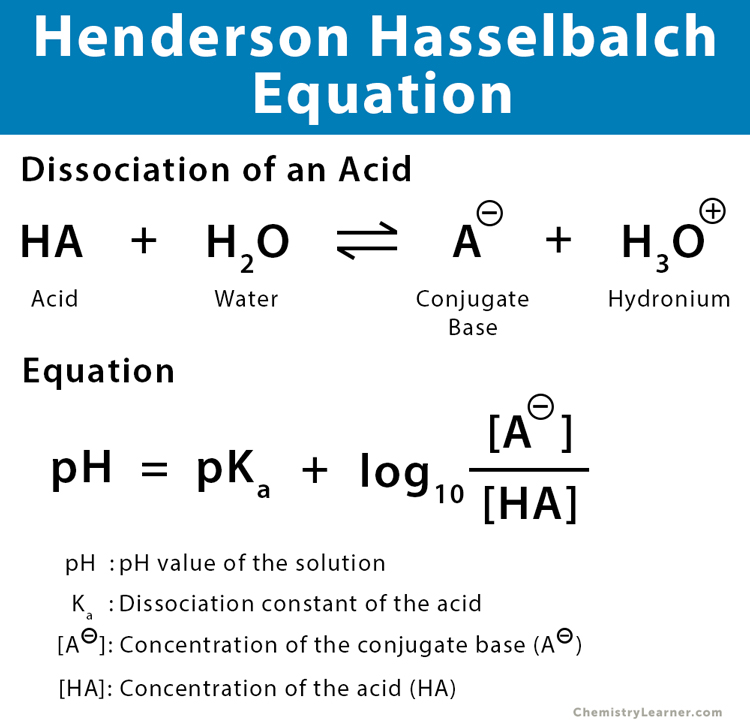
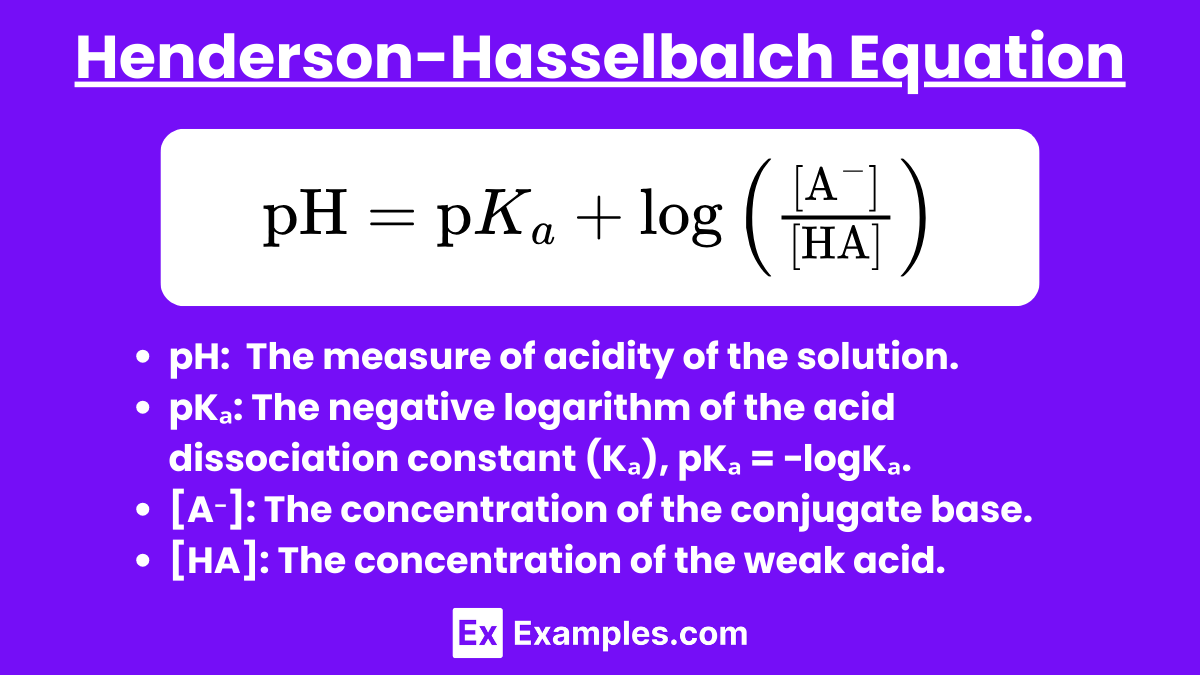
Most people encounter the Handerson And Hasselbalch Equation in their first biochemistry or analytical chemistry class and never really understand what it's doing for them until they're standing over a beaker at 11pm trying to make a buffer that doesn't drift two pH units by morning. The equation itself is straightforward enough: pH = pKa + log([A-]/[HA]). That's it. It relates the pH of a solution to the pKa of the weak acid and the ratio of conjugate base to the undissociated acid. What the equation doesn't tell you is why your calculated buffer is 0.3 pH units off when you measure it, or why the approximation breaks down completely when you're working in the millimolar range instead of the half-molar range you practiced with in lecture. I've spent years fixing buffer problems that should have been impossible according to the textbook version of this equation, so here's what actually happens.
Where the Handerson And Hasselbalch Equation Comes From
The equation is just the Ka expression rearranged and put into log form. You start with the acid dissociation equilibrium: HA H+ + A-. The equilibrium constant is Ka = [H+][A-]/[HA]. Take the negative log of both sides, rearrange, and you get pH = pKa + log([A-]/[HA]). It assumes activity coefficients are unity, which means concentrations approximate activities. That assumption holds reasonably well in dilute solutions but falls apart fast as ionic strength climbs. For practical purposes, the equation is most useful when you're designing a buffer. Pick your weak acid, look up its pKa at your working temperature, decide what pH you need, and solve for the ratio [A-]/[HA]. Then figure out how much of each component to weigh or mix. If you need pH 7.4 and you're using phosphate, pKa2 of phosphoric acid is about 7.2, so you'd want roughly a 1.6-to-1 ratio of HPO4^2- to H2PO4-. Easy math. The hard part starts when you actually make it.
Practical Application and Common Pitfalls
The biggest mistake I see people make is treating the equation as exact when it's an approximation. The Henderson-Hasselbalch equation ignores the autoionization of water and the contribution of H+ from the acid itself when the concentration is very low. At 1 mM total phosphate, calculating pH from the equation gives you something different from what you'd get solving the full equilibrium system. The discrepancy matters when you need precision. I once had a client who was running an enzyme assay that required pH 6.5 ± 0.05 in a 2 mM citrate buffer. They were calculating everything by Handerson And Hasselbalch Equation and getting results that drifted during the run. The problem wasn't the equation itself. It was that citric acid has three pKa values close together, and the simple two-species approximation the equation relies on breaks down when you have overlapping equilibria. What actually stabilized was switching to the full polynomial approach for polyprotic systems. I wrote a small spreadsheet that solved the complete charge balance and mass balance equations simultaneously. Buffer held stable for hours instead of drifting continuously. Another issue people run into is temperature dependence. The pKa values you look up are typically at 25°C. If your reaction runs at 37°C or room temperature varies between 20 and 25, your pKa shifts. For phosphate, the pKa2 drops about 0.008 units per degree Celsius increase. That's small but measurable when you're working at the edge of your tolerance. I always check the pKa at my actual working temperature, not the stockroom value at standard conditions.
Get the Full Details
The equation also assumes you know the exact concentration of both the acid and conjugate base forms. In practice, you're often starting from a solid acid and adding NaOH to generate the conjugate base in situ. That means you need to account for the volume change from the titrant, the purity of your reagents, and the fact that your NaOH solution has probably absorbed some CO2 from the air and its exact concentration has drifted. I standardize my NaOH against potassium hydrogen phthalate before making any critical buffers. Takes ten minutes and saves hours of troubleshooting.
When the Equation Fails Completely
There are situations where Henderson-Hasselbalch gives you garbage and you need to switch approaches entirely. One is concentrated solutions above about 0.1 M ionic strength, where activity coefficients deviate significantly from one. You can apply the Debye-Hückel or Davies equation to correct for this, but then you've left the simple equation behind anyway. Another is when you're dealing with very dilute buffers below 1 mM, where water's autoionization contributes meaningfully to the proton balance. And polyprotic acids with clustered pKa values, like the citrate example, are just not well served by the two-species model. For those cases, the full equilibrium calculation is the right tool. You set up mass balance equations for each species, the charge balance equation, and solve the resulting polynomial. It's more work upfront but it gives you the right answer where Henderson-Hasselbalch doesn't. I keep a script for this now and only reach for the simple equation when I'm in the comfortable range: moderate concentration, monoprotic or well-separated pKa values, and ionic strength below 0.1 M. The equation is a useful heuristic and a fine starting point for buffer design, but it's not a magic formula that gives you the pH no matter what. Treat it like an approximation with a known range of validity, stay aware of its assumptions, and have the fuller calculation ready when you step outside that range. That's how you stop wasting time chasing phantom pH errors.